Related Experiment Video
Updated: Aug 5, 2026
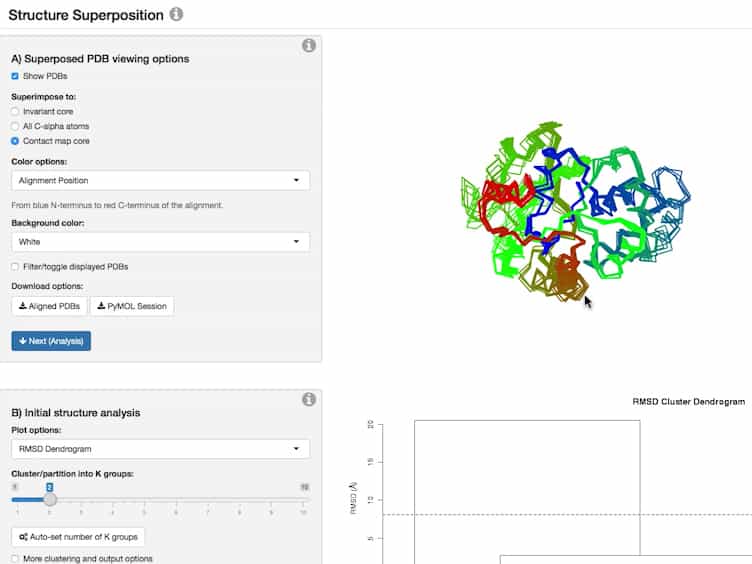

Investigating Protein Sequence-structure-dynamics Relationships with Bio3D-web
Published on: July 16, 2017
Structural heterogeneity defines the allosteric landscape and conformational selection mechanism of the PDZ2 domain
Sreyan Bhowmick1, Krishnendu Sinha1, Suman Chakrabarty1
1Department of Chemical and Biological Sciences, S. N. Bose National Centre for Basic Sciences, Kolkata 700106, India. sumanc@bose.res.in.

Abstract:
The allosteric regulation of PDZ domains is central to cellular signaling, yet whether it is governed primarily by subtle dynamical fluctuations or by structurally distinct conformational states remains unresolved. Here, we address this question for the PDZ2 domain using extensive microsecond-scale molecular dynamics simulations, Markov state models, and complementary state-space clustering analysis. The resulting ensembles reveal substantial conformational heterogeneity in both apo and ligand-bound states, with 14 metastable states identified for the apo ensemble and 9 for the bound ensemble. Quantitative comparison shows that the ligand-bound ensemble occupies a subset of the broader apo conformational landscape, whereas several states are unique to the unbound form, providing strong evidence for conformational selection. Difference contact network and flexibility analyses further show that ligand binding stabilizes the peptide-recognition region while redistributing residue interactions and distal motions across the domain. Together, these findings demonstrate that PDZ2 allostery arises from the coupling between pre-existing structural heterogeneity and ligand-induced network reorganization, providing a mechanistic framework for understanding PDZ regulation and guiding allosteric modulator design.
Related Concept Videos
 01:58
Cooperative Allosteric Transitions
01:58Cooperative Allosteric Transitions
01:58Cooperative Allosteric Transitions
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
01:49Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...