Related Experiment Video
Updated: Aug 5, 2026

Using Phylogenetic Analysis to Investigate Eukaryotic Gene Origin
Published on: August 14, 2018
High-precision binary trait association on phylogenetic trees
Ishaq O Balogun1,2, Christopher P Mancuso1,2, Tami D Lieberman1,2
1Institute for Medical Engineering and Sciences, Massachusetts Institute of Technology, Cambridge, MA 02142, USA.

Abstract:
Traditional methods for identifying associations between genomic features and traits, or between pairs of genomic traits, struggle when applied to bacterial genomes. While several microbial genome-wide association study (mGWAS) methods have been developed to account for the fact that genome-wide linkage in bacteria creates strong evolutionary-induced associations, these methods have high false discovery rates or lack statistical power, have poor performance on negative interactions and face computational limits at the scale required for pangenome-wide study of gene-gene interactions. Here, we present Simulation-based Phylogenetic iNteraction Inference (SimPhyNI), a computationally optimized framework for efficient and rigorous mGWAS studies. SimPhyNI builds null co-occurrence distributions by independently simulating traits using phylogenetically informed parameters, novelly including time to first event. The constrained variation in these simulations, combined with log odds ratio scoring for comparing across traits, robustly identifies both positive and negative associations. Using synthetic datasets mimicking both gene-gene and gene-trait associations, we demonstrate that SimPhyNI achieves high precision and recall for both positive and negative interactions. We demonstrate SimPhyNI's utility by detecting interactions between phage defence systems in Escherichia coli and gene-gene interactions across the entire E. coli pangenome (>9 million tests). Though developed here for binary traits, SimPhyNI's design supports extension to multi-state and continuous traits using generalized models of stochastic simulation. SimPhyNI's performance and scalability enable genome-wide discovery of genetic interactions that drive microbial function, ecology and disease.
Related Concept Videos
 02:54
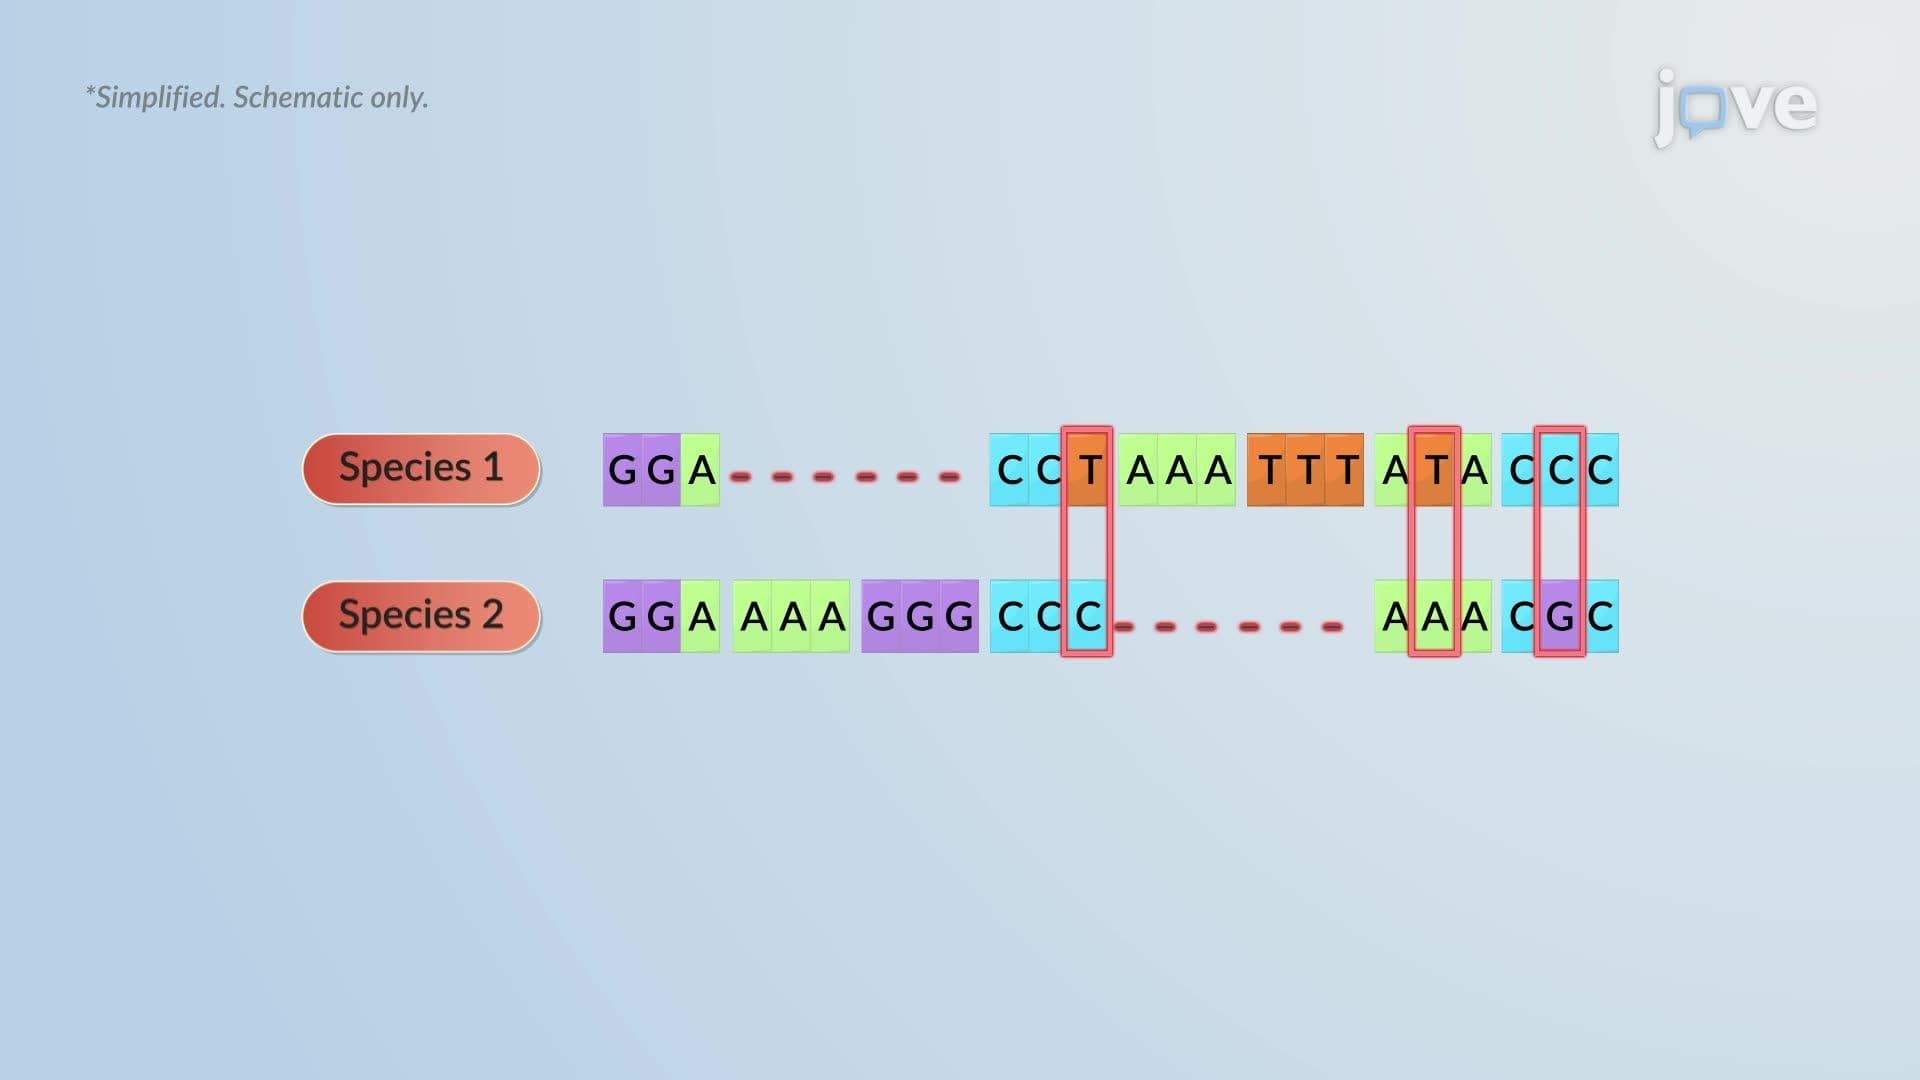
Evolutionary Relationships through Genome Comparisons
 01:28
Microbial Phylogeny
 03:21
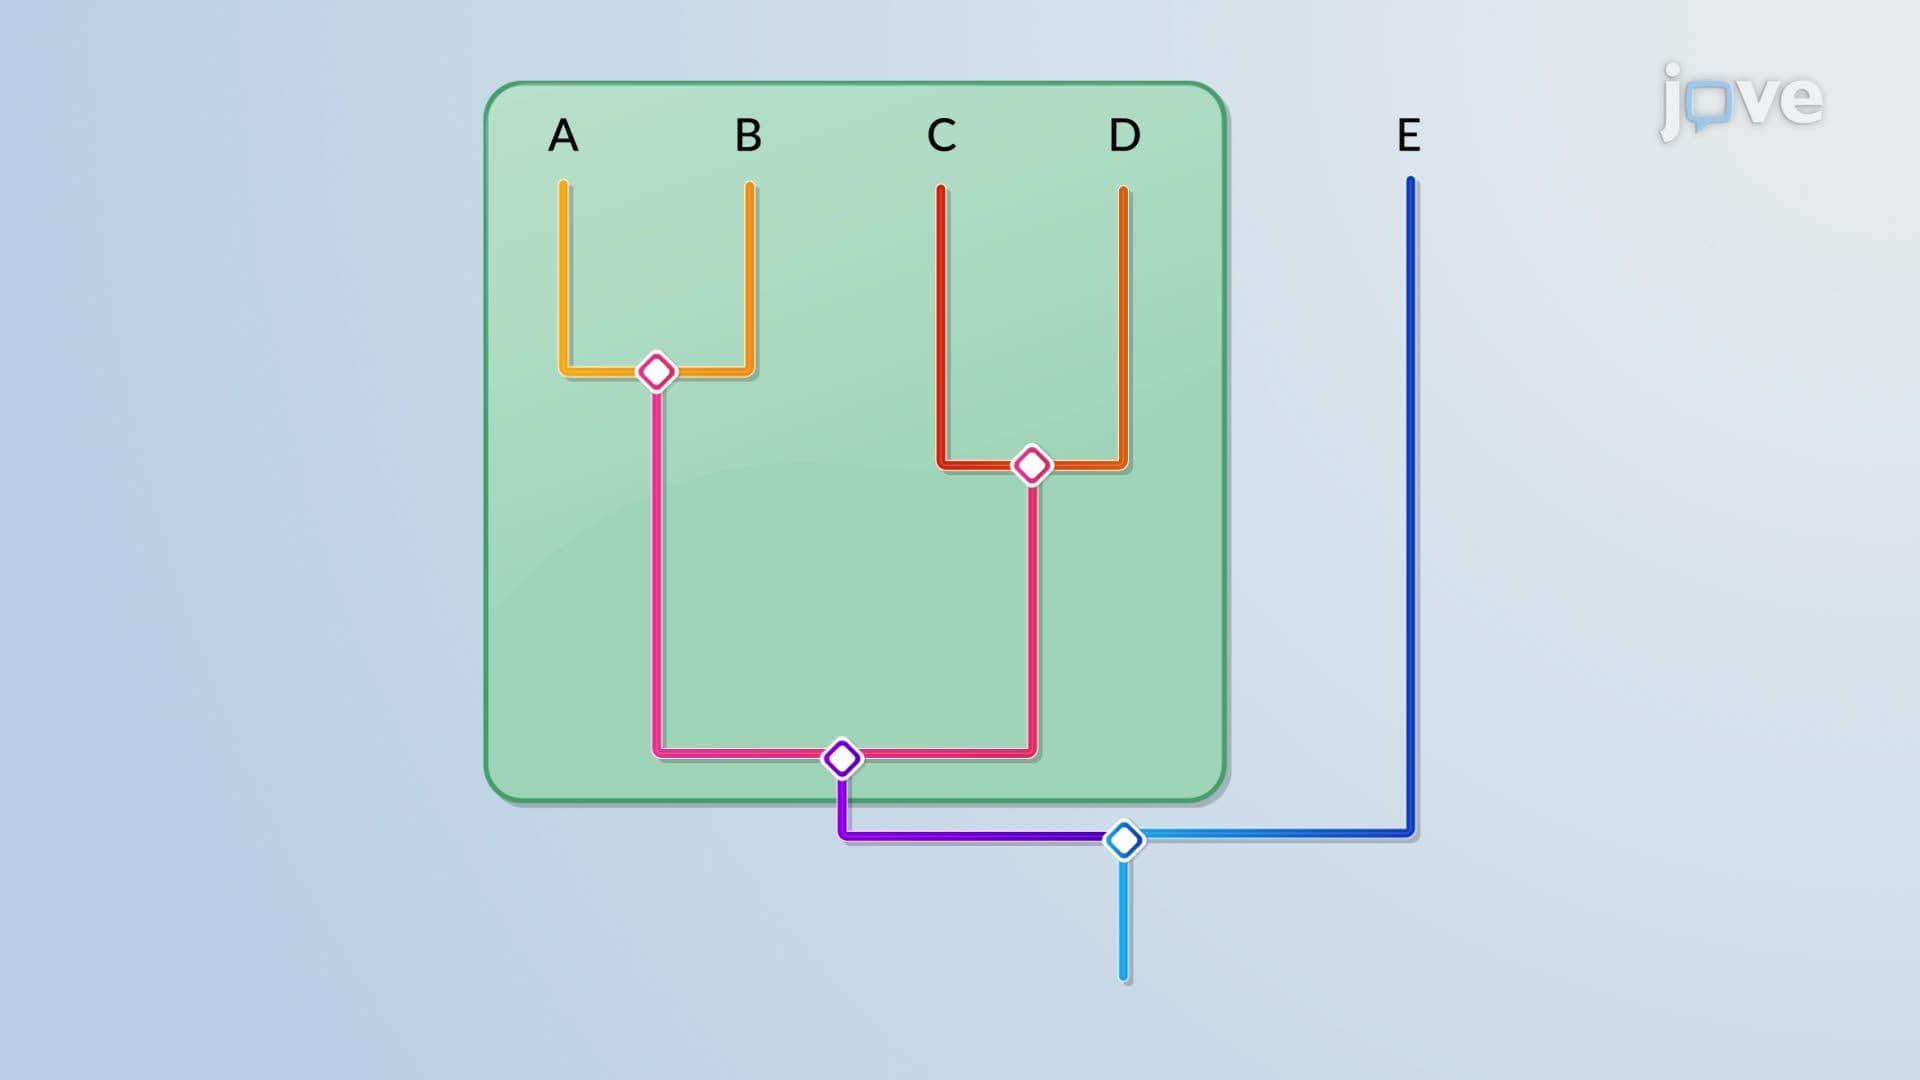
Phylogenetic Trees
03:21Phylogenetic Trees
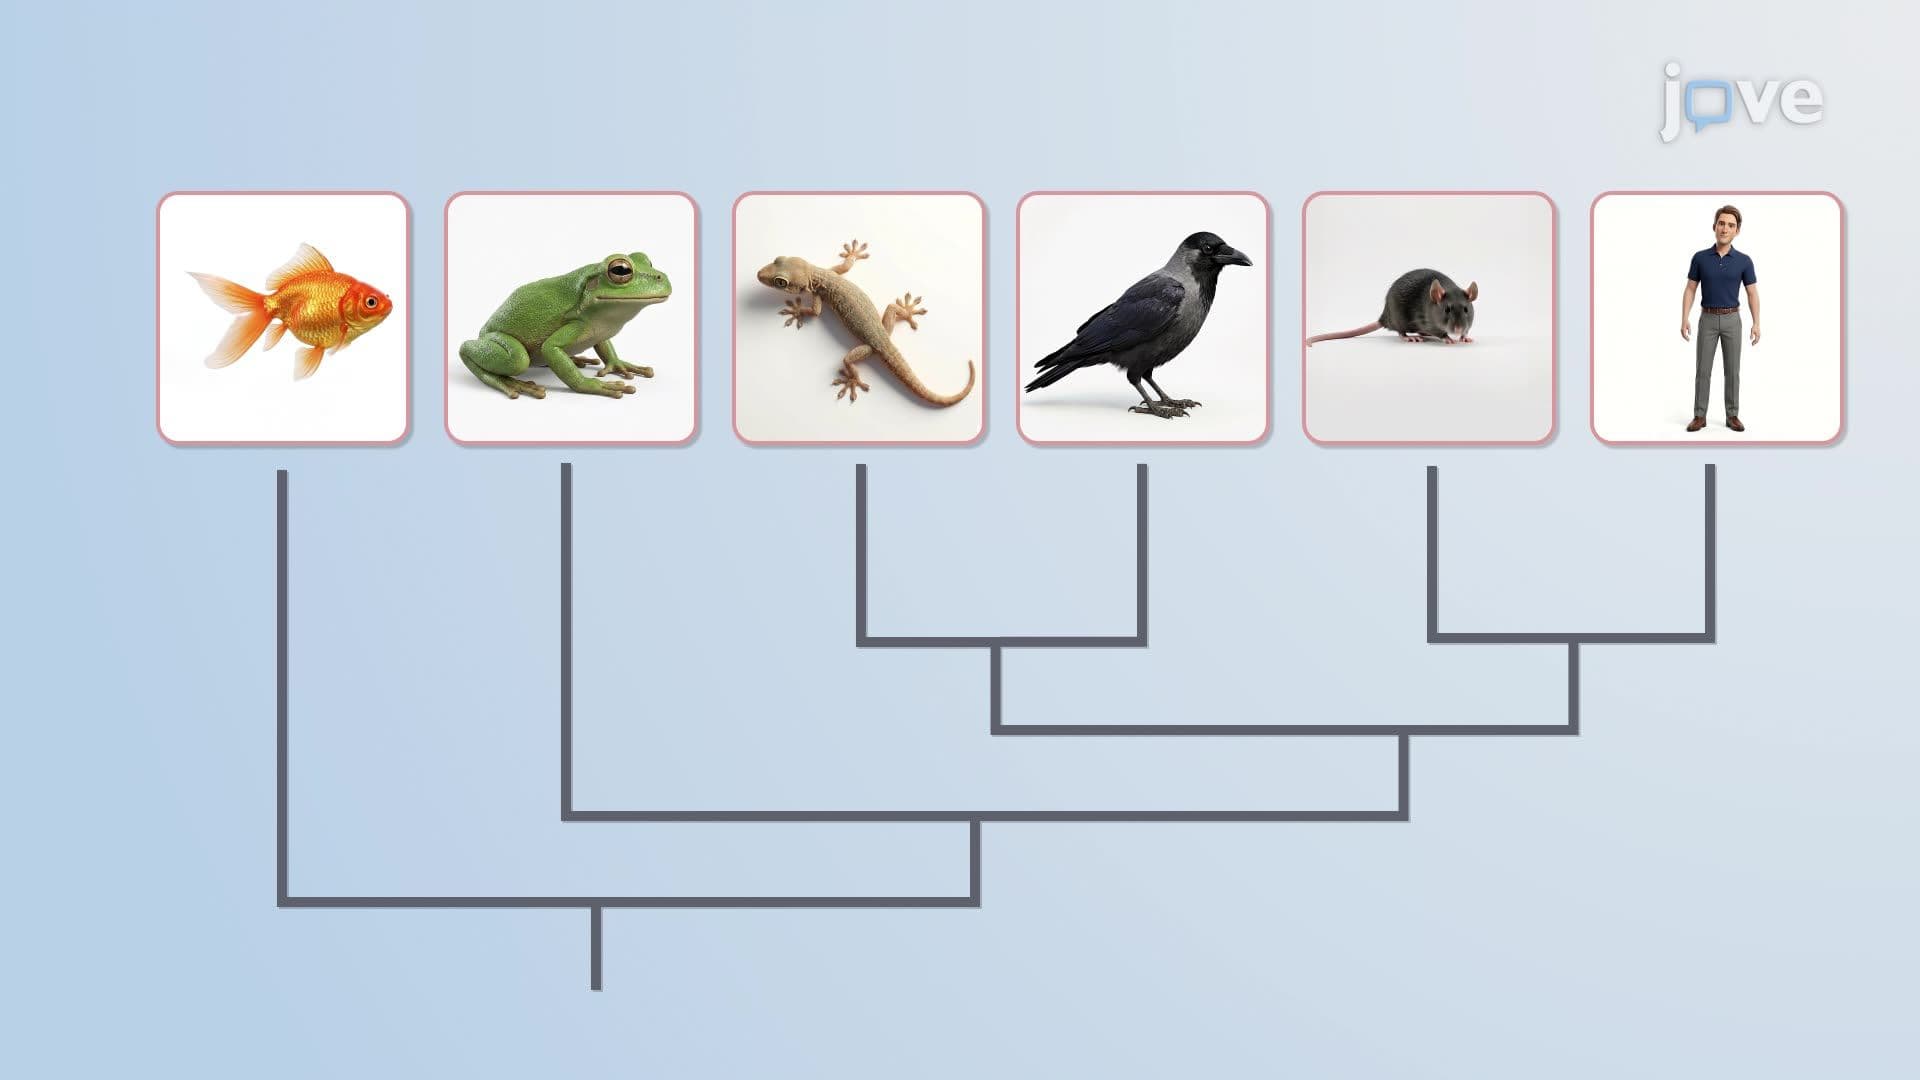 01:23
Phylogeny
 02:05
Gene Evolution - Fast or Slow?
In contrast, regions which code...