Related Experiment Video
Updated: Aug 8, 2026

Rapid in-silico Battery Electrolyte Electrochemical Reaction Generation using 3T-VASP Multi-Scale Energy Minimization
Published on: August 22, 2025
Δ-learning for transferable machine learning interatomic potentials
Nguyen Thien Phuc Tu1, Christopher N Rowley1
1Department of Chemistry, Carleton University, Ottawa, Ontario K1S 5B6, Canada.

Abstract:
Machine-learning interatomic potentials (MLIPs) trained by directly learning the total interatomic interaction energies can suffer from limited transferability, unphysical behavior beyond a finite cutoff, and large errors for out-of-distribution geometries such as transition states and uncommon conformers. We evaluate Δ-learning (delta-learning) as a remedy by training an ANI-style high-dimensional neural network (HDNNP) potential as a correction to predict PBE0/aug-cc-pVTZ energies from a third-order tight-binding density functional theory (DFTB3) baseline model. On a held-out test set derived from the modified ANI-1x training set, the Δ-learning model (named ANIDFTB-Δ) achieves a mean absolute error (MAE) of 0.82 kcal/mol, while the HDNNP model (ANIPBE0-Direct) has an MAE of 2.01 kcal/mol. On selected GMTKN55 benchmarks, the Δ model systematically improves relative energies for conformers and tautomers and avoids catastrophic outliers on challenging structures. The good physical description of DFTB3 significantly reduces the error in proton-transfer transition states in the PX13 benchmark and intermolecular interactions in the DES370K dataset in comparison with reference target PBE0. The long-range electrostatics in DFTB3 also partially correct the long-range behavior of local descriptor MLIPs outside of their predetermined cutoff, reducing the MAE from 0.098 to 0.019 kcal/mol. The computational cost of this method is incrementally more expensive than DFTB3 alone, making it practical for extensive simulations of moderately sized systems, although the DFTB3 step makes the Δ model much more costly than a simple HDNNP alone.

Related Concept Videos
 01:26
Thermodynamic Potentials
 02:27
Predicting Molecular Geometry
 01:10
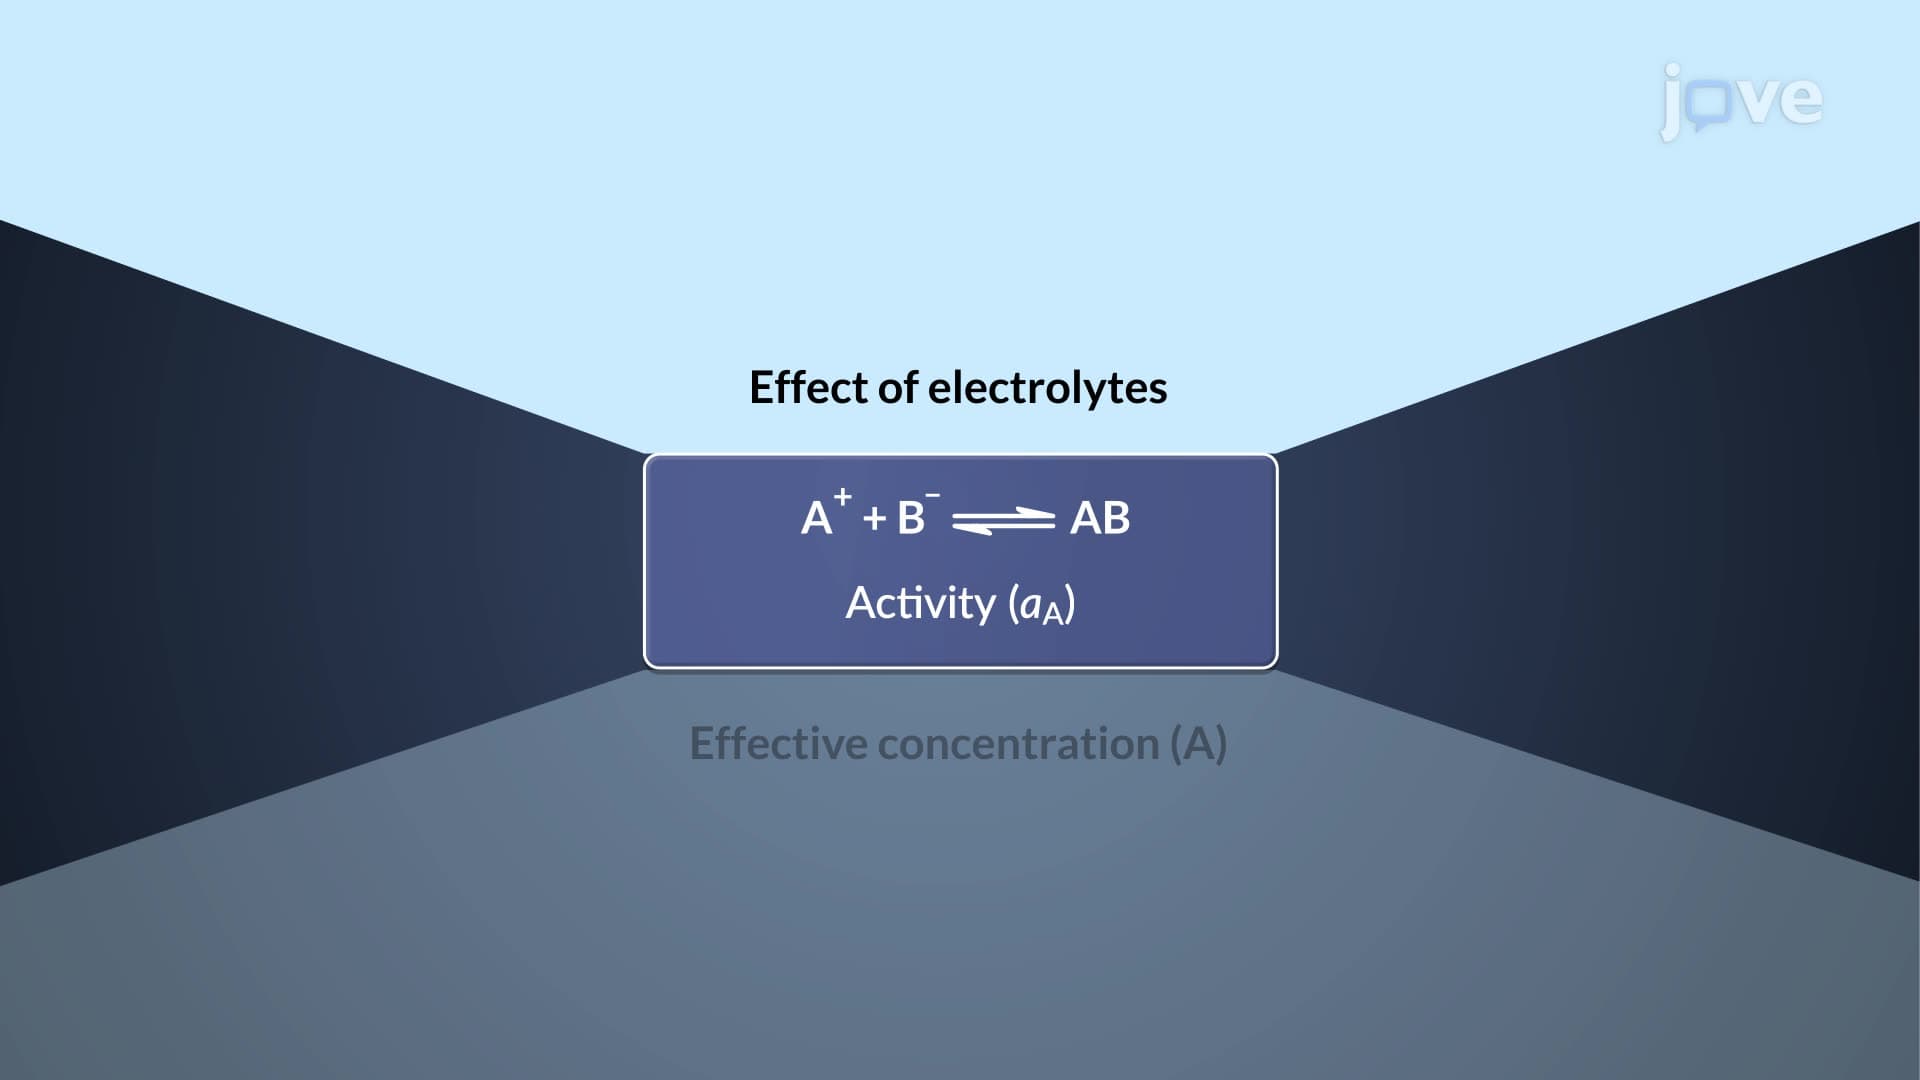
Thermodynamics: Chemical Potential and Activity
The thermodynamic equilibrium constant is more accurately defined in terms of activity rather than concentration.
 03:25
Free Energy Changes for Nonstandard States
 03:24
Hybridization of Atomic Orbitals I
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...