Related Experiment Video
Updated: Aug 10, 2026
High-resolution Respirometry to Measure Mitochondrial Function of Intact Beta Cells in the Presence of Natural Compounds
Published on: January 23, 2018
An epigenetic switch in β-cells links mitochondrial stress to inflammatory fate via HMGB1 acetylation
Yao Cheng1, Luyao Wang2, Xue Bai3
1Department of Hepatobiliary Surgery, College of Basic Medicine, The Second Affiliated Hospital of Chongqing Medical University, Chongqing Medical University, Chongqing, China.

Abstract:
Pancreatic β-cell failure in diabetes is driven by chronic inflammation, yet how metabolic stress determines pro-inflammatory cell fate remains unclear. Here, we report that sublethal oxidative stress activates a β-cell-enriched epigenetic switch that licenses intrinsic inflammation. We identify a β-cell-enriched vulnerability wherein oxidative stress disrupts mitochondrial NAD⁺/acetyl-CoA flux, skewing the nuclear equilibrium between the deacetylase SIRT1 and acetyltransferases p300/CBP. This metabolic-epigenetic imbalance induces hyperacetylation of the alarmin HMGB1 at K96/K128-a modification remarkably prominent in β-cells compared to macrophages or hepatocytes. This site-specific acetylation acts as a molecular gate for HMGB1 nucleocytoplasmic translocation, triggering TLR/RAGE-mediated inflammation. Simultaneously, we discover that oxidative stress co-opts the mechanosensitive Hippo pathway, which sequesters YAP to transcriptionally repress SIRT1, thereby forming a functionally integrated signalling axis that exacerbates HMGB1 acetylation. Therapeutically, reconstruction of mitochondrial retrograde signalling via NAD⁺ supplementation (e.g., NMN) or dual inhibition of mitochondrial ROS and Hippo signalling restored acetylation homeostasis and suppressed HMGB1-driven inflammation. Notably, this combinatorial targeting demonstrates greater efficacy than either intervention alone (~73% reduction in inflammatory markers) in mitigating β-cell failure across murine and porcine models. Further validation in non-human primates was specifically implemented to address the unique translational gap of rodent models and available human single-cell datasets: by leveraging human-like islet anatomy and systemic physiological microenvironment, we verified the pharmacodynamic robustness and in vivo feasibility of this strategy in a clinically recapitulative large-animal setting, rather than merely confirming cross-species molecular concordance. Our work unveils a stress-responsive signalling network in which metabolic and mechanical cues are integrated at the epigenetic level to control the inflammatory fate of β-cells, providing a new mechanistic framework for diabetic pathogenesis and a rationale for combinatorial therapeutic intervention. Oxidative stress triggers mitochondrial dysfunction in pancreatic β-cells, depleting NAD⁺ and accumulating acetyl-CoA. This metabolic crisis skews the SIRT1/p300 balance, inducing β-cell-enriched hyperacetylation of HMGB1 at K96/K128-a molecular switch for its nucleocytoplasmic translocation, which activates TLR/RAGE-mediated intrinsic inflammation. Concurrent Hippo pathway activation further exacerbates this process by repressing SIRT1, forming an integrated signaling axis. To ensure reproducibility, we define a minimal validation workflow using β-TC-6 cells or primary islets under standardized sublethal stress. The central mechanism can be verified by monitoring HMGB1 K96/K128 acetylation and nucleocytoplasmic translocation, while the inflammatory cascade can be effectively blunted through modular rescue approaches, including NAD+ supplementation (NMN), mitochondrial ROS inhibition or hippo pathway inhibition.
Related Concept Videos
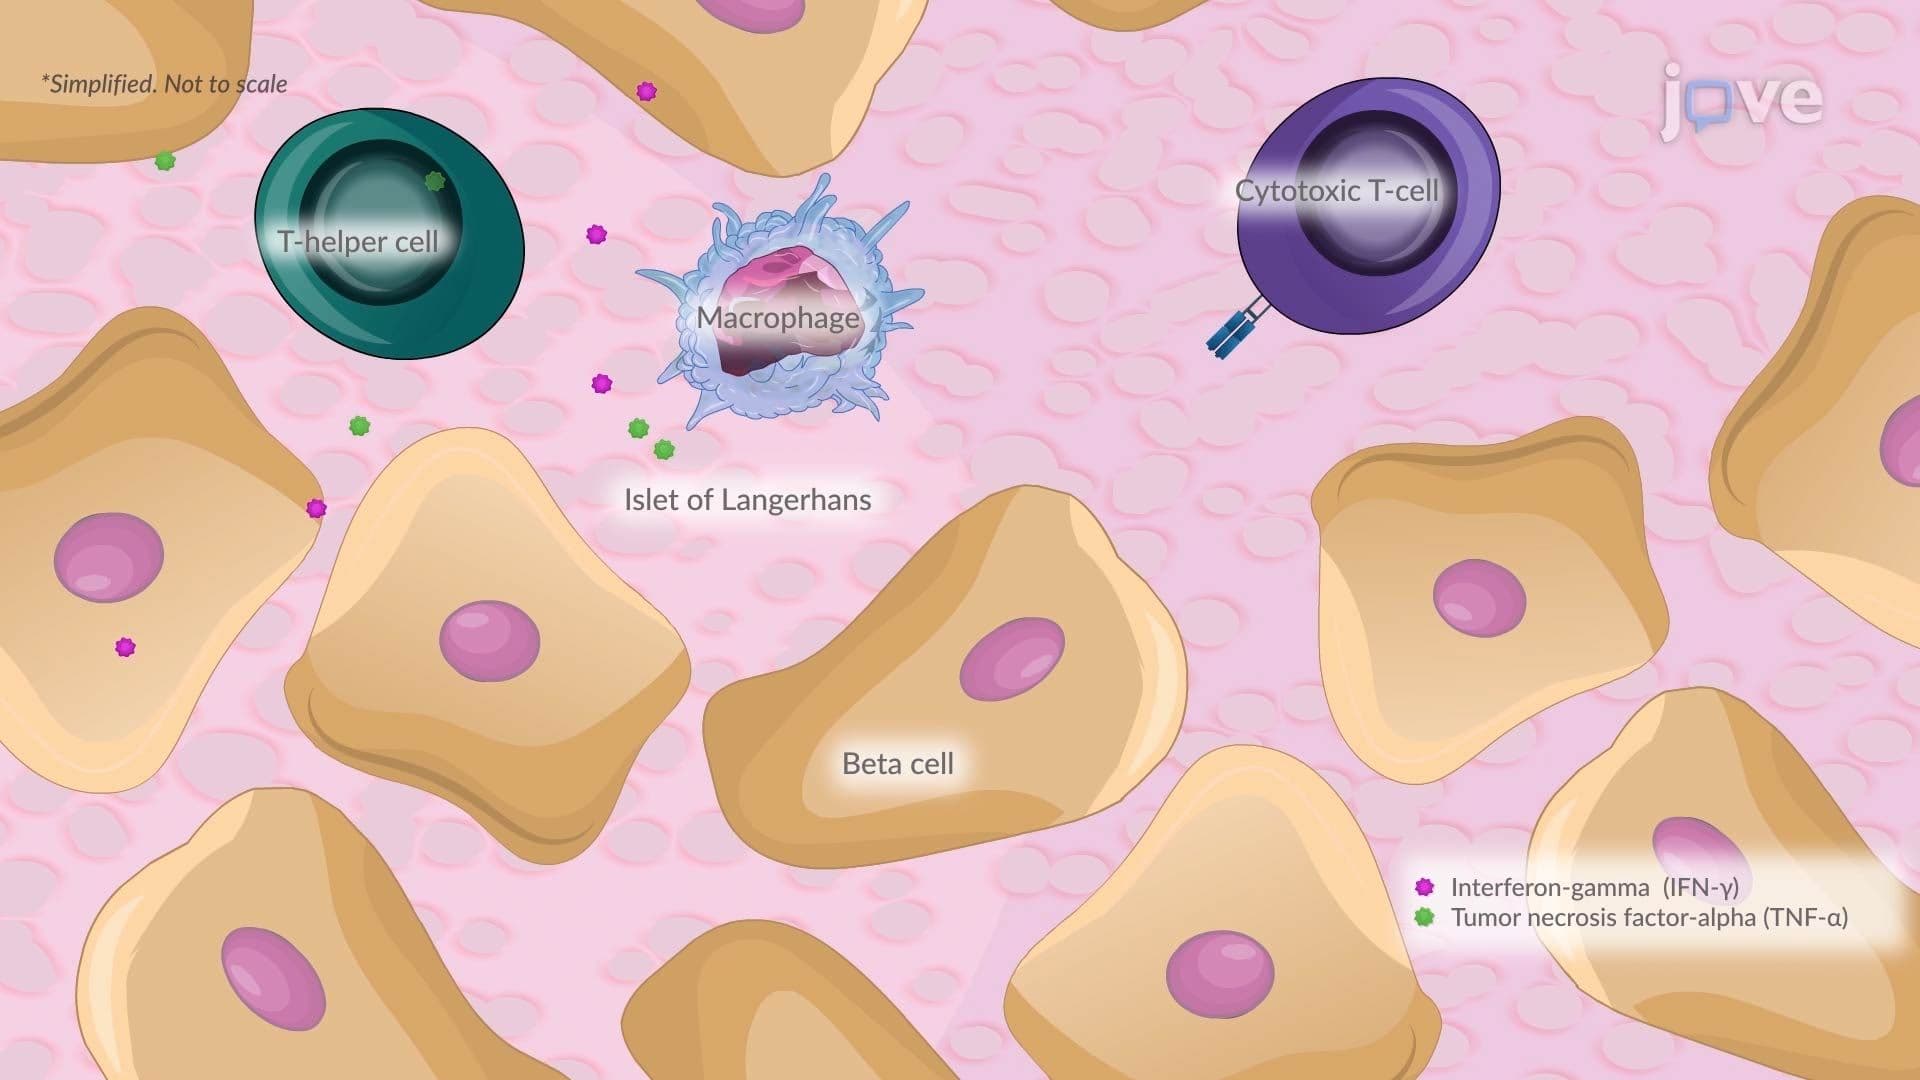 01:26
Type I Diabetes II: Pathophysiology
 01:26
Type II Diabetes I: Introduction
 01:58
Cell Specific Gene Expression
 02:25
Spreading of Chromatin Modifications
Writers
The writer is an enzyme that can...
 01:24
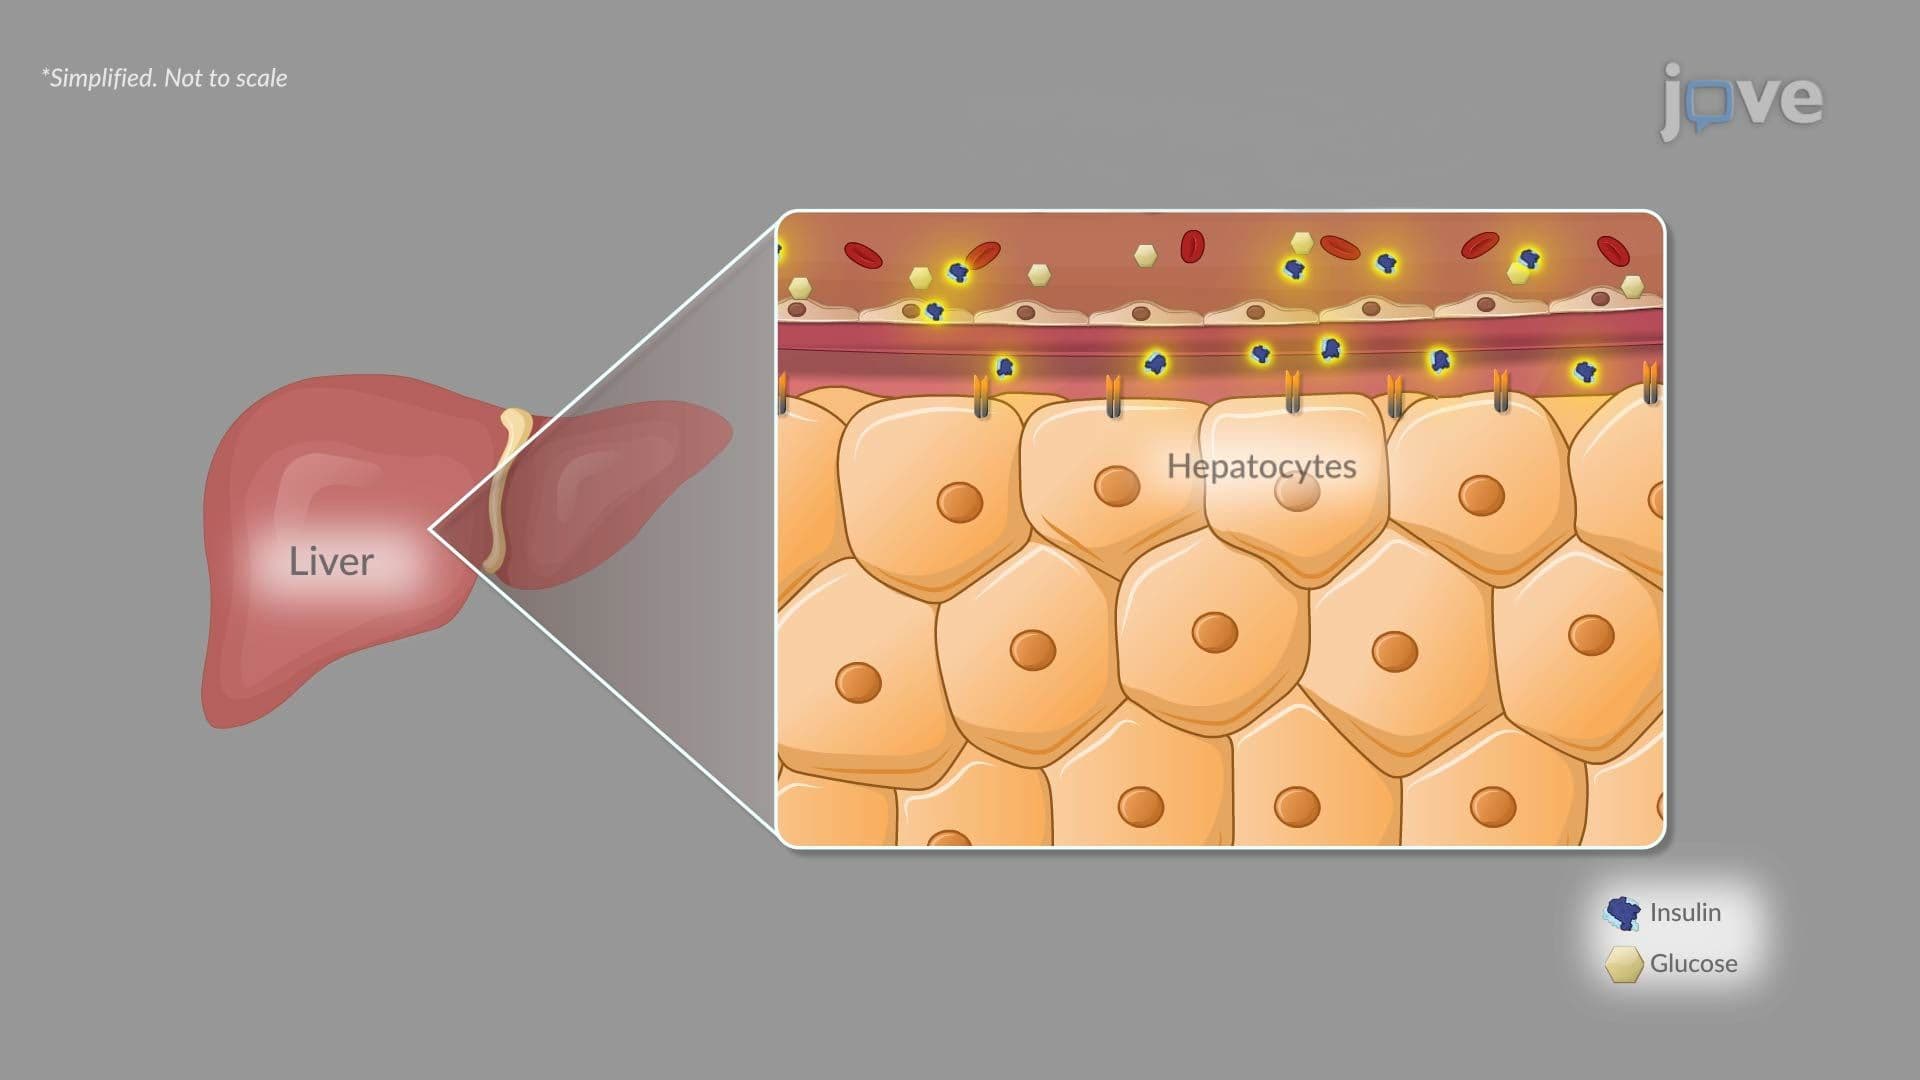
Type II Diabetes II: Pathophysiology
 01:19
Psychoneuroimmunology: Diabetes and Cancer