Related Experiment Video
Updated: Aug 13, 2026
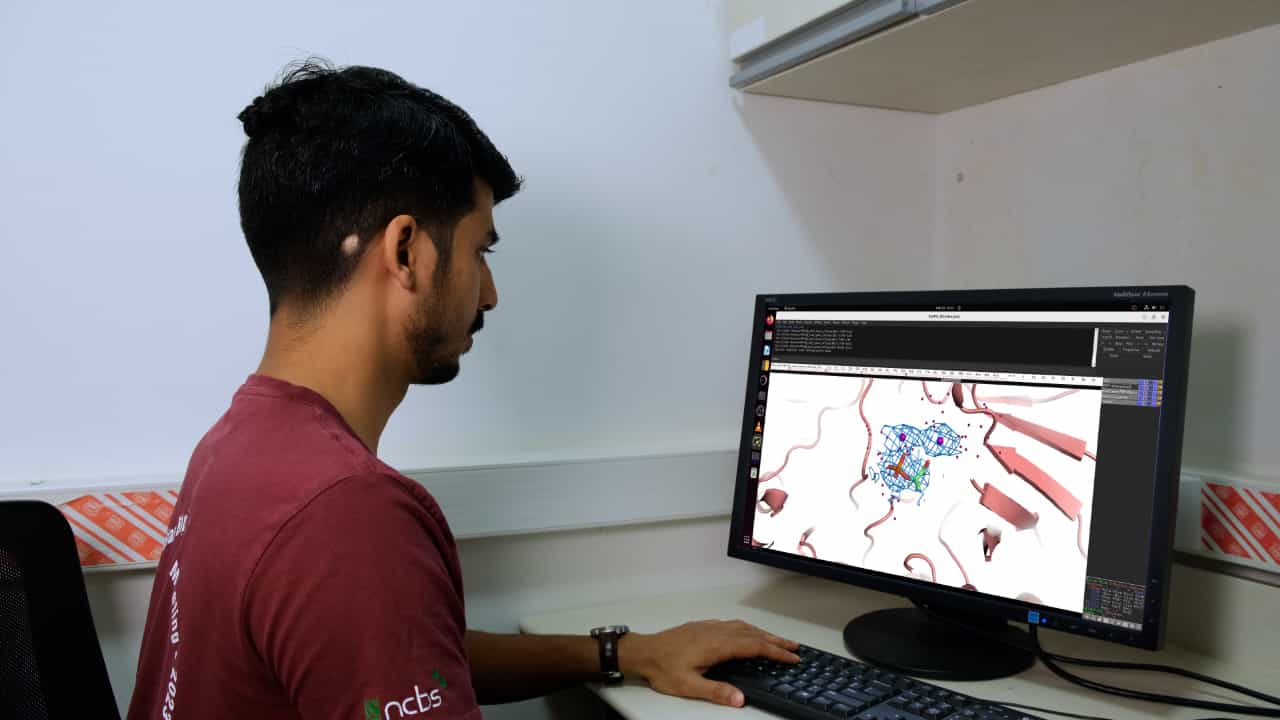

Modeling Ligands into Maps Derived from Electron Cryomicroscopy
Published on: July 19, 2024
Fused framework for predicting binding affinity to ɑ-synuclein using three-dimensional molecular graphs and attention
Xi Wang1, Wenhui Shang1, Fengze Ma1
1School of Science, Dalian Maritime University, Dalian 116026, China.

Abstract:
The aberrant aggregation of α‑synuclein (α‑syn) is a pathological hallmark of synucleinopathies, driving the need for small‑molecule inhibitors that block its oligomerization and fibrillization. In computer‑aided drug discovery, accurate affinity prediction remains critical for early‑stage screening. However, current deep learning models predominantly rely on 1D sequences or 2D topological graphs, failing to exploit stereochemical and geometric cues from 3D space. Moreover, dedicated high‑precision predictors for intrinsically disordered targets such as α‑syn are scarce. To bridge this gap, we compiled a dataset of 9628 α‑syn‑binding compounds from ChEMBL and developed a deep learning framework that integrates Graph Attention Networks (GAT) with Graph Convolutional Networks (GCN). The core innovation lies in using 3D Euclidean interatomic distances as edge features to capture spatial interaction patterns within molecular conformations. We further incorporated an atomic attention layer and a composite loss function combining mean squared error (MSE) with Pearson correlation. The final model achieved an average MSE of 0.1751 and a Pearson correlation of 0.5703 over 10 independent runs, surpassing all baseline GNNs and a recently published α‑syn‑specific predictor. Bootstrap‑calibrated confidence intervals confirmed robustness against noise and conformational perturbations. Under a stringent cluster‑exclusion split, GAT‑GCN exhibited performance degradation comparable to other architectures while retaining the best overall metrics, demonstrating generalization to novel chemical scaffolds. This work provides an accurate, robust, and interpretable computational tool for α‑syn inhibitor screening, and offers methodological insights into 3D structure‑based molecular representation learning for drug discovery.

Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 02:18
The Equilibrium Binding Constant and Binding Strength
 00:49
Ligand Binding and Linkage
 02:26
Protein Networks
These interactions can be represented through maps depicting protein-protein interaction networks, represented as nodes and edges. Nodes are circles that are representative of a protein,...