Related Experiment Video
Updated: Aug 14, 2026

Line Shape Analysis of Dynamic NMR Spectra for Characterizing Coordination Sphere Rearrangements at a Chiral Rhenium Polyhydride Complex
Published on: July 27, 2022
The Role of Terminal Heavy Chalcogenide Ligands in Enabling Hydride Transfer Reactivity of W═E (E = S, Se) Models for
Ananthu V Modappilappally1, Neal P Mankad1
1Department of Chemistry, University of Illinois Chicago, Chicago60607, Illinois, United States.

Abstract:
Whereas molybdenum-dependent hydroxylases in the xanthine oxidase family catalyze site-selective C-H hydroxylation reactions using terminal molybdenum sulfide (Mo═S) active sites, certain variants such as nicotinate dehydrogenase (NDH) replace the sulfide ligand by a selenide (Mo═Se) to enable activation of less reactive C-H bonds. To probe the origins of this dependence on terminal chalcogenide identity, we prepared and crystallographically characterized a synthetic model complex of the NDH active site, [(bdt)W(O)2Se]2-, a dioxotungsten(VI) selenide complex supported by benzene-1,2-dithiolate (bdt), for comparison to the previously reported [(bdt)W(O)3]2- and [(bdt)W(O)2S]2- analogues. Electronic structure analyses combining experimental UV-vis spectroscopy and DFT calculations indicate that replacement of terminal oxo ligands by heavier chalcogenides introduces accessible low-energy acceptor orbitals associated with more spatially distributed W═E (E = S, Se) interactions. Treatment of the terminal selenido and sulfido complexes with BH4- led to reduced tungsten products accompanied by liberation of free hydroselenide (HSe-) and hydrosulfide (HS-), respectively, whereas the corresponding trioxo analogue was unreactive under analogous conditions. These findings provide direct experimental support for the proposal that terminal heavy chalcogenide ligands play an active electronic role in enabling biological hydride-transfer reactivity in molybdenum hydroxylase enzymes.
Related Concept Videos
 02:51
Metal-Ligand Bonds
In these complexes, transition metals form coordinate covalent bonds, a kind of Lewis acid-base interaction in which both of the electrons in the bond are contributed by a donor (Lewis base) to an electron acceptor (Lewis acid). The Lewis acid in...
 01:26
Radical Substitution: Hydrogenolysis of Alkyl Halides with Tributyltin Hydride
The bonds formed in this reaction are stronger than the bonds broken, making it energetically favorable. The reaction follows a radical chain mechanism similar to radical halogenation reactions,...
 01:17
Esters to Alcohols: Hydride Reductions
Lithium aluminum hydride is a source of hydride ions and functions as a nucleophile. The mechanism proceeds in three steps. Firstly, the nucleophilic hydride ion attacks the carbonyl carbon of the ester to form a tetrahedral intermediate. Subsequently, the carbonyl group re-forms,...
 01:19
Acid Halides to Alcohols: LiAlH4 Reduction
The mechanism proceeds in three steps. First, the nucleophilic hydride ion attacks the carbonyl carbon of the acid halide to form a tetrahedral intermediate. Next, the carbonyl group is re-formed, and the halide ion departs as a leaving group, generating an aldehyde. A second nucleophilic attack by the hydride yields an alkoxide ion, which, upon protonation, gives a primary alcohol as...
 02:26
Preparation and Reactions of Sulfides
 01:24
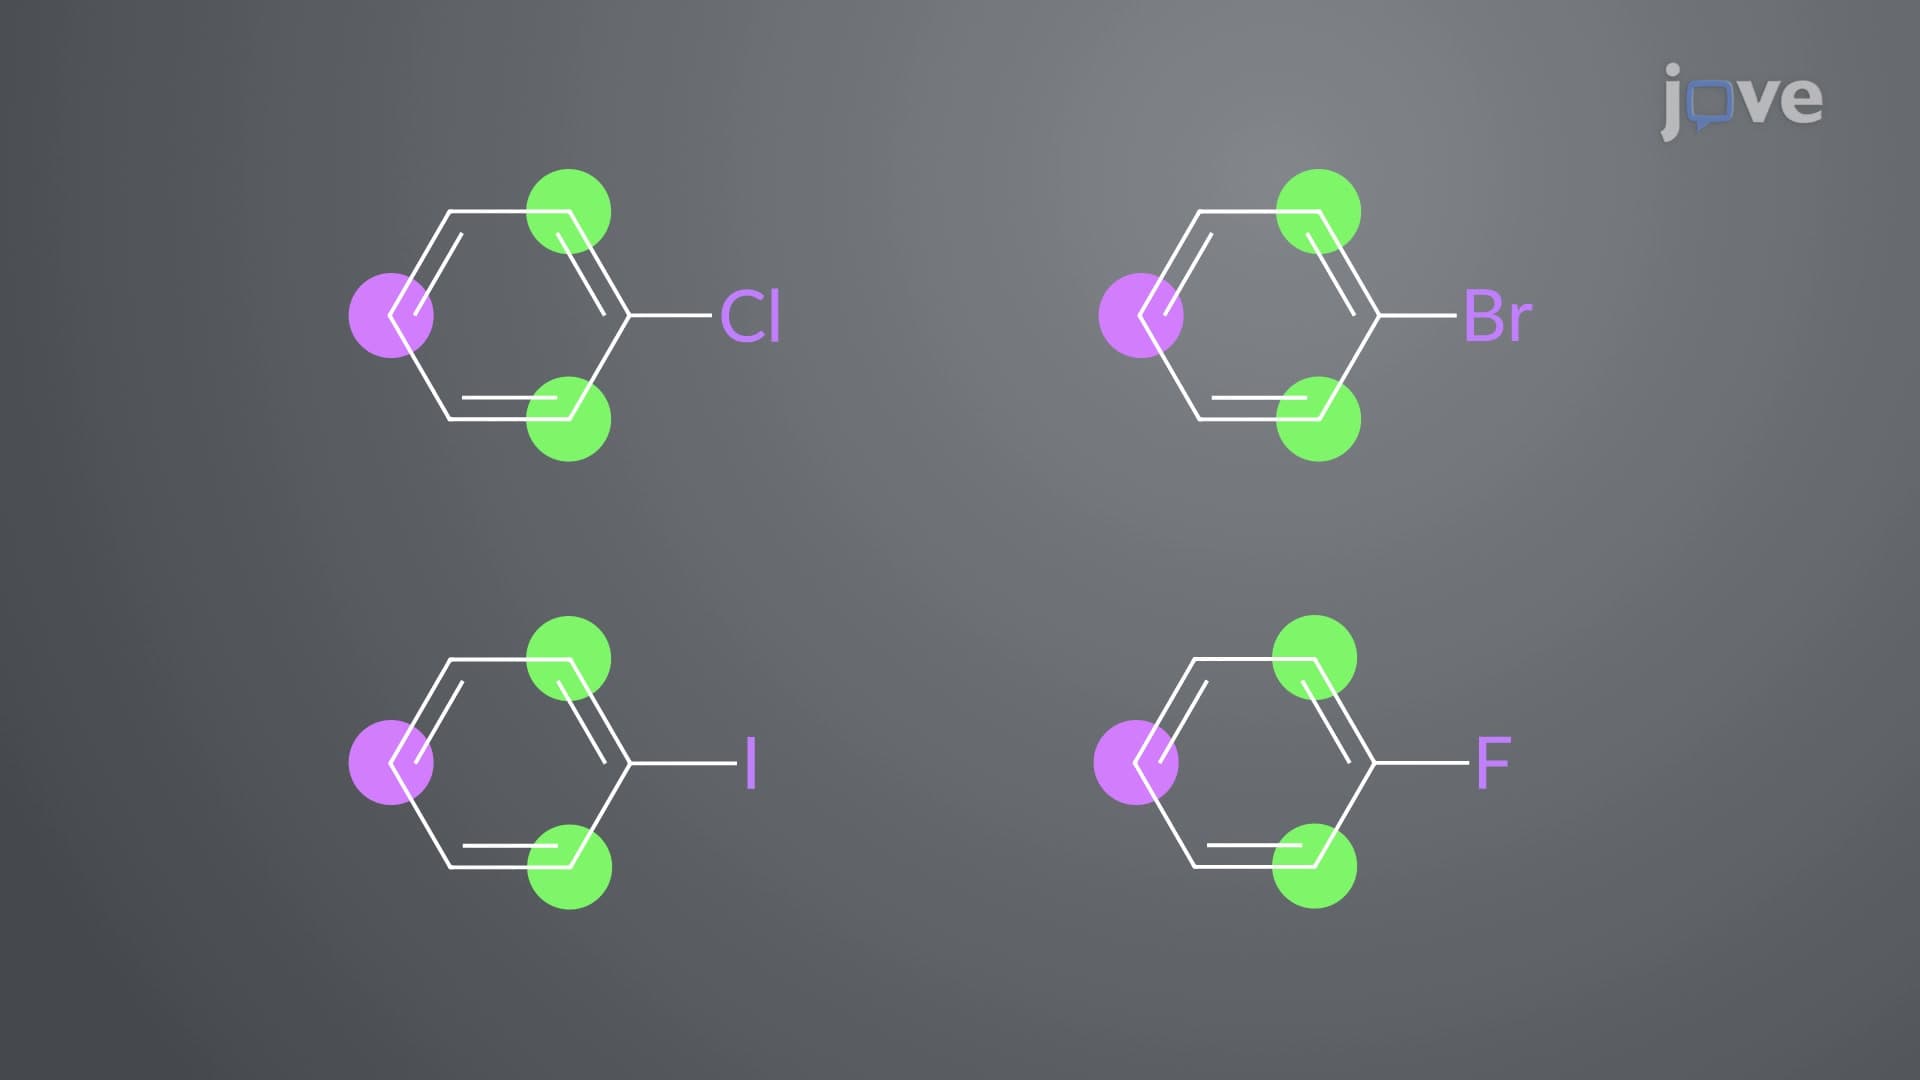
ortho–para-Directing Deactivators: Halogens