Related Experiment Video
Updated: Aug 14, 2026
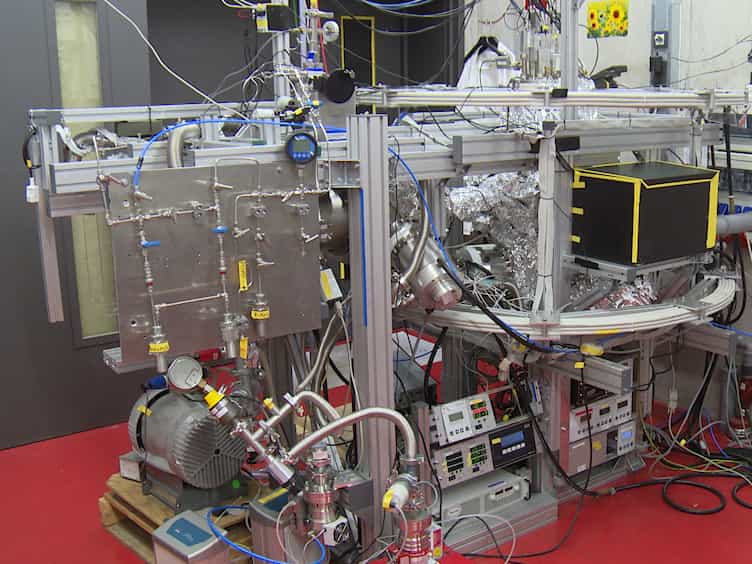

Coulomb Explosion Imaging as a Tool to Distinguish Between Stereoisomers
Published on: August 18, 2017
Error Decomposition in Dissociation Processes within Density-Corrected DFT: An Ensemble Perspective
Alon Zamir1, Nevo Levy1, Tamar Stein1
1Fritz Haber Research Center for Molecular Dynamics and Institute of Chemistry, The Hebrew University of Jerusalem, 9091401Jerusalem, Israel.

Abstract:
Density functional theory (DFT) is widely used for electronic structure calculations but can fail to describe bond dissociation correctly, even for simple systems such as H2+ and H2. These failures are commonly attributed to delocalization error in H2+ and static correlation in H2. More generally, dissociation errors are closely linked to deviations from ideal fractional charge and fractional spin ensemble behavior. Here, we investigate bond dissociation within density-corrected DFT (DC-DFT) for NaCl, H2+, He2+, and H2, using Hartree-Fock (HF) and alternative localized densities. We seek to understand why DC-DFT improves the dissociation behavior of some systems, such as NaCl, while providing little benefit in others, such as H2+. We show that for H2+, a localized density can yield substantially more accurate dissociation energetics than the exact HF density when it is used within DC-DFT. Although formally less accurate, the localized density compensates for the functional's delocalization error and recovers the correct asymptotic behavior. This result demonstrates that an approximate density can sometimes yield a more accurate energy than an exact density when used with an approximate functional. In both NaCl and H2+, localization compensates for the energetic consequences of the convex fractional-charge behavior of the functional by effectively driving the system toward an ensemble end point corresponding to an integer charge. We further show that the connection between dissociation errors and ensemble behavior is preserved within DC-DFT and that HF densities worsen the dissociation of strongly correlated systems such as H2.
More Related Videos


Related Concept Videos
 01:20
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:32
The de Broglie Wavelength
 01:04
¹³C NMR: ¹H–¹³C Decoupling
A broadband decoupling technique is used to simplify these complex, sometimes overlapping, signals. Broadband decoupling relies on a...
 01:28
Debye–Huckel–Onsager Conductance Equation