Related Experiment Video
Updated: Aug 21, 2026
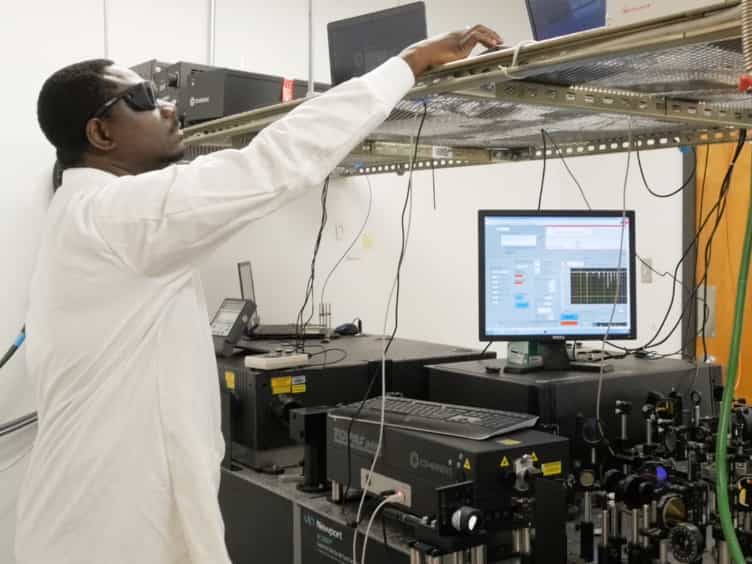

Measurement of Ultrafast Vibrational Coherences in Polyatomic Radical Cations with Strong-Field Adiabatic Ionization
Published on: August 6, 2018
Site-Specific and Mode-Specific Photodissociation of OH bonds in a Pentapeptide by Laser-Driven Tunneling through a
Kendrew Au1, Isaac Ofori2, Chin Lee1
1Gas Phase Chemical Physics, Combustion Research Facility, Sandia National Laboratories, Livermore, California94550, United States.

Abstract:
The prospect of using lasers to selectively break chemical bonds in molecules is often thwarted by the intrinsic anharmonic mixing of the vibrational levels that lead to dissociation with a dense manifold of other vibrational states that do not. Using an ultraviolet-infrared (UV-IR) double resonance excitation scheme, we demonstrate both site- and mode-selective photodissociation of the O-H bond in either of the two tyrosine chromophores (Tyr, Y) in a cryogenically cooled, gas phase protonated pentapeptide ion [YGGYL+H]+ with 231 vibrational modes. The two tyrosine's have unique UV absorptions with electronic origins separated from one another by 324 cm-1, enabling site-specific electronic excitation of the 1ππ* state of the Y1 or Y4 chromophore. Isotopic substitution of 18OH on one of the two tyrosine's phenol moiety (forming Y(18O)GGYL or YGGY(18O)L) produces a shift in the infrared absorption frequency of the OH stretch fundamental of -11 cm-1. Since the OH stretch fundamental shifts by -72 cm-1 in the 1ππ* state relative to the ground state, UV-IR spectra as a function of IR-UV time delay map out the excited state IR spectra and provides a means for promoting either OH group to OH(v = 1) in their 1ππ* excited state. Using UV-IR excitation at short IR-UV delay times, we observe site-selective and mode-selective loss of the OH hydrogen atom, which is efficiently lost only when both UV and IR excitation are on the same Tyr chromophore, with a mode-selectivity of greater than 100:1. No other combination of UV and IR excitation produces any measurable H atom loss. Since the OH stretch coordinate is the reaction coordinate for H atom loss, we identify tunneling underneath the 1ππ*/1πσOH* conical intersection (CI) as responsible for the mode-selectivity of the photofragmentation, and argue that tunneling through the CI following OH(v = 1) excitation is fast enough that it competes favorably over vibrational energy scrambling, making it possible to selectively fragment either OH bond in this large molecule with high fidelity.
Related Concept Videos
 01:26
Photochemical Electrocyclic Reactions: Stereochemistry
Selection Rules: Photochemical Activation
 01:20
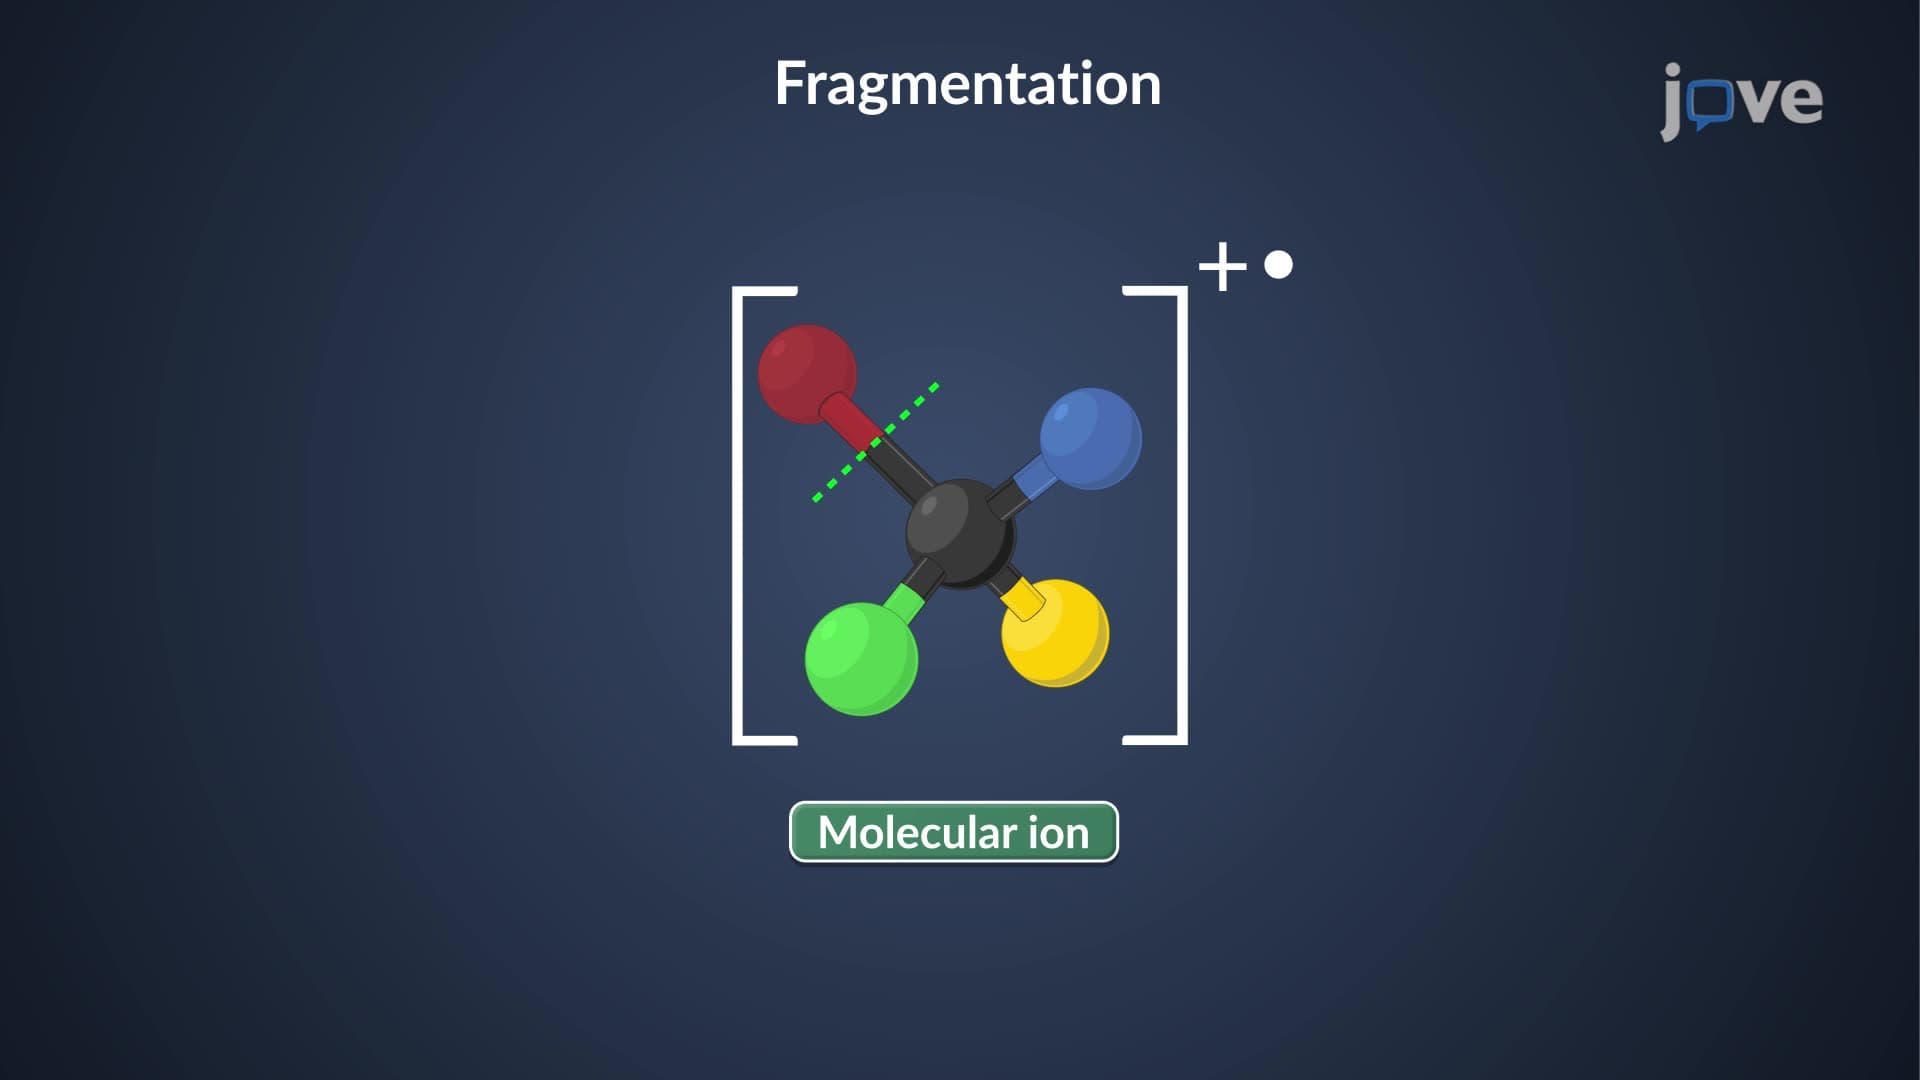
Mass Spectrometry: Molecular Fragmentation Overview
One type of fragmentation pattern is the cleavage of a single bond in the molecular ion. The cleavage leads to a radical and a cation. The cleavage can occur at...
 01:12
Double Resonance Techniques: Overview
Spin decoupling is usually achieved by...
 01:12
Cycloaddition Reactions: MO Requirements for Photochemical Activation
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions
 01:20
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)