Related Experiment Video
Updated: Aug 30, 2026
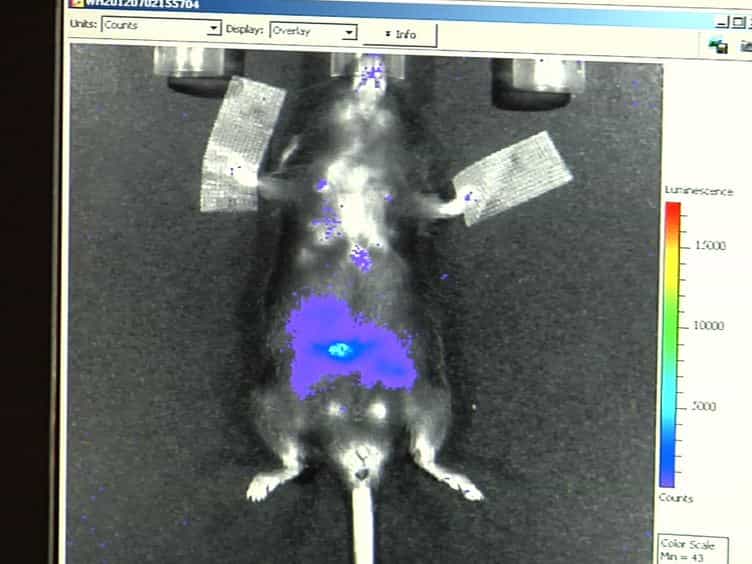

Bioluminescence Imaging of NADPH Oxidase Activity in Different Animal Models
Published on: October 22, 2012
The crosstalk between PFKFB3 and RSPO2 drives sepsis-induced lung injury by coupling metabolic rewiring to macrophage

Abstract:
Aberrantly activated macrophages drive acute lung injury (ALI), but how metabolic reprogramming fuels their dysfunction remains elusive. Here, we investigated PFKFB3, a glycolytic enzyme converting fructose-6-phosphate to fructose-2,6-bisphosphate, in ALI. PFKFB3 was upregulated both in LPS-stimulated macrophages and septic mice, cascading RSPO2 activation. Pharmacological inhibition of PFKFB3 by PFK15 not only mitigated LPS-induced macrophage injury via NF-κB suppression but also protected against ALI in CLP mice. PFK15 effectively blunted PFKFB3-dependent glycolysis and curtailed pro-inflammatory responses by inhibiting RSPO2/β-catenin signaling both in vitro and in vivo. Genetic silencing of PFKFB3 similarly reduced inflammation and suppressed RSPO2/β-catenin signaling. Notably, RSPO2 knockdown ameliorated LPS-induced macrophage dysfunction characterized by hyperglycolysis and excessive inflammation. This protective effect was phenocopied by KYA1797K, an RSPO2/β-catenin inhibitor, which reduced macrophage inflammation and glycolytic activity, thereby alleviating ALI. Collectively, our data support a model wherein a positive feedback loop interconnecting PFKFB3-driven glycolysis and RSPO2 signaling synergizes with the NF-κB pathway to potentiate macrophage inflammation, ultimately exacerbating septic ALI. These findings establish the PFKFB3-RSPO2 circuit as a promising therapeutic target for ALI.