Related Experiment Video
Updated: Sep 2, 2026

EPR Monitored Redox Titration of the Cofactors of Saccharomyces cerevisiae Nar1
Published on: November 26, 2014
Toward Relative Redox Potential Predictions in Flavoproteins: Treatment of a Charge-Changing Mutation in Alchemical
Sarah Elhajj1, Jacopo D'Ascenzi2, Massimo Olivucci2,3
1Department of Chemistry, Georgia State University, Atlanta, Georgia30302, United States.

Abstract:
Redox enzymes play an essential role in nature and in biotechnological applications such as (photo)biocatalysis and biosensing. Understanding how a protein's sequence and structure tune its redox potential is very valuable for engineering proteins with tailored (photo)redox properties. Since protein redox potential measurements are laborious, reliable redox potential computations present an attractive alternative. However, redox calculations come with their own set of challenges, such as ensuring adequate sampling and accurate force fields. Dealing with charge-changing states introduces additional challenges to theory and requires special considerations in the model setup. Here, we report a protocol for computing the change in the proton-coupled one-electron redox potential associated with a D63N charge-changing mutation in a prototypal flavoprotein, Desulfovibrio vulgaris flavodoxin. An automated average protein electrostatic configuration protocol, APEC-F 2.0, was used to construct hybrid quantum mechanics/molecular mechanics (QM/MM) models. These models were used for subsequent alchemical free energy simulations in which a charged surface aspartate residue was gradually converted to an isosteric but neutral asparagine (D63N) over 40 λ windows. A thermodynamic cycle was employed to calculate the redox potential of the mutant relative to the wild-type reference. This calculation was repeated for the same D63N mutation using models prepared under slightly different conditions, focusing primarily on factors that affect the electrostatic environment in the system. Factors tested include (1) the effect of including extra salt ions in the model solution, (2) different protocols to balance the disappearing negative charge associated with the alchemical D63N mutation, and (3) the effect of accounting for the kinetic energy terms in the free energy protocol due to alchemical morphing of the atomic masses. The results indicate that such apparently minor details may have a considerable effect on the random and systematic errors obtained from the free energy simulations. The best models were shown to reproduce the experimental shift in the redox potential due to the D63N mutation with an accuracy of 0.3 kcal/mol. The associated error for this shift is 1.3 kcal/mol, calculated as the total standard deviation of triplicate simulations for each of the oxidized and reduced neutral semiquinone states of flavin. The addition of salt ions in the simulations and the proper treatment of charge-conserving coalchemical counterions in particular are found to be paramount, since less accurate models led to errors more than double the magnitude compared to the best protocol. The sources of those errors are discussed and often found to be associated with medium-range electrostatics due to missing or inaccurate second solvation/ionic shells around the mutation site.
Related Concept Videos
 01:30
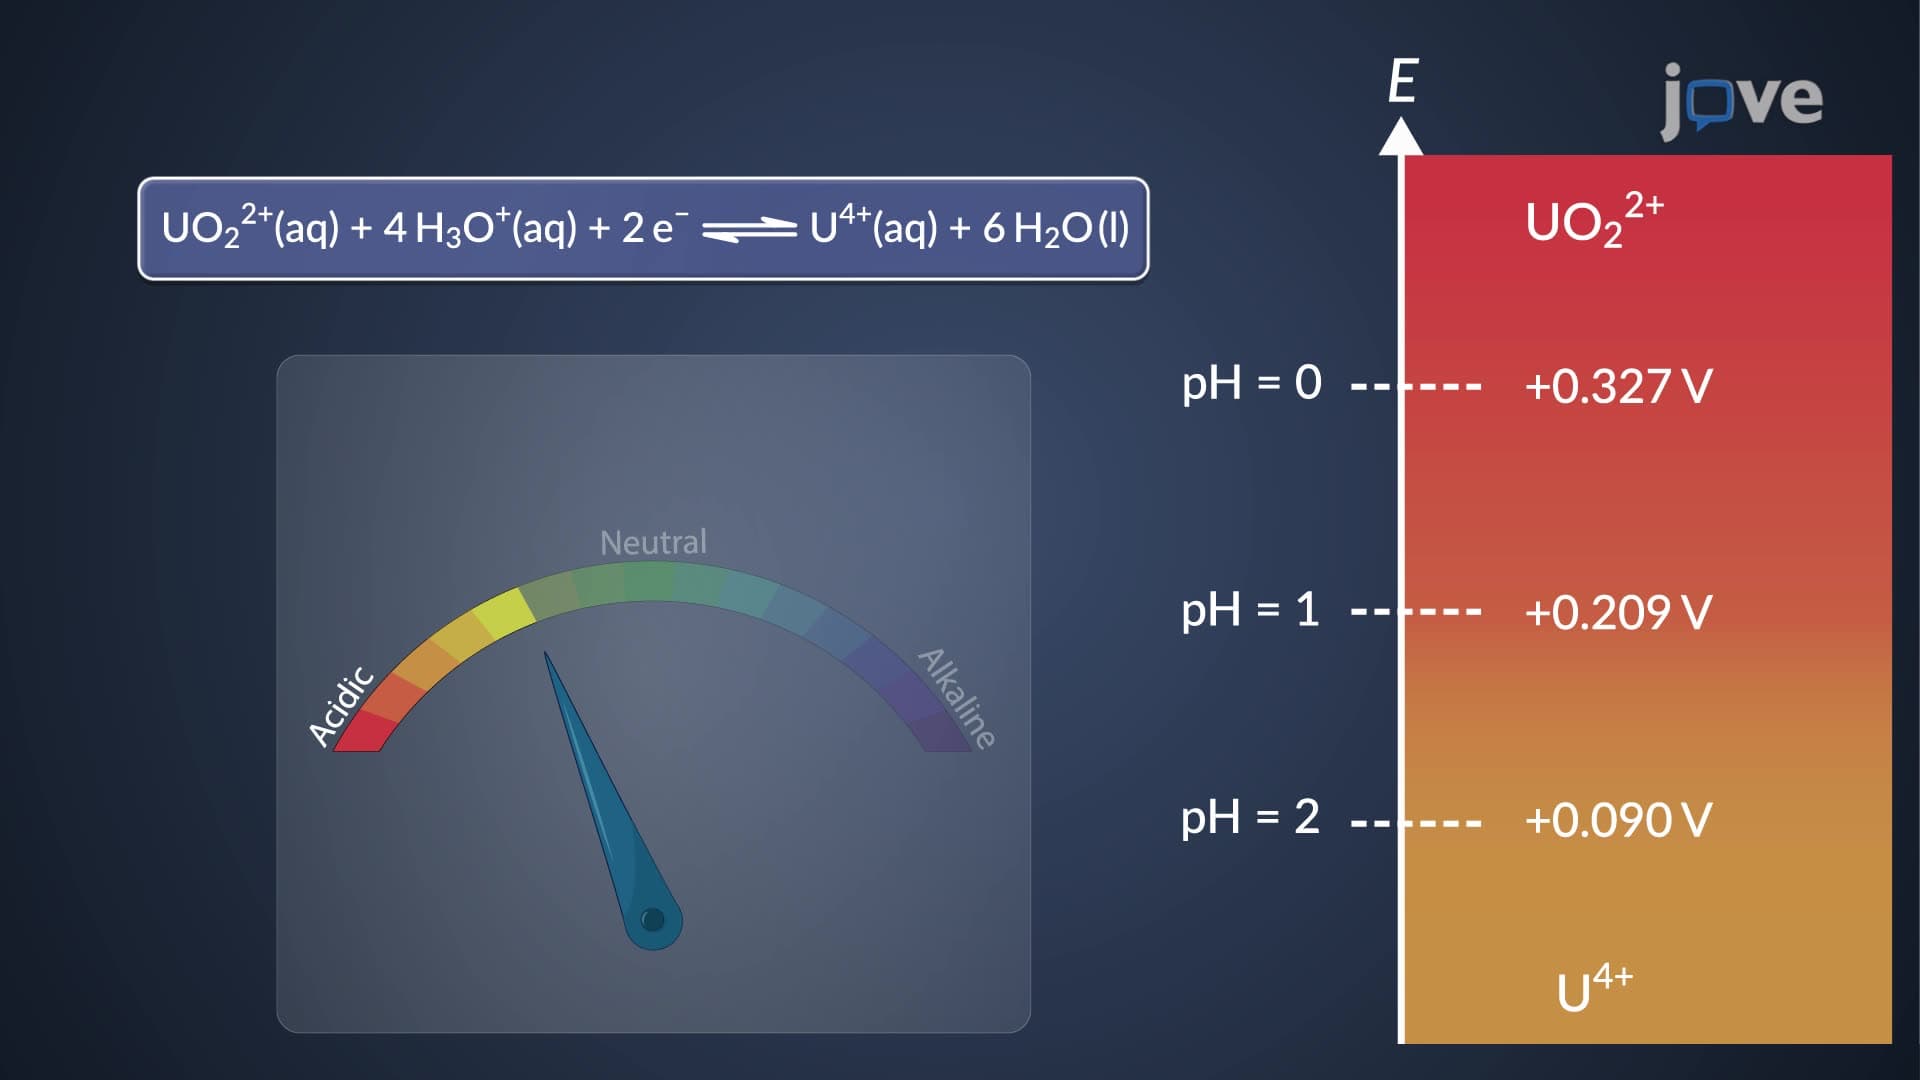
Ladder Diagrams: Redox Equilibria
Consider the Fe3+/Fe2+ half-reaction, which has a standard-state potential of +0.771 V. At potentials more positive than +0.771 V, Fe3+ predominates, whereas Fe2+...
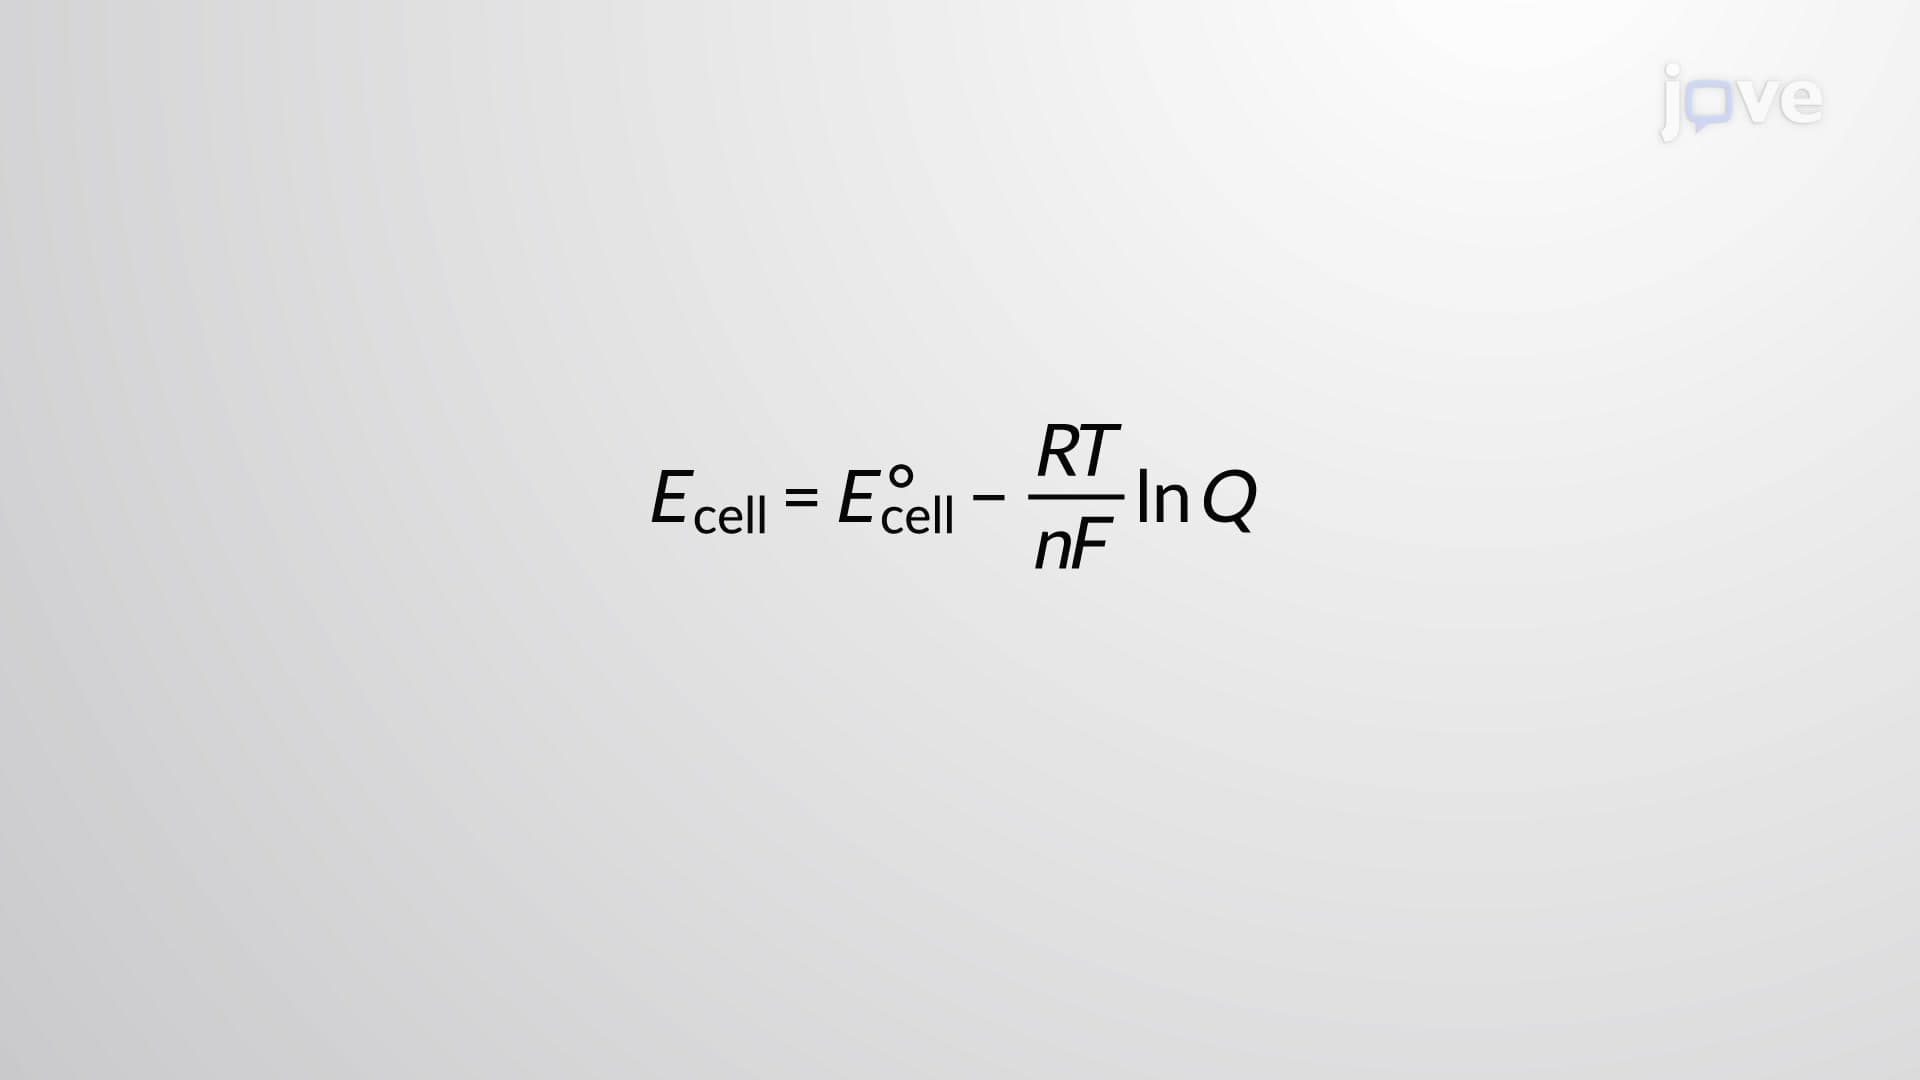 02:59
The Nernst Equation
The interconnection between standard cell potentials and various thermodynamic parameters such as the standard free energy change ΔG° and equilibrium constant K has been previously explored. For example, a redox reaction involving zinc(II) and tin(II) ions at 1 M concentration with Eºcell = +0.291 V and ΔG° = −56.2 kJ is spontaneous.
 01:27
Redox Reactions
 02:58
Balancing Redox Equations
 01:19
Oxidation and Reduction of Organic Molecules
The removal of an electron from a molecule, results in a...
 02:49
Calculating Standard Free Energy Changes