Related Experiment Video
Updated: Sep 11, 2026
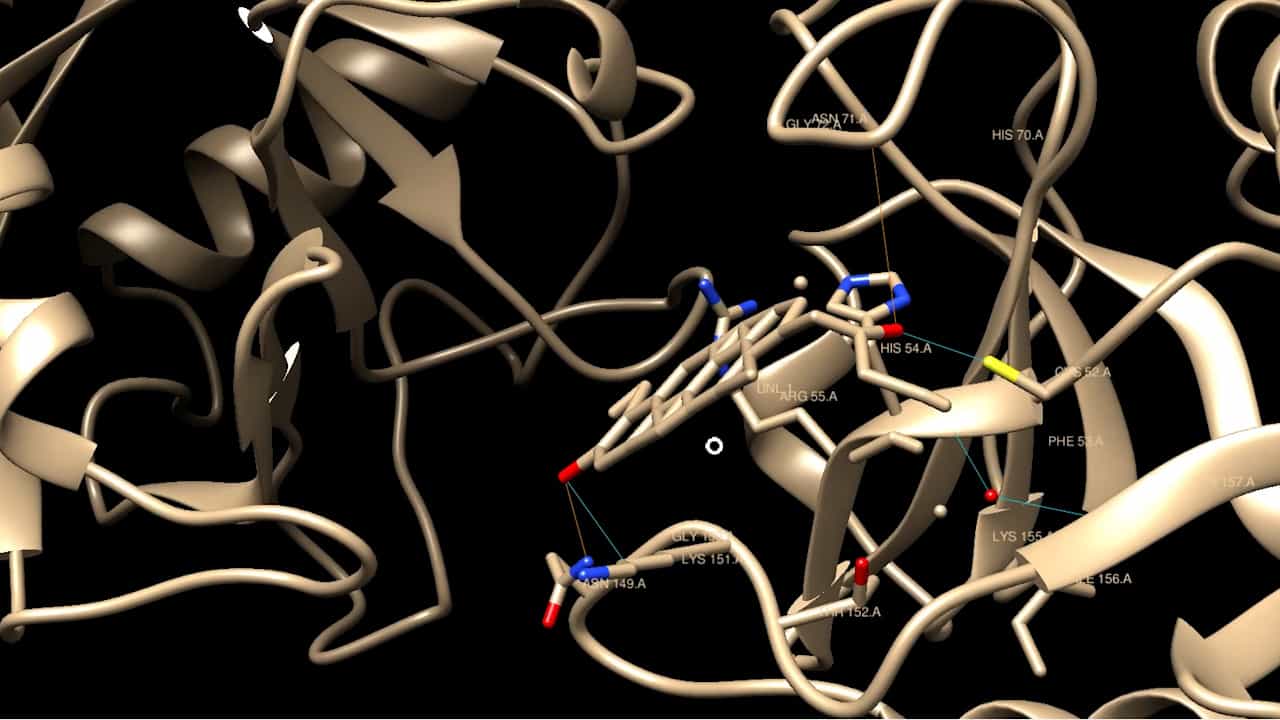

Structure-Guided Design and Development of Novel Cyclophilin A Inhibitors and Ganoderiol-F Derivatives: An In-Silico Approach
Published on: June 23, 2026
Molecular modeling design of antitrypanosomal pyrazolone derivatives targeting Chagas disease: QSAR, docking,
Karen Karla F Sousa1, Ana Beatriz S Barbosa1, Fábio José B Cardoso1
1Laboratório de Modelagem Molecular, Instituto de Ciências Exatas e Naturais, Universidade Federal do Pará, Belém, 66075-110, Brazil.

Context:
Chagas disease, caused by T. cruzi, remains a neglected tropical disease with limited therapeutic options, underscoring the urgent need for novel antitrypanosomal agents. A computer-aided drug design (CADD) protocol was applied to investigate pyrazolone derivatives with reported antitrypanosomal activity and their putative interaction with cruzain (Cz), a cysteine protease essential for parasite survival. A 2D-QSAR model built from a curated set of 33 phenyl-dihydropyrazolone derivatives guided the design of four new candidates (LMM1-LMM4), which showed intermediate predicted antitrypanosomal activity within the chemical space of the original pyrazolone series. Molecular docking identified favorable binding within the Cz active site, with contacts at residues Ser61, Gly66, and Leu67. MD simulations indicated overall structural stability of the protein-ligand complexes and revealed ligand-dependent conformational behavior within the Cz binding site. MM/GBSA calculations suggested that van der Waals interactions are major contributors to the computed binding energies; however, the MM/GBSA ranking did not fully reproduce the phenotypic antitrypanosomal activity trend. Per-residue decomposition and free energy landscape analyses were therefore interpreted qualitatively, suggesting that contacts with S2/S3 subsite residues and conformational adaptability may contribute to Cz recognition but do not, alone, explain whole-cell potency. These findings support pyrazolones as antitrypanosomal scaffolds and provide structural hypotheses for future experimental validation against Cz.
Methods:
Molecular geometries were optimized at the PM6 level (MOPAC2016); descriptors were calculated with ChemDes and UseGalaxy, selected via OPS and a genetic algorithm (QSARINS), and the QSAR model was built using PLS regression (QSAR modeling software). Molecular docking was performed with GOLD, FITTED, and AutoDock Vina 1.2.3 against Cz (PDB: 3KKU). ADMET properties were predicted using OSIRIS Property Explorer, SwissADME, and ADMET-AI. Partial atomic charges were derived at the HF/6-31G* level with RESP fitting (Gaussian 09); MD simulations (3 × 250 ns per system) were run with AMBER20 using GAFF/ff14SB force fields and TIP3P solvent. Binding free energies were calculated by MM/GBSA (MMPBSA.py).
More Related Videos
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
 05:08
Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
Related Concept Videos
 01:22
American Trypanosomiasis
 01:15
Anthelminthic Agents
 01:21
Antiprotozoal Agents