Related Experiment Video
Updated: Sep 11, 2026

Workflow and Tools for Crystallographic Fragment Screening at the Helmholtz-Zentrum Berlin
Published on: March 3, 2021
Addressing the selectivity challenge in matrix metalloproteinases: fragment-based design of MMP-9 inhibitors
Ashish Kumar1, Md Sameer1, Arbaz Sujat Shaikh1
1Department of Chemical Sciences, National Institute of Pharmaceutical Education and Research (NIPER), Hyderabad, 500037, India.

Abstract:
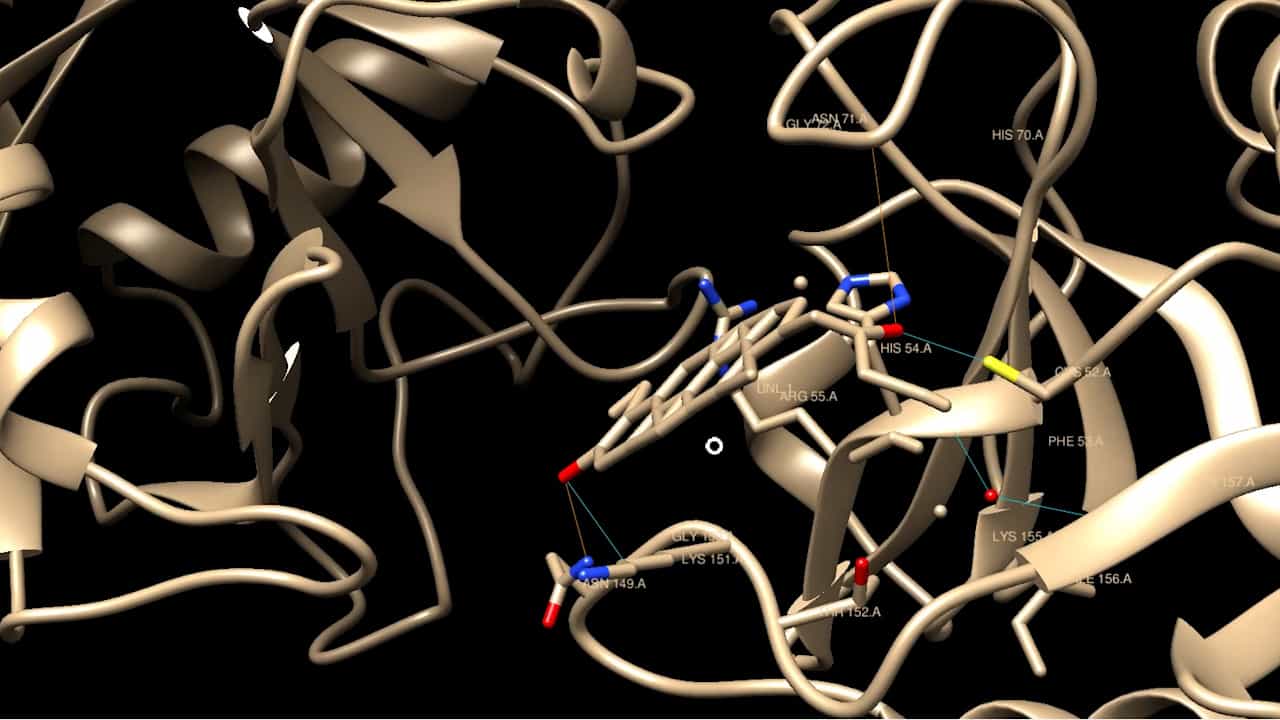
Matrix metalloproteinase-9 (MMP-9) plays a pivotal role in extracellular matrix remodelling and is strongly implicated in tumour progression, metastasis, and angiogenesis, making it an attractive therapeutic target for anticancer drug discovery. However, the development of potent and selective MMP-9 inhibitors has been significantly hampered due to the high structural homology among members of the MMP family, particularly within the catalytic domain. Structural studies revealed a non-conserved S1' pocket of MMP-9 exhibits distinct structural variability compared to other MMP isoforms, providing an opportunity to achieve improved selectivity. To the best of our knowledge, fragment-based drug design (FBDD) approaches specifically targeting the S1' pocket of MMP-9 have not been systematically explored. In the present study, an integrated in silico FBDD workflow was employed to identify and optimize small molecules occupying the S1' pocket of MMP-9. Initial fragment-based virtual screening identified low-molecular-weight fragments with moderate binding affinity toward the target pocket. A promising fragment hit was subsequently optimised through a receptor cavity-guided fragment-growing strategy to extend interactions toward residues in the neighbouring S2' pocket. Induced-fit docking analysis identified several optimized molecules with enhanced binding affinity and favourable orientation within the S1' pocket. The stability of the protein-hit molecule complexes was further validated through 200 ns molecular dynamics simulations, which confirmed stable binding interactions for four molecules. Comparative docking and MM-GBSA analyses against MMP isoforms demonstrated a preferential affinity of hit molecules for MMP-9, suggesting a potential selectivity profile driven by ligand binding free energy and orientation in S1' pocket. In addition, in silico ADME profiling indicated favourable pharmacokinetic properties of all the stable hit molecules, while density functional theory studies helped to gain insights into the electronic distribution and reactivity of the molecules. Collectively, this study establishes an in silico framework for the structure-based design of potentially selective MMP-9 inhibitors targeting the non-conserved S1' pocket.