Related Experiment Video
Updated: Sep 16, 2026

An In Vitro Batch-culture Model to Estimate the Effects of Interventional Regimens on Human Fecal Microbiota
Published on: July 31, 2019
Dietary Fiber-Associated Differences in Gut Microbial Community Composition and Predicted Short-Chain Fatty
1Department of Biological Sciences, College of Sciences and Arts Khulais, University of Jeddah, Jeddah 21959, Saudi Arabia.

Abstract:
Dietary fiber shapes both gut microbial community structure and the fermentable substrates available to resident taxa, yet how fiber deprivation reshapes taxonomic representation and predicted functional potential remains incompletely resolved. We performed an in silico re-analysis of publicly deposited 16S rRNA gene data from 15 female C57BL/6 mice distributed across three dietary cohorts-fiber-replete (F), no-fiber (NF), and no-fiber with exogenous short-chain fatty acid supplementation (NF-SCFA)-using processed taxonomic profiles and PICRUSt-derived KO/EC and KEGG pathway representations. No new animals, sequencing, enzyme assays, metabolomics, or direct SCFA measurements were generated. The F cohort exhibited the highest alpha-diversity summaries (observed features, Shannon, Simpson) and higher read-count representation for fiber-associated taxa, including Clostridium and Bifidobacterium, whereas NF profiles showed elevated representation of Ligilactobacillus in several samples. Beta-diversity analyses (PERMANOVA, PCoA) revealed that F communities were compositionally distinct from both fiber-deprived cohorts, while NF and NF-SCFA remained similar to one another. Among 52 differentially represented KO/EC features mapped across 31 KEGG pathway groups, 19 were F-preferential and 33 NF-preferential, spanning carbohydrate, pyruvate, propanoate, and butanoate metabolism. Critically, the NF-SCFA cohort failed to recapitulate the F-like taxonomic or functional pattern, indicating that exogenous SCFA supplementation alone does not restore fiber-associated community structure. These findings describe associations and predicted functional potential rather than demonstrated enzyme activity, metabolite concentration, or SCFA output, and should be interpreted as hypothesis-generating. The results identify candidate taxa and pathways warranting direct biochemical, transcriptomic, and strain-resolved validation before conclusions about fiber-dependent SCFA biosynthesis can be established.
Related Concept Videos
 01:18
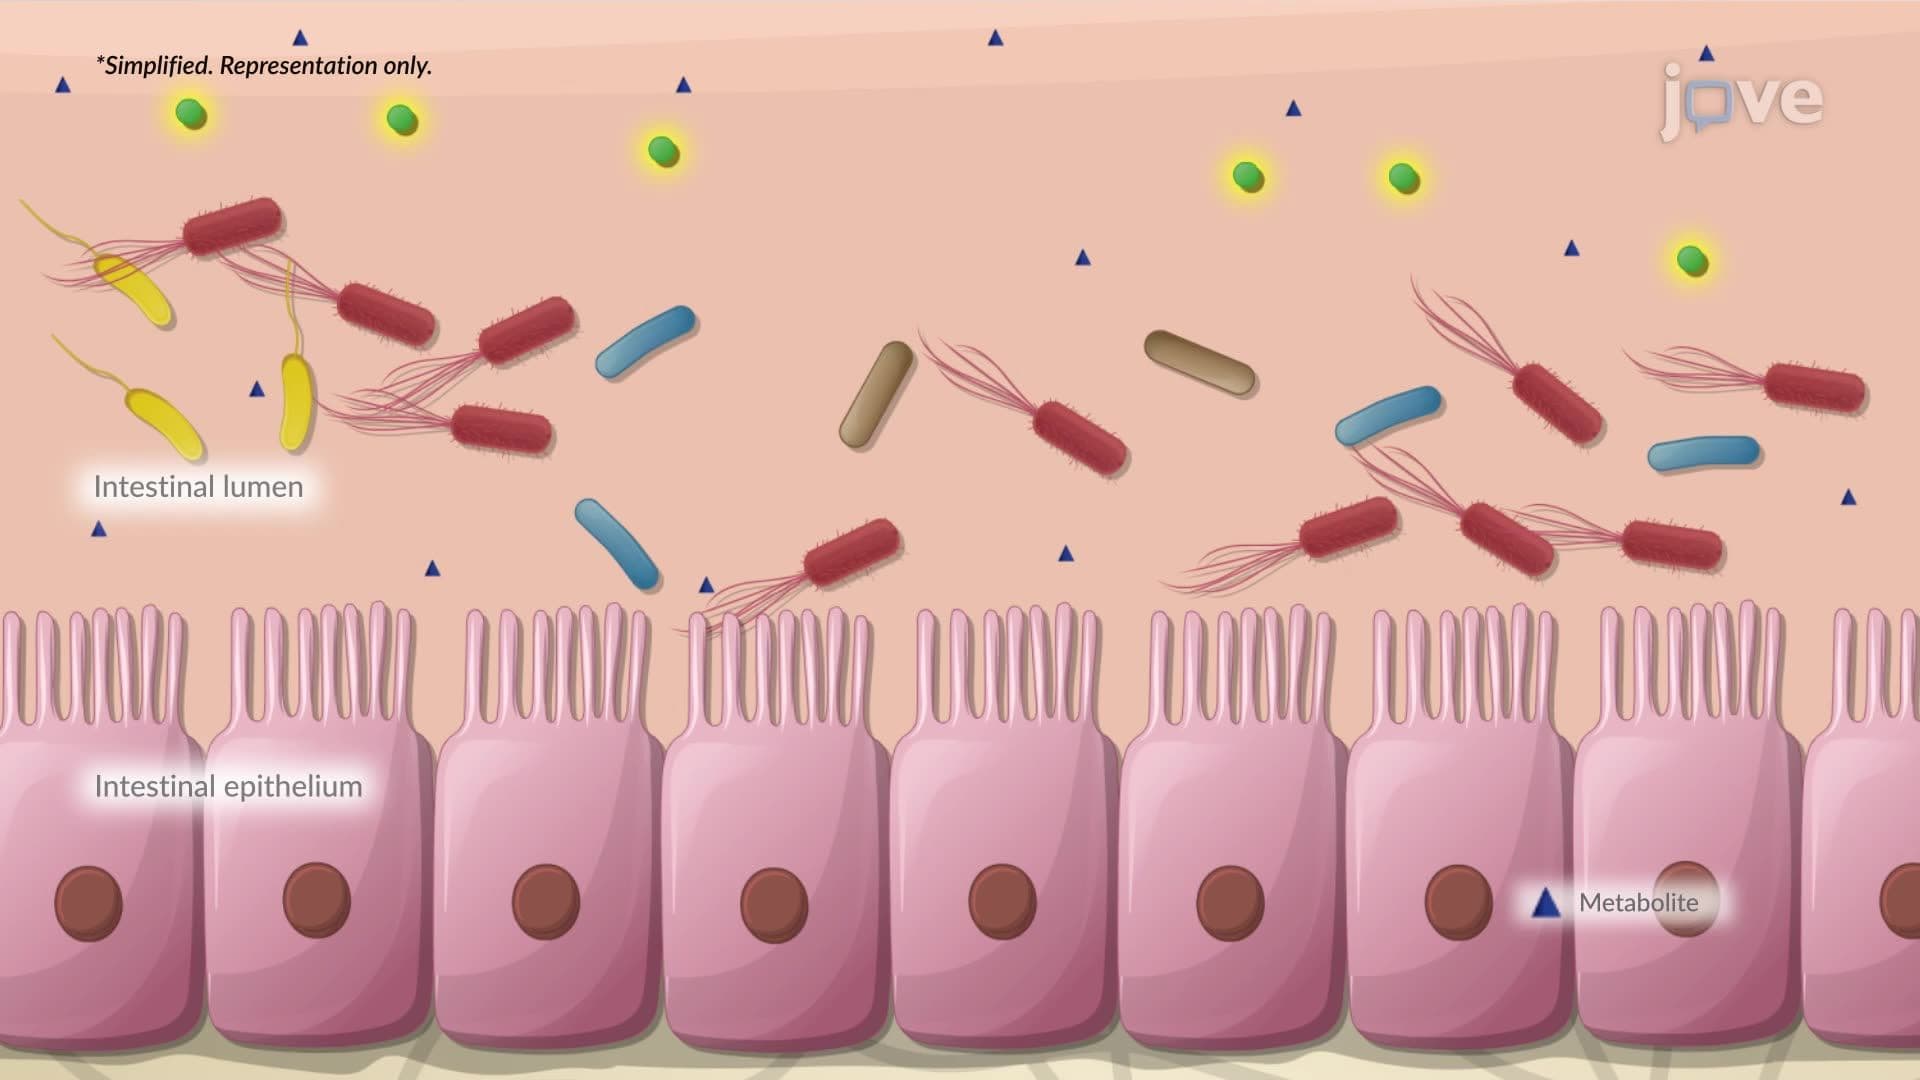
Dysbiosis of the Gut Microbiota
 01:27
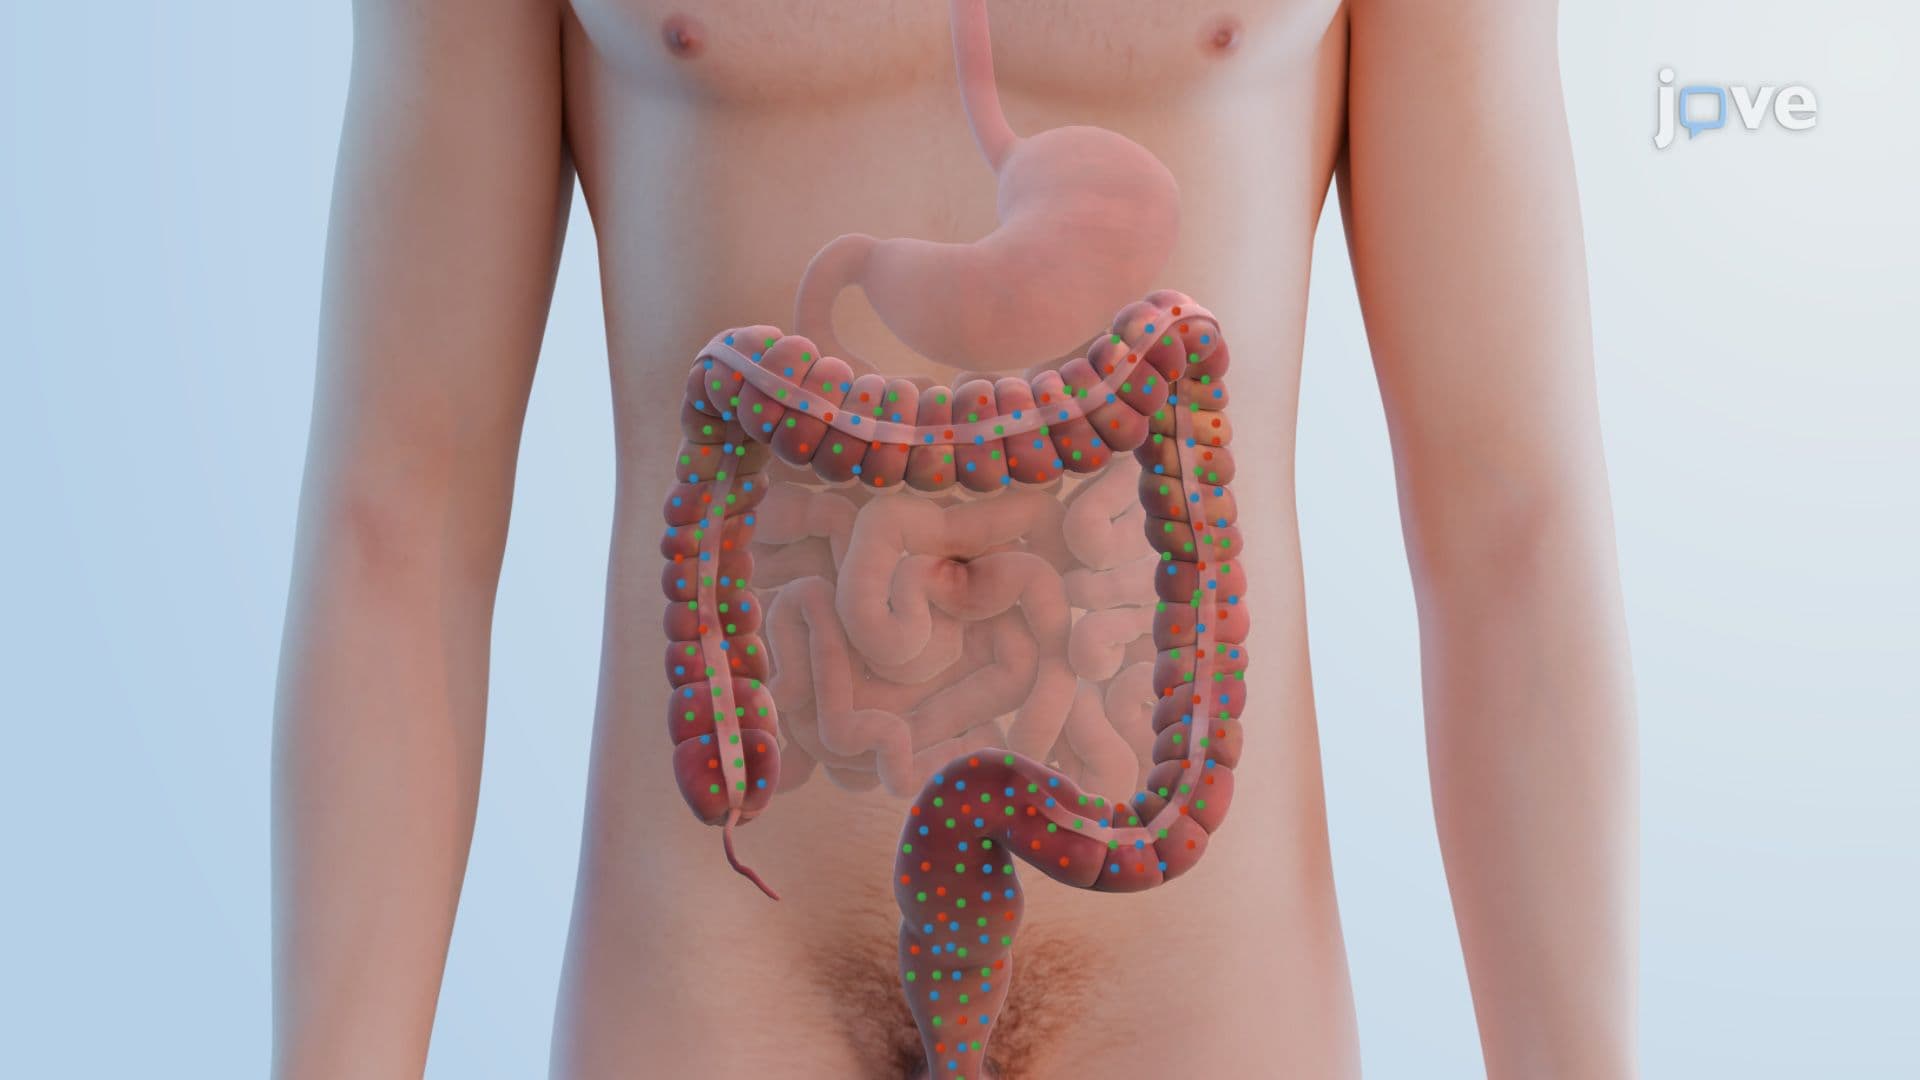
Microbiota of the Large Intestine
 01:18
Functions of the Gut Microbiota
 02:09
Overview of Human Digestion
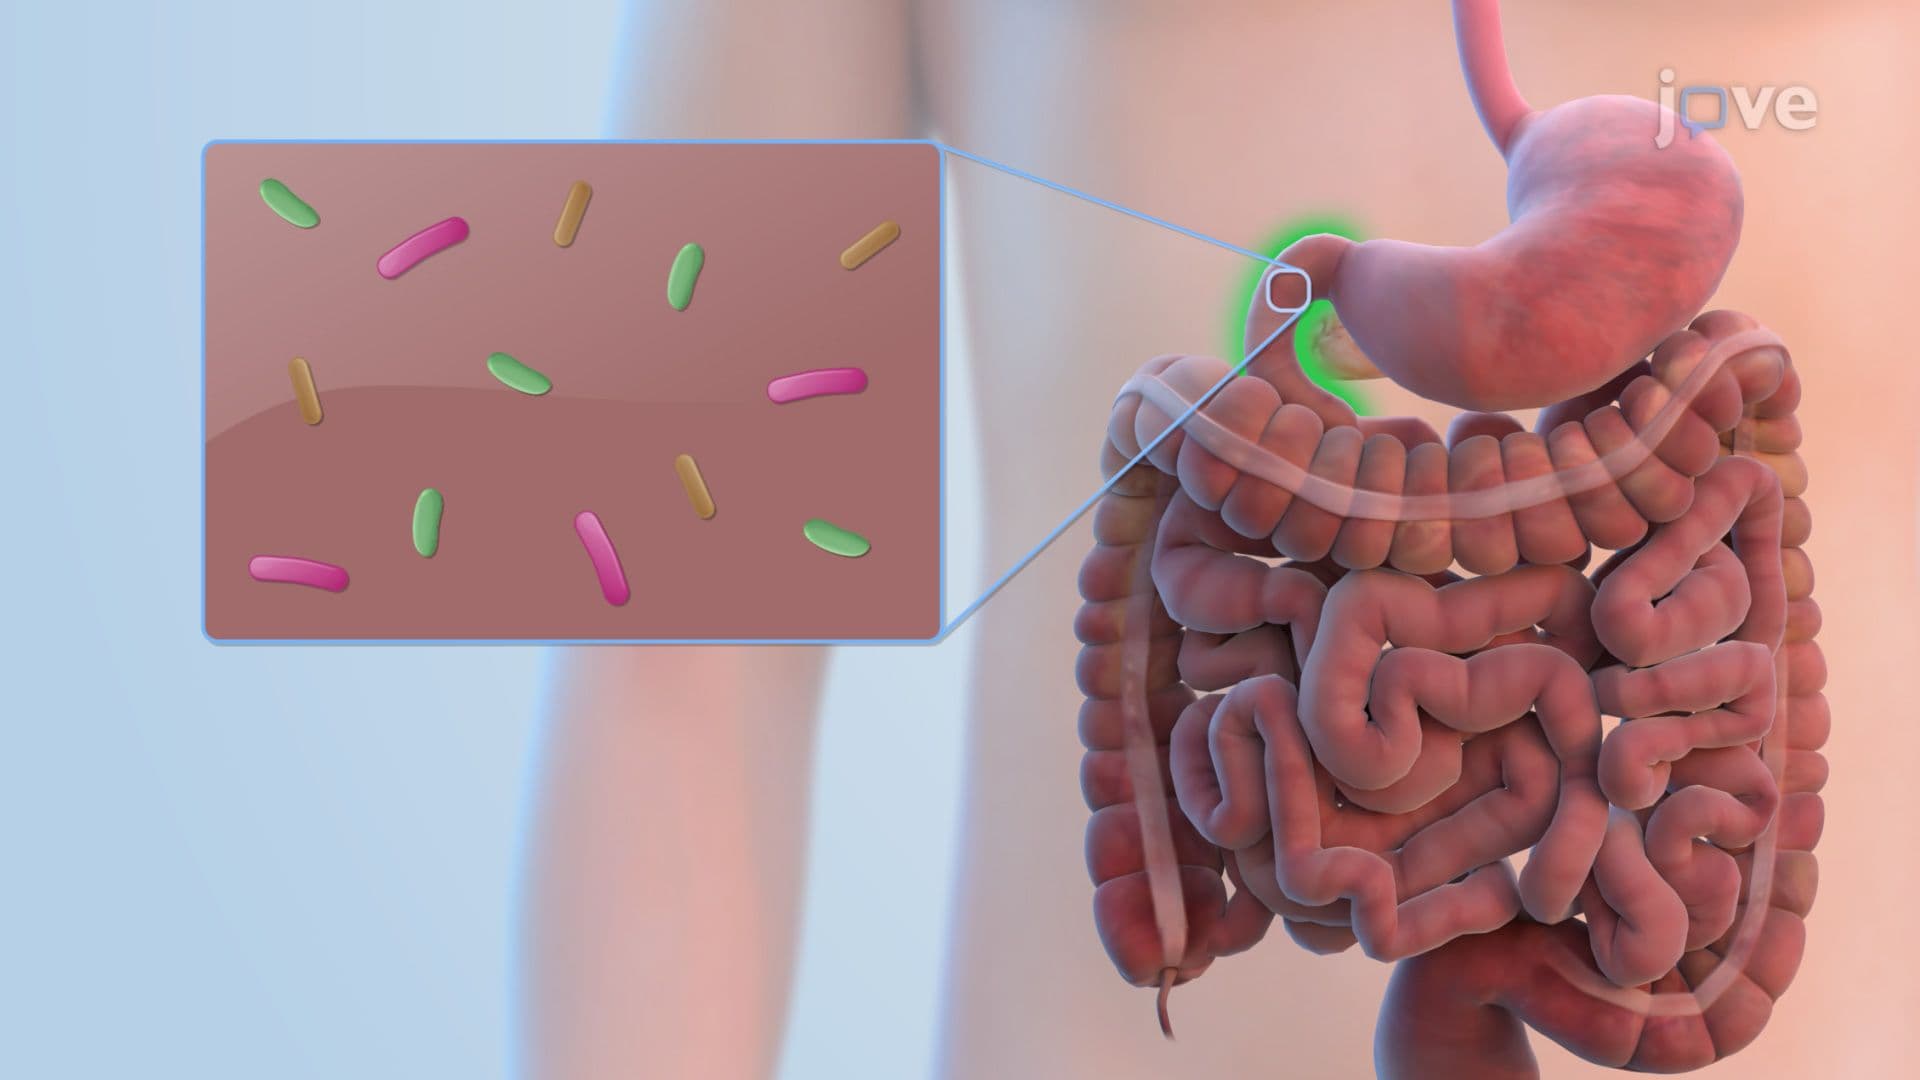 01:27
Microbiota of the Stomach and Small Intestine