Related Experiment Video
Updated: Sep 16, 2026


Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Mutation-Aware Machine Learning Framework for Predicting Binding Affinity of Nirmatrelvir Analogs Targeting
Md Saidur Rahman1, Md Mehedi Hasan1, Shahidul M Islam1
1Department of Chemistry, Delaware State University, Dover, DE 19901, USA.

Abstract:
The emergence of resistance-associated mutations in coronavirus main protease (Mpro) poses a significant challenge to the development of broad-spectrum antiviral therapeutics. In this study, we improved and accelerated a mutation-aware machine learning (ML) framework to predict the binding score of Nirmatrelvir analogue ligands against wild-type and mutant MERS-CoV Mpro. A library of 15,889 Nirmatrelvir derivatives generated through systematic scaffold modification was docked against the wild-type and five variants of the Mpro, producing a total of 95,334 structural and docking score datasets of these protein-ligand complexes. During the ML model development phase, ligand effects were learned from RDKit molecular descriptors and graph-based representations, and the mutation-induced effects were captured through delta-encoded physicochemical properties (hydrophobicity, charge, aromaticity, and polarity) of the active-site residues. Among the evaluated models, the CatBoost regressor tree-based algorithm achieved the lowest mean absolute error (MAE) value of 0.23 Kcal/mol and an R2 of 0.87. Further improvement was achieved by creating a weighted ensemble model combining the CatBoost regressor, XGBoost and LightGBM regressor, resulting in a prediction accuracy with a MAE of 0.19 Kcal/mol and an R2 of 0.90 relative to docking scores. Model robustness was further evaluated through random-, ligand group- and scaffold group- K-fold cross-validation along with their Y-randomization. Moreover, the models were also tested with a new set of 1000 structurally diverse compounds. SHAP analysis was conducted, which identified 20 molecular descriptors critical for accurate predictions. The ensemble model accurately predicted the binding affinities of Nirmatrelvir and its four analogues (E1-E4), reproducing the experimental pIC50 trend and correctly identifying the most potent inhibitors. The ensemble model also showed consistent performance across all MERS-CoV Mpro variants, S147Y, S142G, L144A, S142G/S147Y, and S142G/L144A/S147Y, demonstrating its potential for rapidly discovering mutation-resistant antiviral drugs.
Related Concept Videos
 01:25
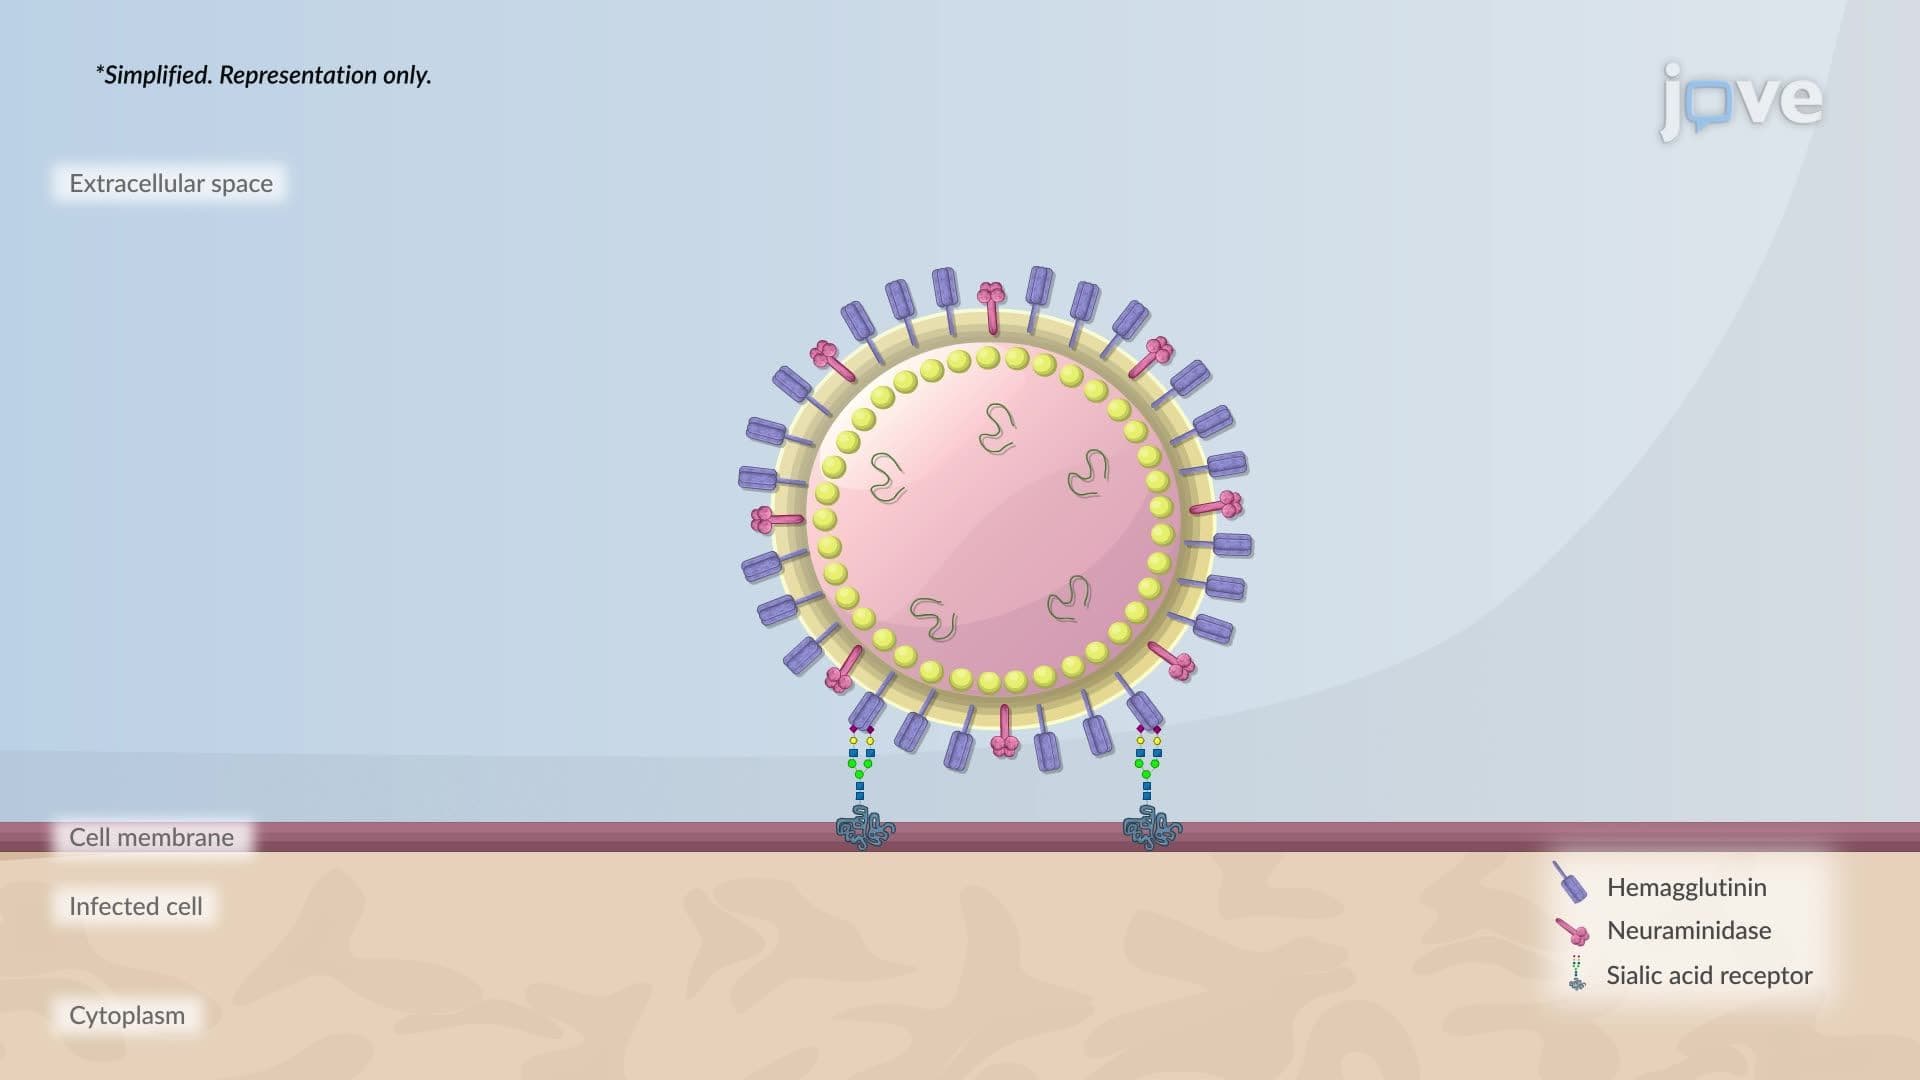
Inhibitors Of Virion Release
 01:19
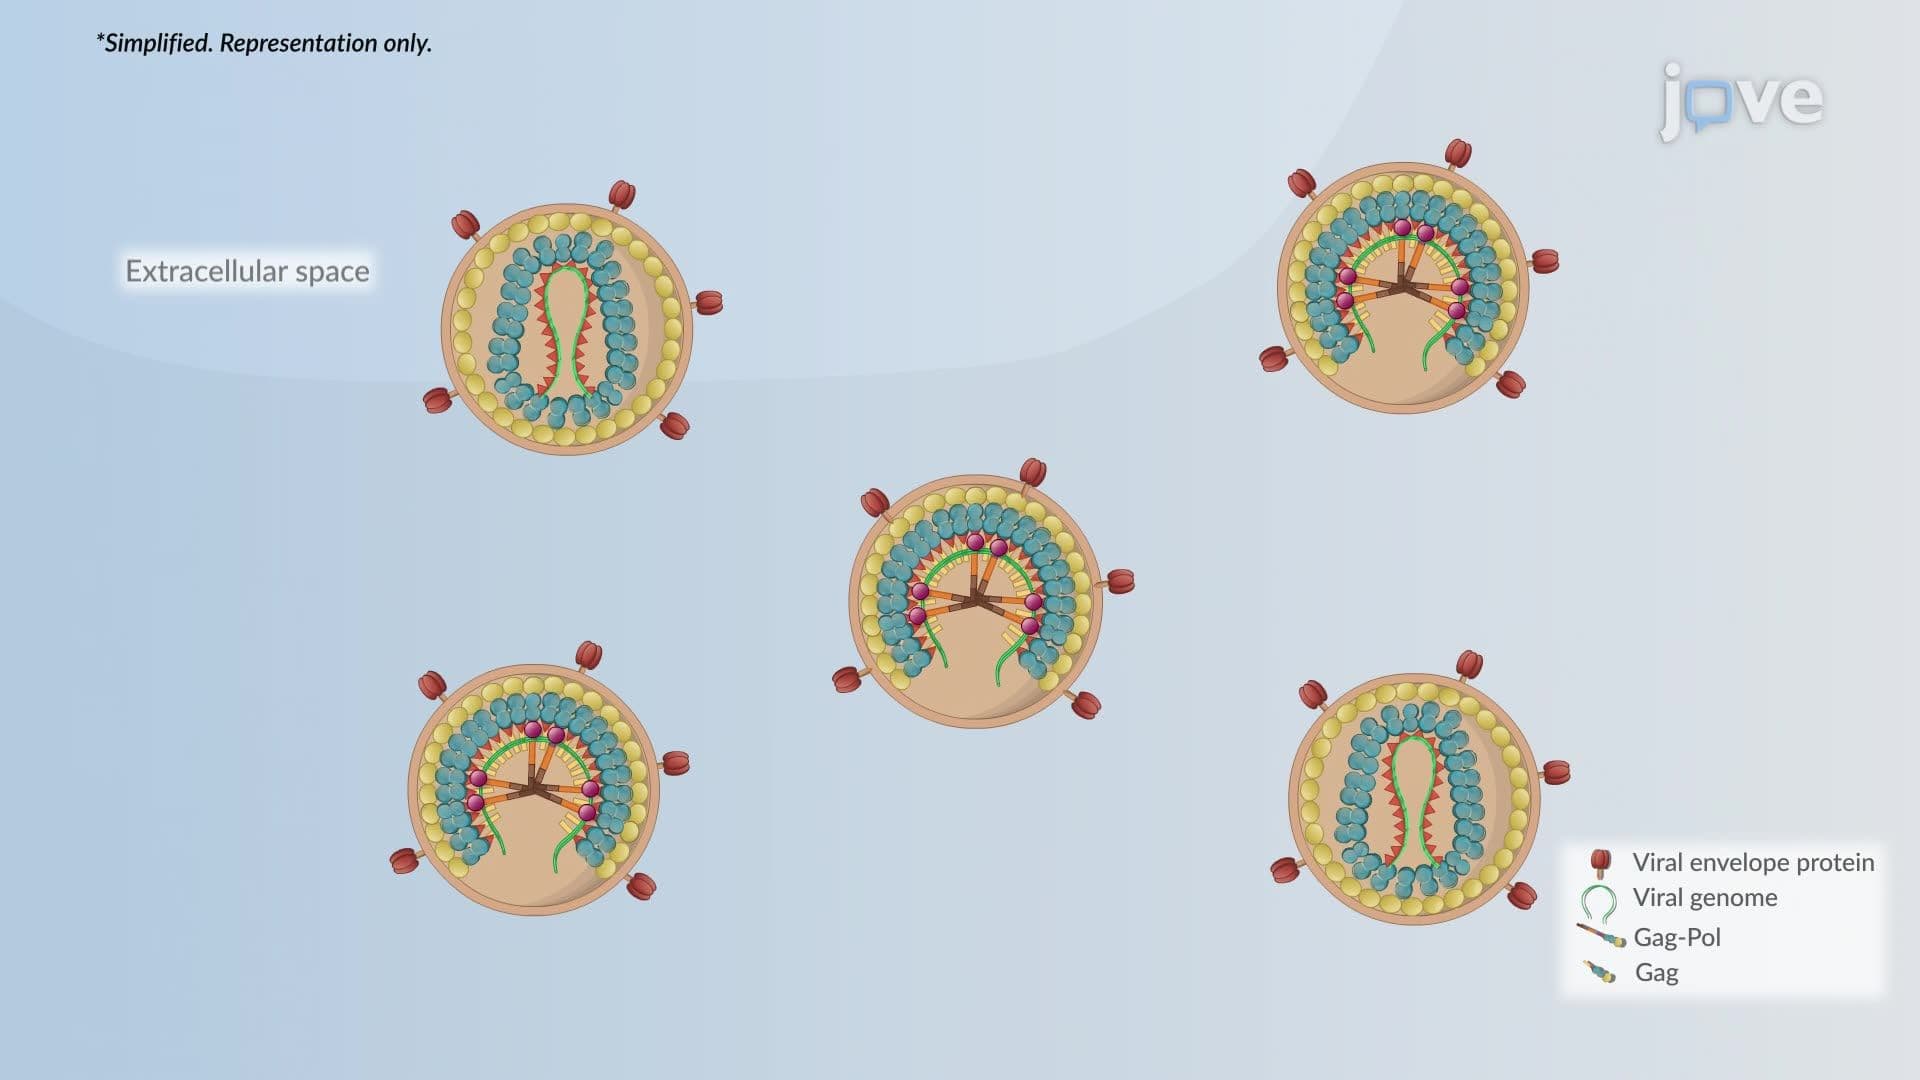
Inhibitors of Virion Maturation and Assembly
 01:30
Inhibitors of Viral Protein Synthesis