Related Experiment Video
Updated: Sep 16, 2026

Quantitative Atomic-Site Analysis of Functional Dopants/Point Defects in Crystalline Materials by Electron-Channeling-Enhanced Microanalysis
Published on: May 10, 2021
Vacancy-induced modifications to the electronic structure of barium lanthanum cobaltite: density functional theory
Francis Oseko1, Aleksandra Mielewczyk-Gryń1, Szymon Winczewski1
1Institute of Nanotechnology and Materials Engineering, Gdańsk University of Technology Gabriela Narutowicza 11/12 80-233 Gdańsk Poland francis.oseko@pg.edu.pl.

Abstract:
This work investigates the influence of oxygen and cobalt vacancies on the structural, electronic and magnetic properties of cubic barium lanthanum cobaltite (BLCO). A novel multi-objective evolutionary algorithm is applied in the generation of dedicated pseudopotentials which are later used in the density functional theory (DFT) calculations performed for the perfect and defective BLCO structures. The Hubbard + U correction is applied to La, Co and O atoms and the accuracy of the developed methodology is assessed through the cohesive energy and equation of state calculations, performed for the perfect BLCO. The results are found to be in agreement with the experimental observations and earlier all electron calculations. The validated methodology is applied in the investigation on defective BLCO, considering three types of defects: a single oxygen vacancy, transverse-oxygen (trans-O) vacancy pair, and complex oxygen-cobalt-oxygen vacancy, all in charged and neutral states. The obtained results reveal that all studied defects considerably affect the structural, electronic and magnetic properties of BLCO, which we comprehensively characterize. Among others, we show that in BLCO the redistribution of charge has a mixed character, with the introduction of defects resulting in both charge localization and delocalization. We attribute this to competing mechanisms originating from the half-metallicity of BLCO, its ferromagnetic ordering, and the tendency of Co atoms to undergo reduction. Our DFT + U calculations show only a mild reduction of the Co atoms near the defect, and this observation is contrary to the strong reduction predicted by defect chemistry models.
More Related Videos
 12:02
Determination of Thermodynamic Properties of Alkaline Earth-liquid Metal Alloys Using the Electromotive Force Technique
Published on: November 3, 2017
 06:49
Radio Frequency Magnetron Sputtering of GdBa2Cu3O7−δ/ La0.67Sr0.33MnO3 Quasi-bilayer Films on SrTiO3 (STO) Single-crystal Substrates
Published on: April 12, 2019
Related Concept Videos
 02:42
Valence Bond Theory
 02:45
Valence Bond Theory
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:54
Trends in Lattice Energy: Ion Size and Charge
 01:27
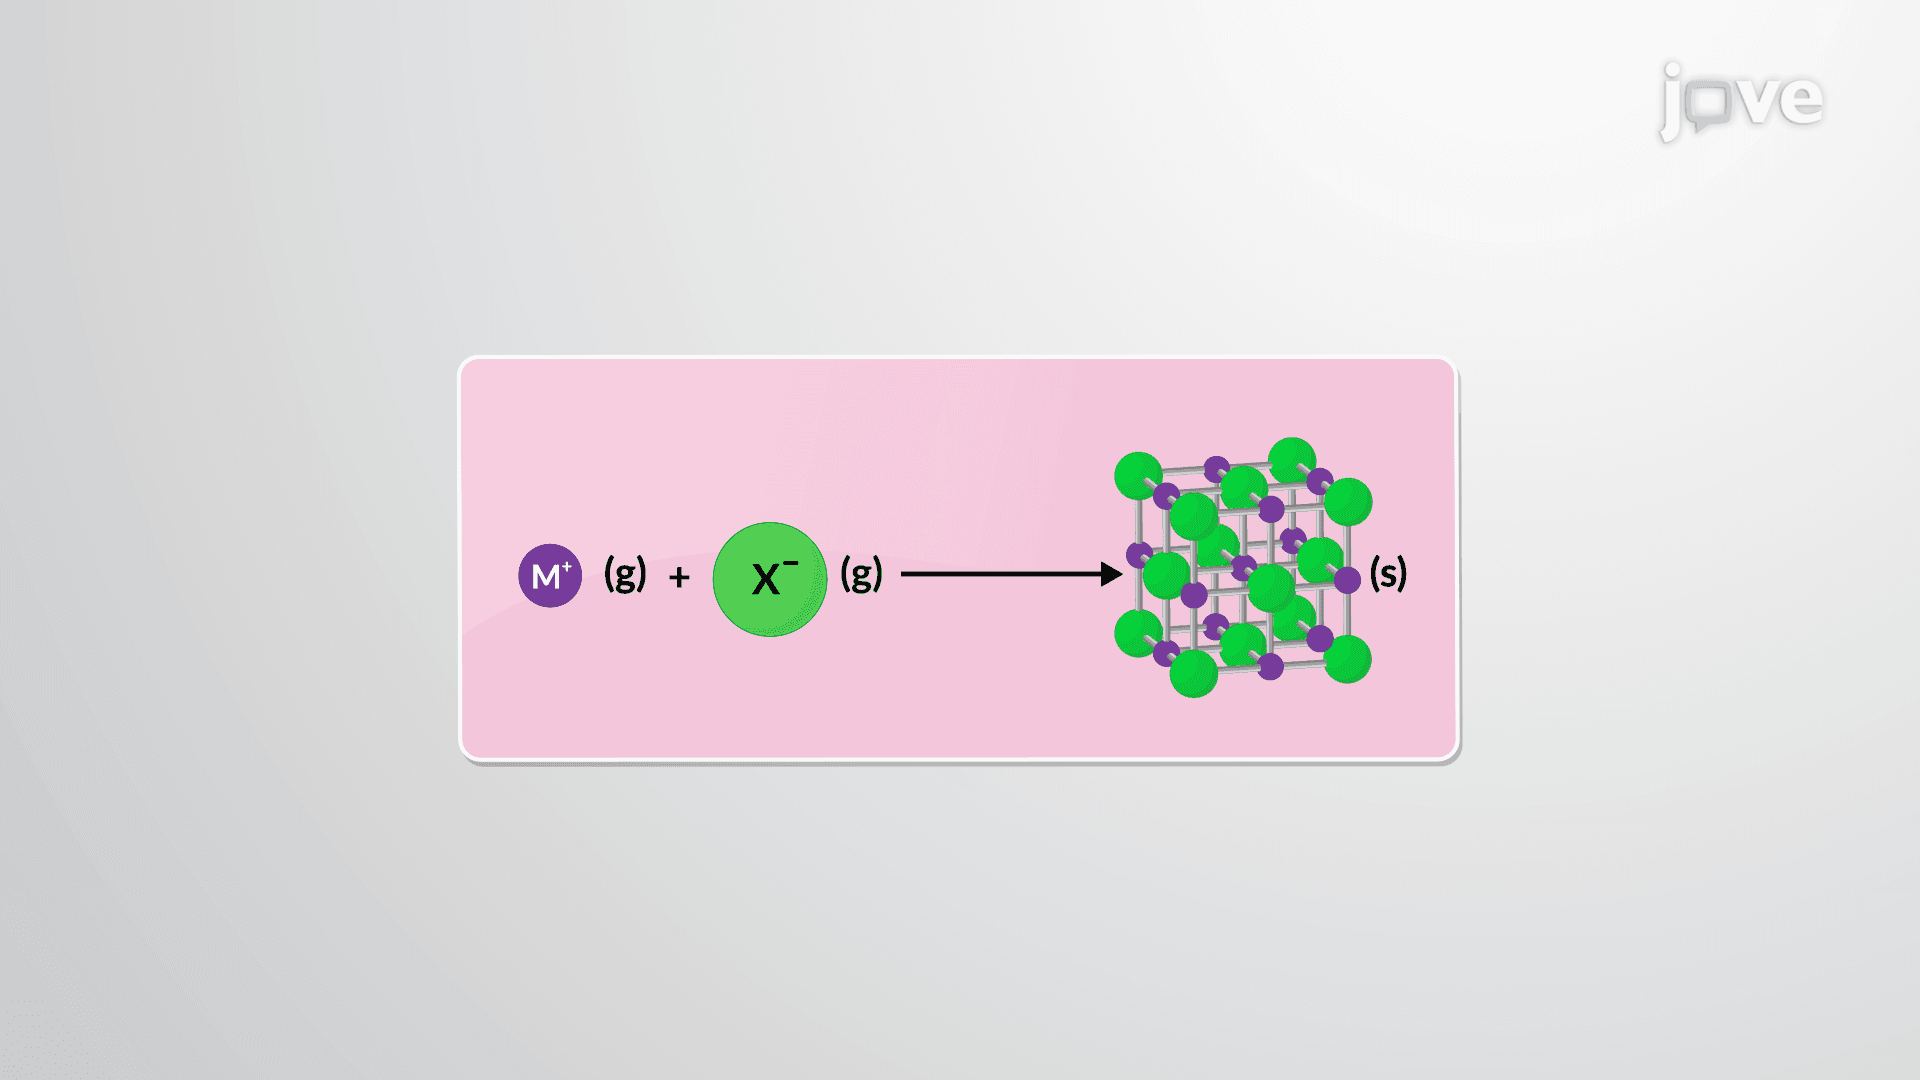
Lattice Energies of Ionic Crystals
 03:02
Colors and Magnetism
When atoms or molecules absorb light at the proper frequency, their electrons are excited to higher-energy orbitals. For many main group atoms and molecules, the absorbed photons are in the ultraviolet range of the electromagnetic spectrum, which cannot be detected by the human eye. For coordination compounds, the energy difference between the d orbitals often allows photons in the visible range to be absorbed and emitted, which is seen as colors by the human eye.