Related Experiment Video
Updated: Sep 16, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Evaluation of the coupled-cluster correction term, δCCSD(T)MP2, calculated using small valence basis sets
Tao Sun1, Weizhou Wang2, Yi-Bo Wang1
1College of Chemistry and Chemical Engineering, and Key Laboratory of Guizhou High Performance Computational Chemistry, Guizhou University, Guiyang 550025, China.

Abstract:
In the focal-point estimation of the coupled cluster with singles, doubles, and perturbative triples [CCSD(T)] interaction energies followed by extrapolation to the complete basis set (CBS) limit, the most critical and challenging step is the rapid and accurate calculation of the coupled-cluster correction term, δMP2CCSDT, which represents the difference between CCSD(T) and MP2 interaction energies at a small basis set. In this study, using calculations with the widely used aug-cc-pVDZ (aVDZ) basis set as the benchmark, we primarily evaluated the reliability of calculating δMP2CCSDT with the much smaller valence basis set vDZP and its diffuse-function-augmented version vDZPD. The results show that with the vDZP basis set, the mean MAD and RMSD of δMP2CCSDT across five benchmark datasets for noncovalent interactions (S22, S66, X40, HSG, and NCIBLIND10) are 0.14 and 0.18 kcal/mol, respectively, indicating relatively large deviations. In contrast, the vDZPD basis set yields much lower average MAD and RMSD of only 0.02 and 0.04 kcal/mol, demonstrating high computational accuracy. Using the vDZPD basis set to calculate noncovalent systems containing around 30-50 atoms, the δMP2CCSDT computation time is saved by about 40%. Furthermore, the results show that the δMP2CCSDT/vDZPD corrections still maintain high accuracy in calculating metal-containing complexes. The CCSD(T)/CBS interaction energy benchmark can be pushed to the hundred-atom scale with vDZPD basis set, holding significant promise for widespread application in the study of large noncovalent systems.
Related Concept Videos
 01:27
π Electron Effects on Chemical Shift: Overview
 01:02
¹H NMR: Interpreting Distorted and Overlapping Signals
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are slanted or...
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 01:08
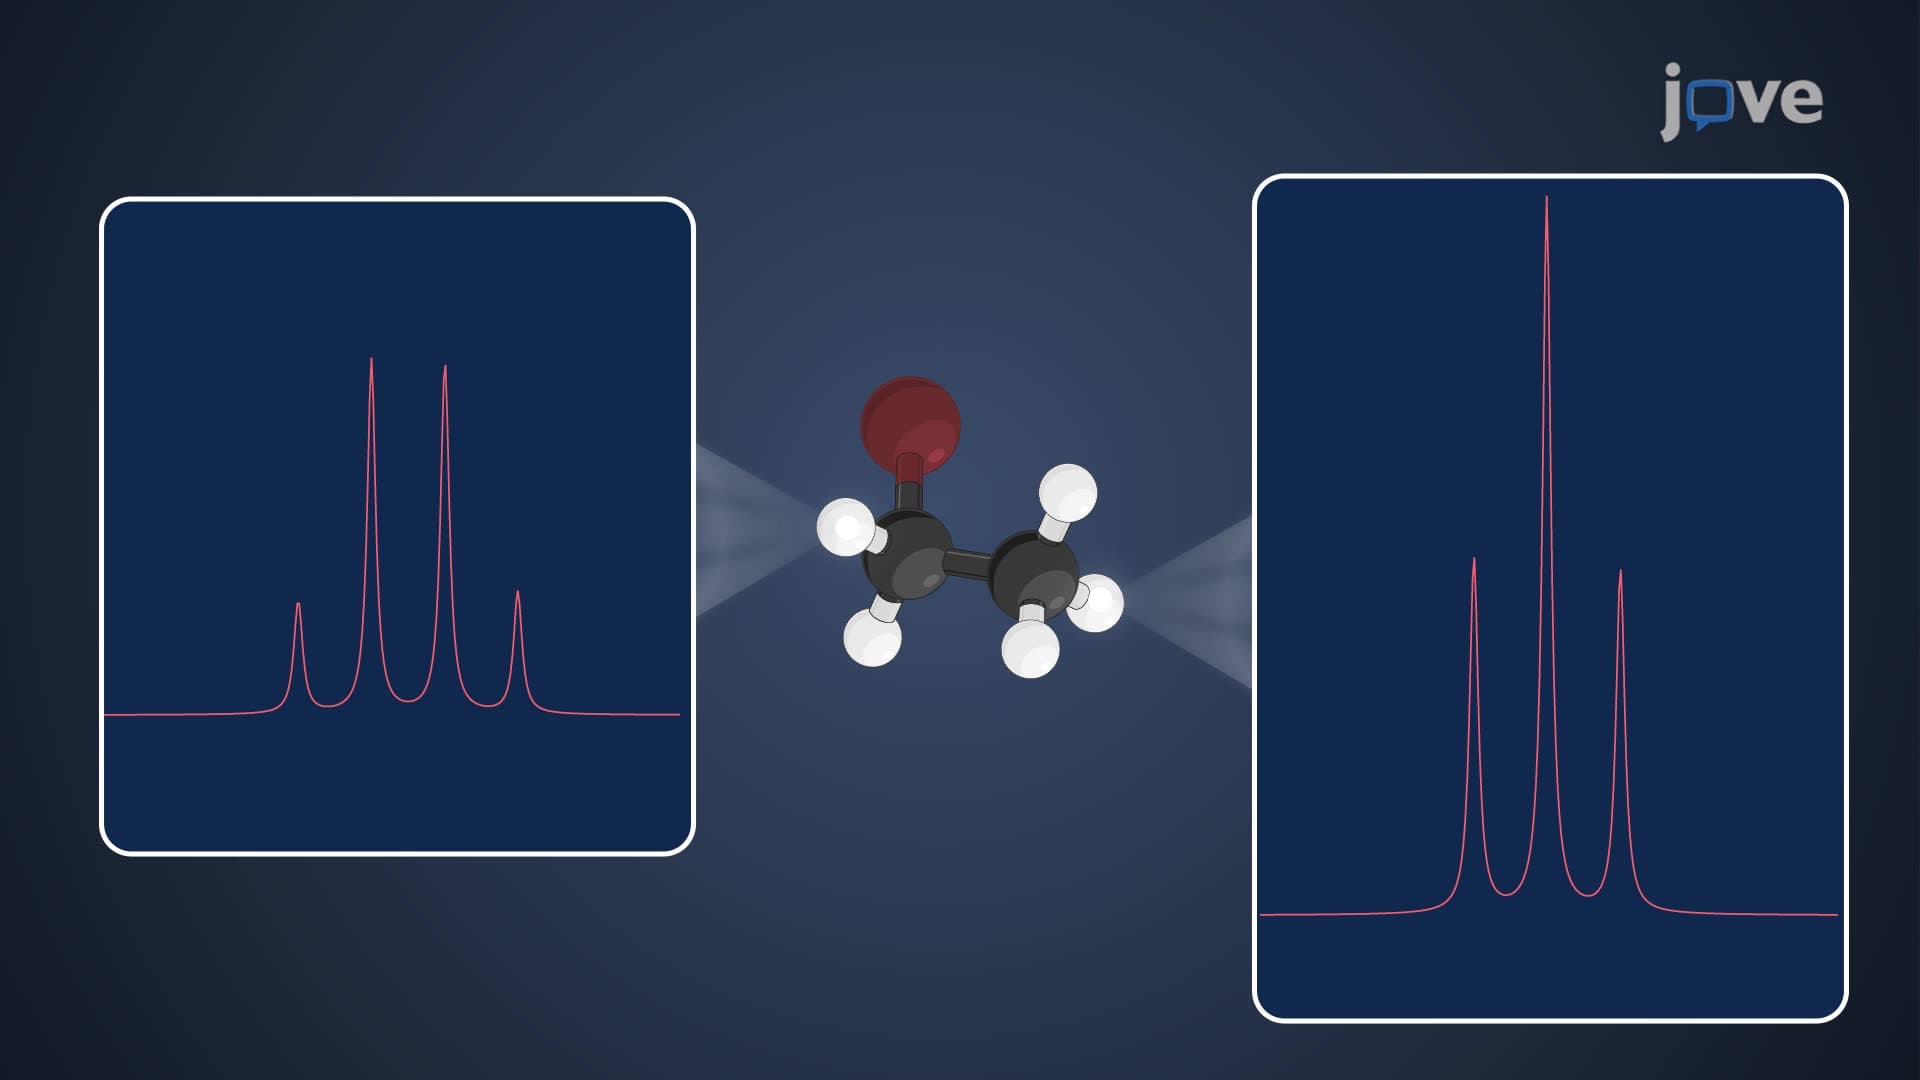
Spin–Spin Coupling Constant: Overview
Qualitatively, any spin plus-half nucleus polarizes the spins of its electrons to the minus-half state. Consequently, the paired electron in the hydrogen–carbon bond must have a...
 01:12
Double Resonance Techniques: Overview
Spin decoupling is usually achieved by...
 01:10
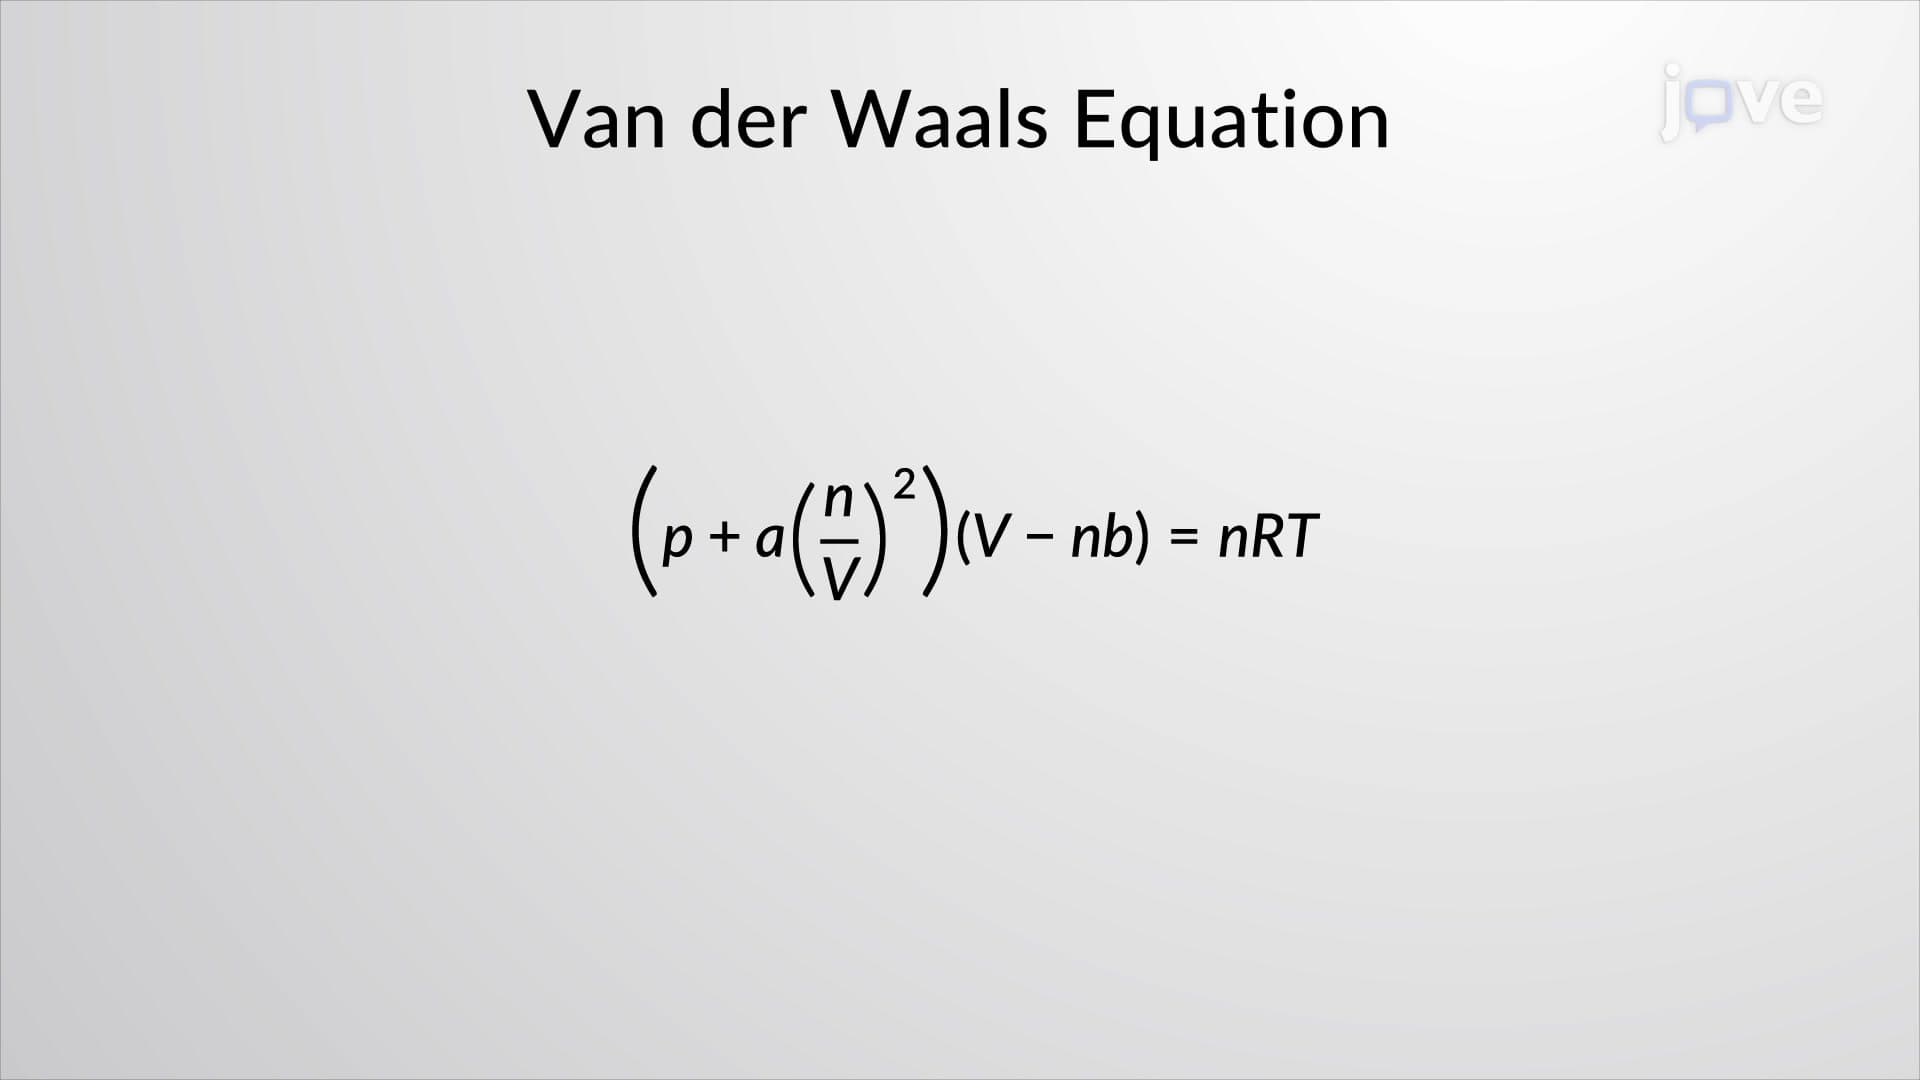
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...