Related Experiment Video
Updated: Sep 16, 2026
Dissecting Innate Immune Signaling in Viral Evasion of Cytokine Production
Published on: March 2, 2014
RABV P protein activates the GSK3β-NF-κB signaling pathway to promote viral replication by interacting with the host
Yujie Bai1, Yongsai Liu1, Xiaoyu Wu1
1State Key Laboratory for Diagnosis and Treatment of Severe Zoonotic Infectious Diseases, Key Laboratory for Zoonosis Research of the Ministry of Education, Institute of Zoonosis, and College of Veterinary Medicine, Jilin University, Changchun, China.

Abstract:
Rabies virus (RABV), a lethal neurotropic pathogen, relies on interaction with the host to sustain neuronal infection; however, how it balances replication with neuronal survival remains unclear. Here, we revealed Snapin, a neuronal trafficking regulator, to be a critical host factor co-opted by RABV to enhance neuroinvasion. We found that RABV infection upregulated Snapin expression in N2a cells, and Snapin could promote the replication of RABV CVS11 and SRV9 strains in a dose-dependent manner. Mechanistic studies demonstrated that RABV P protein interacts with the 37-78 aa domain of Snapin via its C-terminal domain (172-297 aa), recruits glycogen synthase kinase 3β (GSK3β) in a Snapin-dependent manner, and forms a tripartite complex. Upon complex formation, the phosphorylation level at the Tyr216 site of GSK3β is increased, leading to activation of the NF-κB pathway. This activation enhances viral replication by upregulating neuronal stability genes (Septin2, B4gat1, and Disc1) without inducing interferon responses. In vivo studies show that knockdown of Snapin in infected mice delays clinical symptom onset and improves host survival. This study uncovers a unique strategy wherein RABV exploits Snapin as a molecular scaffold linking P to GSK3β/NF-κB signaling, coupling viral replication to neuronal homeostasis. Our work identifies a host-dependent axis for therapeutic targeting in rabies and offers broader implications for understanding neurotropic pathogens utilizing analogous mechanisms.
Importance:
Rabies virus (RABV) depends on host factors to maintain neuronal infection while avoiding cell death, but the mechanisms balancing viral replication and neuronal survival are unclear. This study identifies Snapin as a critical host factor commandeered by RABV to enhance neuroinvasion. Upon RABV infection, Snapin is upregulated and subsequently hijacked by the C-terminal domain of the RABV P protein. This interaction recruits GSK3β in a Snapin-dependent manner, forming a tripartite complex that increases phosphorylation of GSK3β at Tyr216 and activates the NF-κB pathway. Such activation upregulates neuronal stability genes (Septin2, B4gat1, and Disc1), thereby promoting viral replication without eliciting interferon responses. In vivo functional validation further confirms that Snapin deficiency attenuates RABV pathogenicity and prolongs host survival. Collectively, our study highlights a Snapin-dependent GSK3β/NF-κB axis as a potential therapeutic target for rabies treatment.
More Related Videos
 09:15
Tyramide Signal Amplification for the Immunofluorescent Staining of ZBP1-Dependent Phosphorylation of RIPK3 and MLKL After HSV-1 Infection in Human Cells
Published on: October 20, 2022
 09:38
Identification of Nucleolar Factors During HIV-1 Replication Through Rev Immunoprecipitation and Mass Spectrometry
Published on: June 26, 2019
Related Concept Videos
 01:30
Inhibitors of Viral Protein Synthesis
 01:03
Rous Sarcoma Virus (RSV) and Cancer
RSV is a retrovirus that contains two copies of a plus-strand RNA genome. Its genome consists of four main open...
 01:14
Rab Proteins
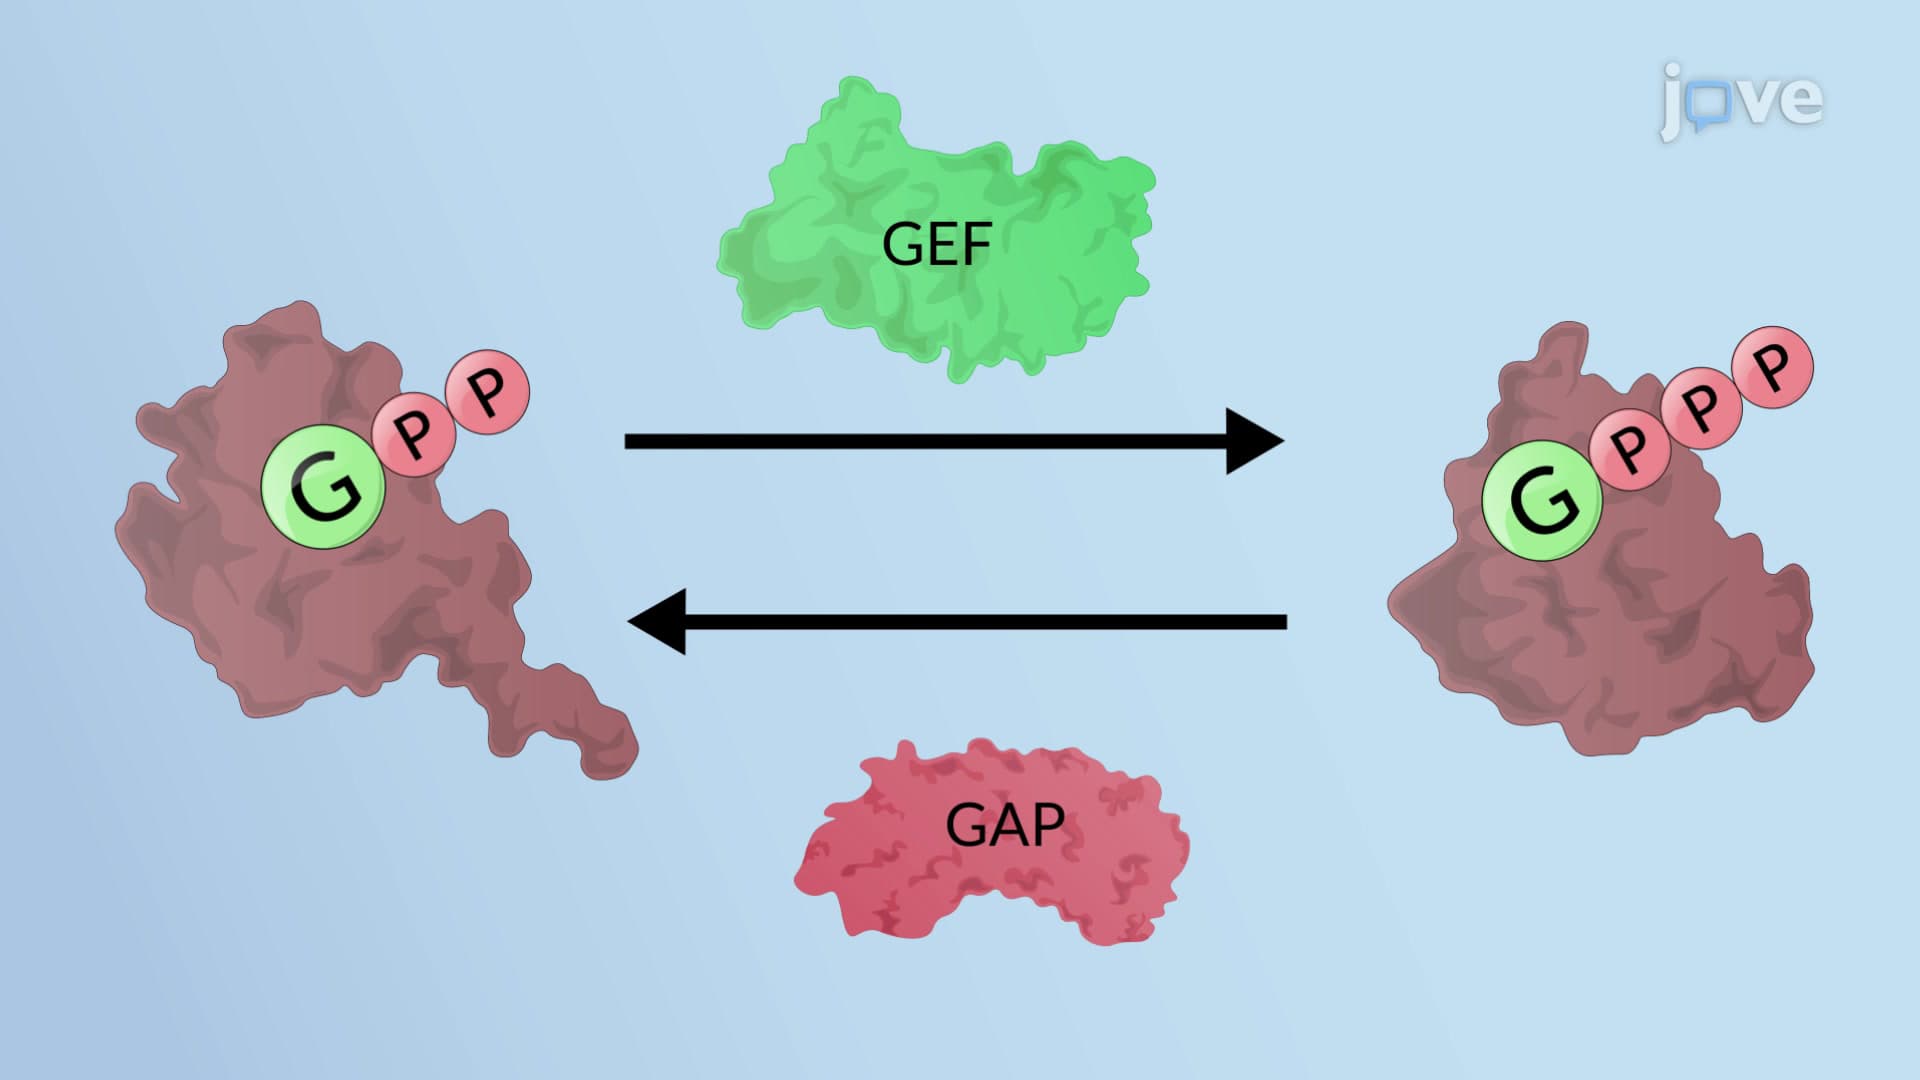
Rab proteins switch between a cytosolic, GDP-bound inactive state and a membrane-anchored, GTP-bound active state. By themselves, Rabs show slow rates of GDP/GTP exchange and GTP hydrolysis. Thus, Rab proteins are considered...
 01:25
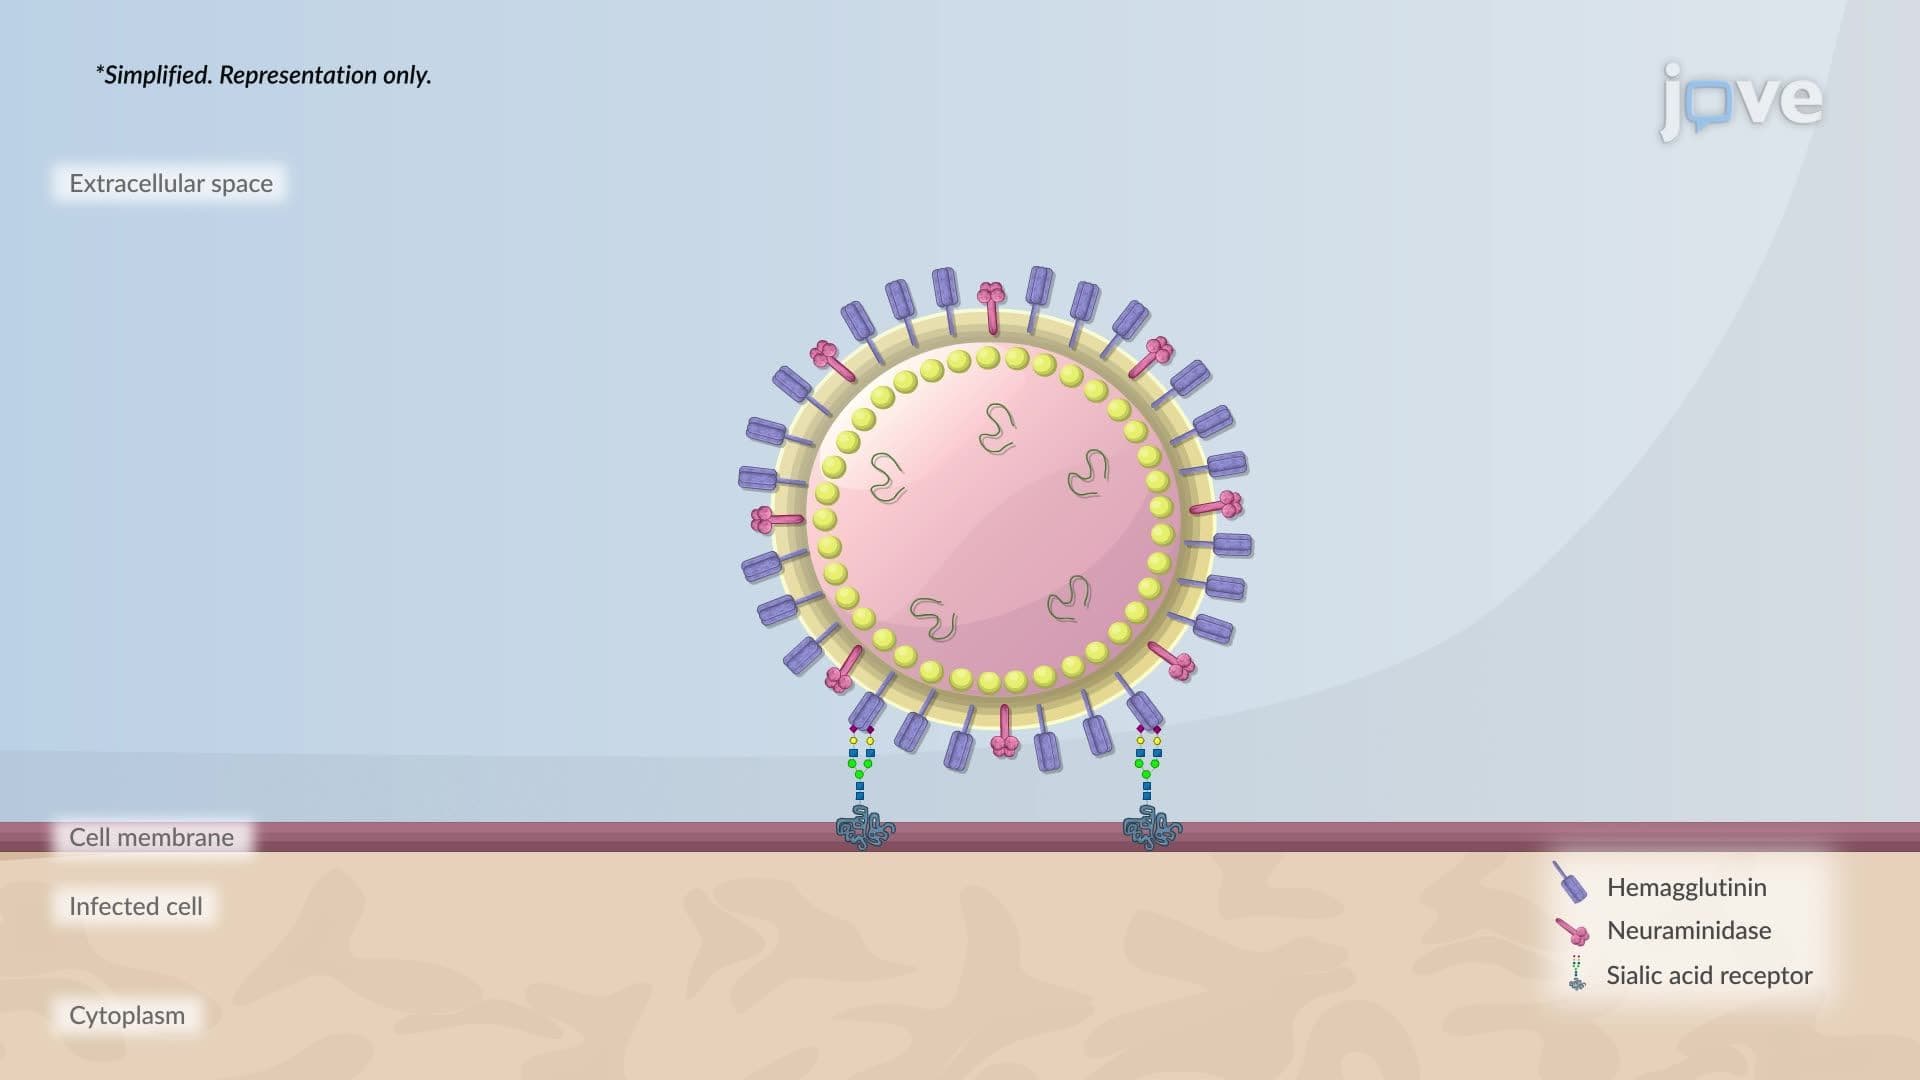
Inhibitors Of Virion Release
 01:10
Retrovirus Life Cycles
 01:51
Mechanisms of Retrovirus-induced Cancers