Cannabis as a source of novel therapeutics: A computational study combining LBDD and structure-based docking/dynamics to identify novel drug-like compounds for colorectal cancer
Related Experiment Video
Updated: Sep 18, 2026
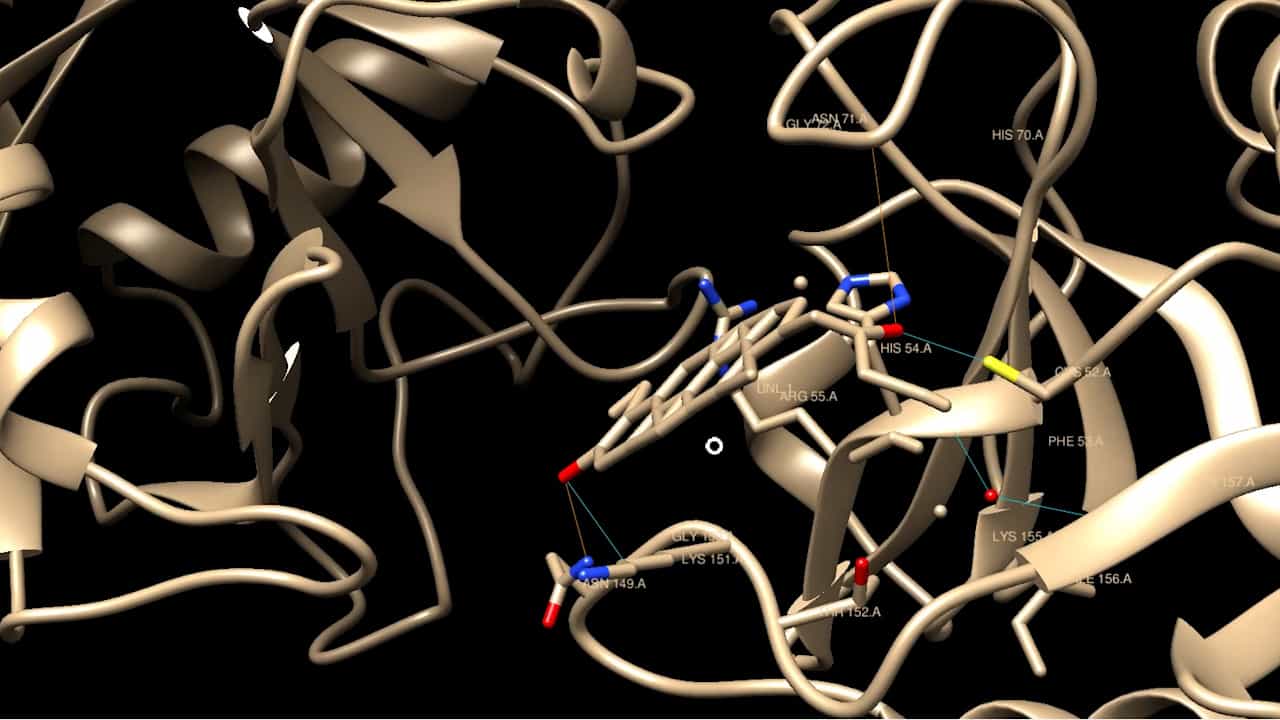
Structure-Guided Design and Development of Novel Cyclophilin A Inhibitors and Ganoderiol-F Derivatives: An In-Silico Approach
Published on: June 23, 2026
Khadija Khaddam Allah1, Khaoula Mkhayar1, Rachid Haloui1
1Laboratory of Engineering, Systems and Applications, National School of Applied Sciences, Sidi Mohamed Ben Abdellah-Fez University, Fez, Morocco.

Abstract:
Cannabis represents a valuable source of bioactive compounds with significant therapeutic potential, thanks to its rich phytochemical profile, making it a prime candidate for cancer research. Our study investigates the potential of cannabis-derived compounds to inhibit cyclin-dependent kinase 2 (CDK2), a crucial regulator of cell-cycle progression and colorectal cancer cell proliferation. A comprehensive computational workflow integrating 3D-QSAR modeling, molecular docking, molecular dynamics simulations, drug-likeness assessment, and ADMET prediction was employed to investigate a dataset of 33 cannabis-derived compounds. The robustness and predictive performance of the generated CoMFA and CoMSIA models were confirmed through internal and external validation, leave-one-out cross-validation (LOOCV), and Y-randomization tests, yielding excellent statistical parameters for CoMFA (Q2=0.651, R2=0.986, SEE=0.069) and CoMSIA (Q2=0.665, R2=0.985, SEE=0.071). Using contour map analysis, twenty new molecules (V1-V20) with enhanced antiproliferative activity against the HCT-116 colorectal cancer cell line were developed, and outperformed the original dataset's most active one. Following drug-likeness screening and ADMET profiling, compounds V2, V5, V7, and V8 emerged as the most promising candidates, exhibiting favorable pharmacokinetic properties and drug-like characteristics. Molecular docking studies revealed that V7 and V8 exhibit high stability within the CDK2 active site and possess stronger binding affinity than the reference compound. Furthermore, 100 ns molecular dynamics simulations demonstrated that both protein-ligand complexes reached stable conformational states, as confirmed by converged backbone RMSD profiles, low residue fluctuations (RMSF), persistent protein-ligand interactions, and complementary structural descriptors including radius of gyration (Rg), solvent-accessible surface area (SASA), and molecular surface area (MolSA). Collectively, these analyses confirmed the structural stability and compactness of the investigated complexes throughout the simulation period, with V8 exhibiting the most favorable dynamic behavior.

Related Concept Videos
 01:29
Pharmacogenomics: Identification of New Drug Targets
 01:26
Drug Discovery: Overview
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence its...