Related Experiment Video
Updated: Sep 23, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Exploring the chemical space of transition-metal cluster thermodynamics via automated first-principles calculations
Ning-Zheng Li1,2, Zi-Yue Wang2,3, Zi-Yu Li4,5,6
1State Key Laboratory for Structural Chemistry of Unstable and Stable Species, Institute of Chemistry, Chinese Academy of Sciences, Beijing, PR China.

Abstract:
Atomic clusters serve as the embryos of materials, yet their enormous and compositionally complex chemical space has long hindered systematic exploration of thermodynamic stability. Here, we show that a unified first-principles and machine-learning framework enables large-scale mapping of transition metal cluster thermodynamics. By integrating a progressive sampling strategy with a composition-based deep-learning model, we alleviate the intrinsic sampling bottleneck associated with exponentially expanding chemical spaces. Based on ≈ 93,000 global-minimum structures obtained via automated first-principles calculations, our model-the cluster-transformer-encoder network-enables reliable predictions of atomization energies (≈ 40 meV per atom accuracy) for 9.13 million metal cluster compositions, covering 30 d-block metals and 4 chemically relevant ligand elements (C, N, O, and S). The resulting thermodynamic trends show strong correlations between cluster atomization energies and bulk cohesive energies, and identify heteronuclear stabilization and the stability of noble-metal-doped oxide clusters. This work establishes a scalable, composition-driven strategy for efficient first-pass exploration of chemically complex small-cluster spaces, providing insights into collective stability trends of metal systems.

Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:27
Predicting Molecular Geometry
 01:10
Thermodynamics: Chemical Potential and Activity
The thermodynamic equilibrium constant is more accurately defined in terms of activity rather than concentration.
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:26
Thermodynamic Potentials
 01:21
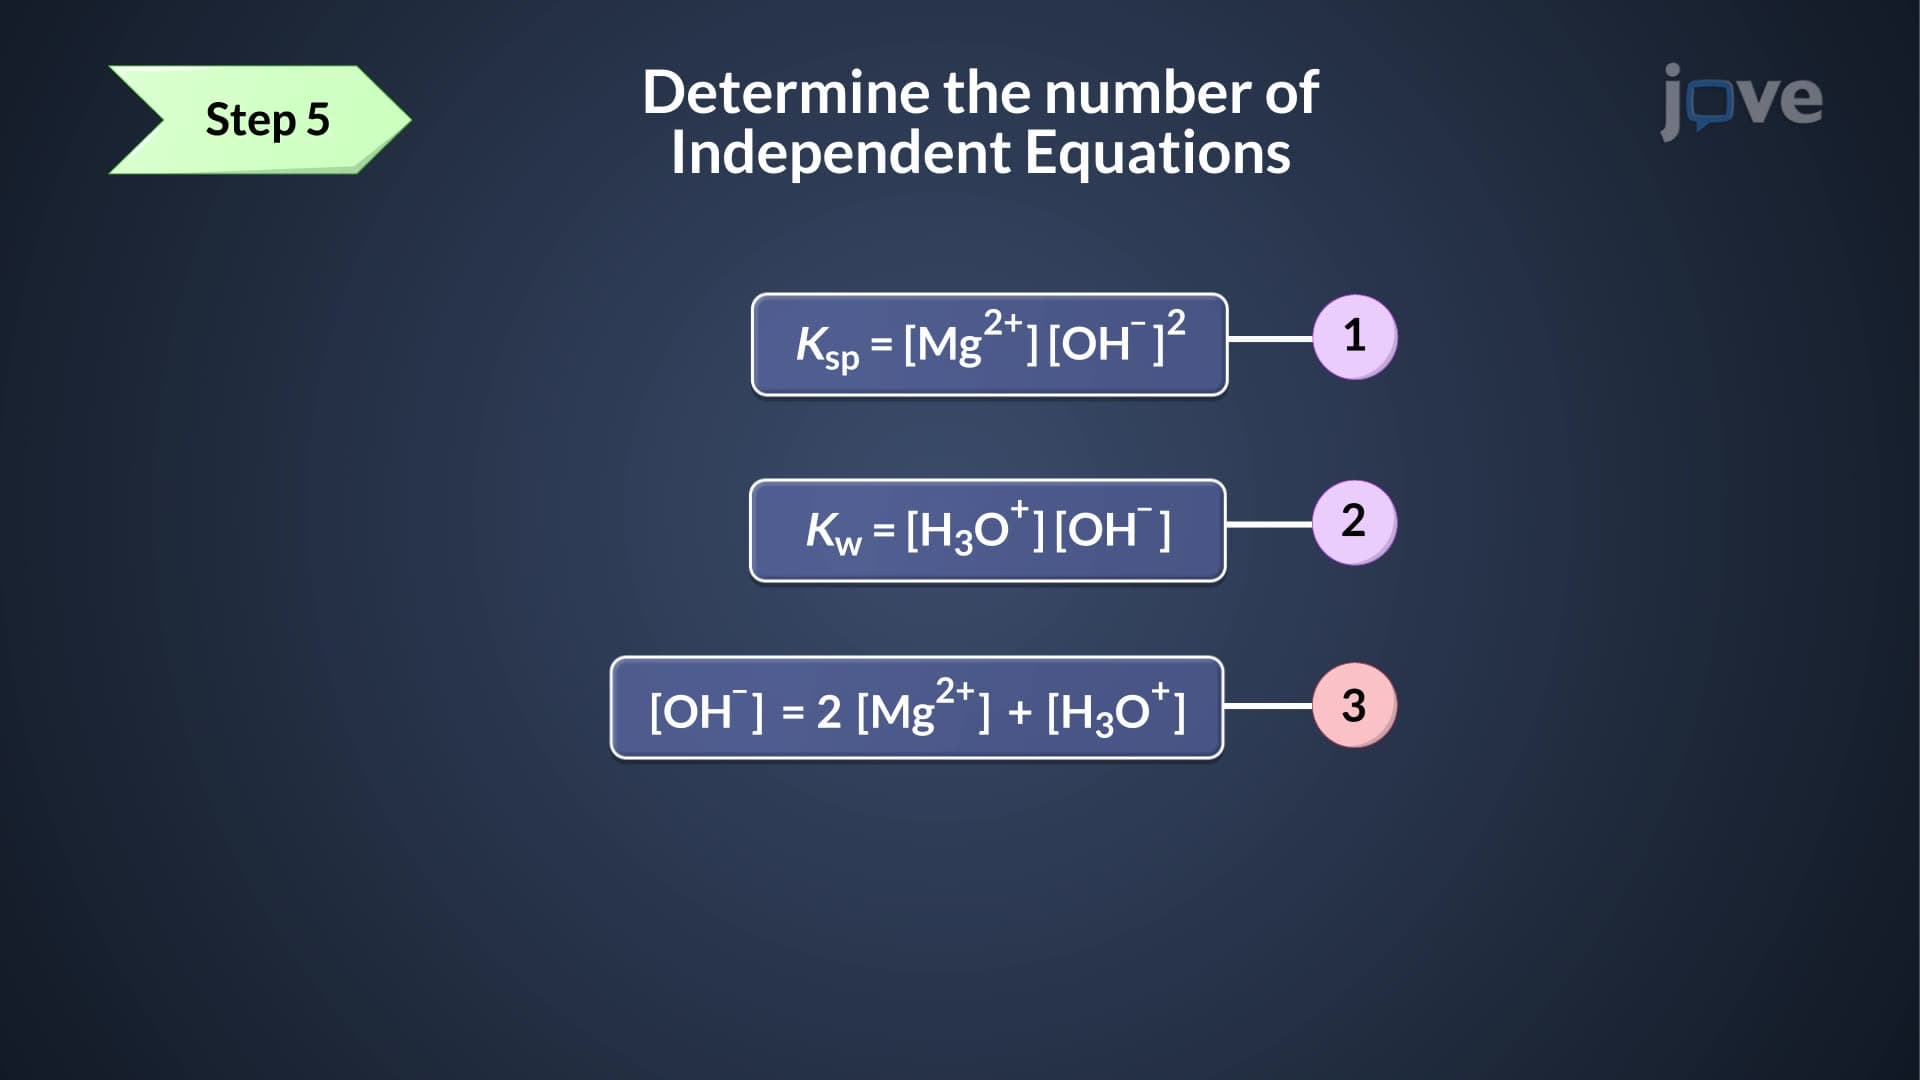
Chemical Equilibria: Systematic Approach to Equilibrium Calculations
The first step is to identify all the chemical reactions involved, The...