Related Experiment Video
Updated: Sep 23, 2026

Quantifying Tissue-Specific Proteostatic Decline in Caenorhabditis elegans
Published on: September 7, 2021
Calpain-Dependent Protein Degradation Contributes to CASQ2-R33Q CPVT
Laura Ramos-Hernández1, Demetrio J Santiago1, Rocío F Serrano-Blanco1
1Centro Nacional de Investigaciones Cardiovasculares (CNIC), Madrid, Spain (L.R.-H., D.J.S., R.F.S.-B., A.C., L.E.-M., E.C., M.L.-O., A.G.-C., J.M.V., M.T., S.G.P., E.L.-P.).

Background:
Catecholaminergic polymorphic ventricular tachycardia (CPVT) is an inherited arrhythmogenic disease associated with cardiac arrest in children and young adults. The recessive form of CPVT is caused by loss-of-function mutations in the genes encoding CASQ2 and TRDN, which, together with JCTN, regulate the release of calcium ions from the sarcoplasmic reticulum through the RyR2 (ryanodine receptor). The CASQ2 R33Q mutation causes calcium mishandling and ventricular tachycardia through triggered activity. The homozygous mouse model Casq2R33Q/R33Q closely replicates the clinical phenotype of patients with CPVT. In this model, CASQ2 (calsequestrin 2), TRDN (triadin), and JCTN (junctin) levels are markedly reduced; however, the mechanisms underlying this reduction and their contribution to CPVT are incompletely understood.
Methods:
We used a combination of proteomics, animal models, and experimental methods to identify the molecular mechanisms responsible for the decrease of couplon proteins in Casq2R33Q/R33Q mice and to explore their functional contribution to the disease.
Results:
Casq2R33Q/R33Q hearts showed activation of endoplasmic reticulum stress, the unfolded protein response, and alterations in major proteolytic pathways. Inhibition of the proteasome or autophagy in neonatal cardiomyocytes and adult mice partially restored CASQ2 but not TRDN levels, suggesting distinct degradation mechanisms. In contrast, calpain inhibition restored CASQ2 and TRDN protein levels, decreased triggered activity in isolated cardiomyocytes, and reduced ventricular tachycardia episodes in vivo in Casq2R33Q/R33Q mice. Biochemical assays showed that TRDN is a direct calpain substrate. TRDN degradation preceded CASQ2 loss, and TRDN overexpression using modified RNA in cardiomyocytes or an adeno-associated viral vector in vivo increased CASQ2 levels in mutant mice, supporting an upstream role for TRDN destabilization.
Conclusions:
Couplon protein loss in CPVT associated with CASQ2-R33Q is mediated by coordinated but distinct proteolytic pathways, with calpain-dependent TRDN degradation representing a key pathogenic event that promotes CASQ2 destabilization and arrhythmogenesis.
Related Concept Videos
 01:24
Caspases
 02:58
Regulated Protein Degradation
Protein degradation plays two important roles in the cells. It helps to protect cells from misfolded or damaged proteins before they lead to a...
02:58Regulated Protein Degradation
Protein degradation plays two important roles in the cells. It helps to protect cells from misfolded or damaged proteins before they lead to a...
 01:32
Export of Misfolded Proteins out of the ER
 01:13
The Proteasome
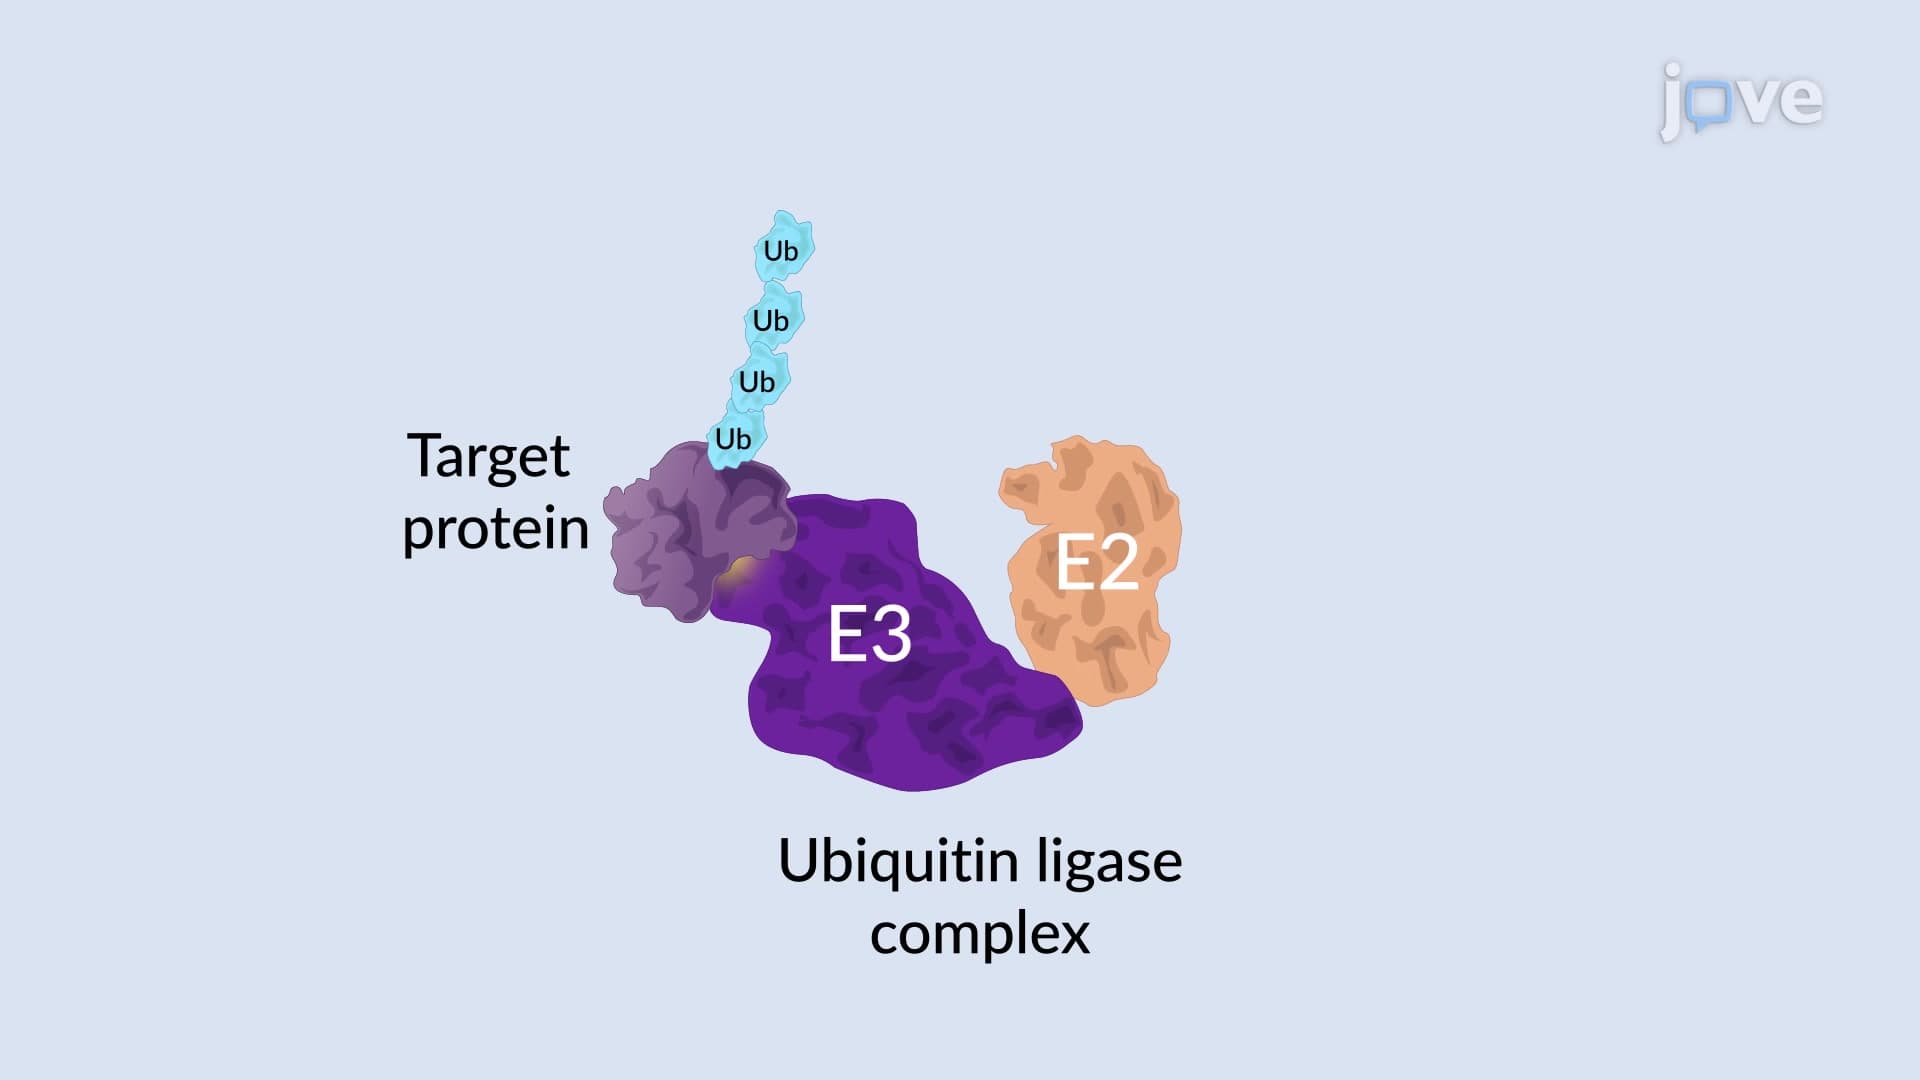
In this pathway, the target proteins are first tagged with small proteins called ubiquitin. This involves participation of a series of enzymes including— E1 (ubiquitin-activating enzyme), E2 (ubiquitin-conjugating enzyme), and E3 (ubiquitin...
 02:18
The Proteasome
In this pathway, the target proteins are first tagged with small proteins called ubiquitin. A series of enzymes carry out the ubiquitination of the target proteins - E1 (ubiquitin-activating enzyme), E2 (ubiquitin-conjugating enzyme), and E3...