Related Experiment Video
Updated: Sep 26, 2026

Microbiota Analysis Using Two-step PCR and Next-generation 16S rRNA Gene Sequencing
Published on: October 15, 2019
A data-driven universal gut microbiome health assessment: a machine learning framework trained on large metagenomic
Bablu Kumar1,2, Erika Lorusso2,3, Bruno Fosso2
1Department of Oncology and Hematology-Oncology, Università degli Studi di Milano, Milan, Italy.

Abstract:
The gut microbiota is essential to maintain host physiology, and its disruption (dysbiosis) is associated with a wide range of diseases. Machine learning (ML) offers a powerful tool to model species-level microbiome profiles, but classifiers that reliably separate healthy from diseased individuals across independent cohorts are still lacking. In this study, we developed a ML classifiers trained on 7,452 publicly available stool metagenomes spanning 32 studies and 12 diseases, designed to distinguish healthy individuals (absence of a clinically diagnosed disease) from non-healthy individuals (presence of a clinically diagnosed disease) based on species-level gut microbiome profiles. We trained 16 supervised models combining four algorithms (RF, SVM-LIN, SVM-RBF, and LR-ElasticNet) combined with all feature sets and three feature-selection algorithms. Performance was assessed by F1 score and ROC-AUC on held-out test data and externally validated on 642 samples from six independent cohorts, including previously unseen diseases. On the test set, all models achieved F1 scores of 78-86% and ROC-AUC values of 89-95%. An SVM-RBF model using permutation-based feature selection performed best (F1 = 86.6%, ROC-AUC = 95.5%; healthy F1 = 86.6%, non-healthy F1 = 88.7%). Importantly, external validation confirmed the generalizability of the full-feature SVM-RBF model (overall F1 = 70.6%; ROC-AUC = 84.7%), including unseen disease types such as Clostridioides difficile infection (F1 = 90.3%) and type 2 diabetes (F1 = 77.4%). Feature-importance and multivariate analyses revealed both shared and disease-specific microbial signatures, suggesting that the model captures biologically meaningful patterns rather than cohort-specific artifacts. Disease-associated taxa included Klebsiella pneumoniae, Raoultella ornithinolytica, Sutterella wadsworthensis, Gemmiger formicilis, and Lactobacillus crispatus. In contrast, healthy status was consistently associated with commensal species such as Extibacter hylemonae and Ruthenibacterium lactatiformans. These results show that models trained on pooled metagenomes predict gut health status accurately and transfer to independent cohorts, providing a scalable, non-invasive framework and a set of candidate microbial biomarkers for further clinical evaluation.
Related Concept Videos
 01:22
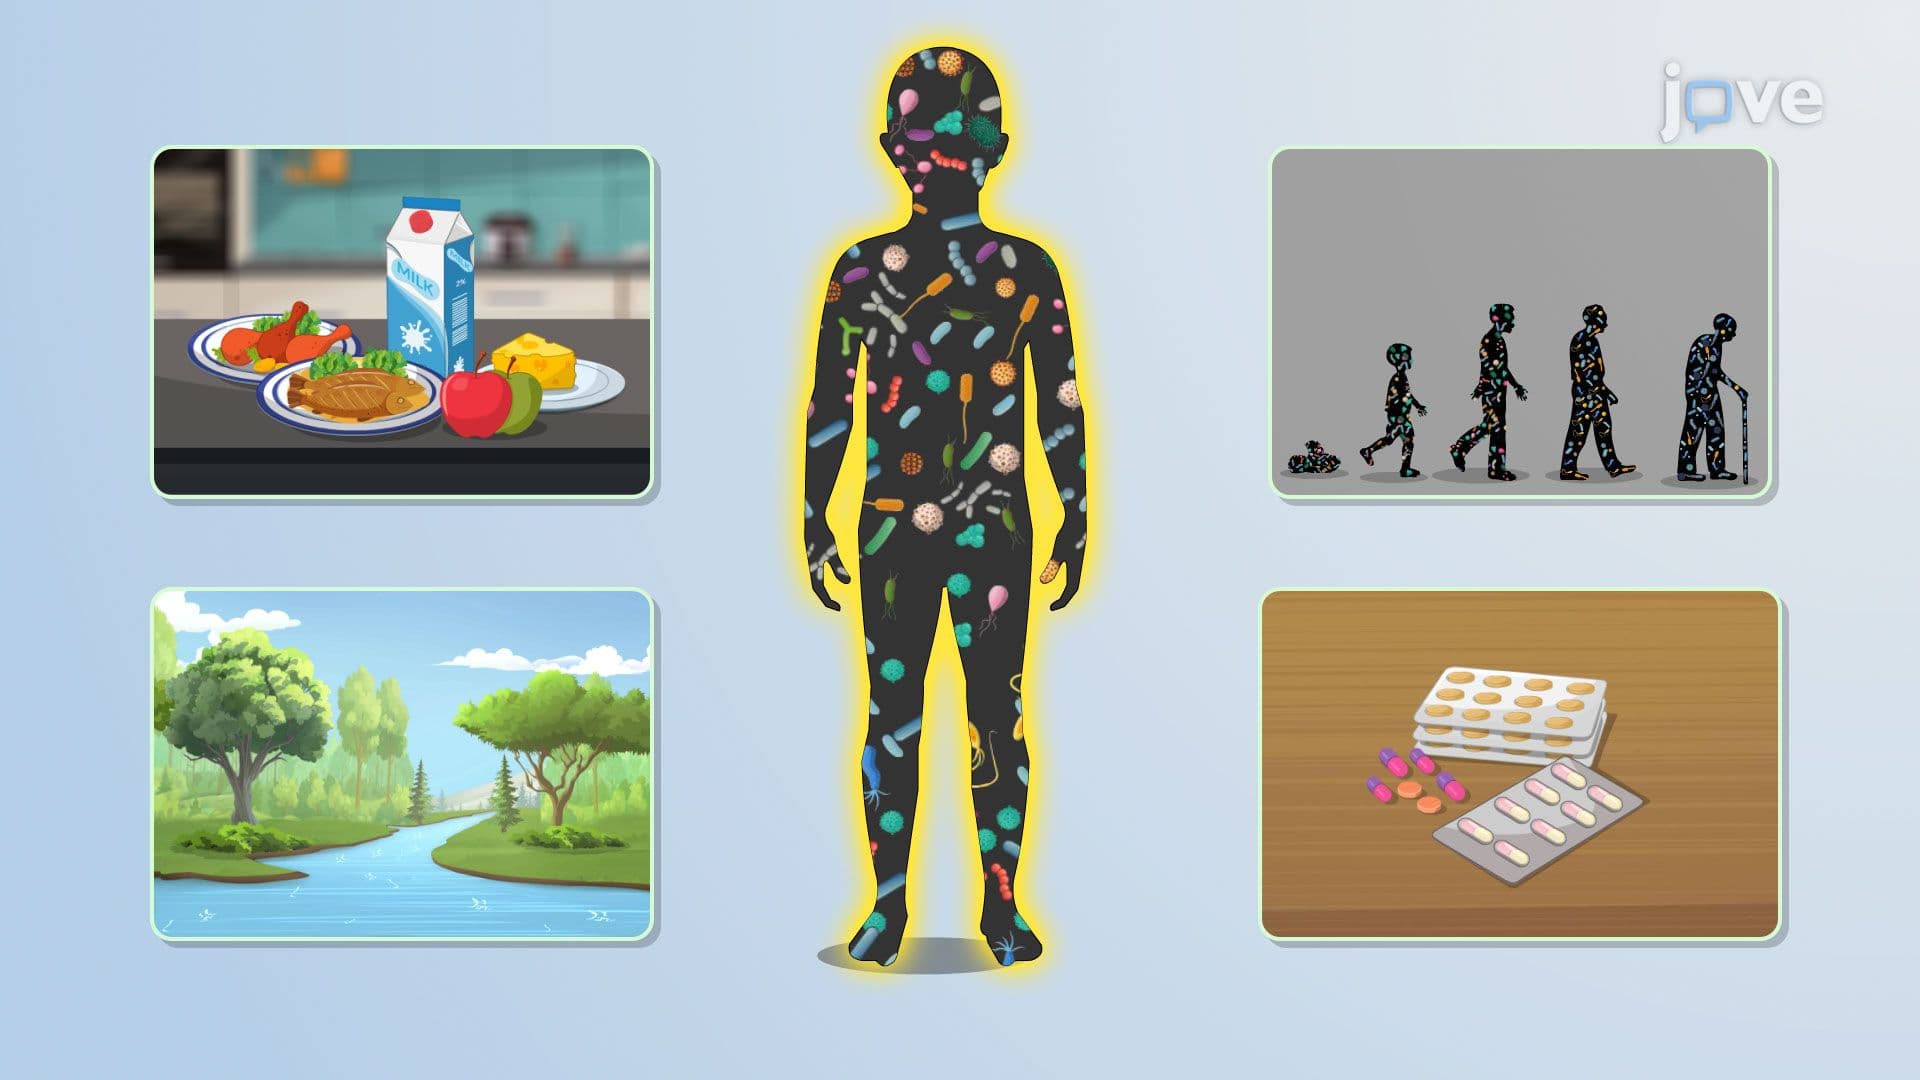
Introduction to the Human Microbiota
 01:19
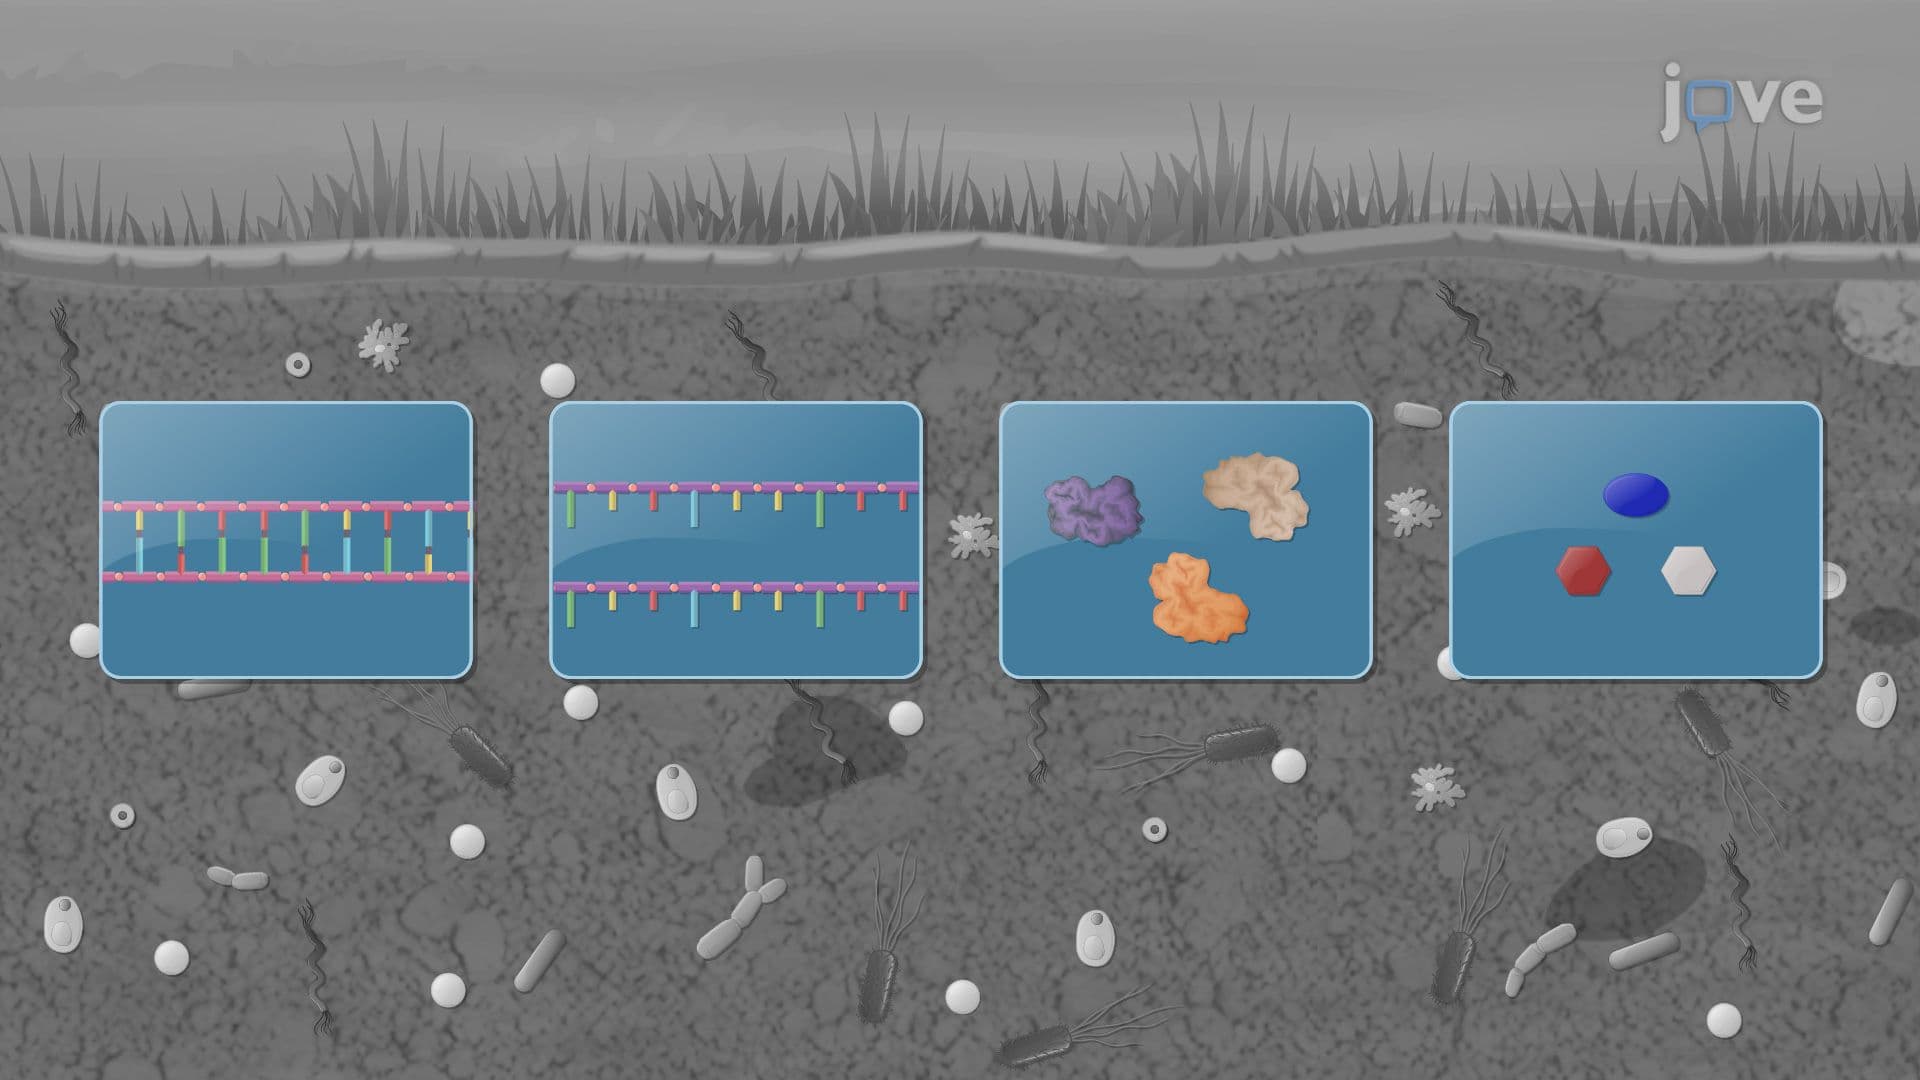
Methods to Assess Microbial Communities