

Photoacidity and ESIPT in luteolin: acidity-vibrational frequency relationships across electronic states
Siramol Photiganit1, Ananda Thongyu1, Pannipa Panajapo1
1School of Chemistry, Institute of Science, Suranaree University of Technology Nakhon Ratchasima 30000 Thailand kritsana@sut.ac.th.

Abstract:
The photochemical antioxidant properties of small organic molecules in electronically excited states have become an important topic in modern photochemistry and redox biology. Photoexcitation can trigger ultrafast proton and hydrogen transfer processes, leading to substantial changes in acidity and redox behavior. This study systematically explores the excited-state proton/hydrogen transfer pathways, acidity, and photophysical characteristics of luteolin (LU), a compound containing multiple potential deprotonation sites, using density functional theory (DFT) and time-dependent DFT (TD-DFT) calculations. Validation against previously reported theoretical and experimental data demonstrates that the employed DFT and TD-DFT/B3LYP/aug-cc-pVDZ methodology provides a reliable and consistent description of the molecular structure, energetics, and photophysical behavior of LU in both the ground (S0) and excited (S1) states. To evaluate acidity, the pK a and values are determined using three distinct yet complementary computational models: gas phase (isolated LU molecule), an implicit solvation model, and an explicit solvation model (LU-H2O hydrogen-bonded complexes). The close agreement among these models confirms that they can be used in a complementary manner to reliably investigate systems with multiple deprotonation sites, offering a comprehensive picture of acid-base properties across different environments. The DFT and TD-DFT calculations reveal that the intramolecular hydrogen bond and the associated excited-state proton transfer (ESIPT) involving the benzopyran-ring O-H group (LU-C5) directly influence the vibrational properties and acidity of the most acidic catecholic O-H group (LU-C4'). Additionally, vibrational analysis reveals that near-linear correlations between O-H stretching frequencies and both pK a and persist in all three models. To the best of our knowledge, these results provide the first evidence that such vibrational descriptors remain valid across different solvation environments and electronic states, offering a practical approach for predicting photoacidity. Overall, this study provides detailed insights into the excited-state deprotonation behavior and photophysical properties of LU while establishing transferable computational strategies for investigating excited-state proton transfer in complex photoactive systems.
Related Concept Videos
 01:32
UV–Vis Spectroscopy of Conjugated Systems
One of the factors influencing λmax is the extent of conjugation in the...
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions
 01:28
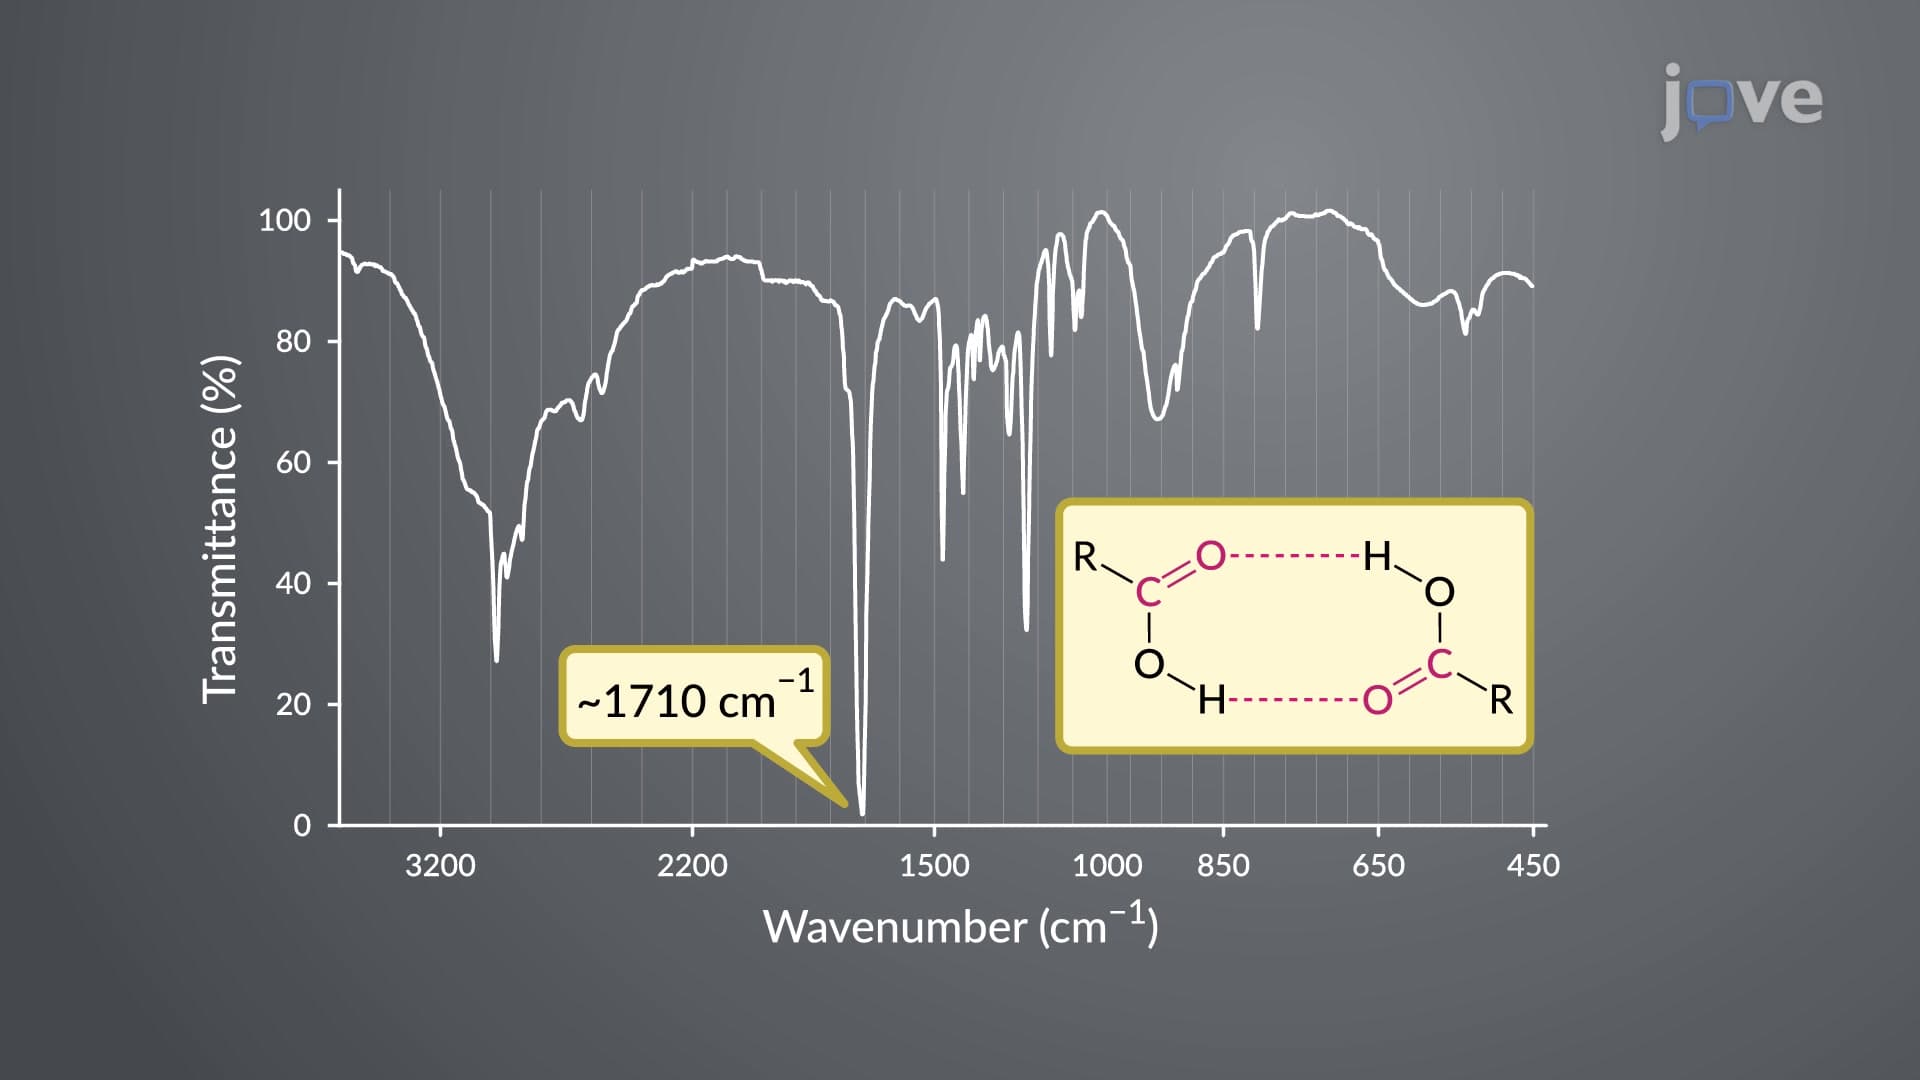
IR and UV–Vis Spectroscopy of Carboxylic Acids
However, the stretching absorptions for the C=O bond vary depending on the structure of carboxylic acids. The C=O bond of the free carboxylic acids shows a higher stretching frequency, 1760 cm−1, while H-bonded carboxylic acids (dimers) exhibit stretching absorptions at a lower frequency, 1710 cm−1. The C=O bond of the...
 01:30
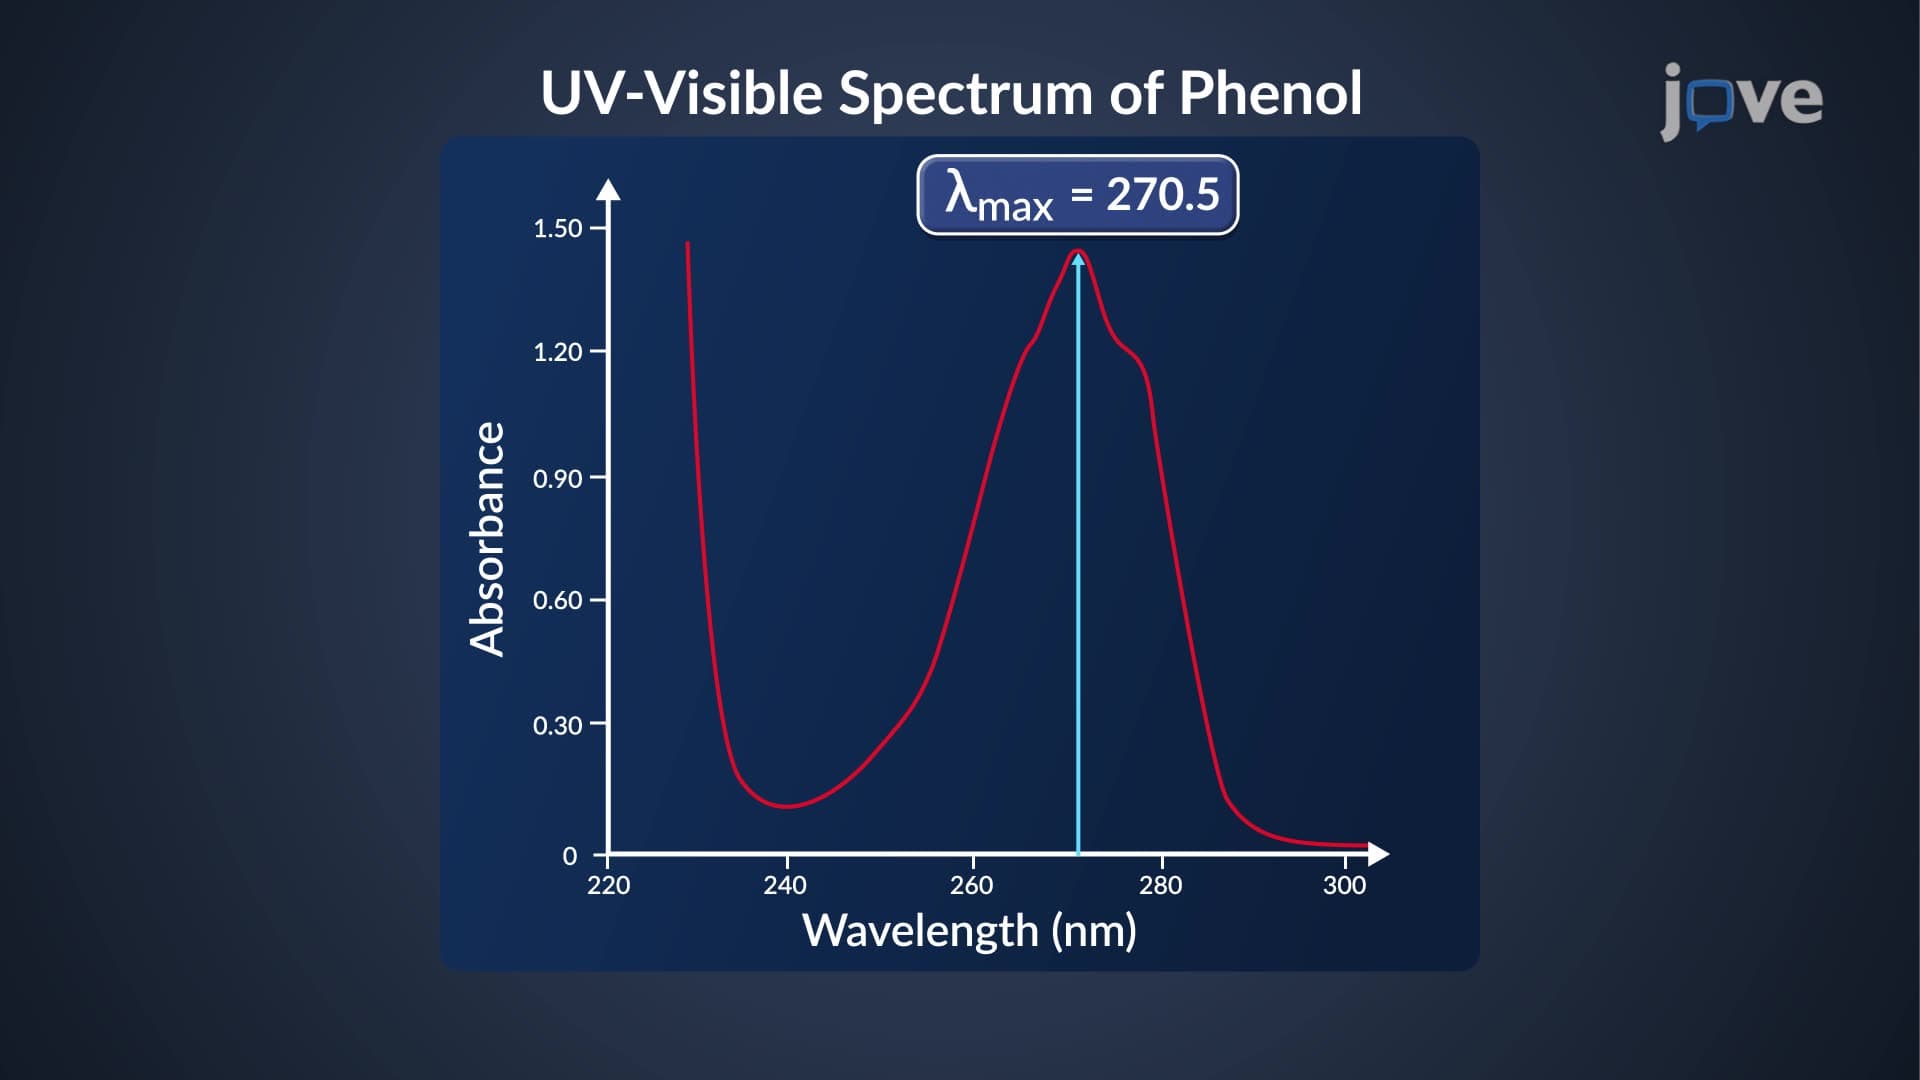
UV–Vis Spectrum
The UV–Vis spectrum of a molecule is the plot of its absorbance versus wavelength. The plot is drawn by taking molar absorptivity (ε) or log ε on the y-axis (ordinate)...
 01:14
Molecular Spectroscopy: Absorption and Emission
 01:26
Photochemical Electrocyclic Reactions: Stereochemistry
Selection Rules: Photochemical Activation