Related Experiment Video
Updated: Sep 27, 2026
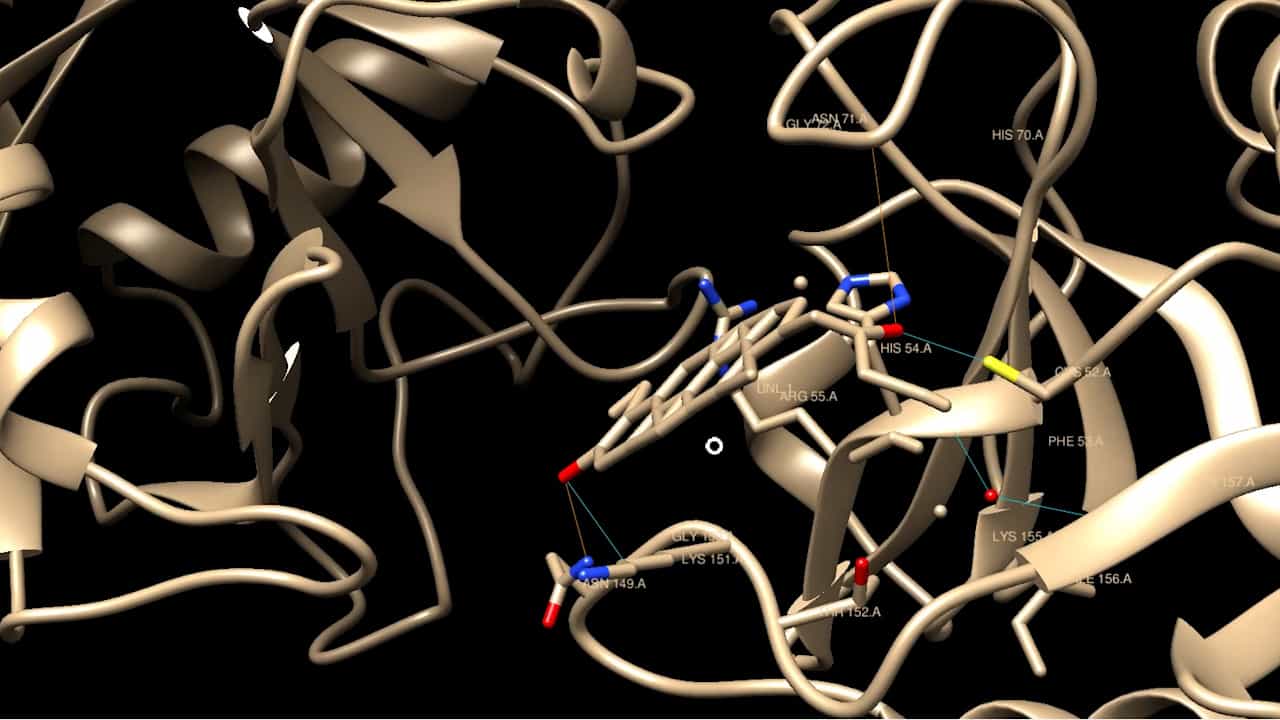

Structure-Guided Design and Development of Novel Cyclophilin A Inhibitors and Ganoderiol-F Derivatives: An In-Silico Approach
Published on: June 23, 2026
Cracking ERα Y537S Resistance: Explainable Machine Learning-Guided Discovery and Molecular Dynamics Validation of
1Department of Medical Laboratories, College of Applied Medical Sciences, Qassim University, Buraydah 51431, Saudi Arabia.

Abstract:
Background/Objectives: Endocrine resistance due to activating mutations in the estrogen receptor alpha (ERα), especially Y537S, is still a big challenge in the treatment of hormone receptor-positive breast cancer, and there is a need to develop new small-molecule inhibitors that can overcome ligand-independent receptor activation. We used an integrated machine learning and molecular dynamics (MD)-based virtual screening (VS) approach to identify potential computationally prioritized hits of ERα Y537S in this study. Methods: The dataset (~3353 compounds) was characterized using molecular fingerprints and graphically represented by PCA and t-SNE to show structurally different clusters linked to potency (pIC50). To guarantee robust downstream model training, the diagnosis of structural and influential outliers was completed through the application of rigorous data quality control methods, including the Williams plot, Mahalanobis distance, and Cook's distance diagnostics. This was followed by multiple benchmarks of machine learning classifiers (random forest, SVM, KNN, and CNN) for activity classification, yielding the highest AUC of 0.95 for the random forest classifier. Interpretable structure-activity insights were gained by the identification of key substructures of the fingerprint by SHAP and feature importance analysis that influence the predicted potency. Three lead candidates (Hit-1, Hit-2, and Hit-3) were screened and tested with a 200 ns all-atom MD simulation in comparison with a reference control to evaluate the binding stability. Comprehensive trajectory analysis, such as PCA, FEL, RDF, salt bridges, and DCCM, was completed following the structure-based identification of the most stable ligands. Results: Three candidates were identified as Hit-1, Hit-2, and Hit-3. Hit-3 was found to be bound in the most stable binding pose and had the most similar conformational behavior to the control, whereas Hit-2 showed transient pose instability associated with increased anti-correlated domain-level motions and an alternate high-salt-bridge conformational state. As no computational metric can be an absolute measure of experimental affinity, Hit-1 has the most favorable calculated end-point binding-energy estimate, while Hit-3 showed conformational stability in MD simulations. Conclusions: The computationally prioritized candidates can be subjected to further experimental testing, as the conducted docking, MD simulations, and end-point free energy analyses cannot establish overall compound binding, cell-based activity, pharmacological mechanism, or therapeutic efficacy.
More Related Videos

Related Concept Videos
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
02:40Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 02:18
The Equilibrium Binding Constant and Binding Strength
 00:49
Ligand Binding and Linkage
00:49Ligand Binding and Linkage
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...