Related Experiment Video
Updated: Sep 30, 2026
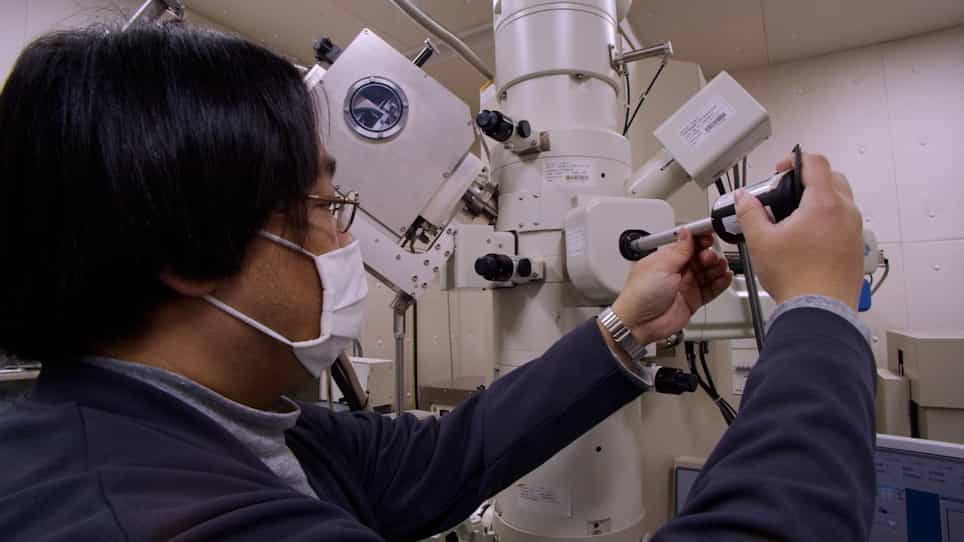

Quantitative Atomic-Site Analysis of Functional Dopants/Point Defects in Crystalline Materials by Electron-Channeling-Enhanced Microanalysis
Published on: May 10, 2021
defermi: A tool for computing and visualizing point-defect equilibria, bridging computational and experimental users
1Fachgebiet Materialmodellierung, Institut für Materialwissenschaft, TU Darmstadt, Otto-Berndt-Straße 3, D-64287 Darmstadt, Germany.

Abstract:
Computational methods provide a rigorous alternative to assumption-based defect chemistry for studying defect thermodynamics. However, existing tools for calculating and visualizing point-defect equilibria are predominantly designed around high-throughput density functional theory workflows, which can limit their accessibility to experimentally focused researchers. Here, we present defermi, a Python package with an integrated graphical user interface for analyzing point-defect equilibria. The interface enables efficient and reproducible workflows for plotting defect formation energies and charge transition levels and for computing Brouwer and doping diagrams. In parallel, the Python API provides the same high-level functionality while allowing advanced customization, including user-defined routines for defect formation energies and defect concentrations. This flexibility enables extensions beyond standard ab initio approaches, such as the inclusion of temperature and volume dependence in defect formation energies. By combining an accessible graphical interface with a customizable programmatic framework, defermi connects computational defect thermodynamics with experimentally oriented defect chemistry.
Related Concept Videos
 01:26
Imperfections in Crystal Structure: Stoichiometric Point Defects
 01:25
Imperfections in Crystal Structure: Point, Line and Plane Defects