Related Experiment Video
Updated: Oct 1, 2026

Guided Protocol for Fecal Microbial Characterization by 16S rRNA-Amplicon Sequencing
Published on: March 19, 2018
Comparative gut microbiome in diarrheal and non-diarrheal children: an individually matched case-control study
Jialiang Xu1, Yufeng Fan2, Geruo Qu1,3
1School of Light Industry Science and Engineering, Beijing Technology and Business University, Beijing, China.

Abstract:
Diarrhea, a leading cause of under-five mortality in developing countries, drives therapeutic challenges amid global antibiotic resistance. This study aimed to compare gut microbiota, antibiotic resistance genes (ARGs), mobile genetic elements (MGEs), and virulence factors (VFs) in diarrheal and non-diarrheal children, explore reasons for asymptomatic pathogen carriage, and develop a microbiome-based model to predict the potential etiology of diarrhea. This individually matched case-control study analyzed 42 paired fecal samples (selected from 716 diarrhea/non-diarrhea cases) to compare gut microbiome profiles, ARGs, MGEs, and VFs using metagenomic sequencing and pathogen-specific PCR. Diarrheal children showed reduced alpha diversity, increased Proteobacteria, and elevated VFs and MGEs. Surprisingly, ARGs were more abundant in non-diarrheal children, suggesting ARG colonization in healthy hosts. Actinomyces was enriched in non-diarrheal pathogen carriers and may be associated with asymptomatic bacterial pathogen carriage. A random forest (RF) model incorporating non-pathogenic bacteria achieved high accuracy in predicting diarrheal status and pathogen carriage. This study reveals distinct microbial ecologies between diarrheal and healthy children. The enrichment of specific bacterial taxa in asymptomatic carriers may be associated with pathogen tolerance. The unexpected ARG abundance in healthy children highlights a hidden antimicrobial resistance reservoir. These findings inform microbiome-based diagnostics and antibiotic stewardship in pediatric diarrhea.IMPORTANCEThis study is significant because it provides robust, individually matched case-control evidence linking childhood diarrhea to gut microbiome dysbiosis and the distribution of pathogenic and resistance-related genetic elements. By integrating metagenomic sequencing with pathogen-specific polymerase chain reaction (PCR), it offers a comprehensive comparison of microbial composition, virulence factors (VFs), mobile genetic elements (MHEs), and antibiotic resistance genes (ARGs) between diarrheal and non-diarrheal children. The findings demonstrate reduced microbial diversity and enrichment of Proteobacteria, virulence factors, and mobile genetic elements in diarrheal cases, highlighting microbiome instability during infection. Importantly, the unexpectedly higher abundance of antibiotic resistance genes in non-diarrheal children underscores the underestimated role of healthy populations as reservoirs of resistance. Furthermore, the establishment of predictive models for diarrhea status and pathogen carriage enhances the translational value of the study. Overall, this work advances the understanding of pediatric diarrheal disease and informs prevention, surveillance, and treatment strategies in the context of global antimicrobial resistance.
Related Concept Videos
 01:30
Development of Human Microbiota
 01:21
Microbiota Modulation by Antibiotics
 01:53
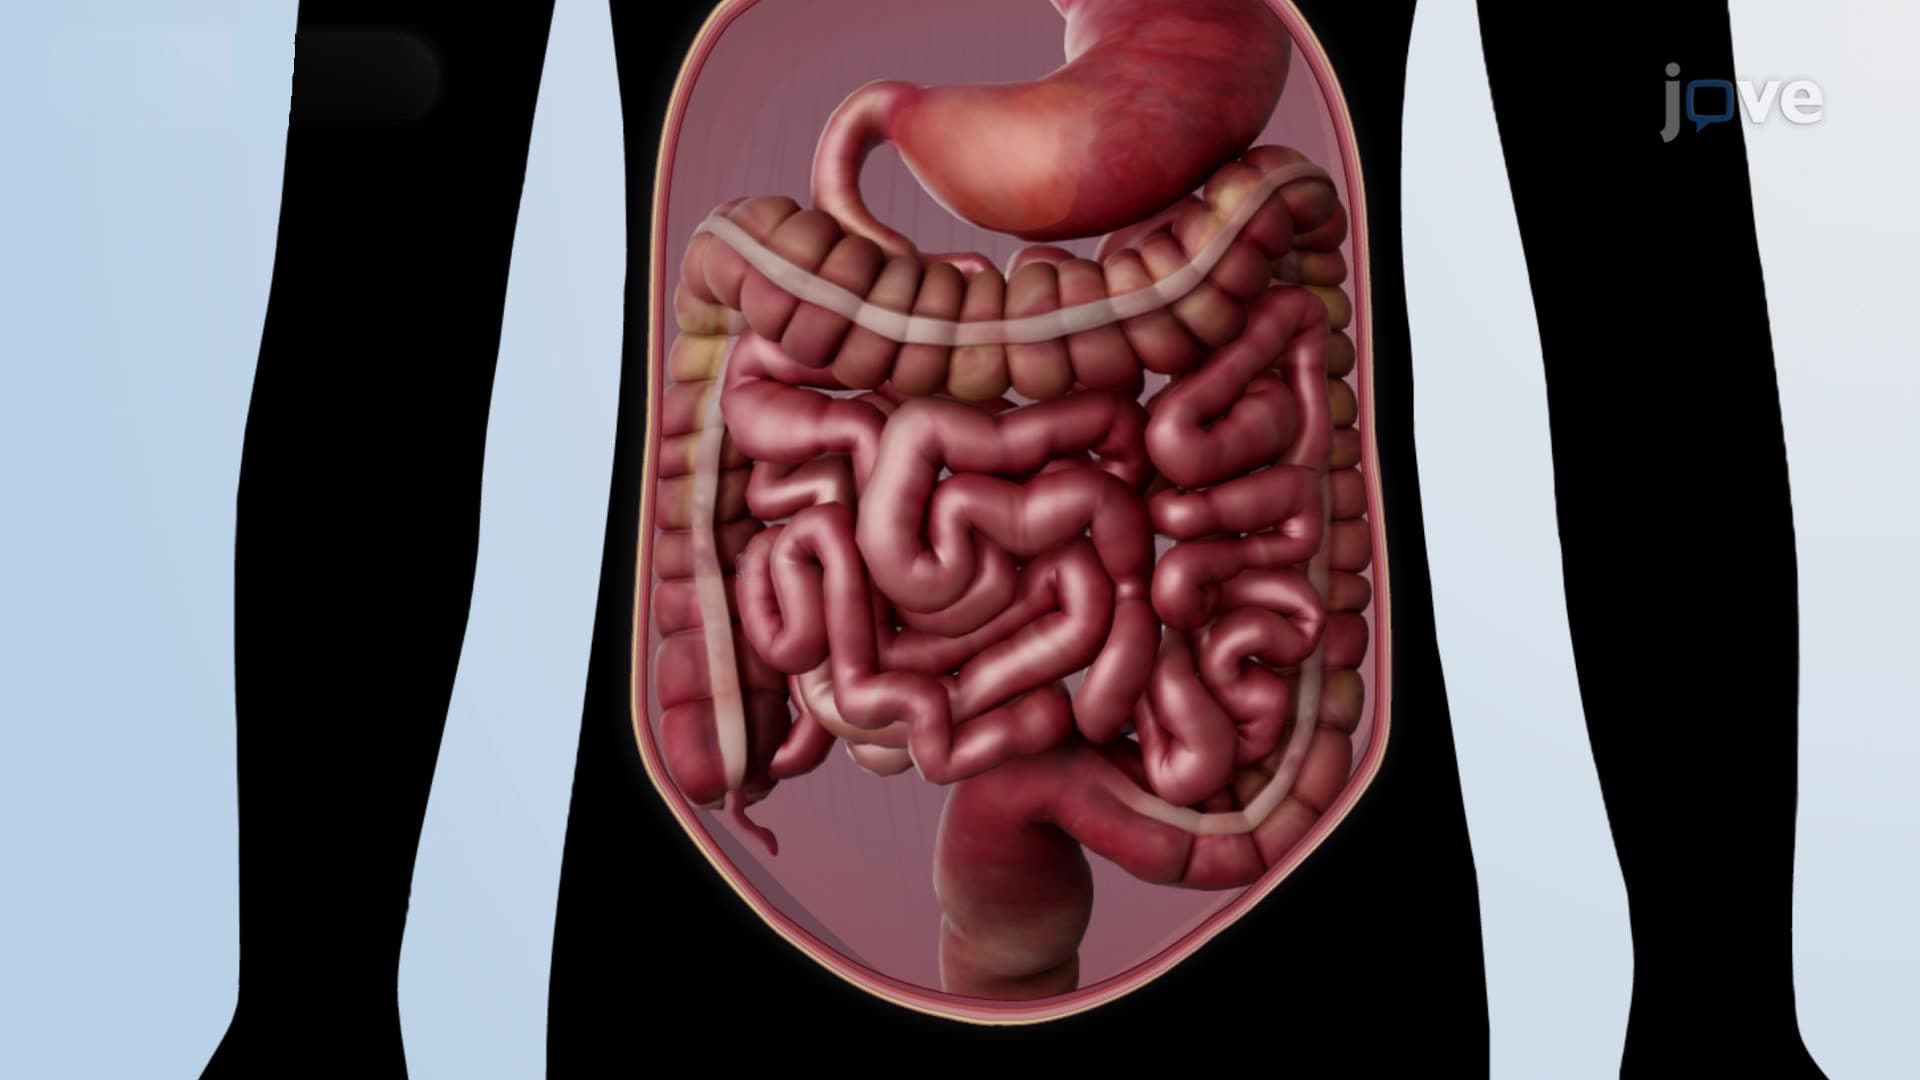
Anatomy of the Intestines
 01:27
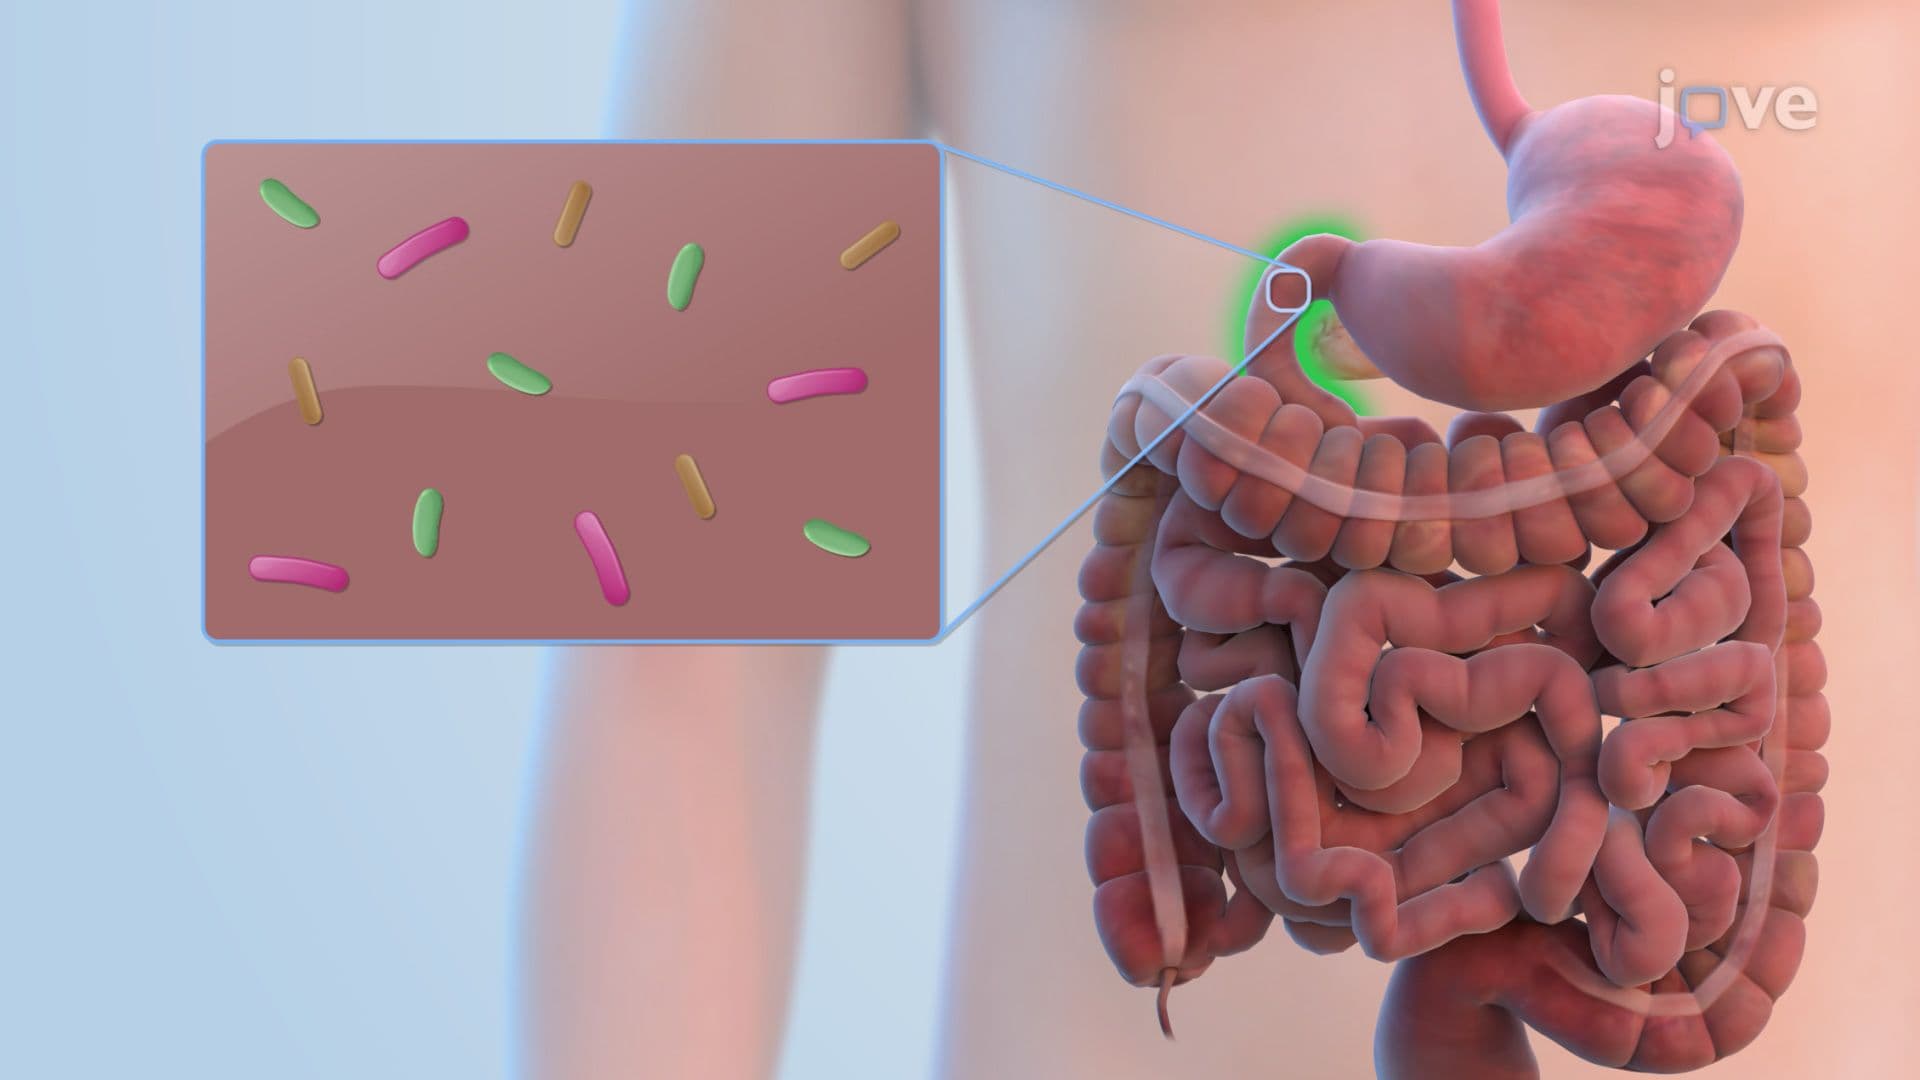
Microbiota of the Stomach and Small Intestine
 01:22
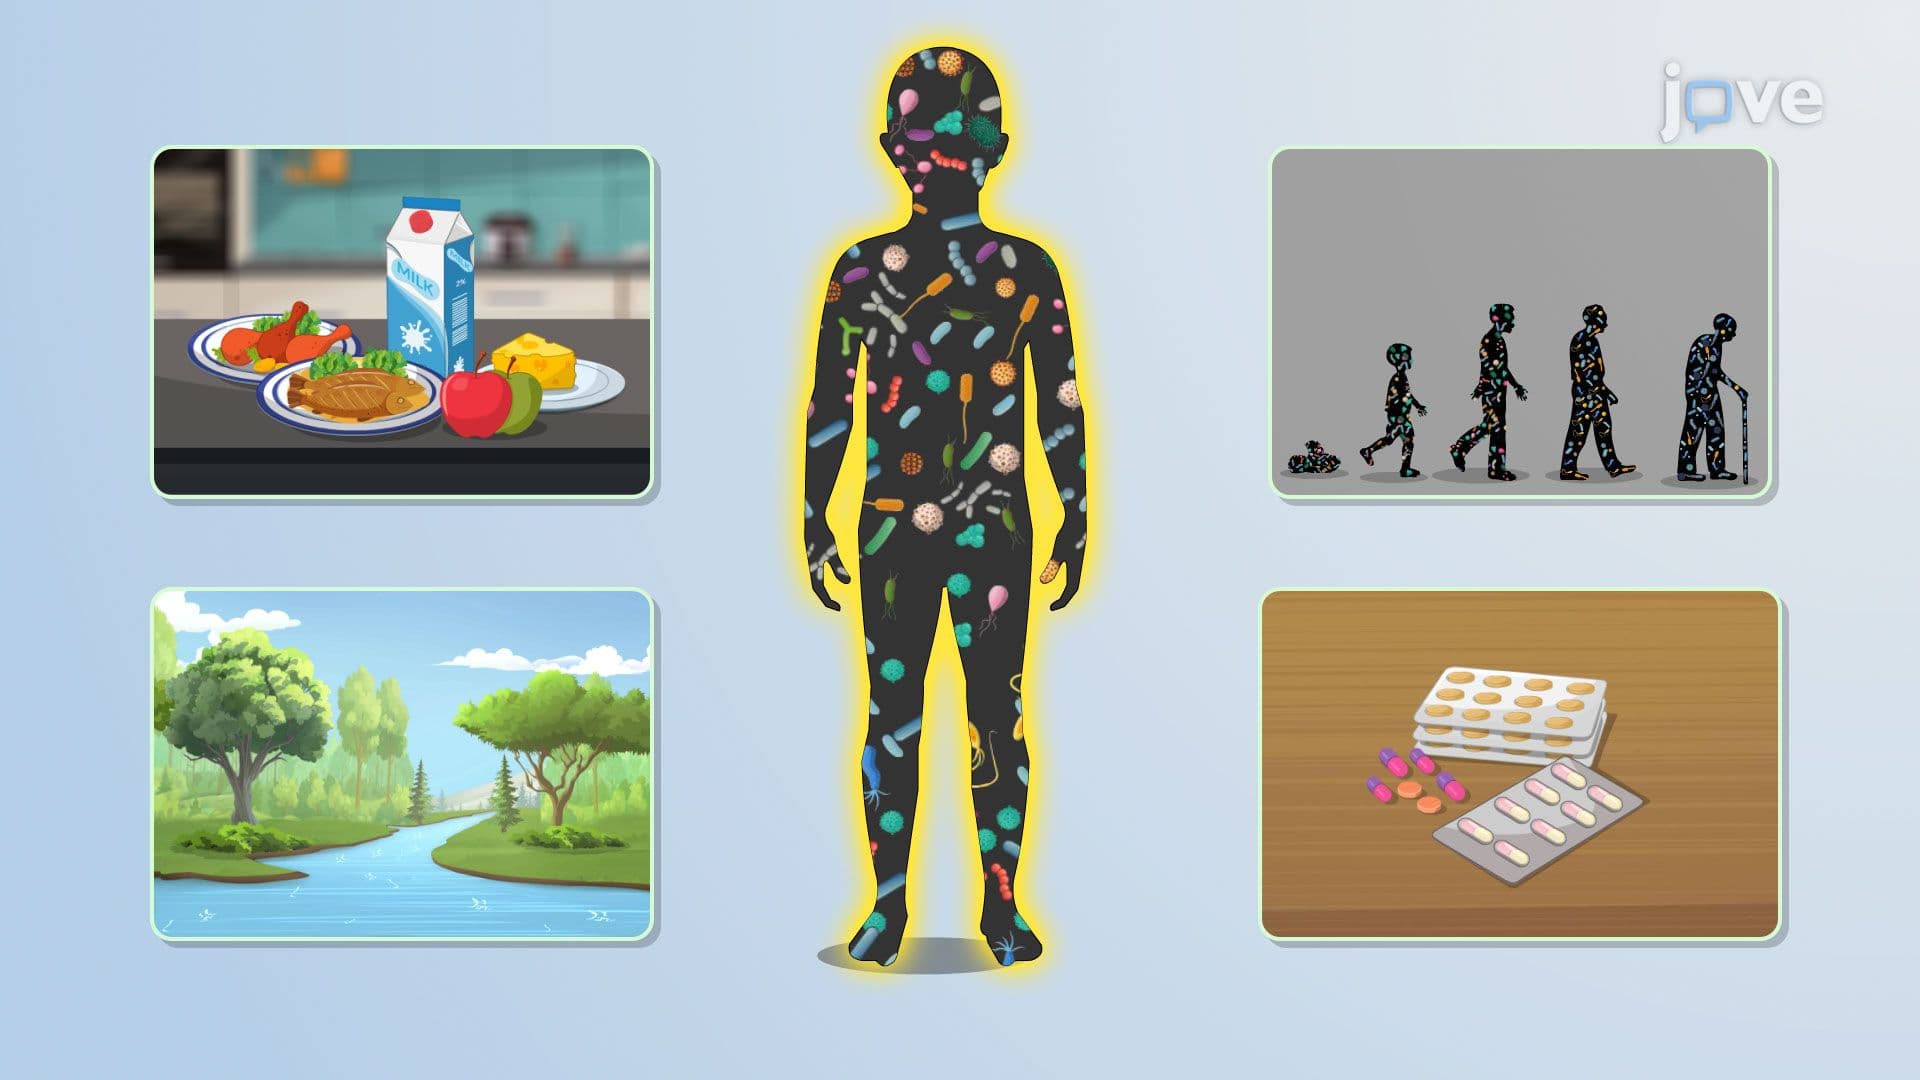
Introduction to the Human Microbiota
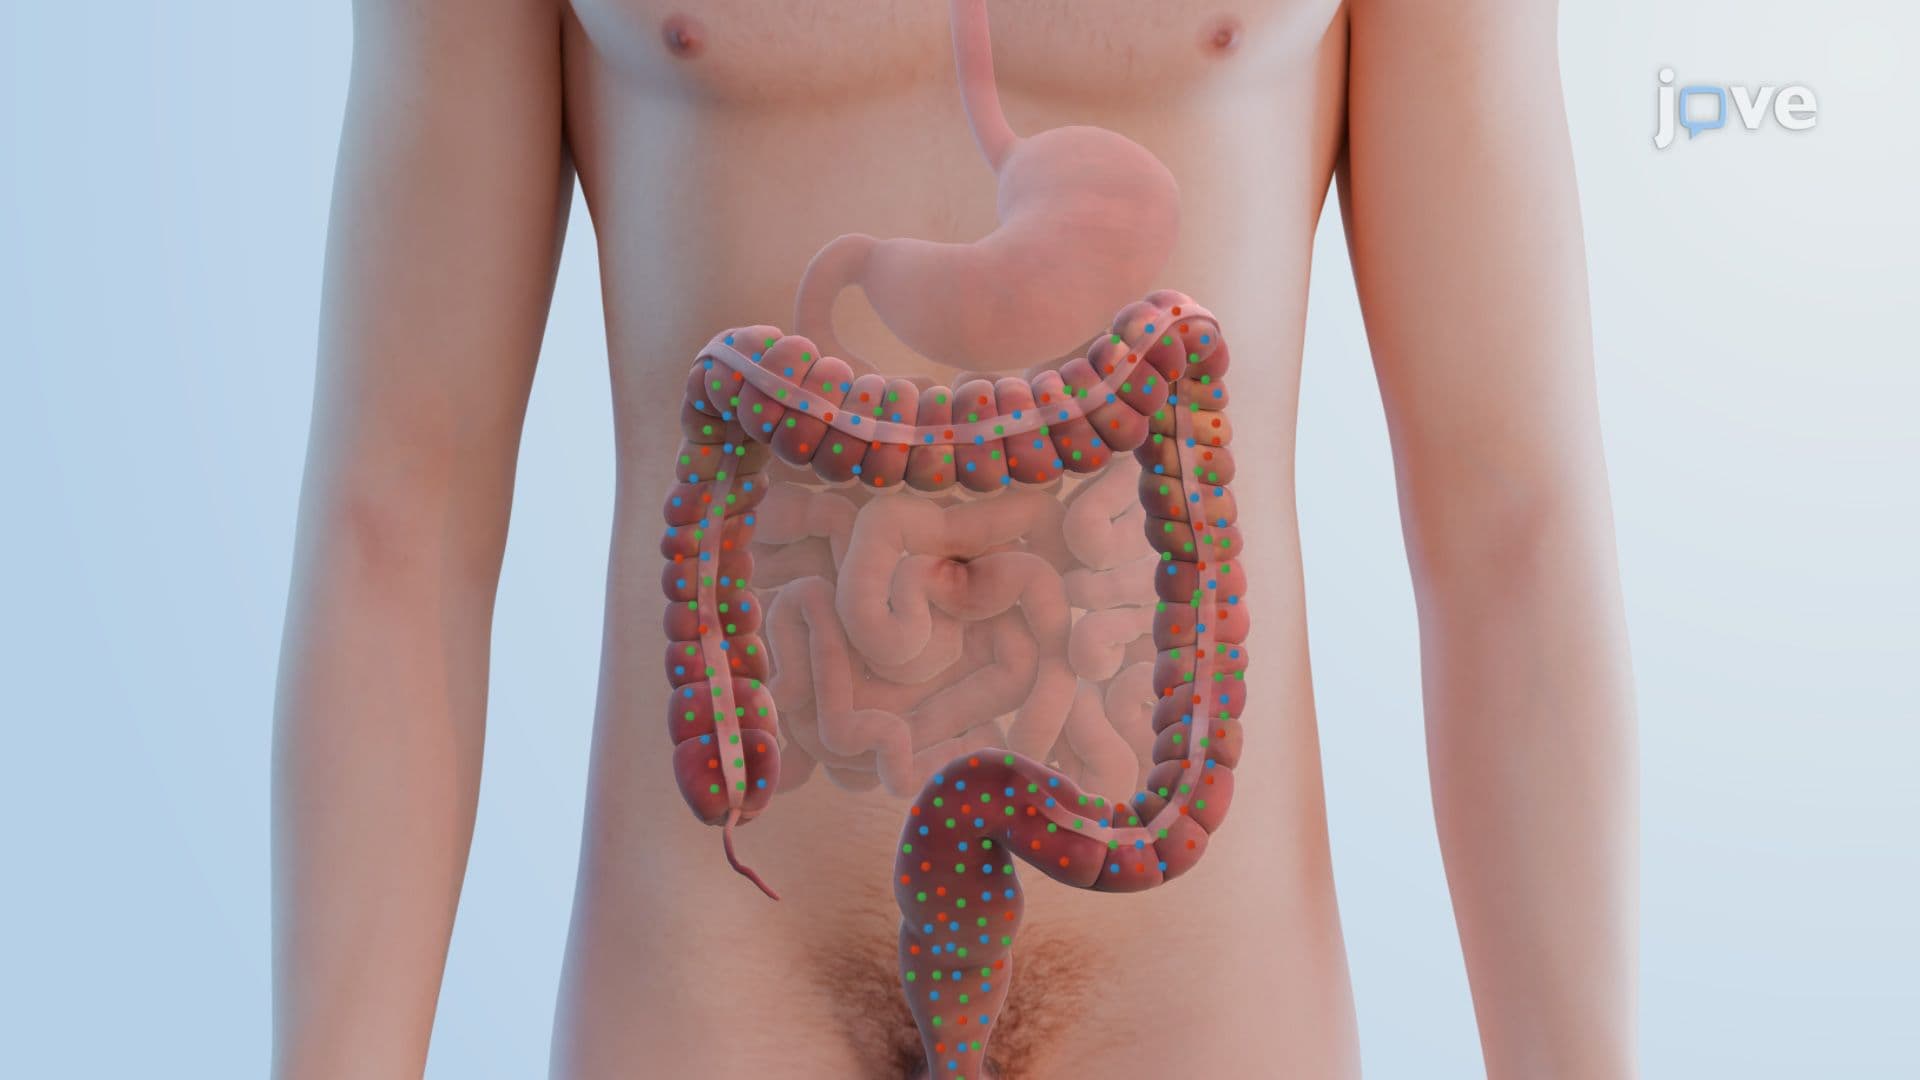 01:27
Microbiota of the Large Intestine