Related Experiment Video
Updated: Oct 2, 2026

On-line Analysis of Nitrogen Containing Compounds in Complex Hydrocarbon Matrixes
Published on: August 5, 2016
Substituent effects on the thermal stability and initial decomposition of methylene bis(dinitropyrazole) derivatives
Yuqin Chu1, Peng Zhang1, Yang Zhu1
1College of Safety Science and Engineering, Nanjing Tech University, Nanjing, 210009, China.

Context:
Elucidating the role of functional-group substitution in the thermal stability of energetic materials is of considerable importance for the rational development of advanced energetic compounds. In this study, density functional theory (DFT) calculations, together with electronic-structure, weak-interaction, crystal-packing, and transition-state analyses, were performed to systematically explore the influence of amino (A), hydroxyl (B), and azido (C) substituents on methylene-bridged bis(3,5-dinitropyrazole) (MBDNP) derivatives. The results reveal that functional-group substitution induces distinct electronic redistribution, leading to significant differences in key bond stability, intra- and intermolecular interactions, and crystal packing. Among the three derivatives, A exhibits the strongest electron delocalization, the most favorable interaction network, and the strongest intermolecular interactions, whereas C shows the weakest molecular and crystal stability. Transition-state calculations further demonstrate that different substituents alter the preferred initial decomposition pathways, and the calculated activation free energies follow the order A > B > C, consistent with the experimentally observed decomposition temperatures. These findings demonstrate that the thermal stability of MBDNP derivatives is governed by the synergistic effects of electronic structure, molecular interactions, crystal packing, and initial decomposition kinetics, providing molecular-level insights into the substituent-dependent thermal decomposition mechanism of energetic materials.
Methods:
Density functional theory (DFT) calculations at the M06-2X/def2-TZVPP level were performed to investigate the electronic structures and initial decomposition behavior of compounds A-C. Natural bond orbital (NBO), electrostatic potential (ESP), localized bond order (LBO), interaction region indicator (IRI), and quantum theory of atoms in molecules (QTAIM) analyses, together with Hirshfeld surface, energy framework, and transition-state analyses, were further performed to clarify how different substituents influence the electronic characteristics, intermolecular interactions, crystal-packing behavior, and thermal stability of the studied compounds.
Related Concept Videos
 01:18
Directing and Steric Effects in Disubstituted Benzene Derivatives
 01:13
meta-Directing Deactivators: –NO2, –CN, –CHO, –CO2R, –COR, –CO2H
 02:30
Stability of Substituted Cyclohexanes
The two chair conformations of cyclohexanes undergo rapid interconversion at room temperature. Both forms have identical energies and stabilities, each comprising equal amounts of the equilibrium mixture. Replacing a hydrogen atom with a functional group makes the two conformations energetically non-equivalent.
For example, in...
 01:09
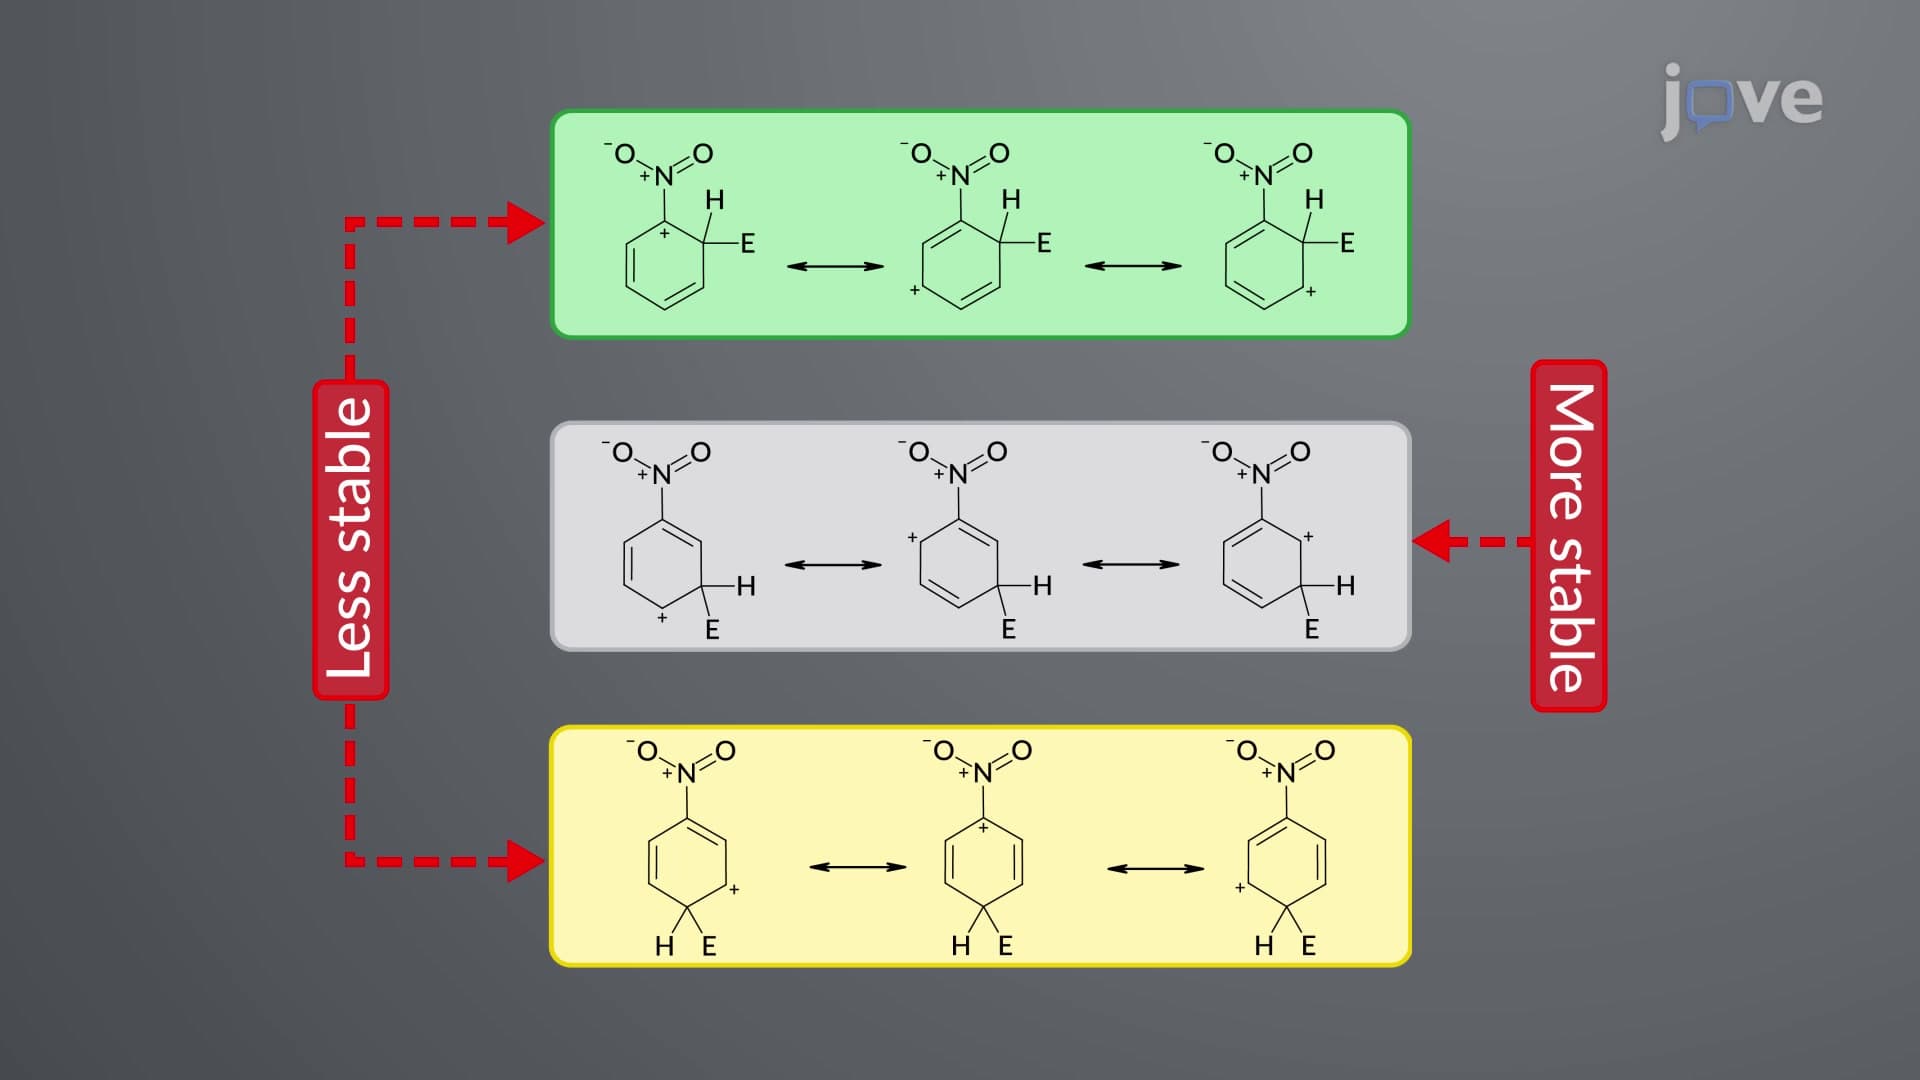
Directing Effect of Substituents: meta-Directing Groups
 01:11
ortho–para-Directing Activators: –CH3, –OH, –NH2, –OCH3
 01:14
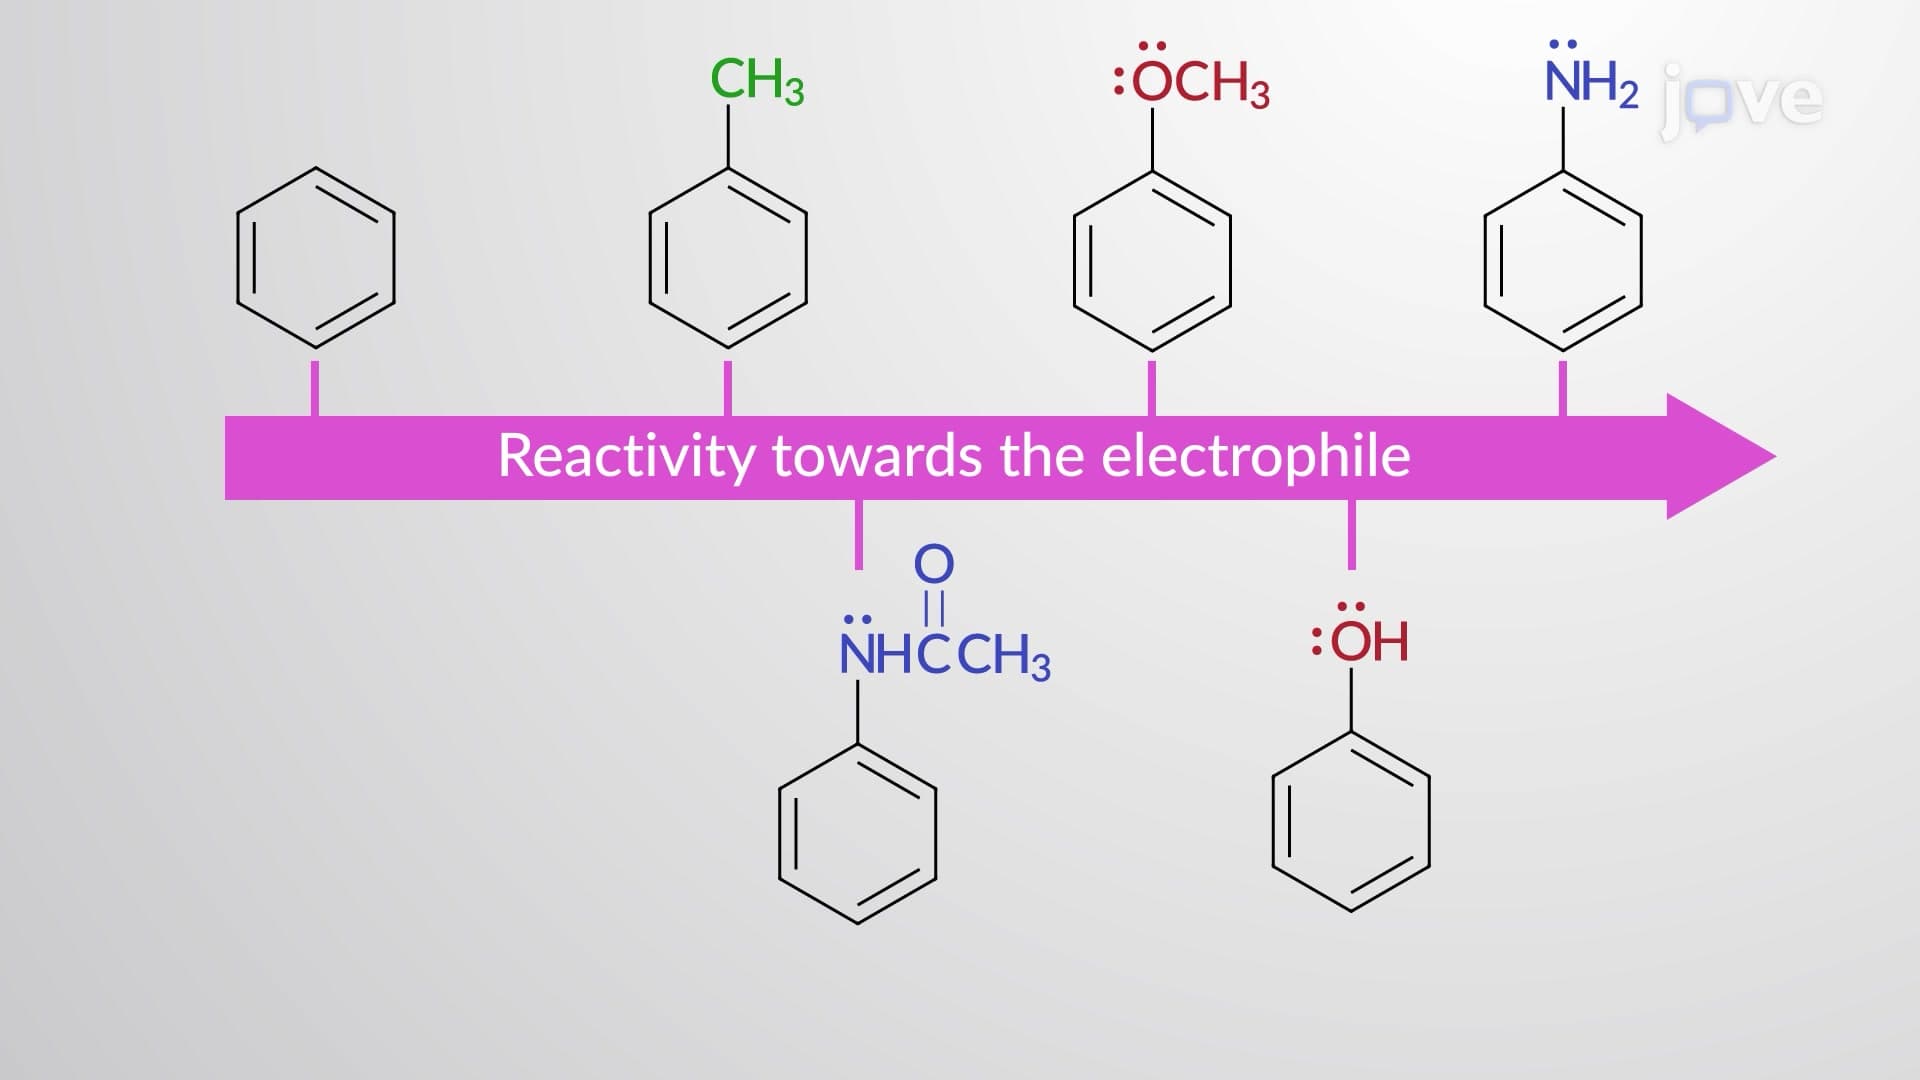
Directing Effect of Substituents: ortho–para-Directing Groups