Related Experiment Video
Updated: Oct 3, 2026


Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Accurate electron affinities from anion HOMO energies within the self-consistent random phase approximation
Egor Trushin1,2, Raviraj Mandalia1, Andreas Görling1,2
1Lehrstuhl für Theoretische Chemie, Universität Erlangen-Nürnberg, Egerlandstr. 3, D-91058 Erlangen, Germany.

Abstract:
Density functional calculations of small anions using the self-consistent random phase approximation (scRPA) optimized effective potential method are presented. scRPA calculations provide bound anions with the negative of the highest occupied molecular orbital (HOMO) energy closely matching the electron affinity of the corresponding neutral systems. For a benchmark of small atomic and molecular anions, electron affinities calculated as the negatives of HOMO energies from scRPA calculations are substantially more accurate than those from calculations with local, semilocal, global hybrid, and even range-separated hybrid exchange-correlation functionals, such as the well-established LC-ωPBE functional considered as an example. The superior performance of the scRPA method compared to conventional Kohn-Sham (KS) approaches lies in its ability to provide a closer approximation to the exact KS potential. Specifically, scRPA exchange-correlation potentials exhibit the correct -1/r asymptotic behavior and closely resemble the exact exchange-correlation potential in the molecular region. In addition to the standard scRPA method, we also explored the hybrid scRPA (hscRPA) approach, which incorporates a fraction of nonlocal Hartree-Fock exchange. The hscRPA method addresses convergence challenges that may arise in standard scRPA calculations associated with a vanishing HOMO-LUMO gap. At the same time, hscRPA also provides bound anions with accurate HOMO energies.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Related Concept Videos
 03:07
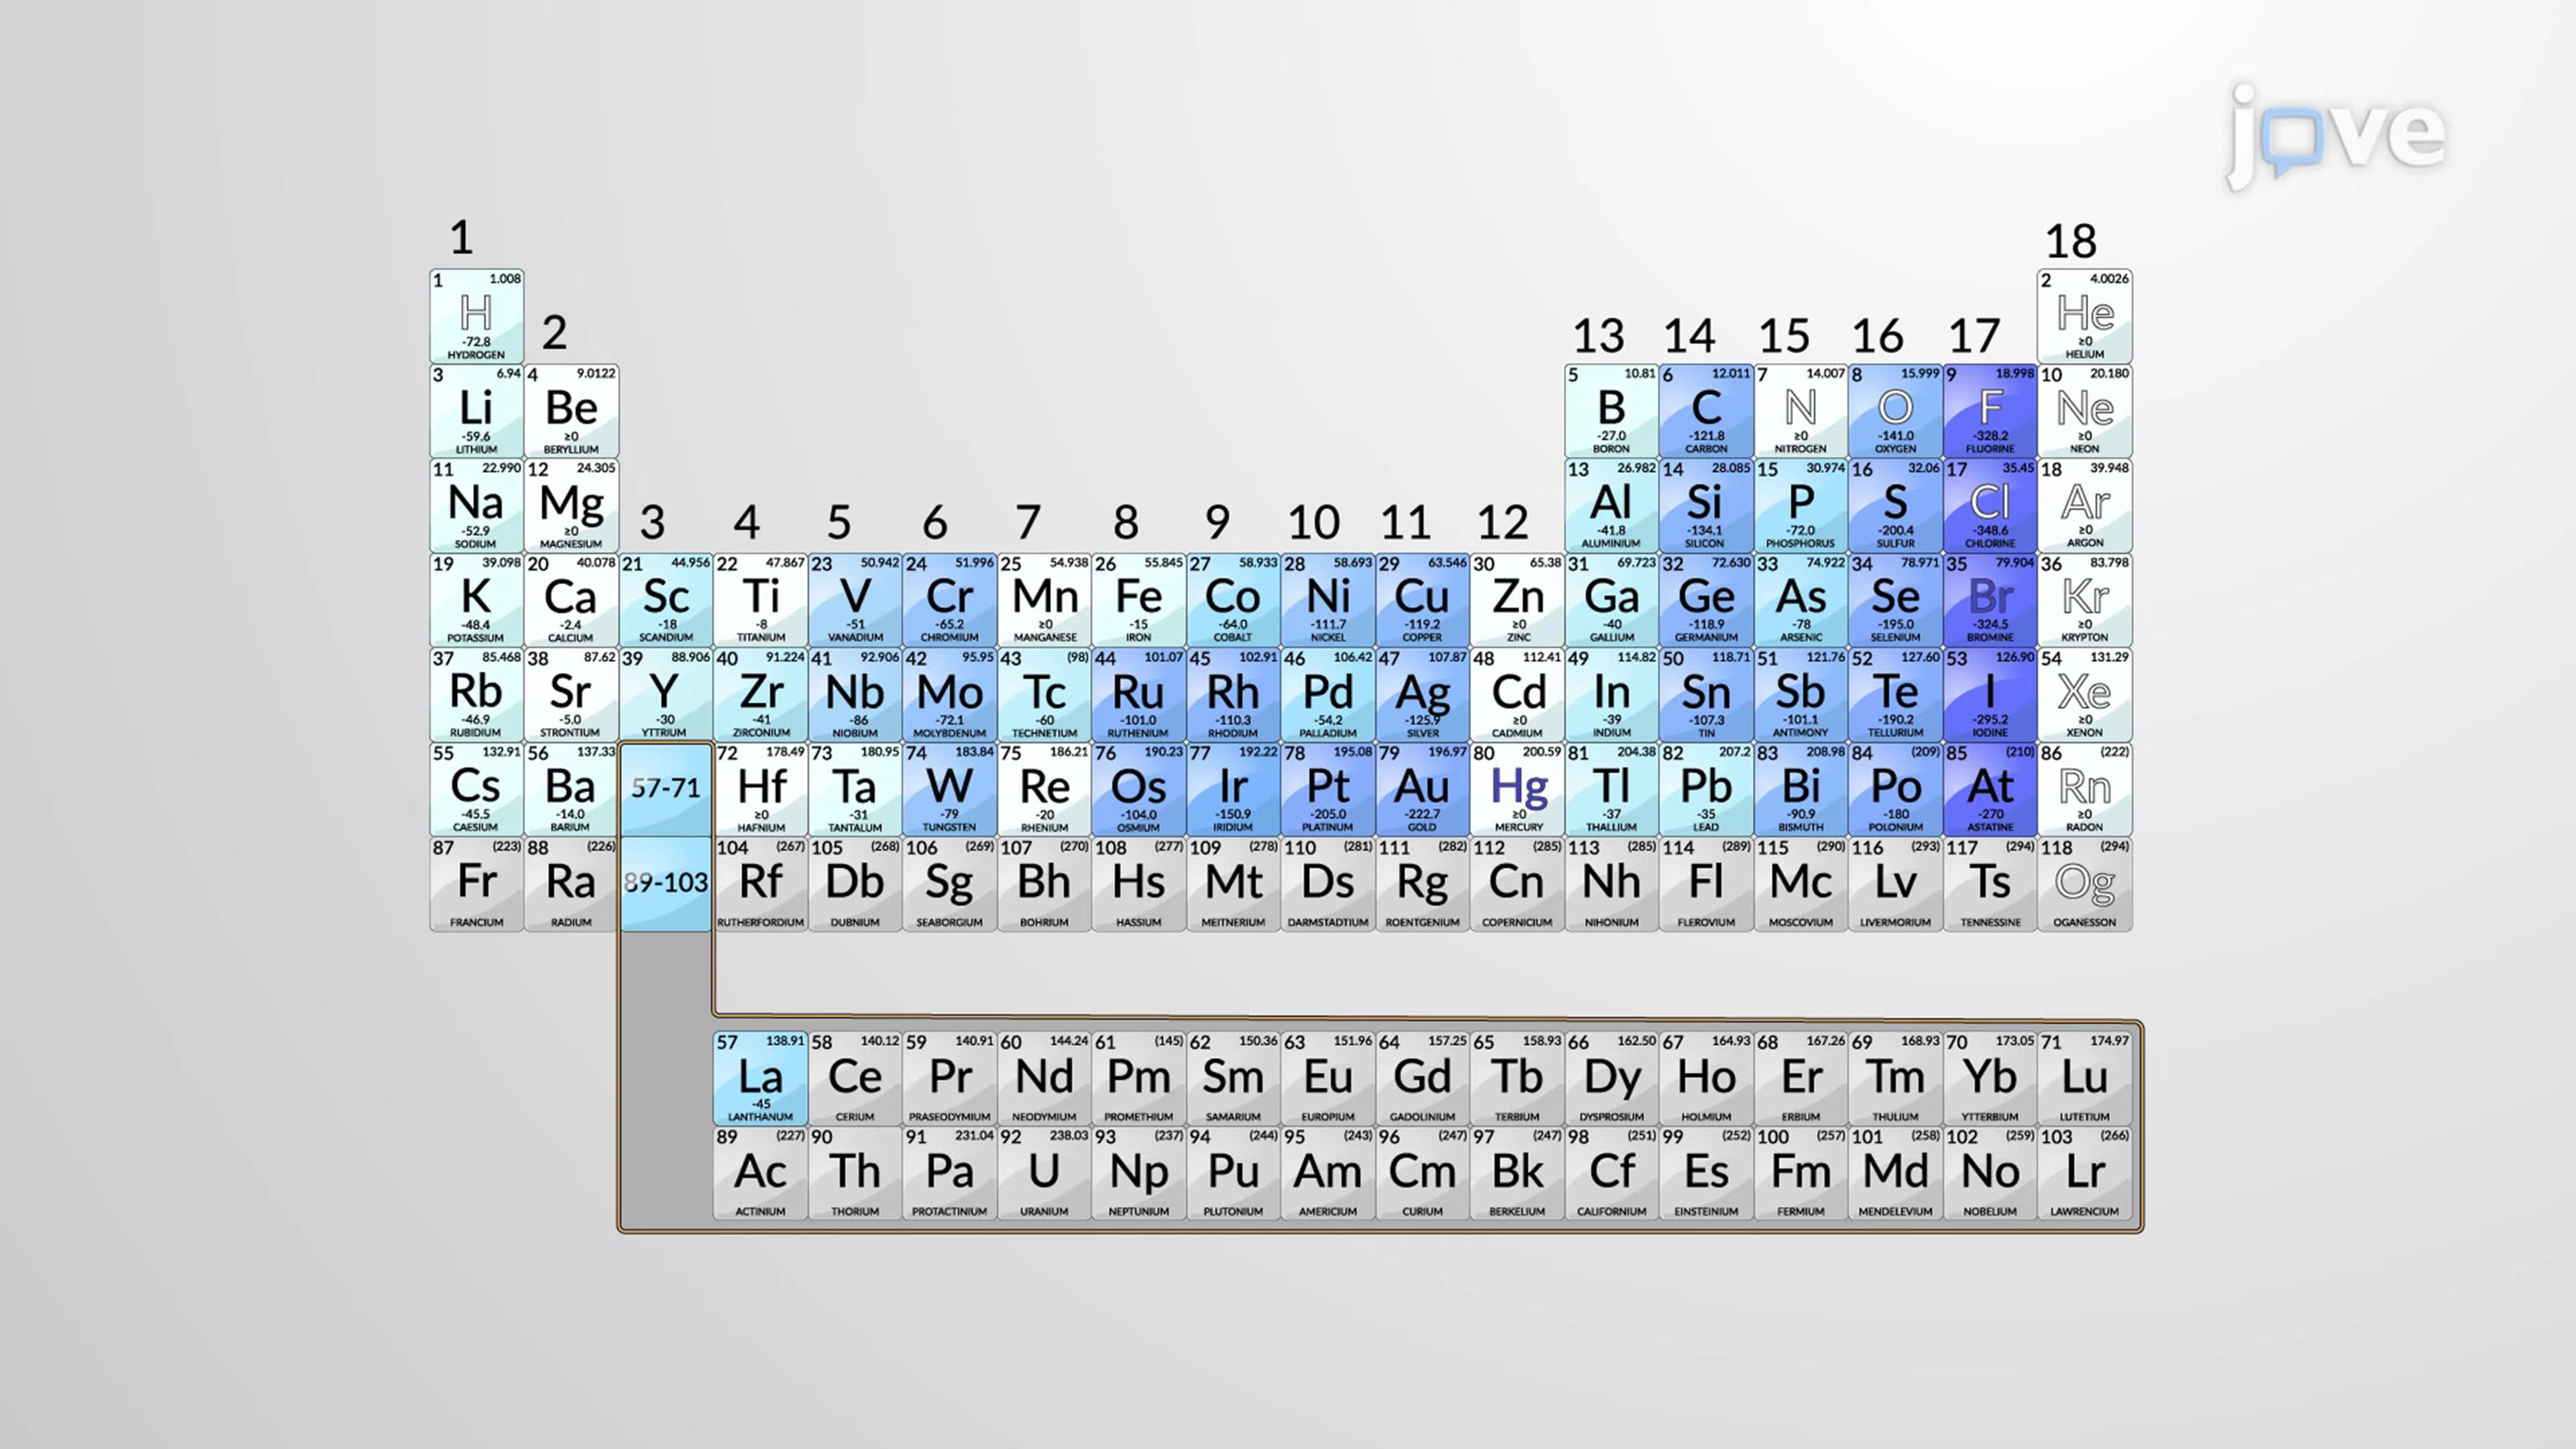
Electron Affinity
 03:21
The Energies of Atomic Orbitals
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 03:12
Ionization Energy
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:28
Ionic Association