Related Experiment Video
Updated: Oct 6, 2026
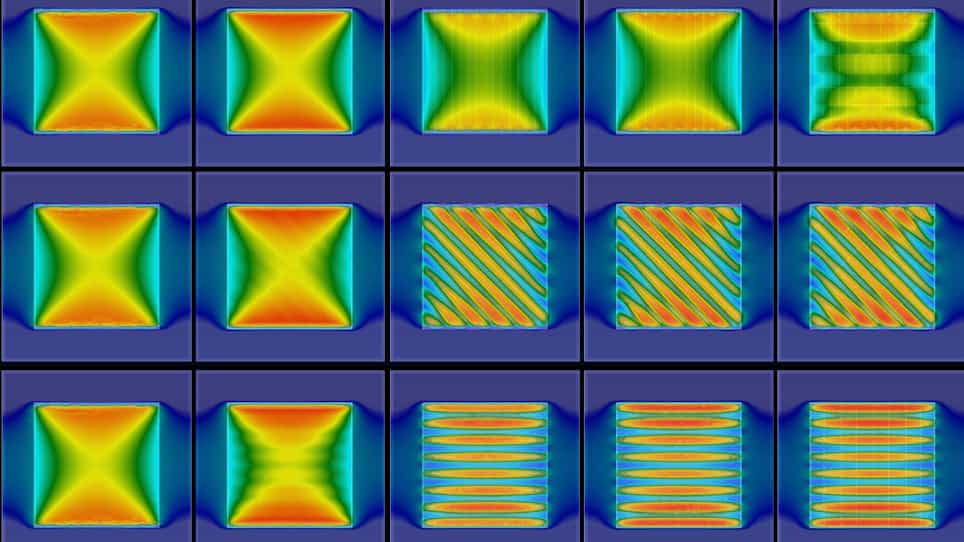

Finite Element Modelling of a Cellular Electric Microenvironment
Published on: May 18, 2021
Electronic origin of ion-specific structural responses: The role of charge density and self-interaction error
Weiyu Li1, Yankai Ding1, Zhaoru Sun1
1School of Physical Science and Technology, ShanghaiTech University, 393 Middle Huaxia Road, Shanghai, China.

Abstract:
Ab initio molecular dynamics provides a first-principles framework for aqueous systems but is limited in accuracy by self-interaction error (SIE) in exchange-correlation functionals. While SIE corrections are known to reduce the over-structured hydrogen bonding in bulk water, their impact on ion hydration remains unclear. Here, we employ machine learning molecular dynamics to systematically compare the Perdew-Burke-Ernzerhof functional with Tkatchenko-Scheffler-van der Waals dispersion correction and the strongly constrained and appropriately normed (SCAN) functional, thereby investigating the effects of different treatments of exchange-correlation (including SIE) on the hydration structure and electronic properties of Na+ and Cl- in aqueous solutions. We find that the SCAN functional yields a more ordered and compact first hydration shell of Na+, while leaving the orientational distribution of water molecules largely unchanged. In contrast, for Cl-, the Cl-water correlation of the first hydration shell remains nearly unchanged, whereas the orientational distribution of surrounding water molecules is significantly modified. Charge density analysis reveals that these differences originate from ion-dependent electronic responses: the charge density is more localized around Na+ under the SCAN description, strengthening Na+-water interactions, whereas Cl- is comparatively less affected. These results reveal that ion-specific hydration responses are governed by electronic structure effects, particularly charge-density differences associated with reduced SIE, and highlight the importance of accurate treatment of electronic structure in aqueous simulations.
More Related Videos
 11:04
Ion Mobility-Mass Spectrometry Techniques for Determining the Structure and Mechanisms of Metal Ion Recognition and Redox Activity of Metal Binding Oligopeptides
Published on: September 7, 2019
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 01:30
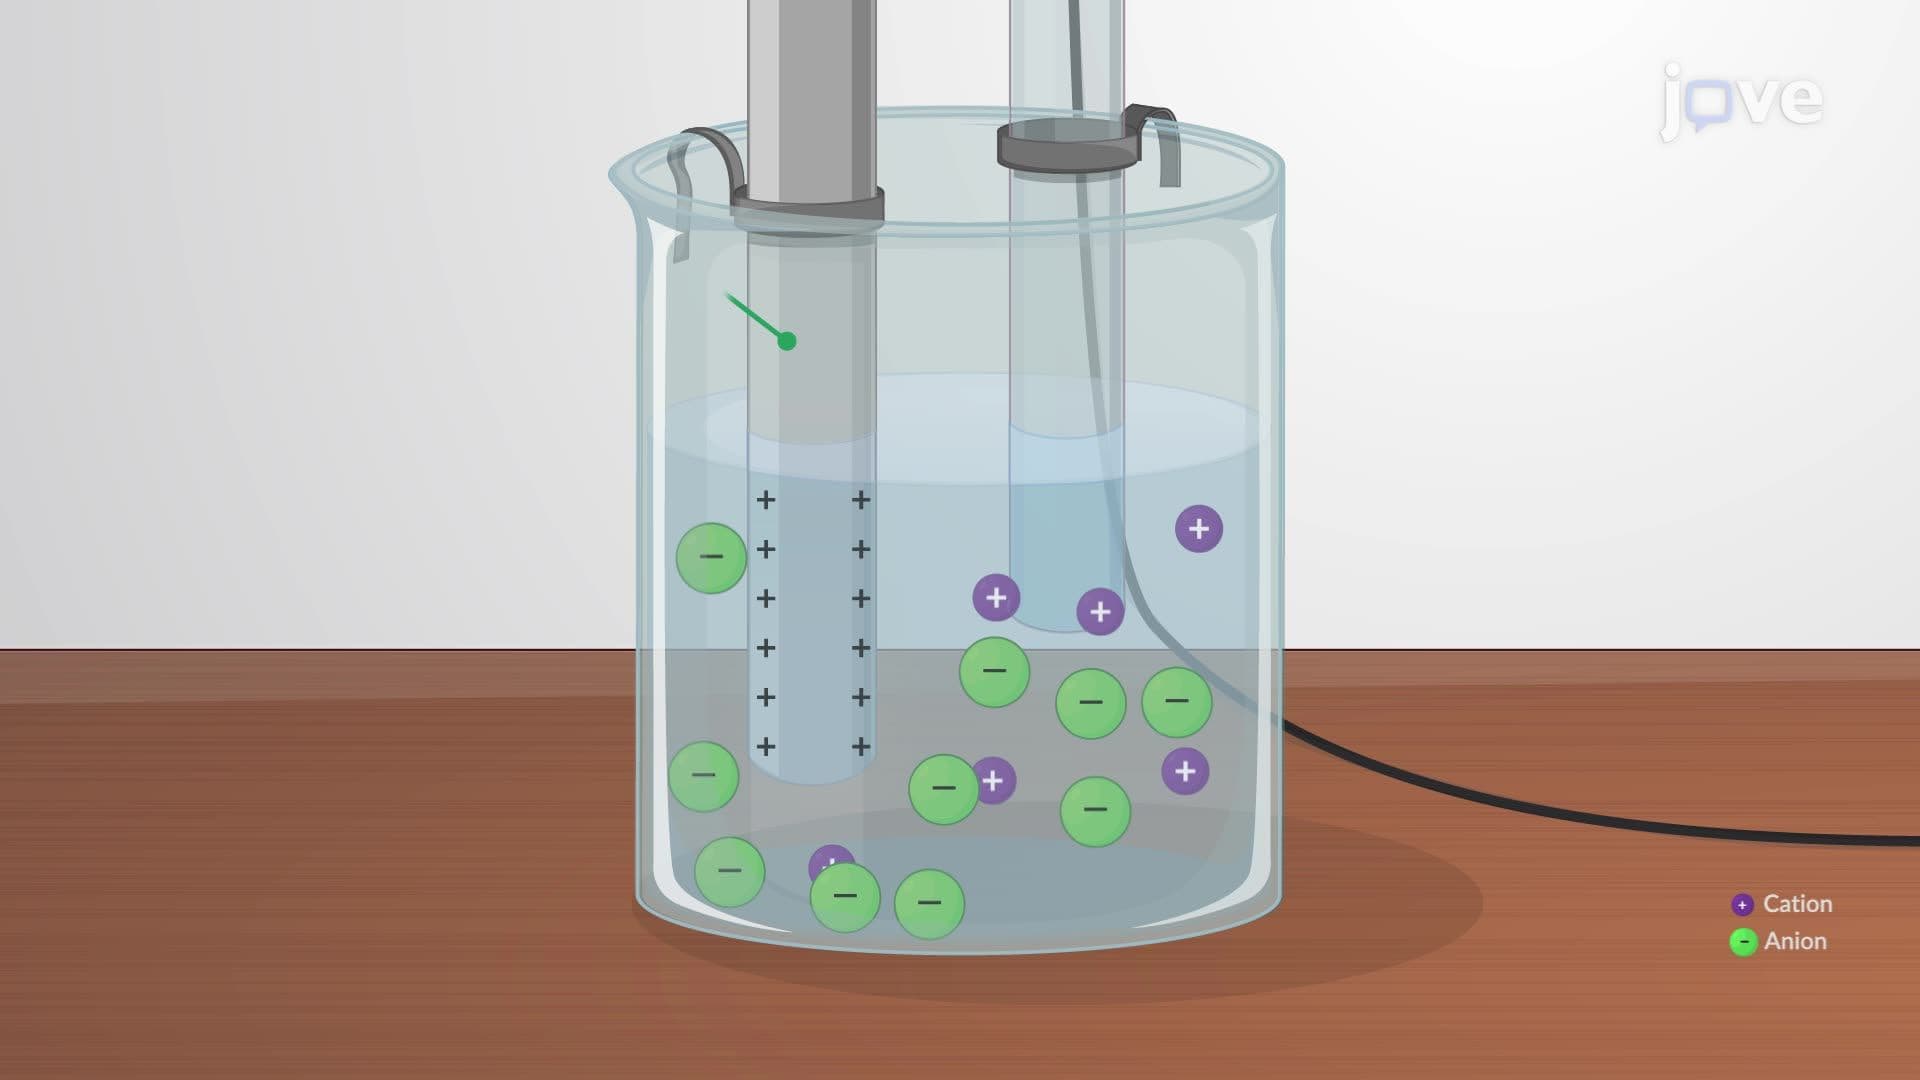
The Electrical Double Layer
 01:24
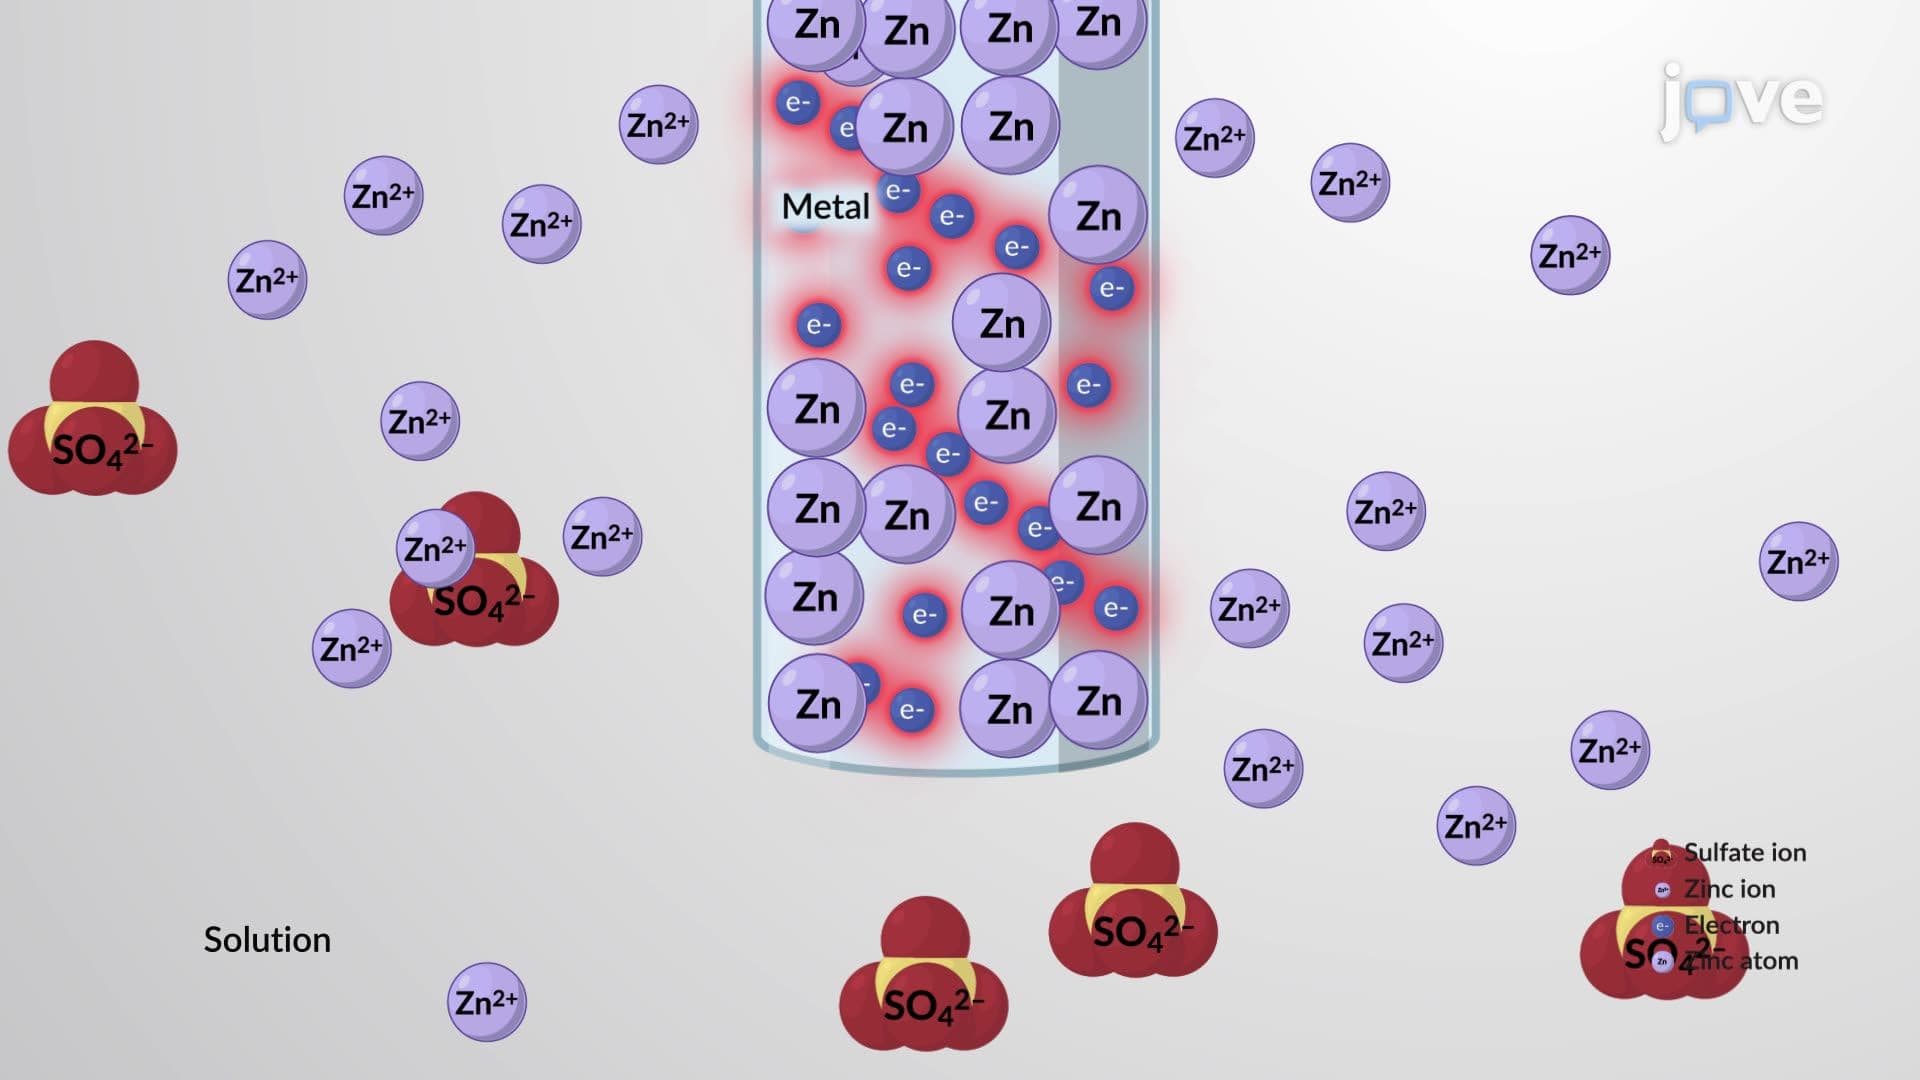
Electrochemical Systems
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:23
Theory of Strong Electrolytes
 01:27
Electrostatic Boundary Conditions in Dielectrics
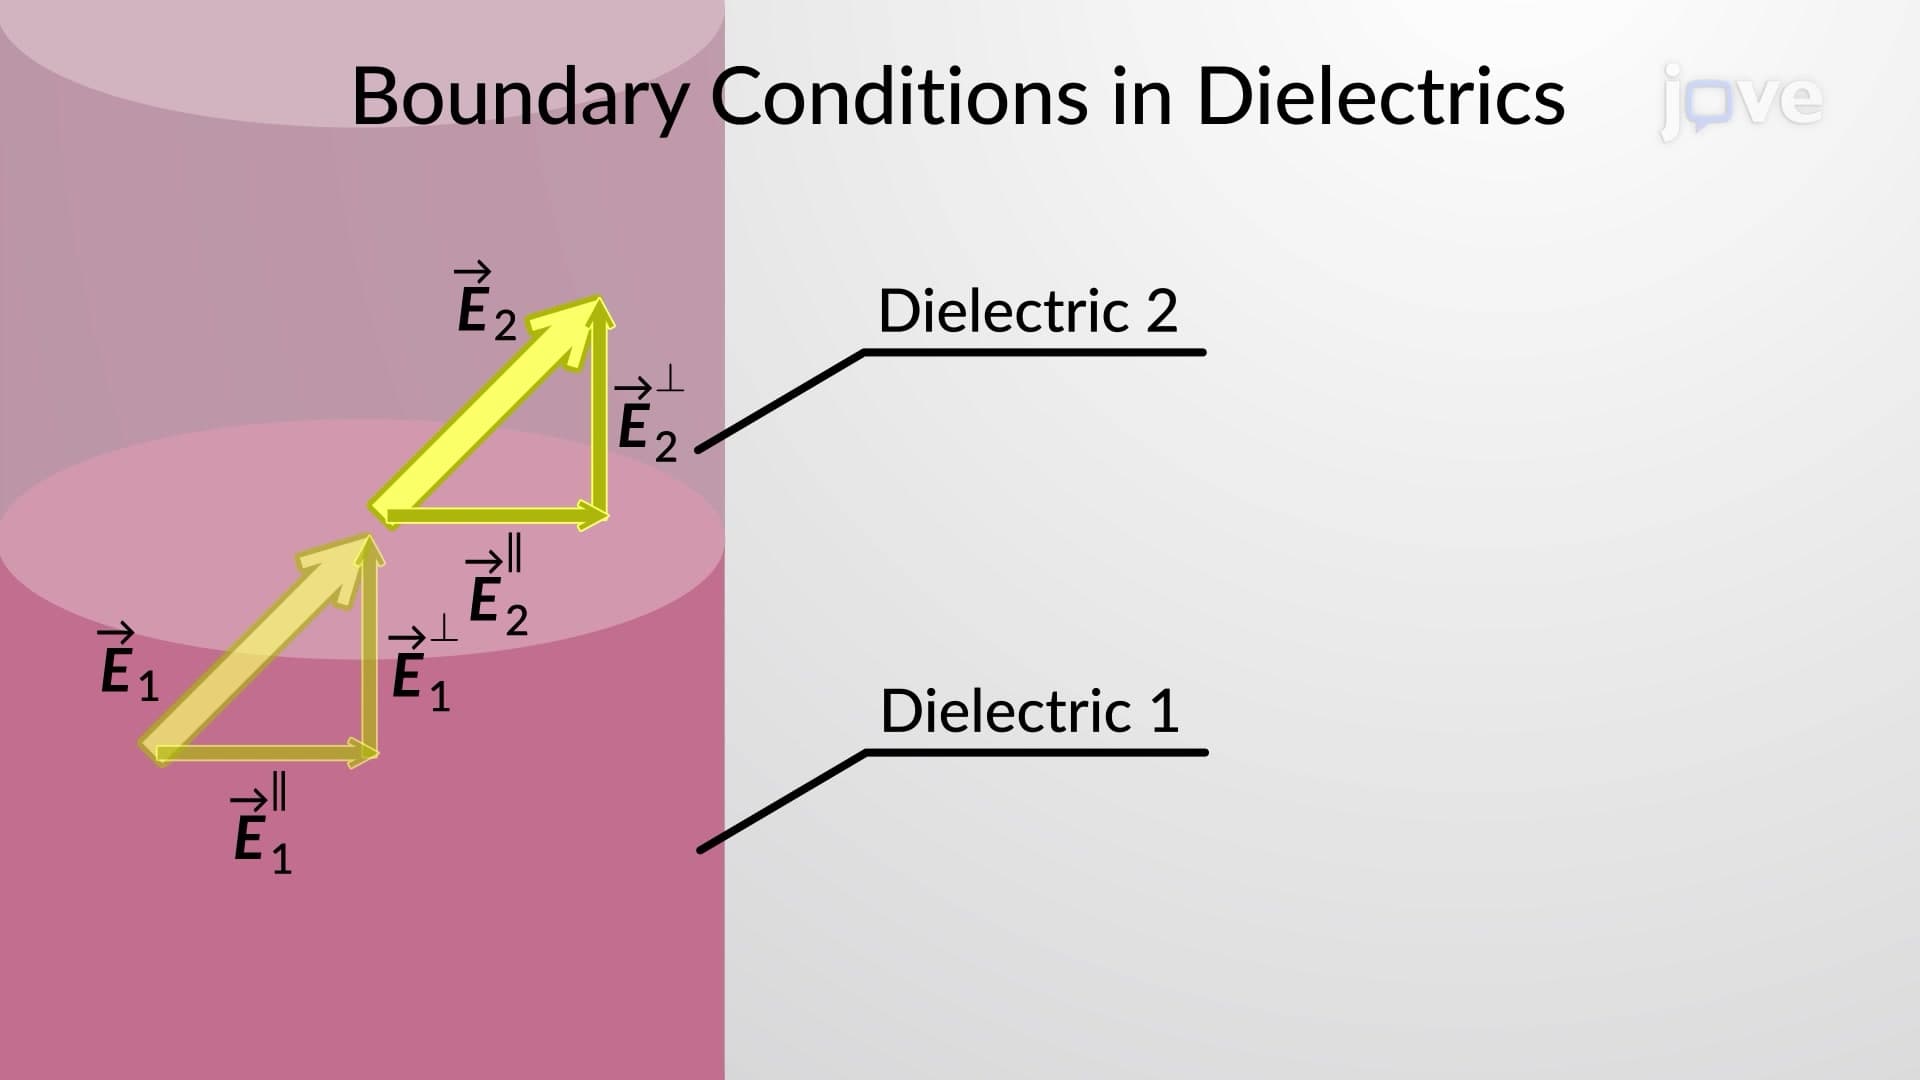
Consider a case where both the mediums across a boundary are two different dielectric materials. Recall that the electric field and electric displacement are proportional and related through the material's permittivity.
 02:48
Ionic Bonding and Electron Transfer