Related Experiment Video
Updated: Oct 7, 2026
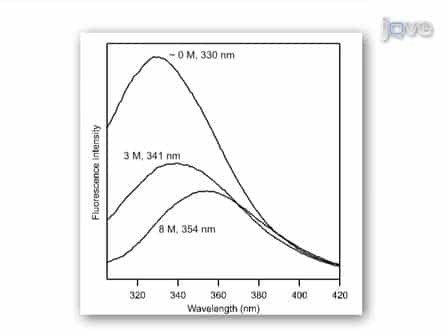

Thermodynamics of Membrane Protein Folding Measured by Fluorescence Spectroscopy
Published on: April 28, 2011
MutTMP Stab-pred: Predicting thermodynamic stability changes upon mutations in α-helical and β-barrel membrane
PRamakrishna Reddy1, A Kulandaisamy1, MMichael Gromiha1
1Department of Biotechnology, Bhupat and Jyoti Mehta School of Biosciences, Indian Institute of Technology Madras, Chennai 600036, Tamil Nadu, India.

Abstract:
Membrane proteins (TMPs) are integral components of cell membranes that adopt two principal structural folds, transmembrane α-helical proteins (TMHs) and transmembrane β-barrel proteins (TMBs). Mutations in these proteins can substantially alter their native structure, folding, and function, and are often implicated in human diseases. Although several computational approaches have been developed to predict the effects of mutations on protein stability, their performance for TMPs remains limited. To address this problem, we compiled a dataset of 379 membrane protein mutations with experimentally measured stability changes (ΔΔG0) from the MPTherm database and developed two machine-learning models for predicting ΔΔG0 of TMHs and TMBs using sequence, structure, and energetic features. Our method, MutTMH Stab-pred could predict ΔΔG0 of TMHs with a correlation and mean absolute error (MAE) of 0.70 and 0.92 kcal/mol, respectively on the hold-out test set. Further, the MutTMB Stab-pred method could predict ΔΔG0 of TMBs with a correlation and MAE of 0.92 and 0.97 kcal/mol, respectively, on the hold-out test set. In leave-one-protein-out and leave-one-homology-cluster-out cross-validation, the MAEs are 1.36 and 1.37 kcal/mol, respectively for MutTMH Stab-pred and 1.31 and 1.35 kcal/mol for MutTMB Stab-pred. Both models are freely available at https://web.iitm.ac.in/bioinfo2/MutTMP%20Stab-pred/. As a large-scale application, we analyzed the stability changes of 33,449 disease-associated and benign missense mutations from the MutDPAL dataset and found that disease-associated mutations were predicted to be destabilizing more frequently than benign mutations. These computational tools provide a valuable resource for understanding mutation-induced stability changes in membrane proteins at a large scale.

Related Concept Videos
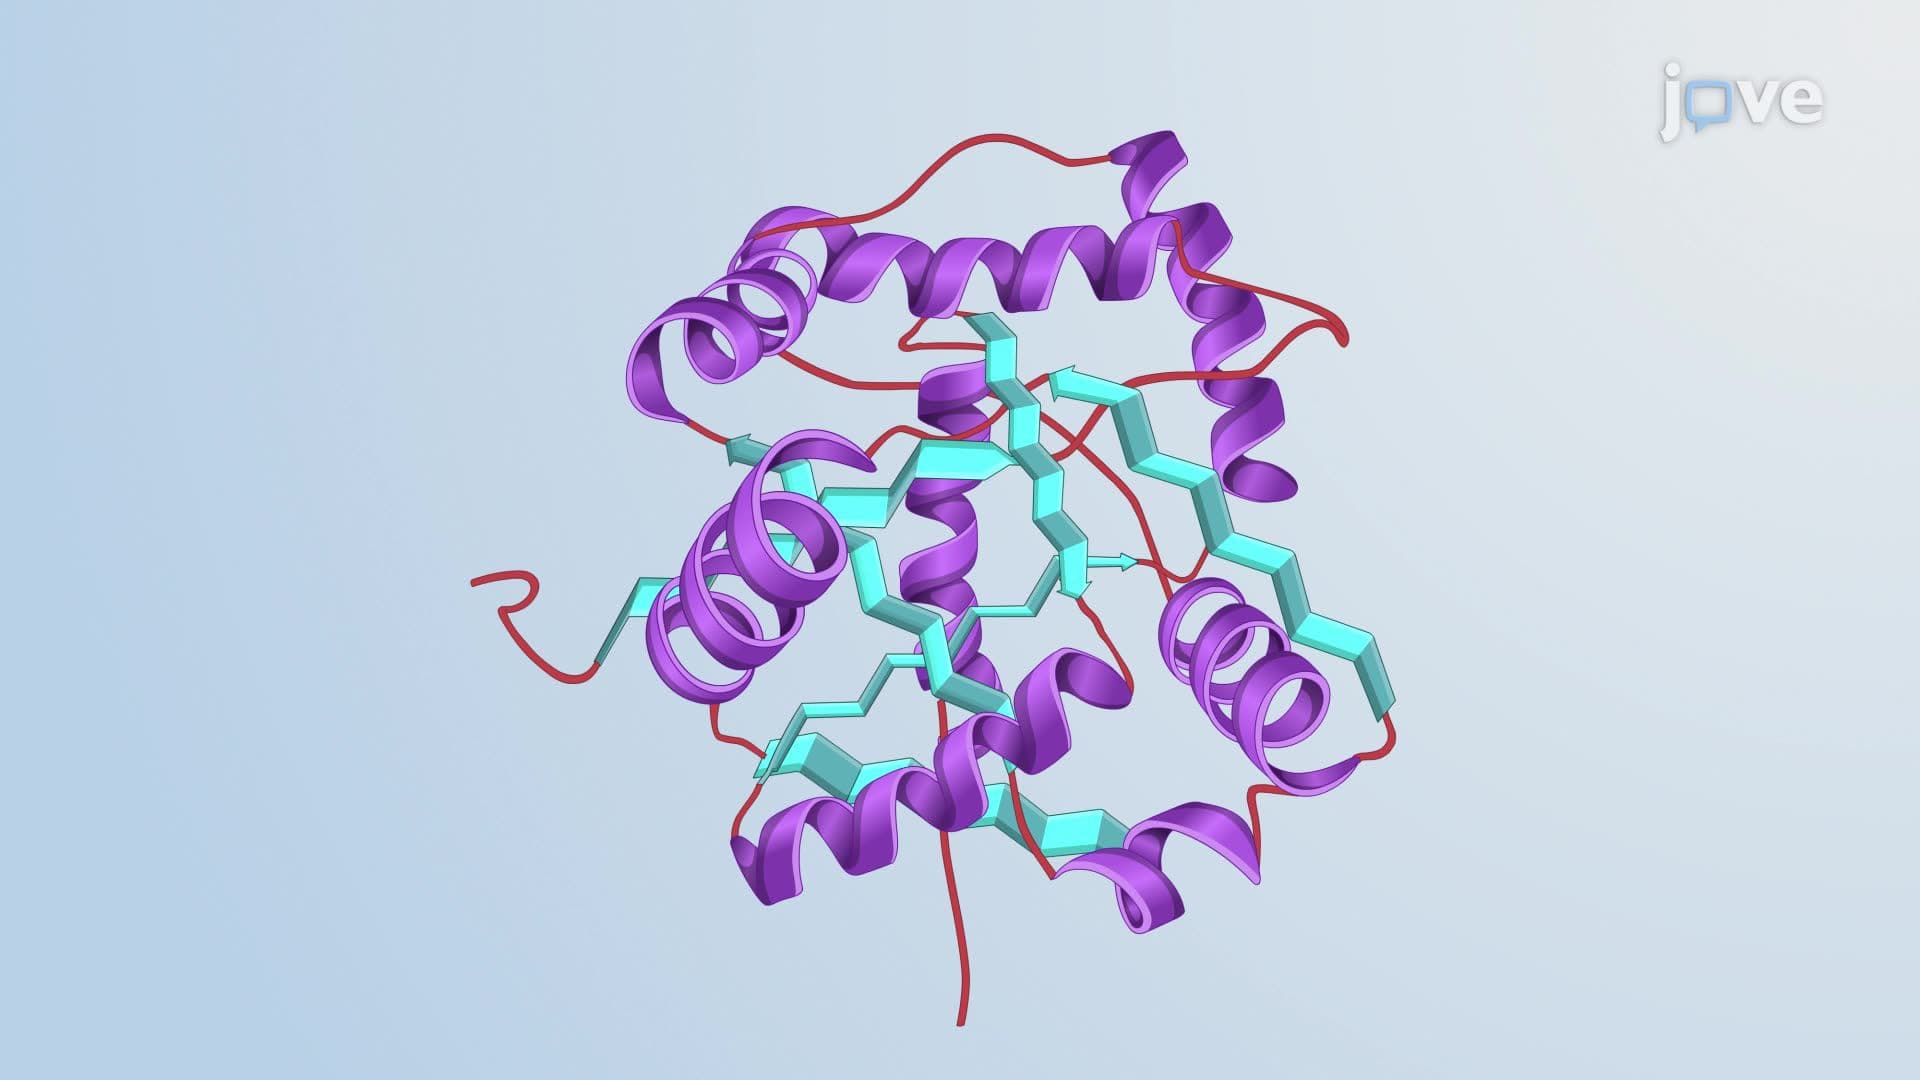 01:49
Protein Folding
 01:25
Protein Folding
Protein Structure Is Critical to Its Biological Function
Proteins perform a wide range of biological functions such as catalyzing chemical reactions, providing...
01:49Protein Folding
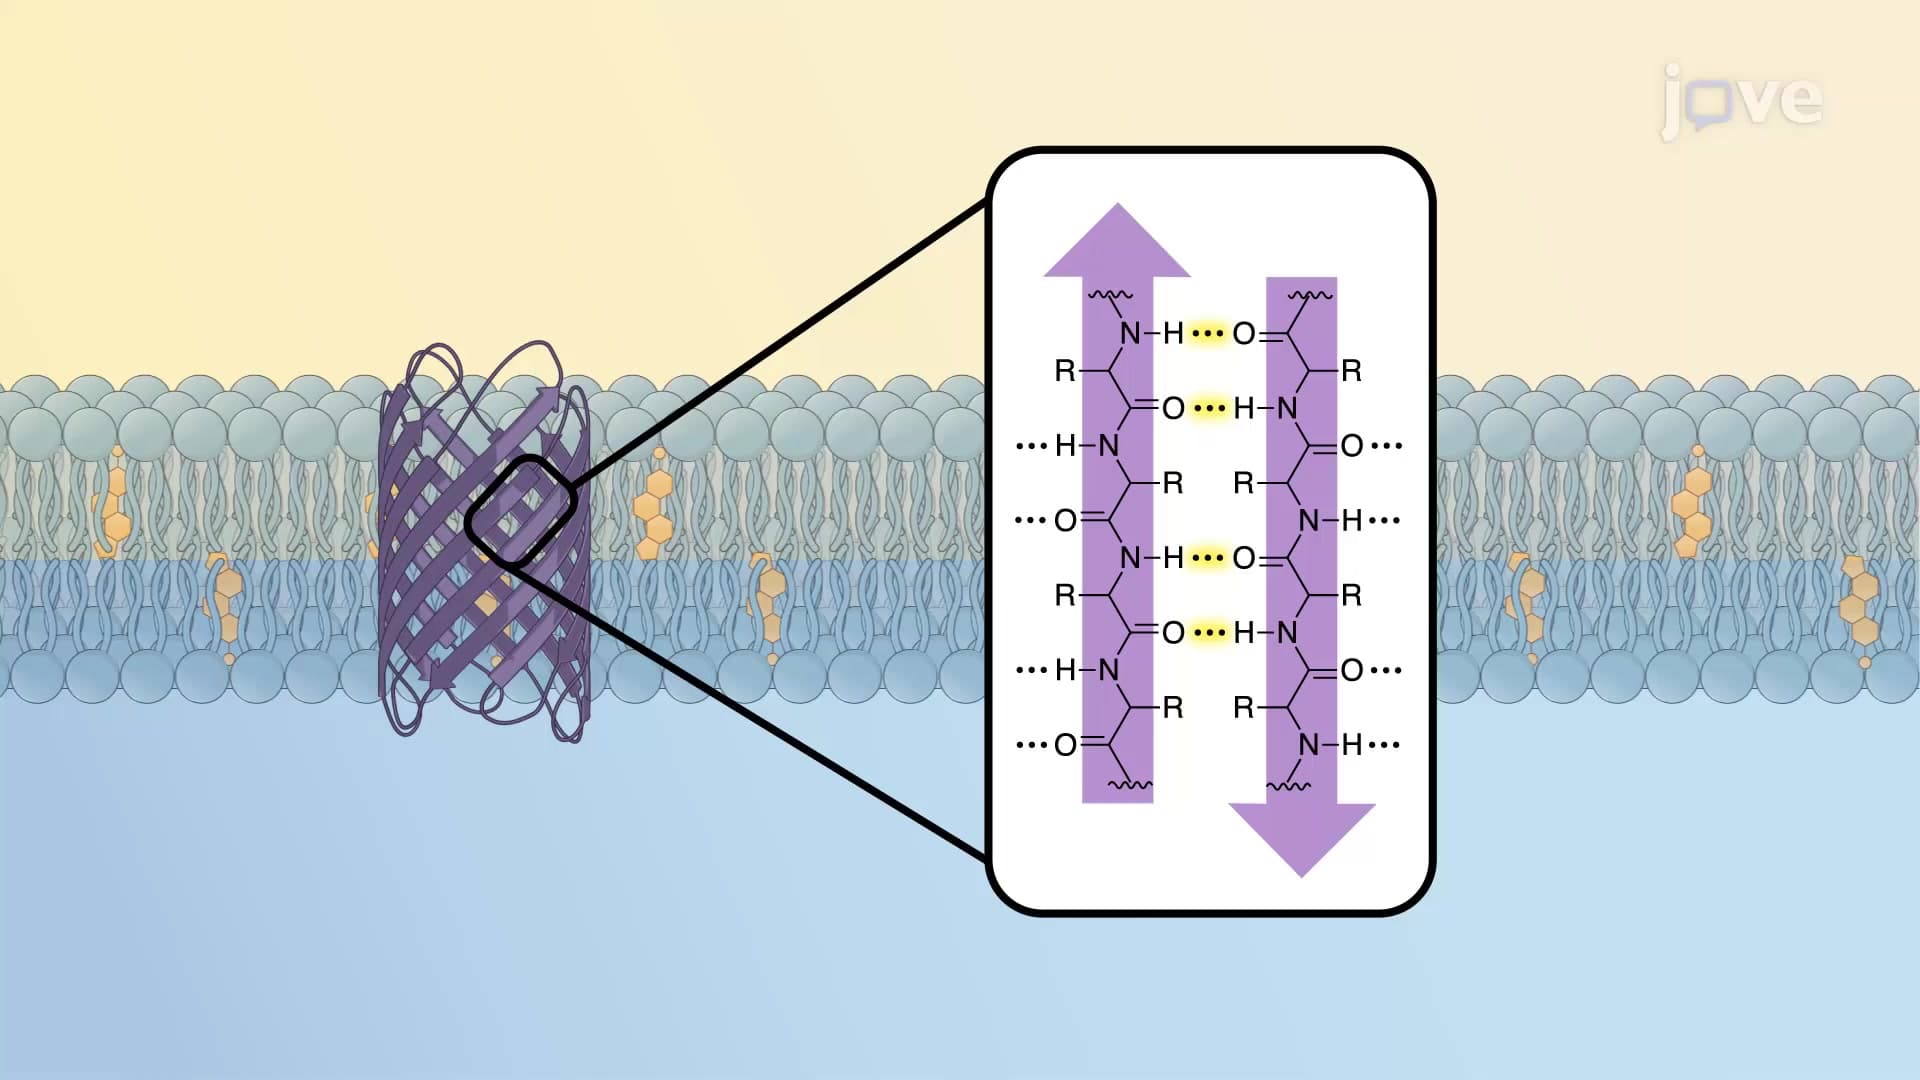 01:09
Multi-pass Transmembrane Proteins and β-barrels
α-Helix containing multi-pass transmembrane proteins
Multi-pass transmembrane proteins such as G-protein-linked receptors (GPCRs) and...
 01:28
Protein Denaturation
 03:00
Molecular Chaperones and Protein Folding
The...