Related Experiment Video
Updated: Oct 7, 2026

Microbiota Analysis Using Two-step PCR and Next-generation 16S rRNA Gene Sequencing
Published on: October 15, 2019
Deciphering the gut microbiota's role in infection risk: Mendelian randomization analysis of microbial taxa and
Fengning Chen1, Chi Zhang2, Guangyue Yao3
1Department of Clinical Laboratory, Shandong Provincial Hospital Affiliated to Shandong First Medical University, Jinan, China.

Background:
Observational studies have linked the gut microbiota (GM) to infectious diseases, but confounding, reverse causation, and disease-related microbiome changes limit causal interpretation. We combined Mendelian randomization (MR) with focused experimental analyses to investigate microbiota-infection relationships.
Methods:
Two-sample MR evaluated genetically predicted microbial taxa across 75 infectious outcomes. Inverse-variance weighted (IVW) estimates were used as the primary analysis, with weighted-median and MR-Egger estimates as complementary methods. Benjamini-Hochberg false discovery rate (FDR) and Bonferroni corrections were applied across all IVW tests. Instrument strength, Cochran's Q, MR-Egger intercepts, leave-one-out analysis, MR-PRESSO, and Steiger directionality testing were used to assess robustness. Exploratory association patterns were used to prioritize a Coriobacteriia-related respiratory signal for experimental investigation using Collinsella aerofaciens cell-free supernatant and A549 epithelial cells.
Results:
Among 15,728 IVW tests, 659 associations had nominal P < 0.05. After global correction, three taxonomically related associations with urinary tract infection (UTI) remained significant. Higher genetically predicted abundance of genus Bifidobacterium was associated with lower UTI risk (odds ratio [OR] 0.29, 95% confidence interval [CI] 0.17-0.49; FDR-adjusted P = 0.029), while order Bifidobacteriales and family Bifidobacteriaceae showed similar estimates (both OR 0.25, 95% CI 0.14-0.45) and met the Bonferroni threshold. Sensitivity analyses showed no significant heterogeneity or MR-PRESSO-detected outliers, and leave-one-out and Steiger analyses were supportive, although MR-Egger intercepts suggested possible directional pleiotropy. Exploratory analyses identified clustering between Coriobacteriia-lineage taxa and respiratory outcomes. C. aerofaciens cell-free supernatant altered 993 genes and enriched inflammatory pathways in A549 cells. qPCR supported increased CSF2, CXCL8, and CCL20 expression, while ELISA detected IL-8 and MIP-3α but not quantifiable GM-CSF. Untargeted metabolomics identified 201 differential metabolites between bacterial supernatant samples and matched BHI controls.
Conclusion:
The corrected Bifidobacterium-related UTI association was the principal MR finding, while the broader analysis identified exploratory microbiota-infection patterns. The C. aerofaciens model linked extracellular metabolic remodeling during bacterial growth with epithelial transcriptional and inflammatory responses, providing focused directions for further investigation.
Related Concept Videos
 01:22
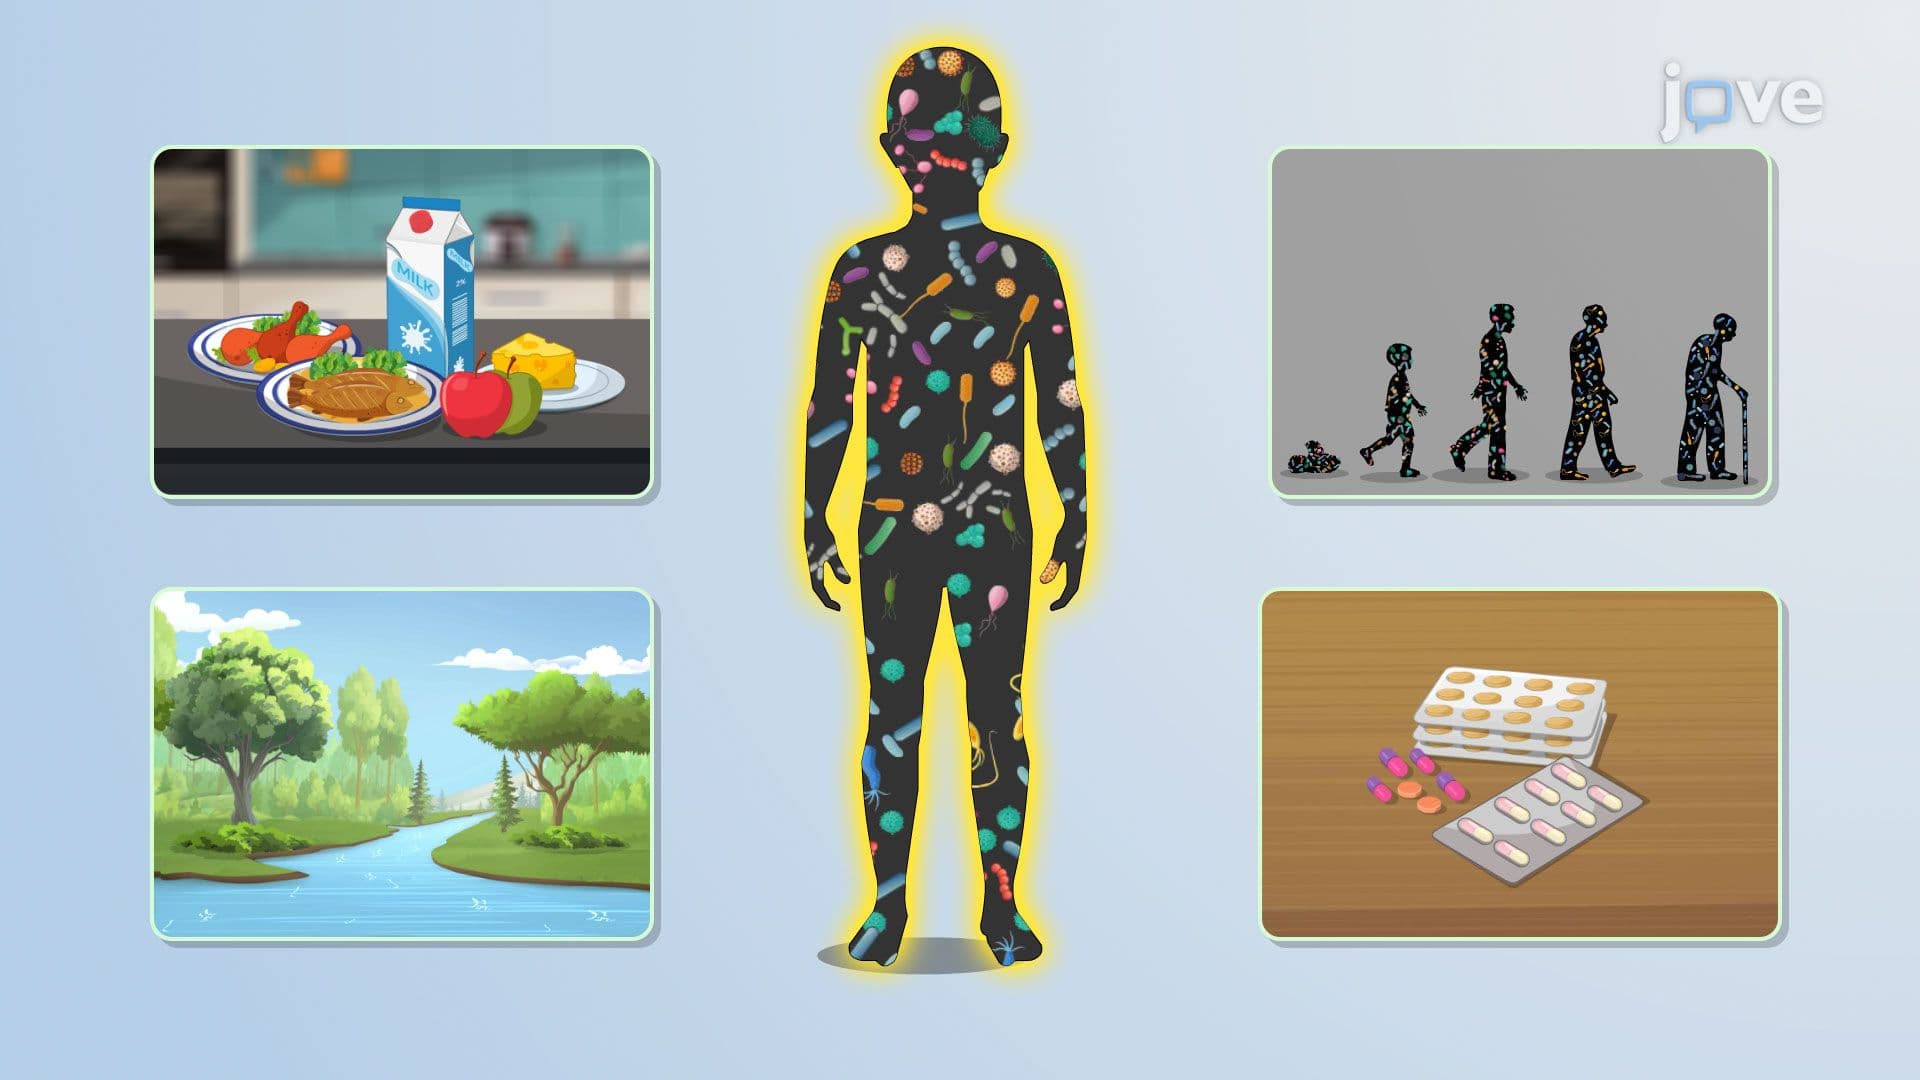
Introduction to the Human Microbiota
 01:18
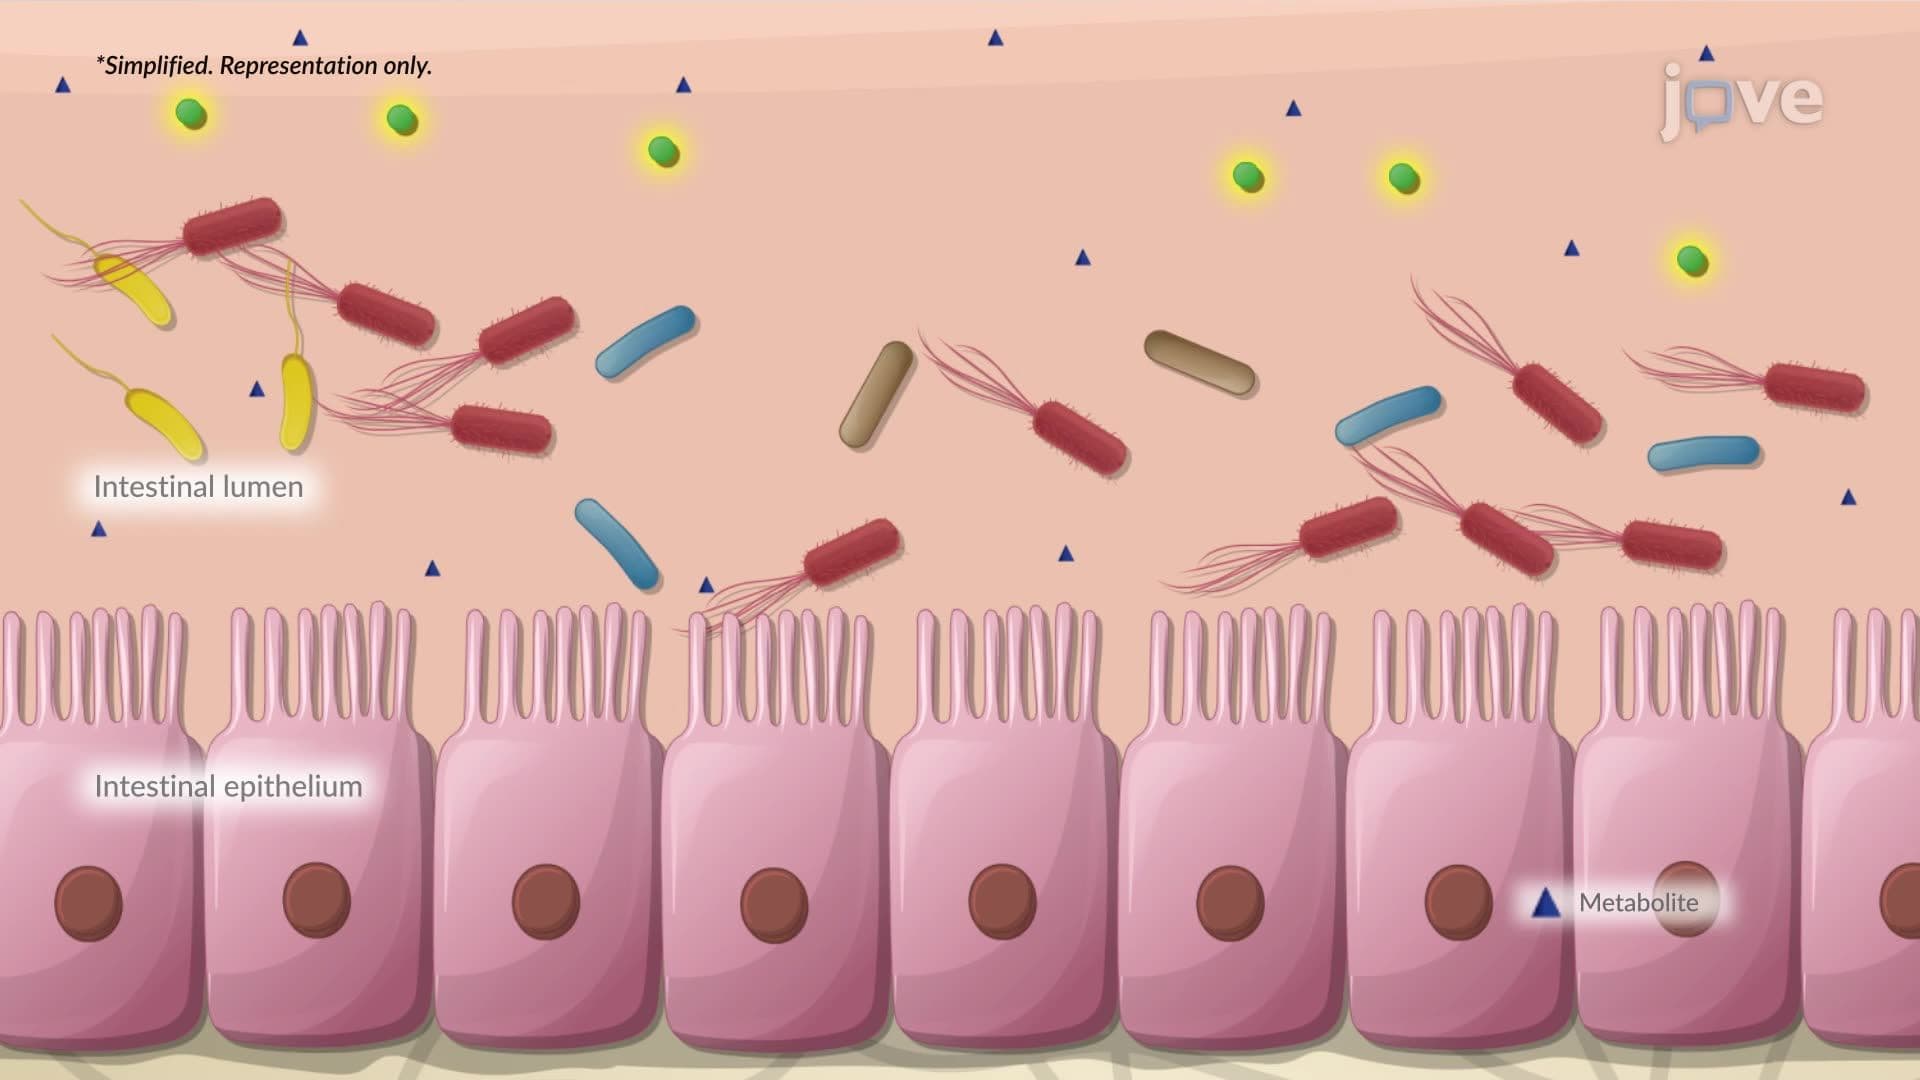
Dysbiosis of the Gut Microbiota
 01:21
Microbiota Modulation by Antibiotics
 01:18
Functions of the Gut Microbiota
 01:30
Development of Human Microbiota
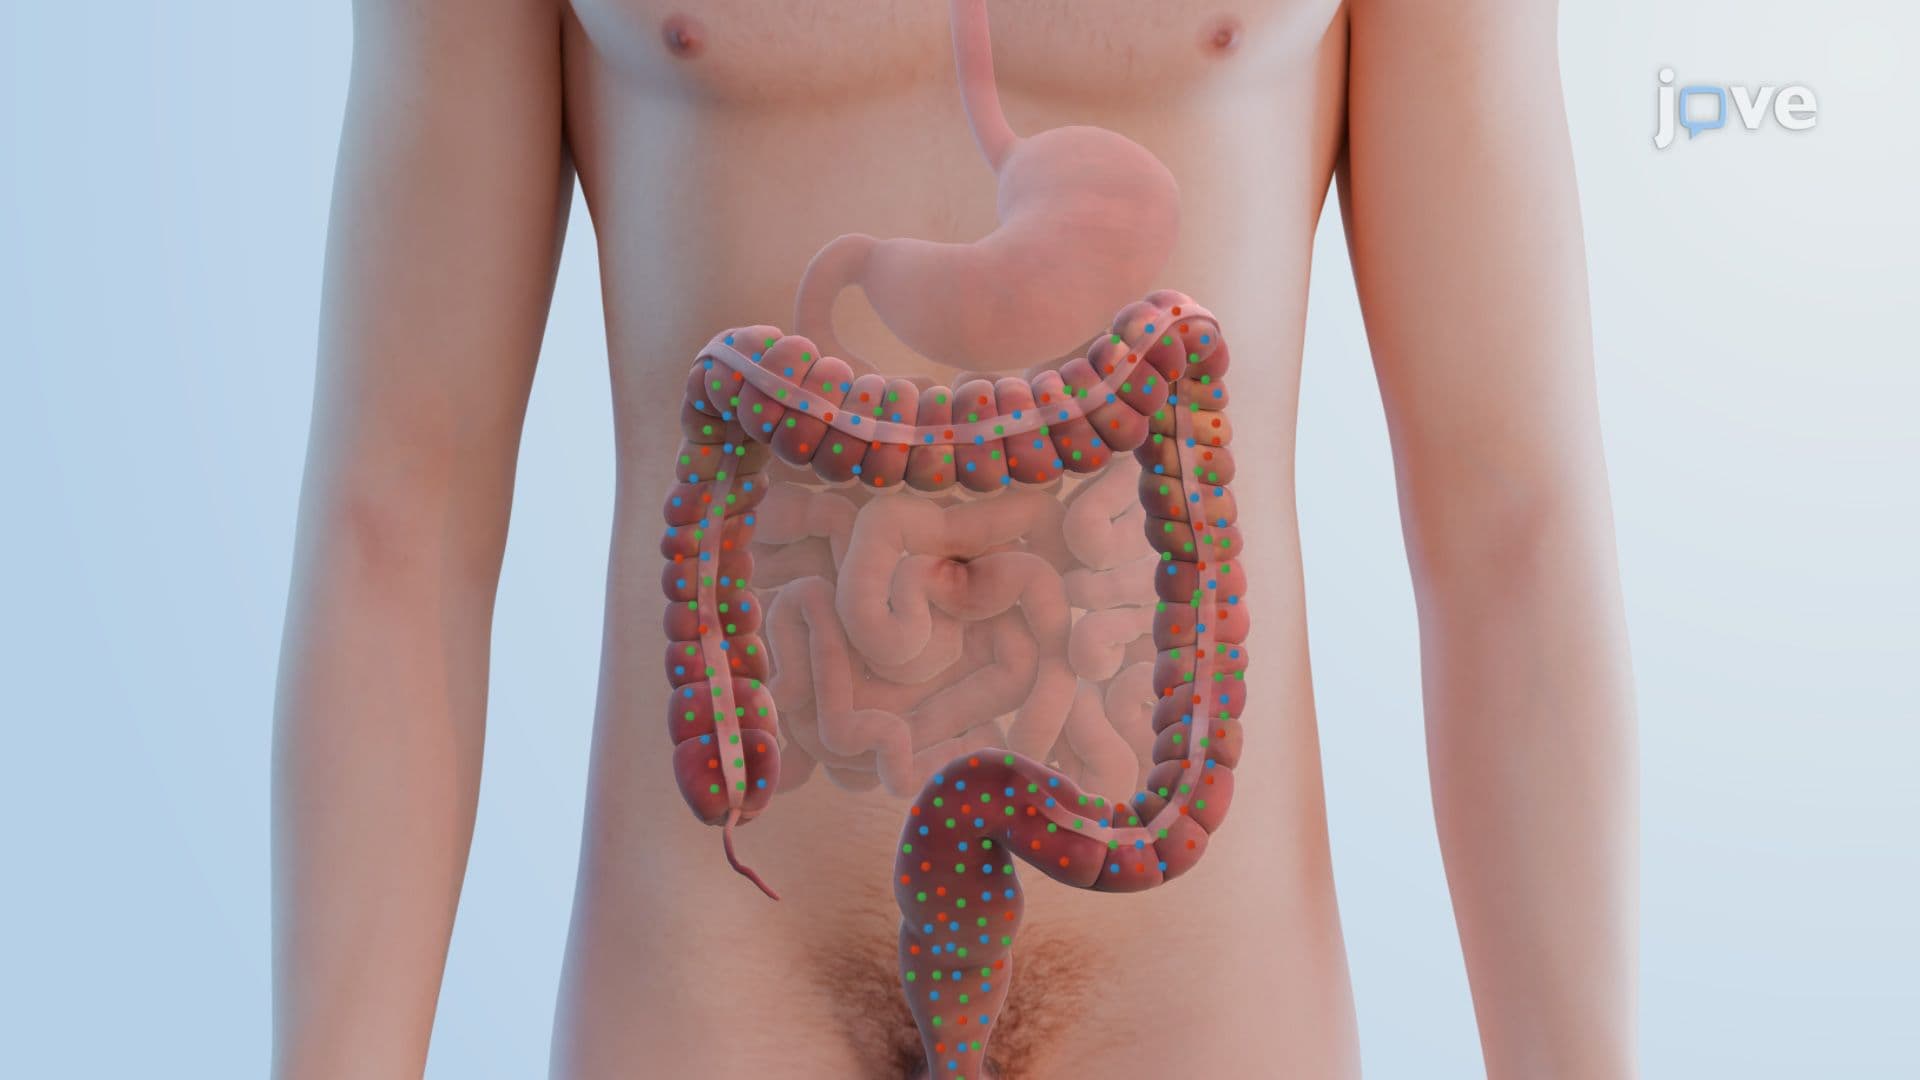 01:27
Microbiota of the Large Intestine