Related Experiment Video
Updated: Oct 8, 2026


Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Virtual screening, structural optimization, computational studies, and biological evaluation of quinoline derivatives
1School of Pharmaceutical Sciences, Jilin University, Changchun 130021, China; Department of Clinical Pharmacy, The First Hospital of Jilin University, Changchun 130021, China.

Abstract:
Butyrylcholinesterase (BChE) has been recognized as an important therapeutic target for neurodegenerative diseases, including Alzheimer's disease (AD). In this study, a molecular docking-based virtual screening strategy led to the identification of a novel BChE inhibitor, Hit 5 (eqBChE IC50 = 4.1 ± 0.19 μM, hBChE IC50 = 6.2 ± 2.1 μM). Thirty-one compounds were designed and synthesized via rational structural modification guided by molecular docking. The quinoline core scaffold was constructed by the Pfitzinger reaction, and the target compounds were afforded via amide condensation. All compounds were unambiguously characterized by 1H/13C NMR, FT-IR and high-resolution mass spectrometry (HRMS), with HPLC-verified purities >95%. Structure-activity relationship (SAR) analysis indicated that the type, position and number of substituents on the terminal phenyl ring had a marked influence on BChE inhibitory activity. Among the synthesized compounds, compound 6j emerged as the most potent inhibitor, with IC50 values of 0.20 ± 0.09 μM against eqBChE and 0.35 ± 0.11 μM against hBChE. Further biological evaluation demonstrated that compound 6j possessed low cytotoxicity, anti-inflammatory, antioxidant, and metal ion chelating activities, as well as promising blood-brain barrier permeability predicted by in vitro assay. The interaction mechanism between compound 6j and BChE was comprehensively investigated by isothermal titration calorimetry (ITC) and molecular simulations. ITC assays experimentally confirmed a strong binding affinity between compound 6j and BChE, with a dissociation constant Kd of 1.37 μM. Furthermore, molecular docking results indicated that compound 6j occupied the active pocket of BChE through hydrophobic interactions, π-π stacking, and hydrogen bonds. Molecular dynamics (MD) simulations and MM-GBSA binding free energy analysis suggested that the stable interaction with the VAL288 residue may account for the high BChE selectivity of compound 6j. In addition, pharmacokinetic studies in mice revealed that compound 6j displayed favorable drug-like properties, with a half-life (t1/2) of 4.09 h and a time to peak concentration (Tmax) of 1.33 h. Compound 6j may serve as an important lead compound for subsequent structural optimization.
Related Concept Videos
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence its...
 01:29
Indirect-Acting Cholinergic Agonists: Chemistry and Structure-Activity Relationship
Reversible inhibitors display short to medium durations of action. Short-acting agents include simple alcohols with...
 01:29
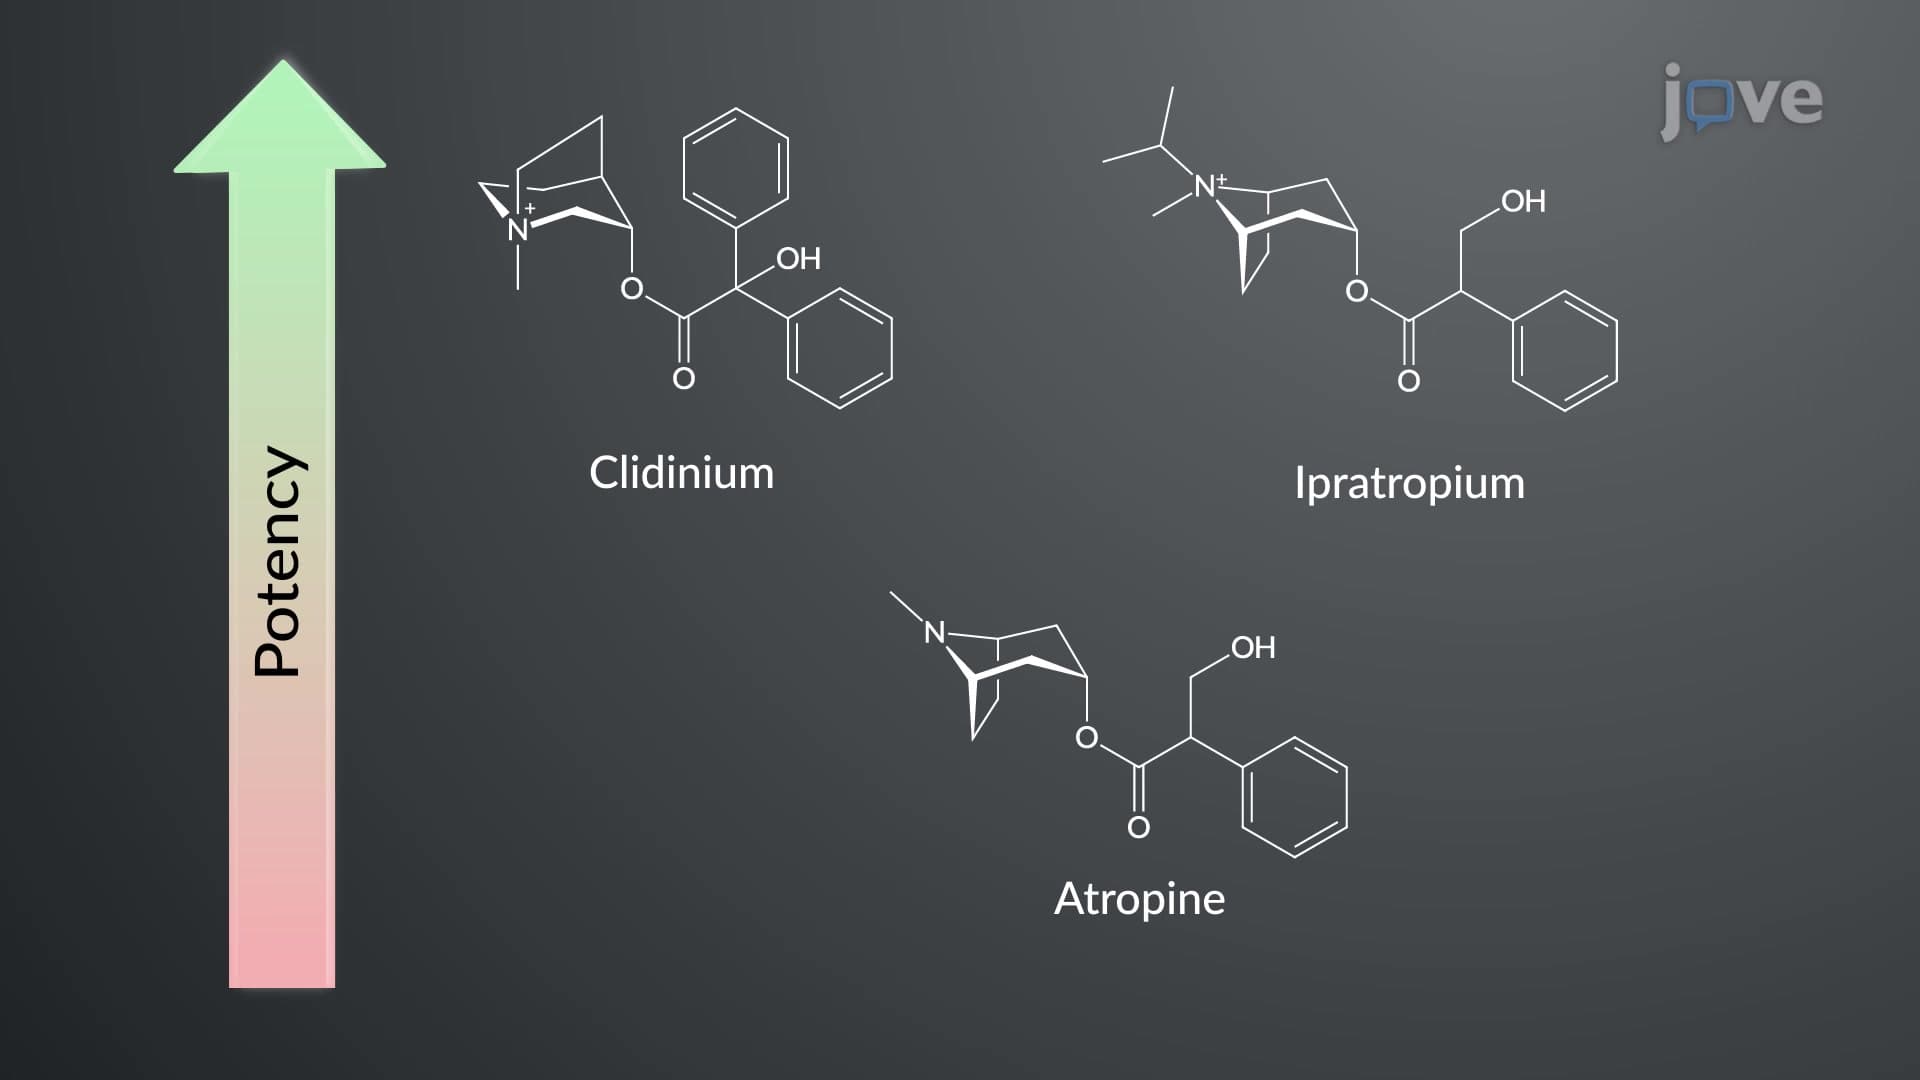
Cholinergic Antagonists: Chemistry and Structure-Activity Relationship
 01:18
Indirect-Acting Cholinergic Agonists: Mechanism of Action
Reversible inhibitors like edrophonium bind to a specific part of the enzyme called the anionic catalytic site. They form noncovalent bonds, which means they are not strongly attached to the enzyme. This creates a temporary and less stable enzyme–inhibitor complex, leading to...
 01:30
Enzyme Inhibition