Related Experiment Video
Updated: Oct 10, 2026

Investigation of Microbial Cooperation via Imaging Mass Spectrometry Analysis of Bacterial Colonies Grown on Agar and in Tissue During Infection
Published on: November 18, 2022
Gut microbiome diversity, genus-level variation, and associations with clinical features in critically ill patients
Yibing Wang1, Peng Guo1, Jie Shen1
1Center of Emergency and Critical Medicine, Jinshan Hospital of Fudan University, Shanghai, China.

Background:
Gut microbiota alterations are common in critical illness, but differences between critically ill patients with and without sepsis and their clinical relevance remain incompletely understood. We investigated gut microbial diversity, genus-level differences and their associations with inflammatory profiles and clinical outcomes in critically ill patients.
Methods:
This single-centre case-control study enrolled adult critically ill patients grouped by Sepsis-3 criteria. Gut microbiota was profiled by 16S rRNA sequencing. The primary M2 model, adjusted for age, intra-abdominal infection, pre-sampling anti-anaerobic antibiotic exposure, APACHE II score, and chronic kidney disease, was used for all adjusted analyses. Alpha diversity was analyzed with HC3-robust linear regression; beta diversity with PERMANOVA, PERMDISP, and dbRDA. Genus-level primary discovery used Wilcoxon rank-sum with Benjamini-Hochberg FDR and |log2 fold change| threshold. Complementary sensitivity analyses used fully adjusted MaAsLin2 and ANCOM-BC2 with pseudo-count sensitivity. Associations of the four Wilcoxon-identified genera with clinical variables were assessed via partial Spearman correlations, logistic regression, and longitudinal mixed-effects models.
Results:
Of 103 participants, 68 had sepsis and 35 did not. After M2 adjustment, Shannon, Simpson, and Pielou evenness were lower in sepsis (β = -0.60, -0.54, -0.60; all q = 0.025), whereas Chao1 richness and observed species did not differ. Bray-Curtis and Jaccard group effects did not meet FDR threshold. Wilcoxon identified higher Lactococcus_A and lower Ezakiella, Fenollaria, Peptoniphilus_B in sepsis. ANCOM-BC2 signals were not robust to pseudo-count sensitivity, and MaAsLin2 identified no significant genus (min q = 0.333), indicating genus-level findings inconsistent across methods. No cross-sectional clinical association met significance after FDR correction. Longitudinal analyses found time-by-genus interactions: Fenollaria with CRP (q = 0.010), Peptoniphilus_B with PCT (q = 0.042) and CRP (q = 0.010).
Conclusions:
Sepsis was associated with lower values for selected alpha-diversity measures, whereas differences in overall community composition were modest. Four genera met the primary FDR- and effect-size-adjusted Wilcoxon criteria, but genus-level differences were not consistently supported across Wilcoxon, ANCOM-BC2 and MaAsLin2, indicating these are exploratory signals rather than confirmed candidates. Associations with clinical features and inflammatory markers were limited; therefore, these findings should be considered exploratory and validated using multiple analytical approaches in independent cohorts.
Related Concept Videos
 01:30
Development of Human Microbiota
 01:22
Introduction to the Human Microbiota
 01:18
Dysbiosis of the Gut Microbiota
 01:21
Microbiota Modulation by Antibiotics
 01:18
Functions of the Gut Microbiota
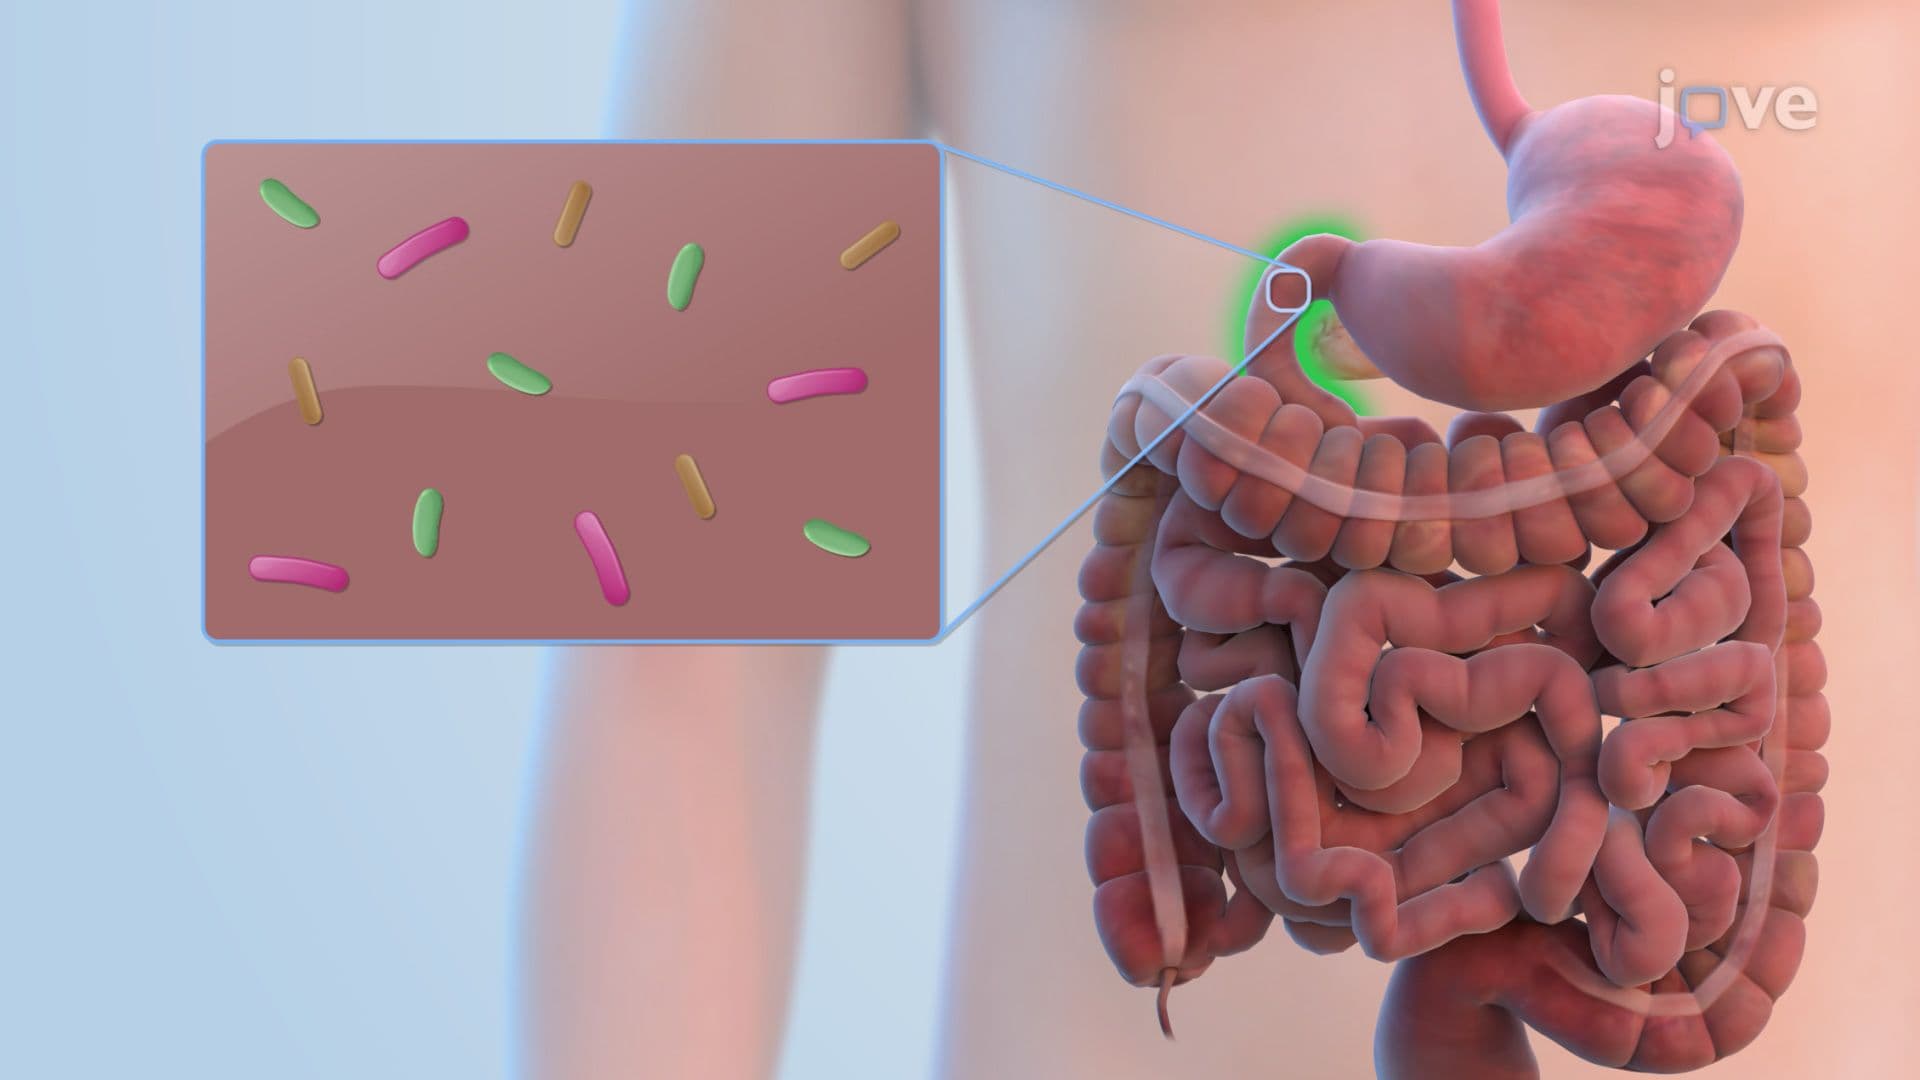 01:27
Microbiota of the Stomach and Small Intestine