Related Experiment Video
Updated: Oct 10, 2026
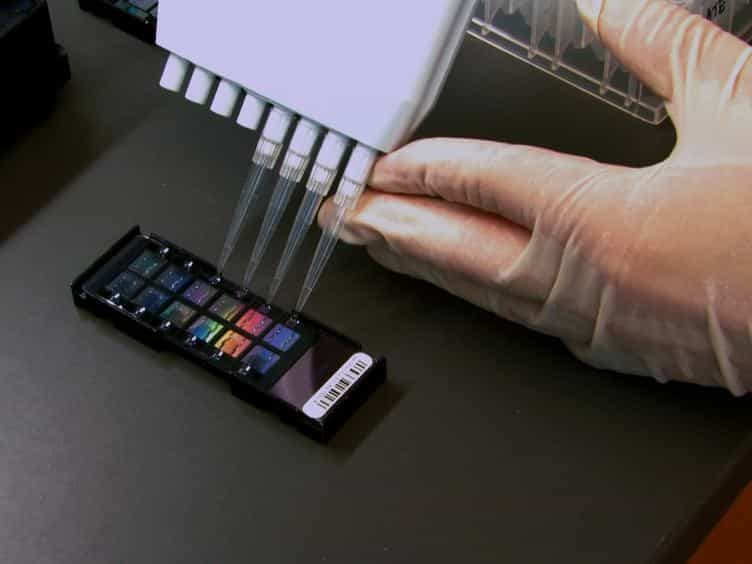

Infinium Assay for Large-scale SNP Genotyping Applications
Published on: November 19, 2013
Characterizing Runs of Homozygosity and Inbreeding in American Mink Using SNP Data and Pedigree Information
Persia Carol Thapa1, Duy Ngoc Do1, Pourya Davoudi2
1Department of Animal Science and Aquaculture, Dalhousie University, Truro, Nova Scotia, Canada.

Abstract:
Inbreeding increases the likelihood of deleterious recessive alleles becoming homozygous and consequently affecting economically important traits. Runs of homozygosity (ROH), the continuous homozygous segments of the genome and the higher than expected homozygosity (HOM) are genomic indicators of inbreeding, quantified as FROH and FHOM, respectively. Previous ROH analyses in American mink (Neogale vison) were restricted to scaffold-level references and could not resolve long autozygous segments. In this study, we characterized ROH in 1413 mink genotyped with the Affymetrix Mink 70K panel (50,902 autosomal SNPs after quality control) on the chromosome-level genome assembly and compared FROH with FHOM and with pedigree-based inbreeding (FPED) derived from 24,633 pedigree records. ROH were detected in all individuals, with a mean of 149.61 ± 13.82 segments per individual. Segments longer than 8 Mb, not detectable in earlier studies of this population, represented 12.86% of all ROH and accounted for 0.10 of the mean FROH of 0.28. Mean FHOM and FPED were -0.03 and 0.02, respectively. ROH were unevenly distributed across chromosomes and chromosomes 1, 2, 3, 4, 6 and 11 harboured 78.59% of all segments. The Pearson correlation coefficient was highest (0.72) between FROH and FHOM and moderate (0.34) between genomic inbreeding and FPED, increasing with ROH length to 0.35 for segments longer than 16 Mb. Because the density of the marker panel limits the resolution of short segments, quantitative inference was restricted to ROH longer than 4 Mb. Our results indicate that recent autozygosity in this closed population is concentrated in a small number of long genomic segments, providing an opportunity to inform mating decisions.
Related Concept Videos
 01:43
Incomplete Dominance
 02:21
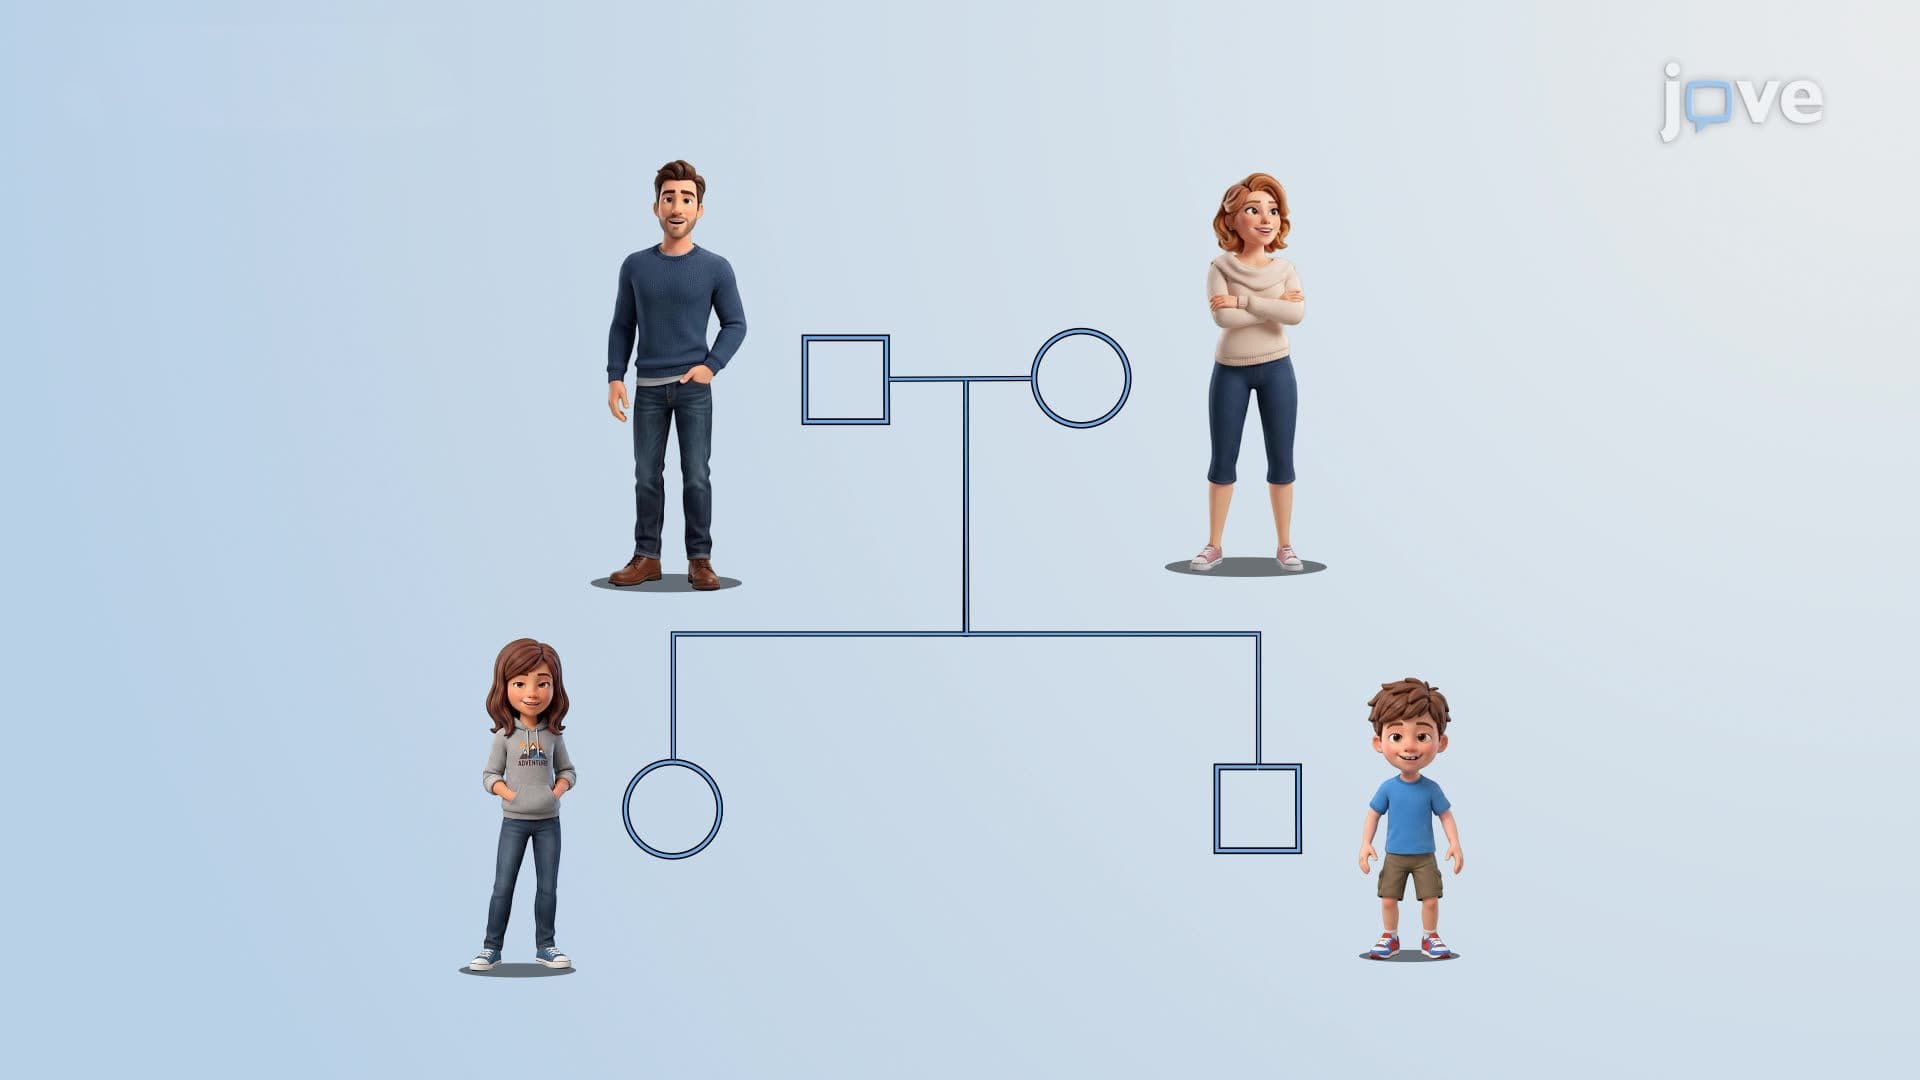
Pedigree Analysis
02:21Pedigree Analysis
 01:05
Single Nucleotide Polymorphisms-SNPs
 01:11
Genome-wide Association Studies-GWAS
GWAS does not require the identification of the target gene involved in...
 02:26
Comparing Copy Number Variations and SNPs
Copy number variations or CNVs are the structural variations that cover more than 1kb of DNA sequence. The single nucleotide polymorphism (SNP), on the other hand, is a single nucleotide change or a point mutation that is found in more than 1%...