Related Experiment Videos
Lightweight 2D-Convolutional Neural Network for Drug-Kinase Binding Affinity Prediction with Minimal Encoded
Hassan Zohrevand1, Saeid Afshar2,3, Mohsen Ahmadi1
1Department of Biomedical Engineering and Medical Physics, School of Medicine, Shahid Beheshti University of Medical Sciences Tehran Iran.

Background:
Drug-kinase binding affinity (DKBA) prediction plays a crucial role in prioritizing compounds and reducing the time and financial resources required for drug discovery. However, many existing deep learning models rely on complex molecular representations, such as molecular graphs or pretrained embeddings, which may increase computational complexity and limit scalability.
Objectives:
This study developed a lightweight two-dimensional convolutional neural network (2D-CNN)-based model to predict DKBA, providing an efficient, low-cost framework for early-stage virtual screening of kinase inhibitors using label-encoded sequences. The primary objective was to determine whether a relatively simple CNN architecture could achieve useful predictive performance without complex molecular representations.
Methods:
Drug SMILES strings and protein amino acid sequences were label-encoded and reshaped into two-dimensional matrix formats (10 × 10 for drugs and 40 × 25 for proteins) without using graph-based representations or pretrained embeddings. The proposed 2D-CNN model was trained on the KIBA dataset containing 118,254 drug-protein interactions and evaluated using mean squared error (MSE), root mean squared error (RMSE), and the concordance index (CI). Dropout, regularization, and early stopping were applied during training.
Results:
The proposed model achieved acceptable predictive performance on the KIBA test set, with an MSE of 0.184, RMSE of 0.429, and CI of 0.870. The model comprised 179,649 trainable parameters and achieved an inference throughput of approximately 1,674 samples per second on the evaluation hardware. The model showed numerically better performance than the literature-reported results for SimBoost and KronRLS, and performance comparable to that reported for DeepDTA and WideDTA. However, these benchmark values were obtained from the original publications rather than reproduced under identical data-splitting and preprocessing conditions and, therefore, do not constitute strictly controlled head-to-head comparisons.
Conclusions:
The proposed framework demonstrates that a lightweight 2D-CNN can provide useful DKBA predictions using simple label-encoded ligand and protein sequences. This approach may serve as a foundation for computational DKBA prediction; however, further validation using independent and disease-specific datasets, improved sequence representations, and more stringent validation strategies is needed to assess its generalizability and practical utility.
Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 01:22
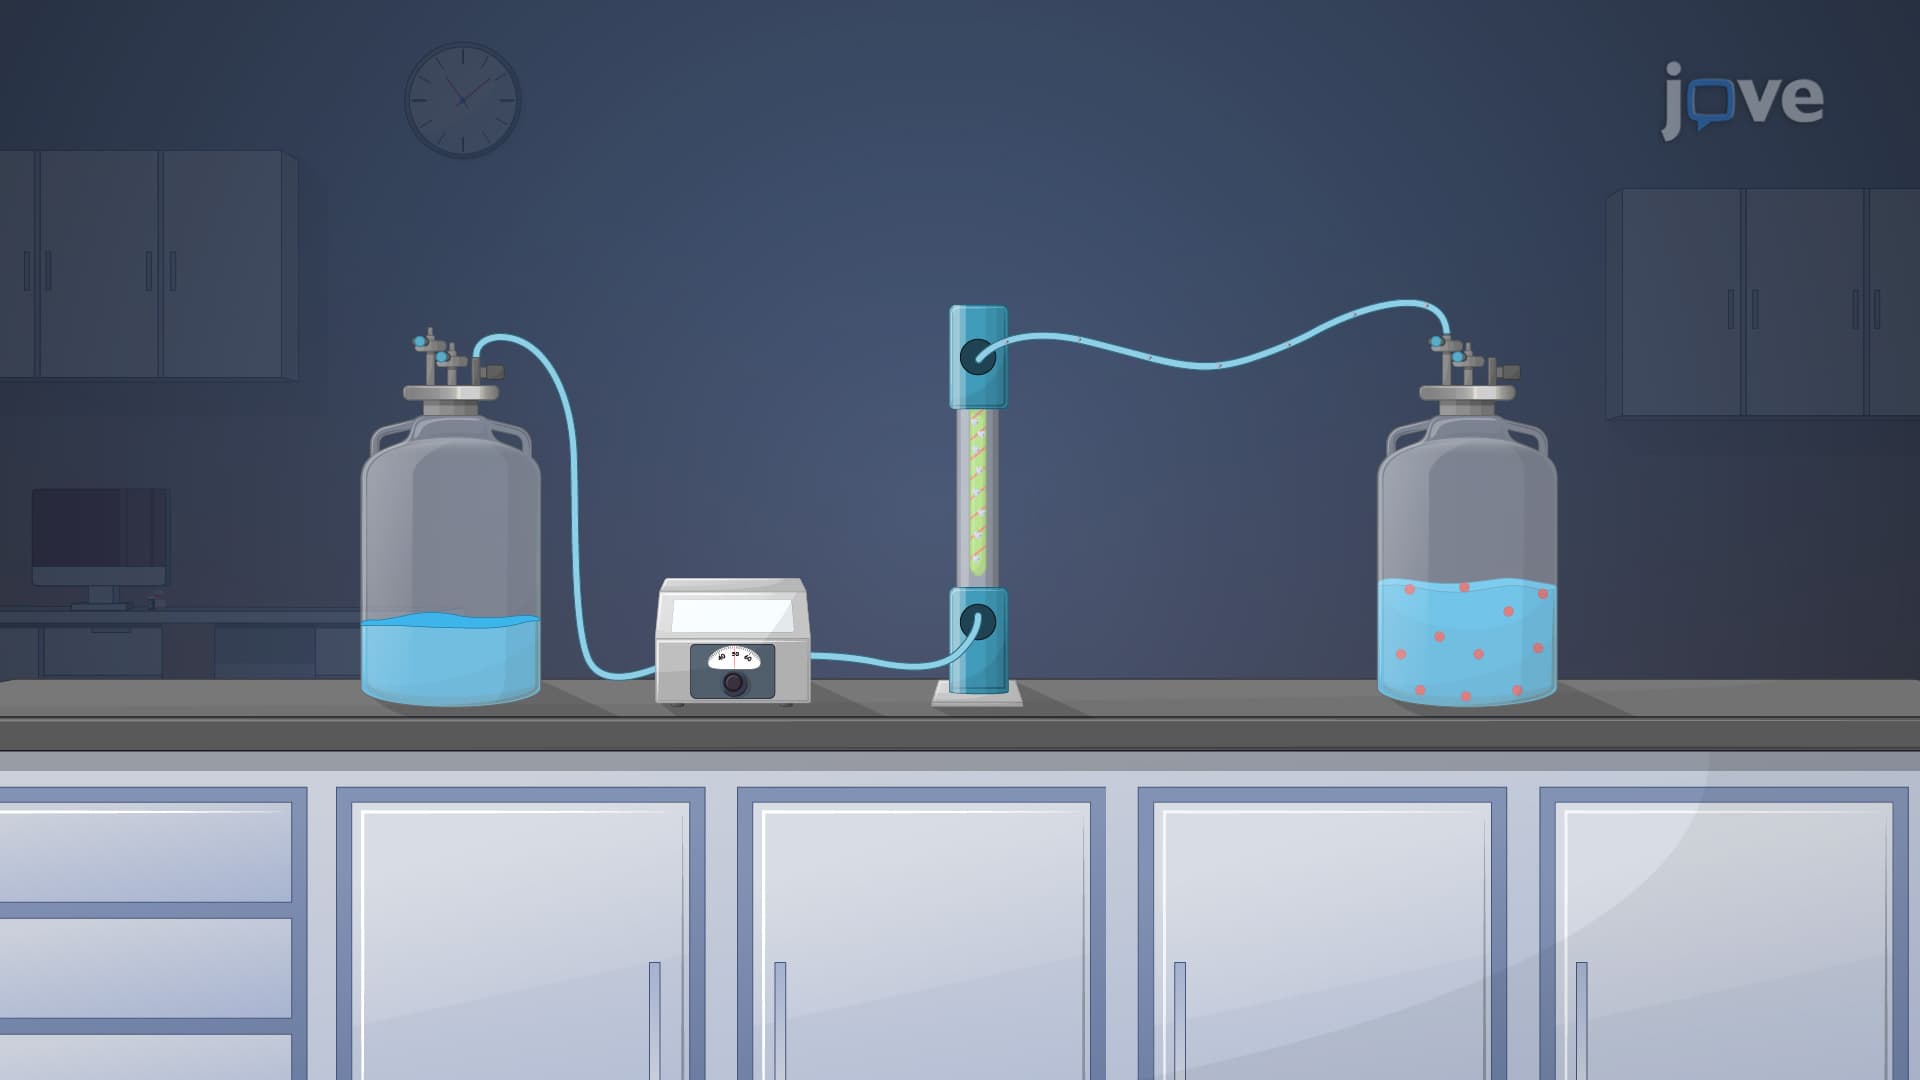
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 01:16
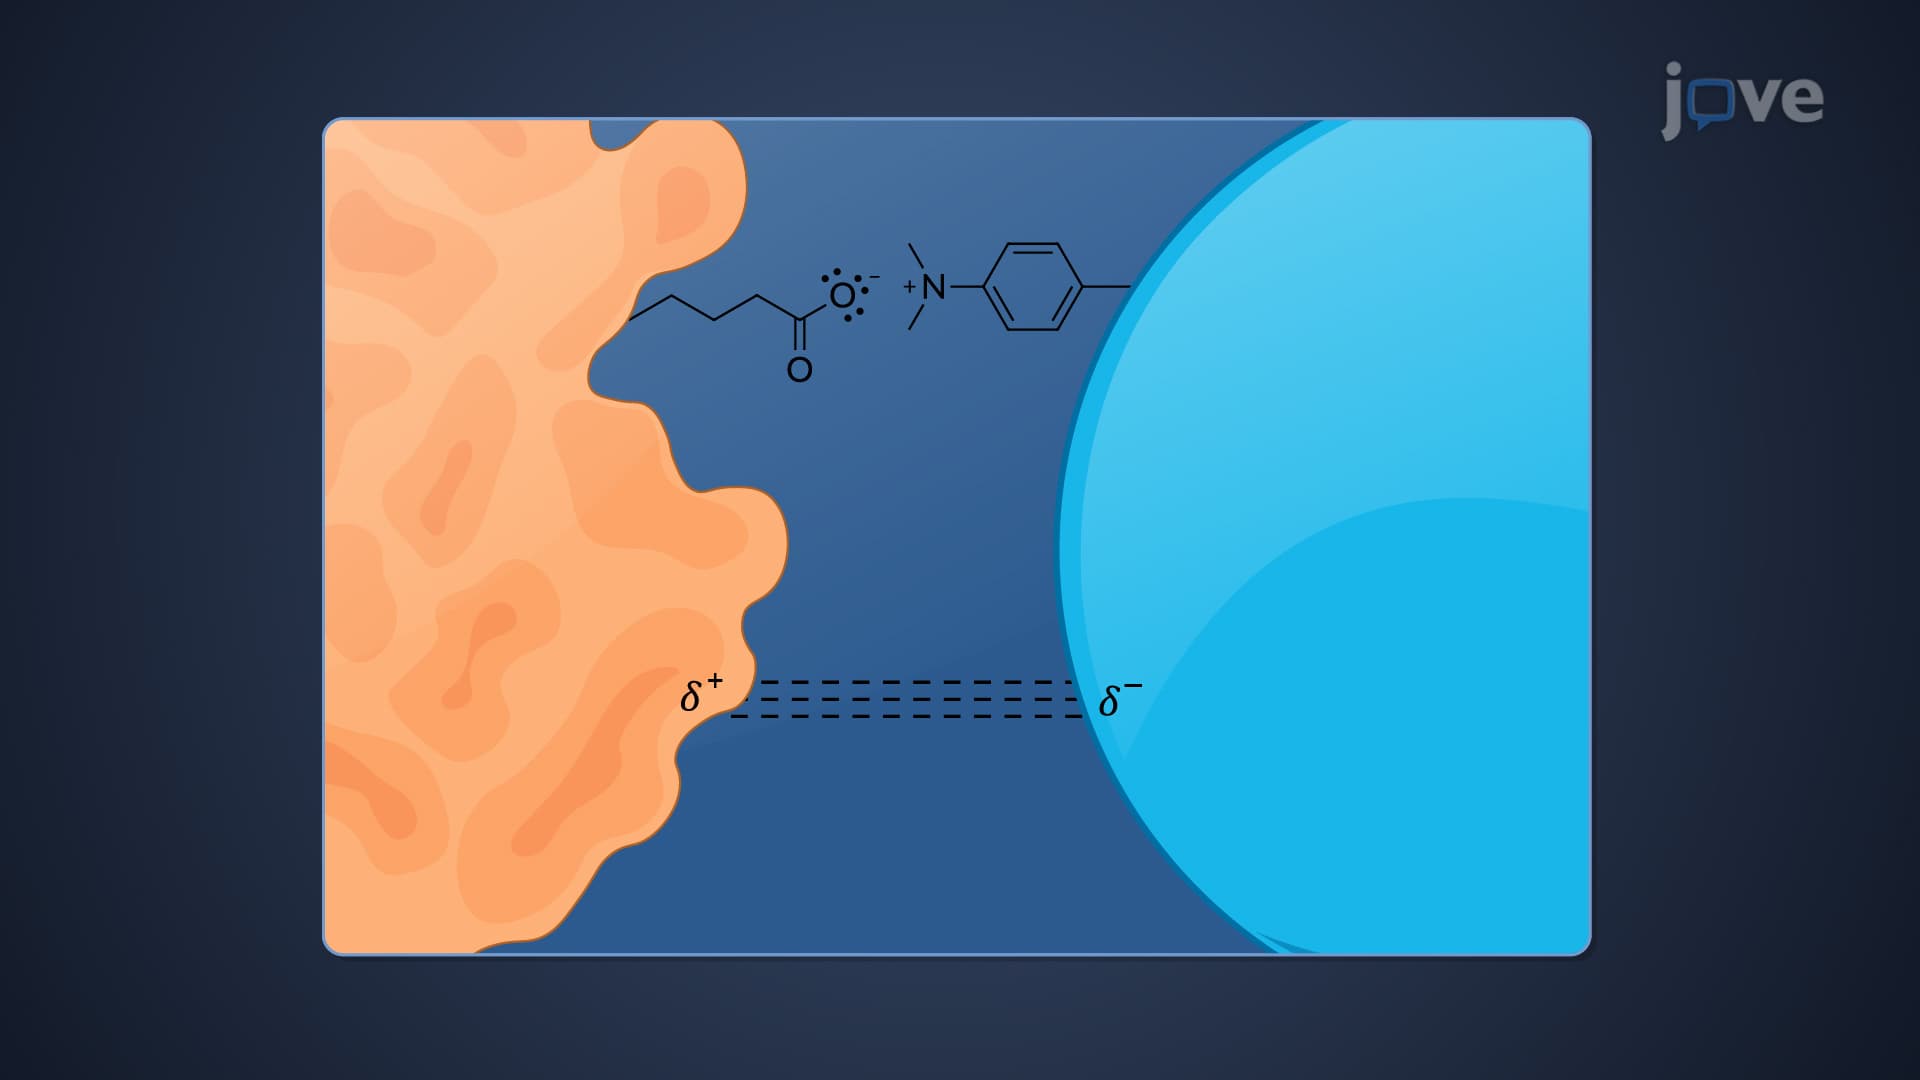
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 01:26
Drug Discovery: Overview
 02:26
Protein Networks
These interactions can be represented through maps depicting protein-protein interaction networks, represented as nodes and edges. Nodes are circles that are representative of a protein,...