Related Experiment Video
Updated: Oct 11, 2026
Preparation of Enantiopure Non-Activated Aziridines and Synthesis of Biemamide B, D, and epiallo-Isomuscarine
Published on: June 13, 2022
Yohimbine derivatives containing carboxylic acid and ester bioisosteres as potent ADRA2A antagonists
Artem Chayka1, Michal Česnek1, Erika Kužmová1
1Institute of Organic Chemistry and Biochemistry of the Czech Academy of Sciences, Flemingovo nám. 2, 6, Prague, 160 00, Czech Republic.

Abstract:
The α2A-adrenergic receptor (ADRA2A) has emerged as a promising target for precision medicine in metabolic and vascular disorders. However, the clinical translation of the prototypical antagonist, yohimbine, is hindered by poor subtype selectivity, leading to ADRA1A-mediated cardiovascular adverse events, and rapid, highly variable CYP2D6-mediated hepatic clearance. To overcome these liabilities, we designed and synthesized a novel series of yohimbine analogues by replacing the metabolically labile methyl carboxylate with potentially stable isosteric five-membered heterocycles or a nitrile group. Structure-activity relationship profiling revealed that while acidic bioisosteres yielded inactive compounds, non-ionizable methyl carboxylate bioisosteres exhibited robust ADRA2A antagonism with successfully abolished ADRA1A affinity. Notably, the 3-methyl-1,2,4-oxadiazole derivative 5 demonstrated highly potent ADRA2A antagonism (IC50 = 34 nM), exceptional off-target selectivity, and improved in vivo systemic exposure. Conversely, the 1,3-oxazole derivative 4 maintained favorable selectivity and metabolic stability while displaying a ∼20-fold reduction in membrane permeability, indicating a peripherally restricted profile. This bioisosteric strategy establishes a versatile chemical roadmap for developing highly selective, pharmacokinetically tunable ADRA2A modulators devoid of off-target cardiovascular risks.
Related Concept Videos
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of the aromatic...
 01:17
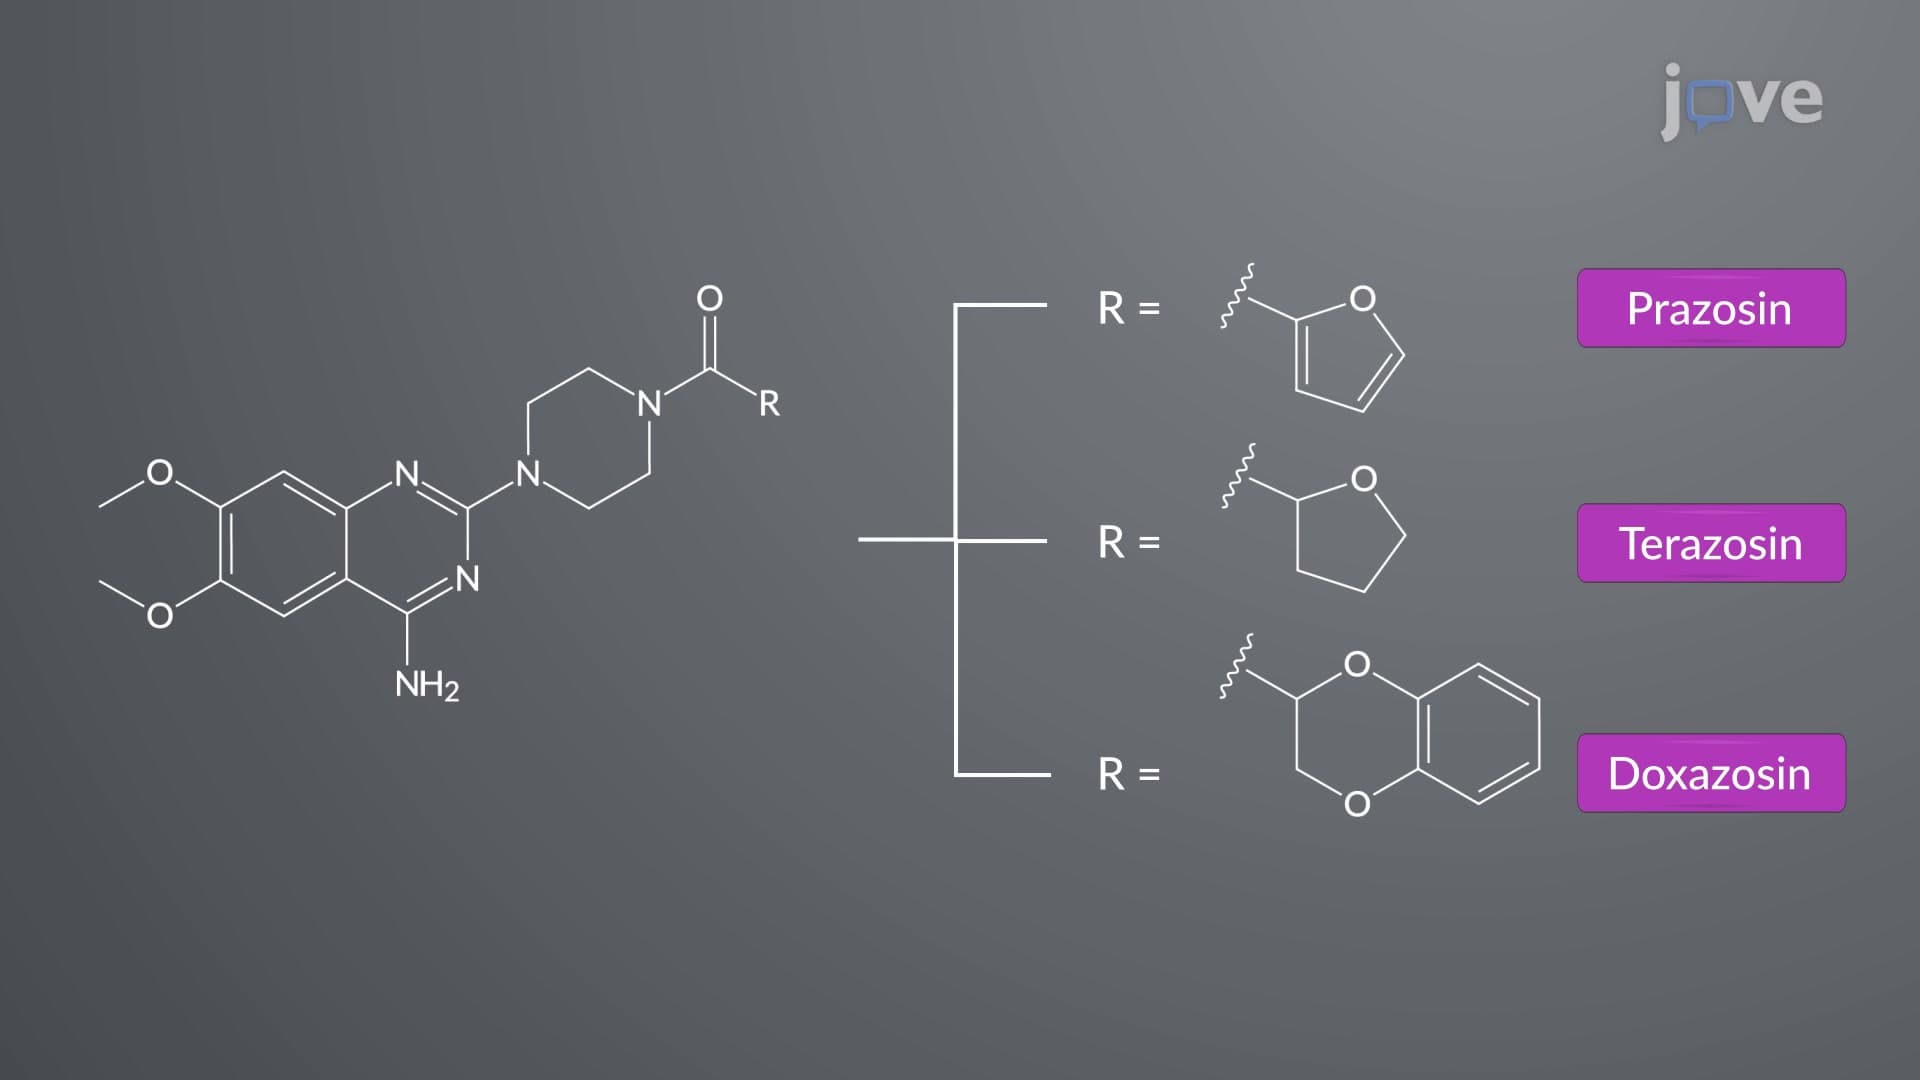
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline moieties. Phenoxybenzamine, with a haloalkylamine...
 01:25
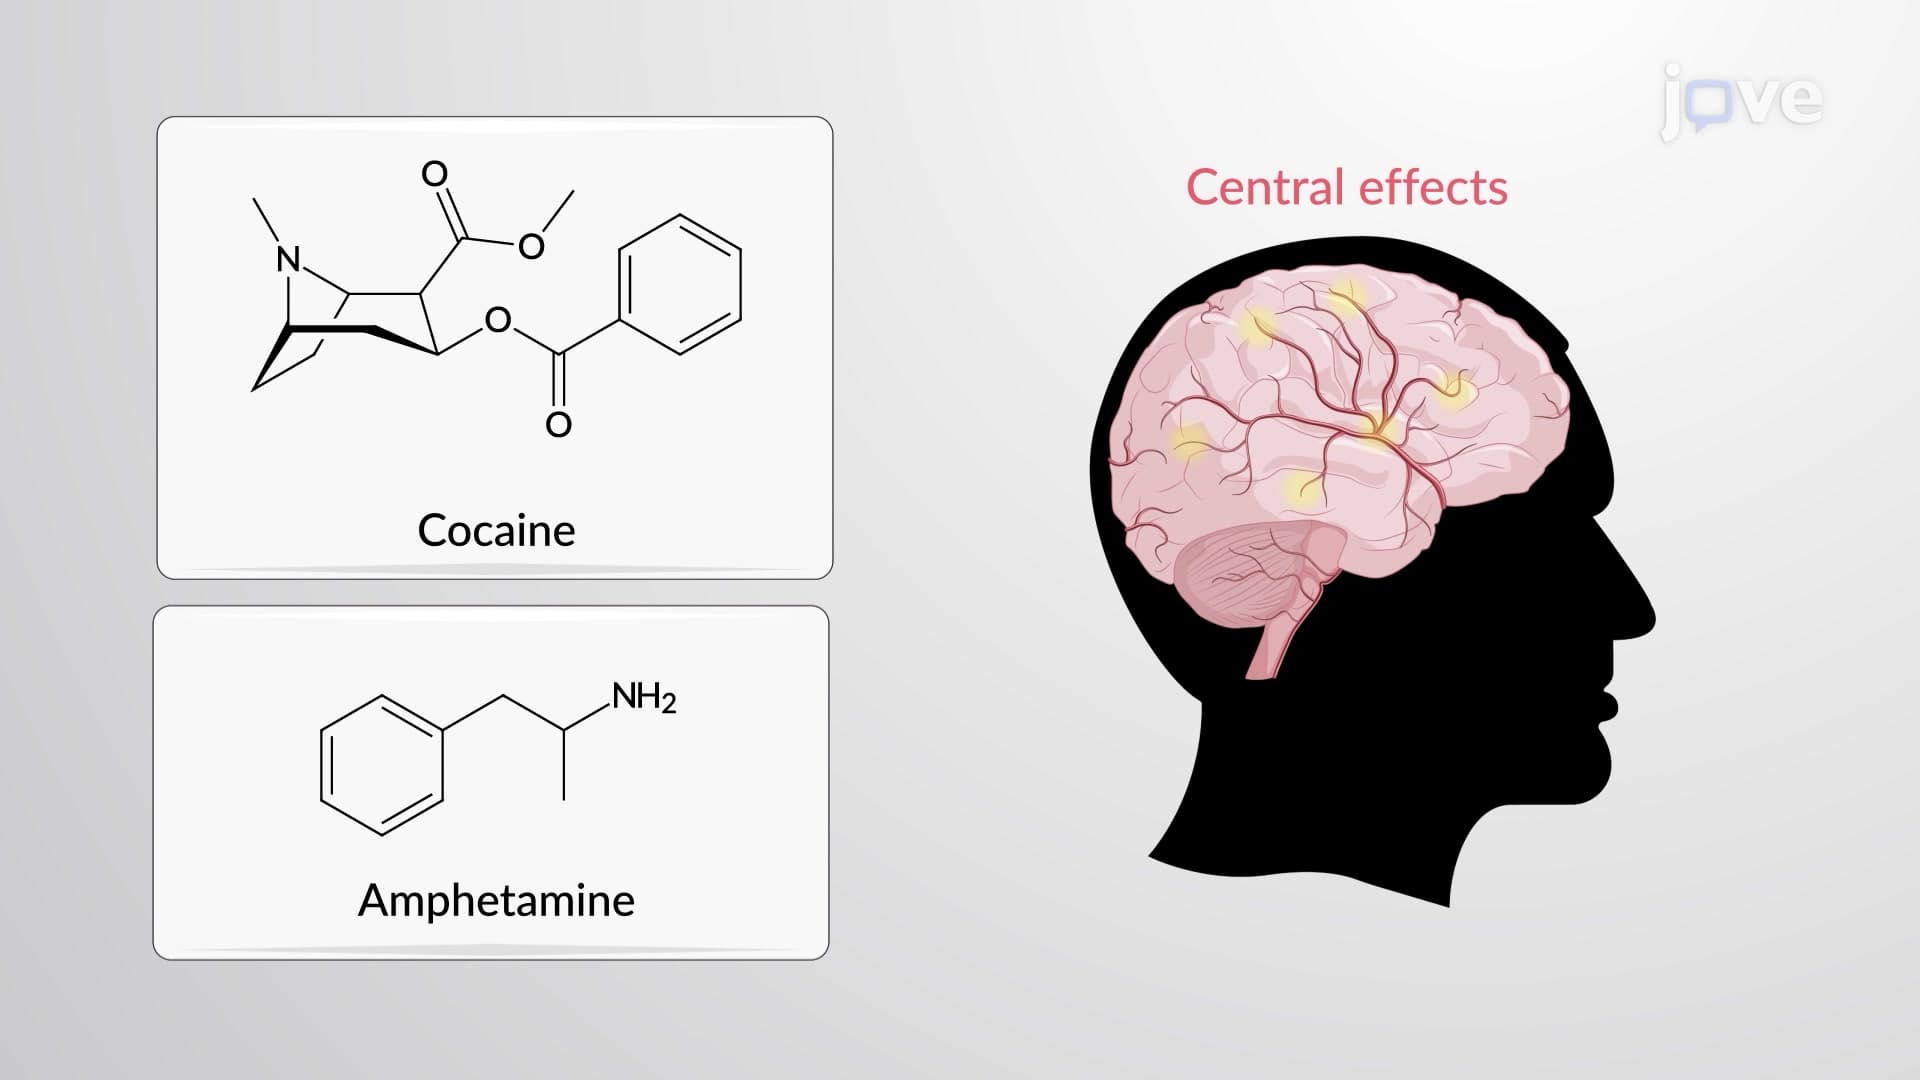
Adrenergic Agonists: Indirect-Acting Agents
One mechanism involves depleting stored catecholamines by displacing them from synaptic vesicles. These agents, known as "displacers," are transported into vesicles at the expense of noradrenaline. Examples include amphetamine and tyramine, which lack a catechol moiety, resulting in prolonged action, improved oral bioavailability, and...
 01:30
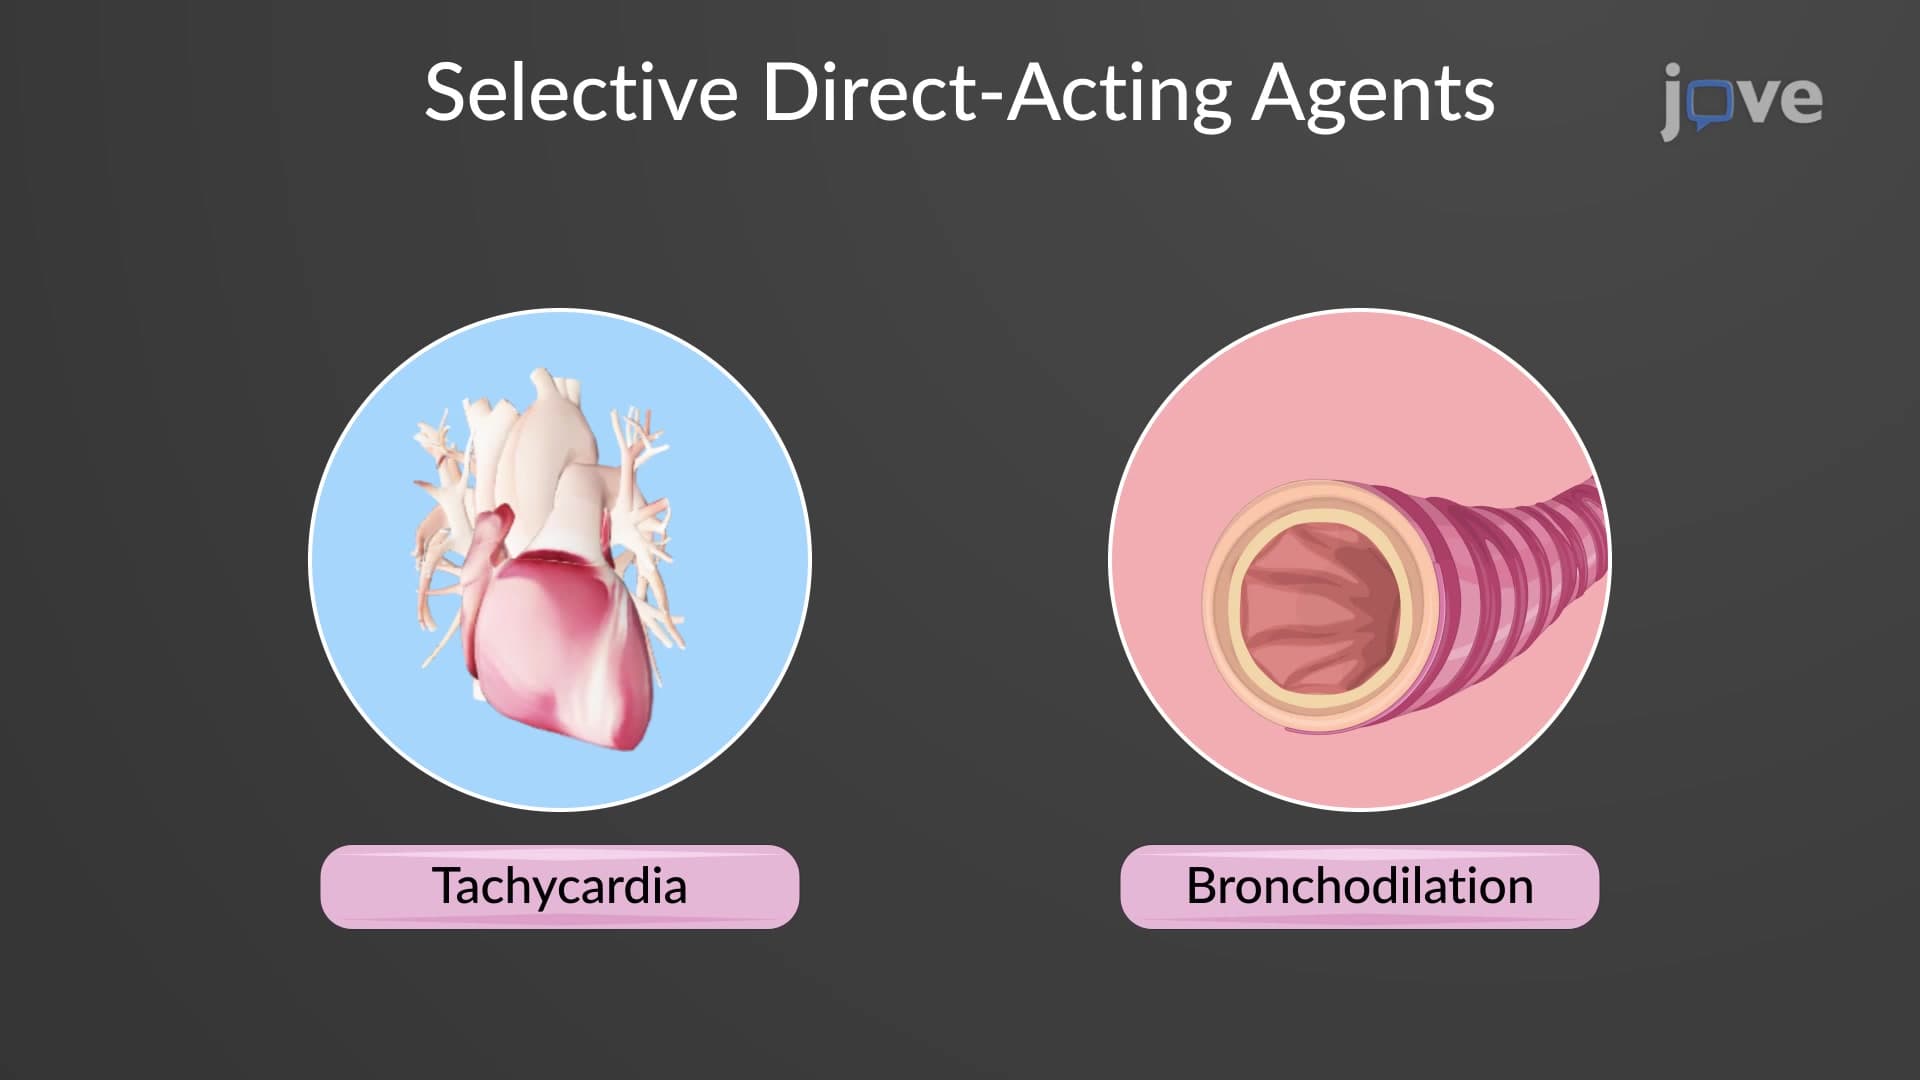
Adrenergic Agonists: Direct-Acting Agents
These agents can be classified...
 01:22
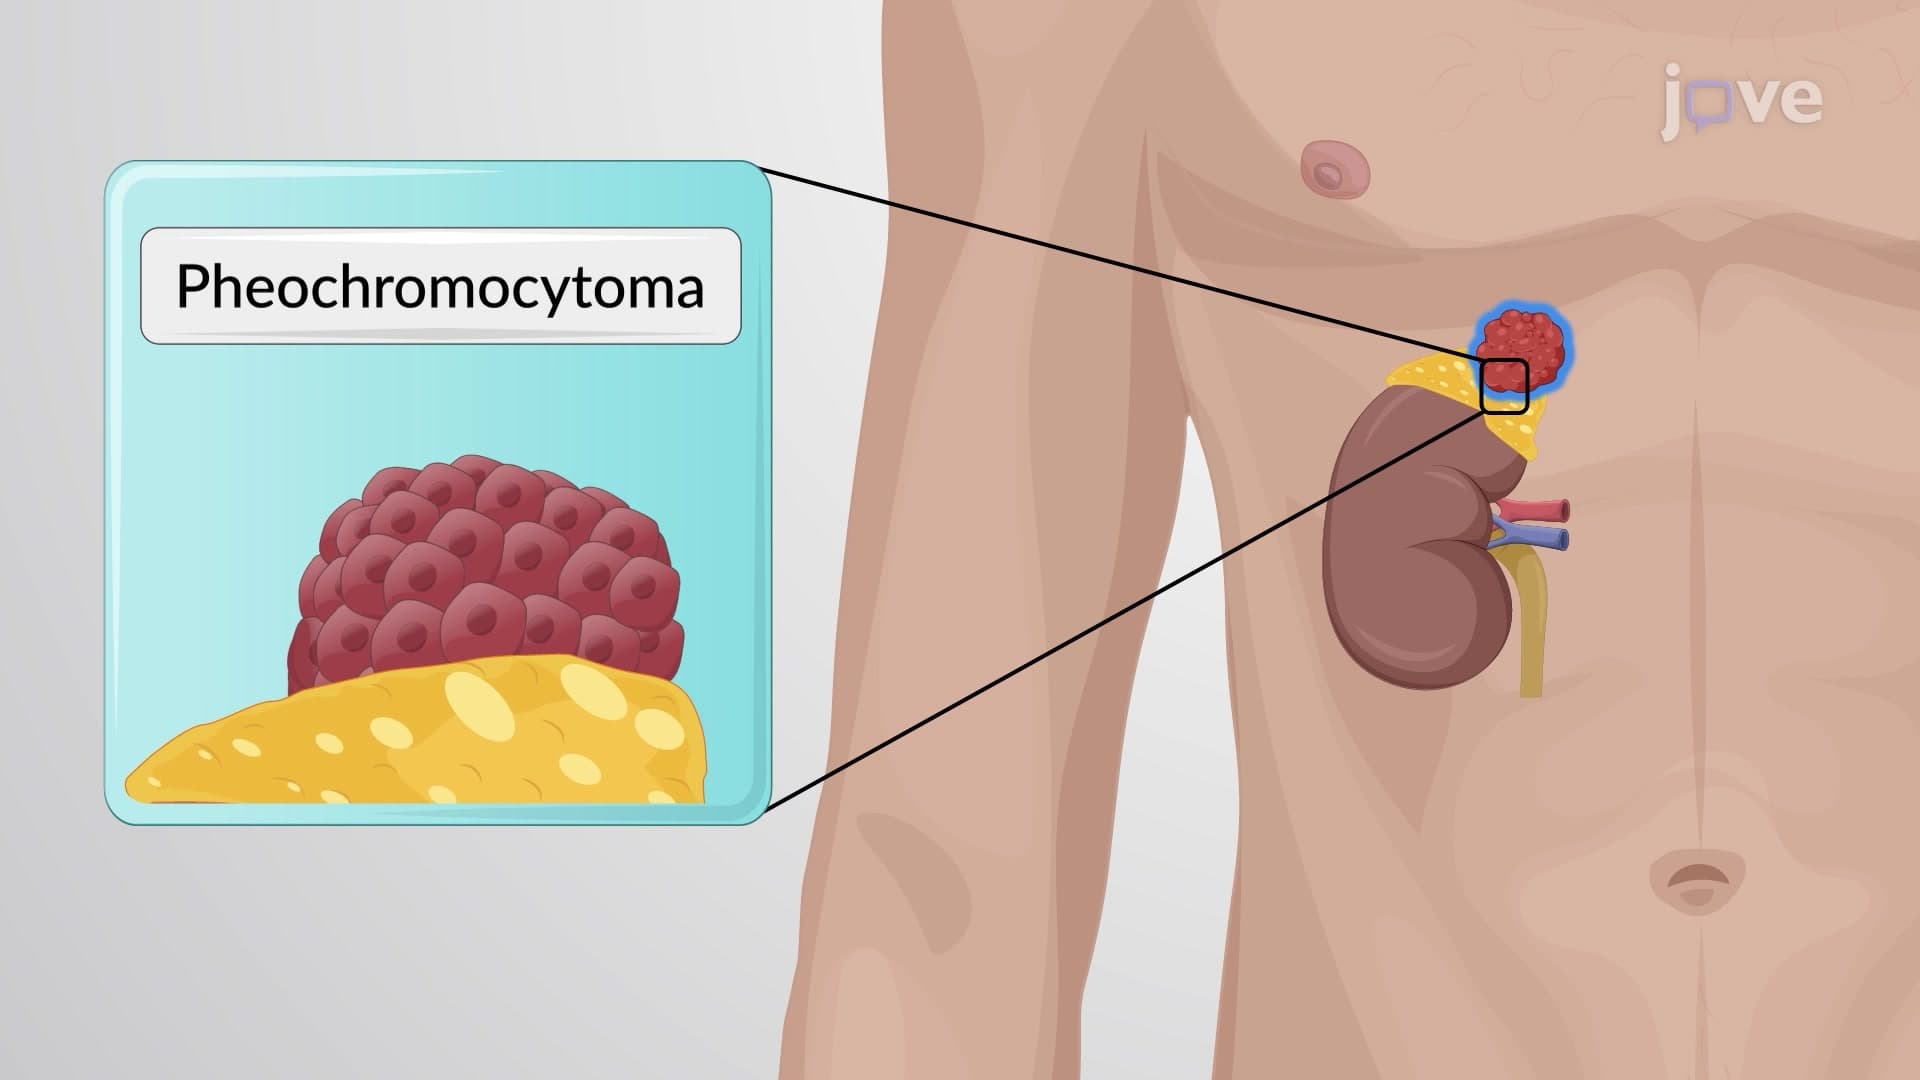
Adrenergic Antagonists: Pharmacological Actions of ɑ-Receptor Blockers
α1-blockers: These drugs inhibit α1-adrenoceptors on smooth muscle cells, resulting in vasodilation. This vasodilation lowers blood pressure, making α1-blockers valuable in treating hypertension. Additionally, α1-blockers effectively address urinary obstruction...
 01:25
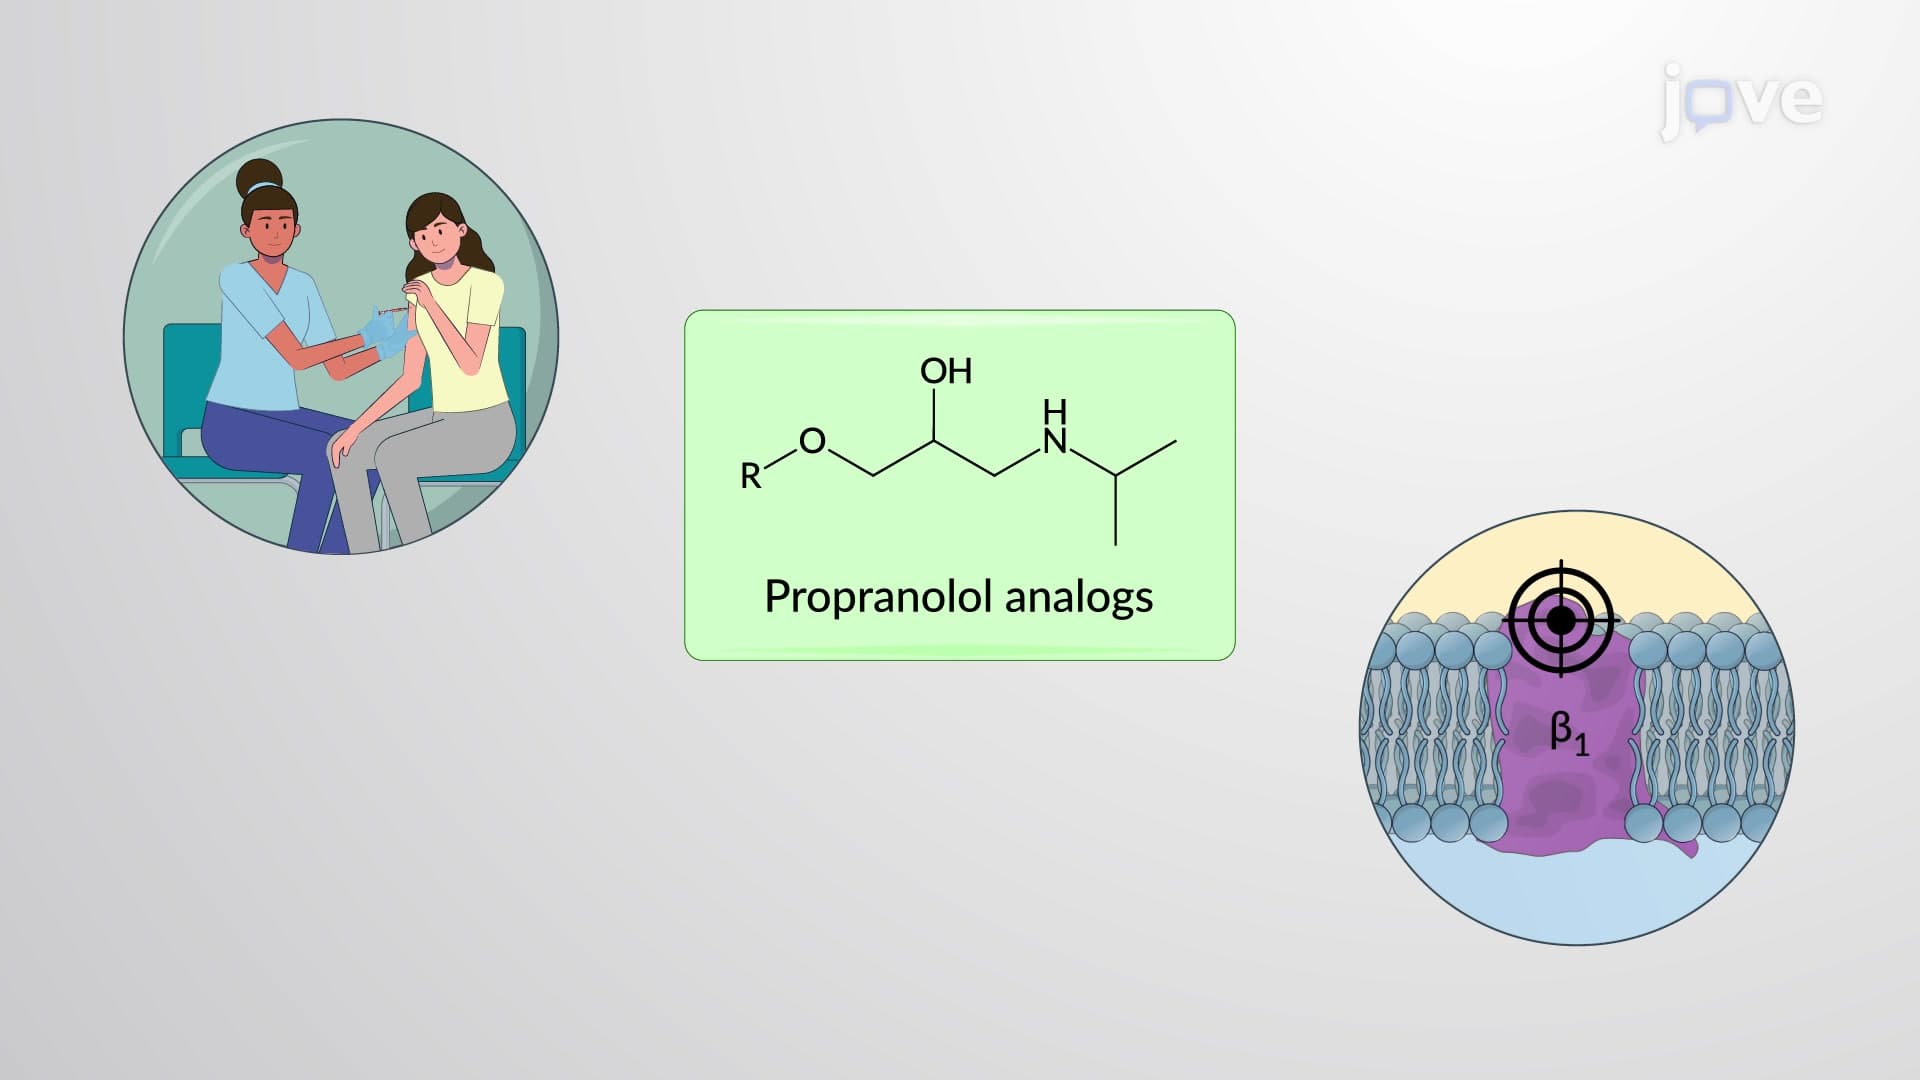
Adrenergic Antagonists: Chemistry and Classification of β-Receptor Blockers