Related Experiment Video
Updated: Oct 13, 2026

Assessing the Viability of a Synthetic Bacterial Consortium on the In Vitro Gut Host-microbe Interface
Published on: July 4, 2018
Integrated Quantitative and Relative Microbiome Profiling Reveals a Bacterial Group Associated With Primary Apical
Yefei Liu1,2, Yufeng Hu1,2, Mi Xu1,2
1Department of Endodontics, Shanghai Stomatological Hospital and School of Stomatology, Fudan University, Shanghai, China.

Aim:
To overcome the confounding effects of taxonomic heterogeneity in primary apical periodontitis (AP), this study aimed to characterize the quantitative and compositional profiles of the root canal microbiota associated with severe clinical features (percussion tenderness, large periapical lesions, and sinus tracts) using integrated relative (RMP) and quantitative microbiome profiling (QMP).
Methodology:
Root canal samples from 52 patients (out of 62 initially enrolled) with primary AP were analysed using spike-in-based 16S rRNA sequencing. Microbial load, diversity, and taxonomic composition were evaluated across these three predefined clinical features (with large lesions defined as ≥ 5 mm). Spearman correlation analysis was applied to assess the associations between differentially abundant genera and clinical severity. Furthermore, PICRUSt2 was used to infer functional profiles and calculate taxonomic contributions to enriched pathways under both RMP and QMP frameworks.
Results:
Samples with percussion tenderness exhibited significantly elevated microbial loads, with similar trends observed for large periapical lesions and sinus tracts. Percussion-positive samples also showed significantly higher Inverse Simpson index and Pielou's evenness index, whereas no significant differences in alpha diversity were observed for large periapical lesions or sinus tracts. Beta diversity did not differ significantly across any of the three clinical features under either QMP or RMP. Spearman correlation analysis identified a core bacterial group-Porphyromonas, Treponema, Tannerella, Peptostreptococcus, Selenomonas, Fusobacterium, and Fretibacterium-whose members showed broadly positive associations with all three clinical features in both relative and absolute abundance. Functional inference revealed these taxa collectively accounted for a shared set of virulence-associated pathways (e.g., lipopolysaccharide/peptidoglycan biosynthesis and protein export) consistently enriched in severe cases.
Conclusions:
Despite taxonomic heterogeneity in the root canal microbiota, severe clinical features of primary AP were associated with elevated microbial load and with increased relative and absolute abundances of a candidate bacterial group, whose members converged on a shared set of virulence-associated pathways, suggesting that compositionally diverse communities may nevertheless converge on common virulence-related functions.
Related Concept Videos
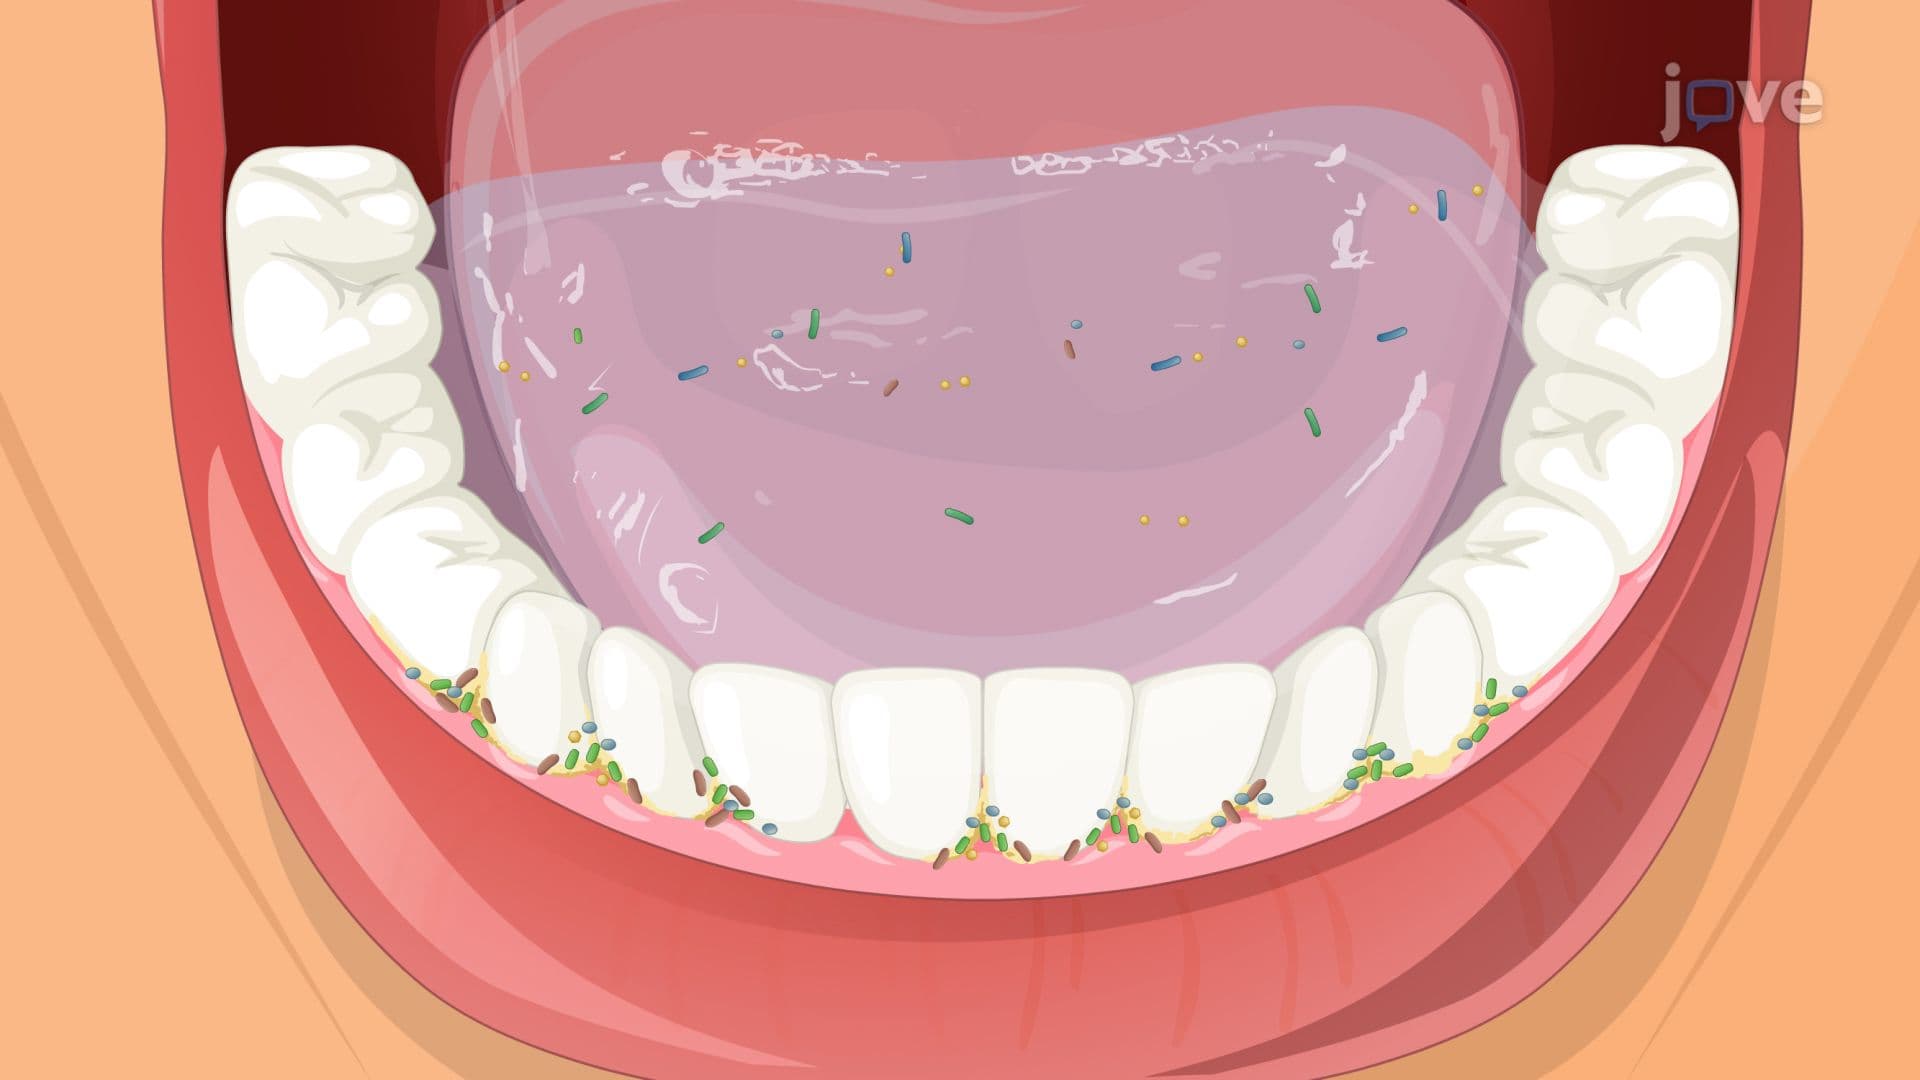 01:27
The Oral Microbiota
 01:20
Determinants of Bacterial Pathogenicity and Virulence
 01:28
Gene Regulation in Microbial Communities: Quorum Sensing
 01:29
Microbiota of the Respiratory Tract
 01:22
Introduction to the Human Microbiota
 01:28
Development of the Oral Microbiota