Related Experiment Video
Updated: Oct 14, 2026
Using Live Cell STED Imaging to Visualize Mitochondrial Inner Membrane Ultrastructure in Neuronal Cell Models
Published on: June 30, 2023
Role of Miro1 in Mitochondrial Dysfunction and Neurodegeneration in Huntington's Disease
Mustak Ahamed1, Khadga Raj Aran1
1Department of Pharmacy Practice, ISF College of Pharmacy, Moga, Punjab, 142001, India.

Introduction:
Huntington's Disease (HD) is an autosomal dominant neurodegenerative disorder that is caused by expansion of Cytosine-Adenine-Guanine (CAG) repeats in the Huntingtin (HTT) gene and leads to progressive motor, cognitive, and psychiatric dysfunction. Mitochondrial dysfunction is a major pathological feature of HD. Mitochondrial Rho GTPase 1 (Miro1) controls mitochondrial trafficking, calcium homeostasis, mitophagy, and neuronal signaling, and its dysregulation may contribute significantly to the progression of HD.
Methods:
The literature search to identify relevant studies on Miro1 biology, mitochondrial dysfunction, mitophagy, neuroinflammation, and emerging therapeutic strategies in HD followed a narrative review of the PubMed, Scopus, ScienceDirect, and Google Scholar databases. Original research articles, review articles, and preclinical studies published in English were screened for relevance, and the retrieved evidence was critically synthesized to provide an updated overview of Miro1's role in HD pathogenesis and its therapeutic potential.
Results:
Mutant Huntingtin (mHTT) perturbs Miro1-dependent mitochondrial transport, which leads to mitochondrial mislocalization, defective mitophagy, oxidative stress, and synaptic dysfunction. Miro1 dysregulation also facilitates neuroinflammatory signaling and neuronal vulnerability. Advances in AI have led to improved early diagnosis of HD through neuroimaging and digital biomarkers. Emerging therapies targeting Miro1, including gene therapy, nanocarriers, and pharmacological modulators, show promising neuroprotective potential in preclinical models.
Discussion And Conclusion:
Miro1 is a potentially important molecular link between mitochondrial dysfunction and neurodegeneration in HD. Targeting Miro1-mediated pathways may restore mitochondrial homeostasis and may be a promising therapeutic strategy for disease modification.
Related Concept Videos
 01:21
Huntington Disease l: Introduction
 01:19
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
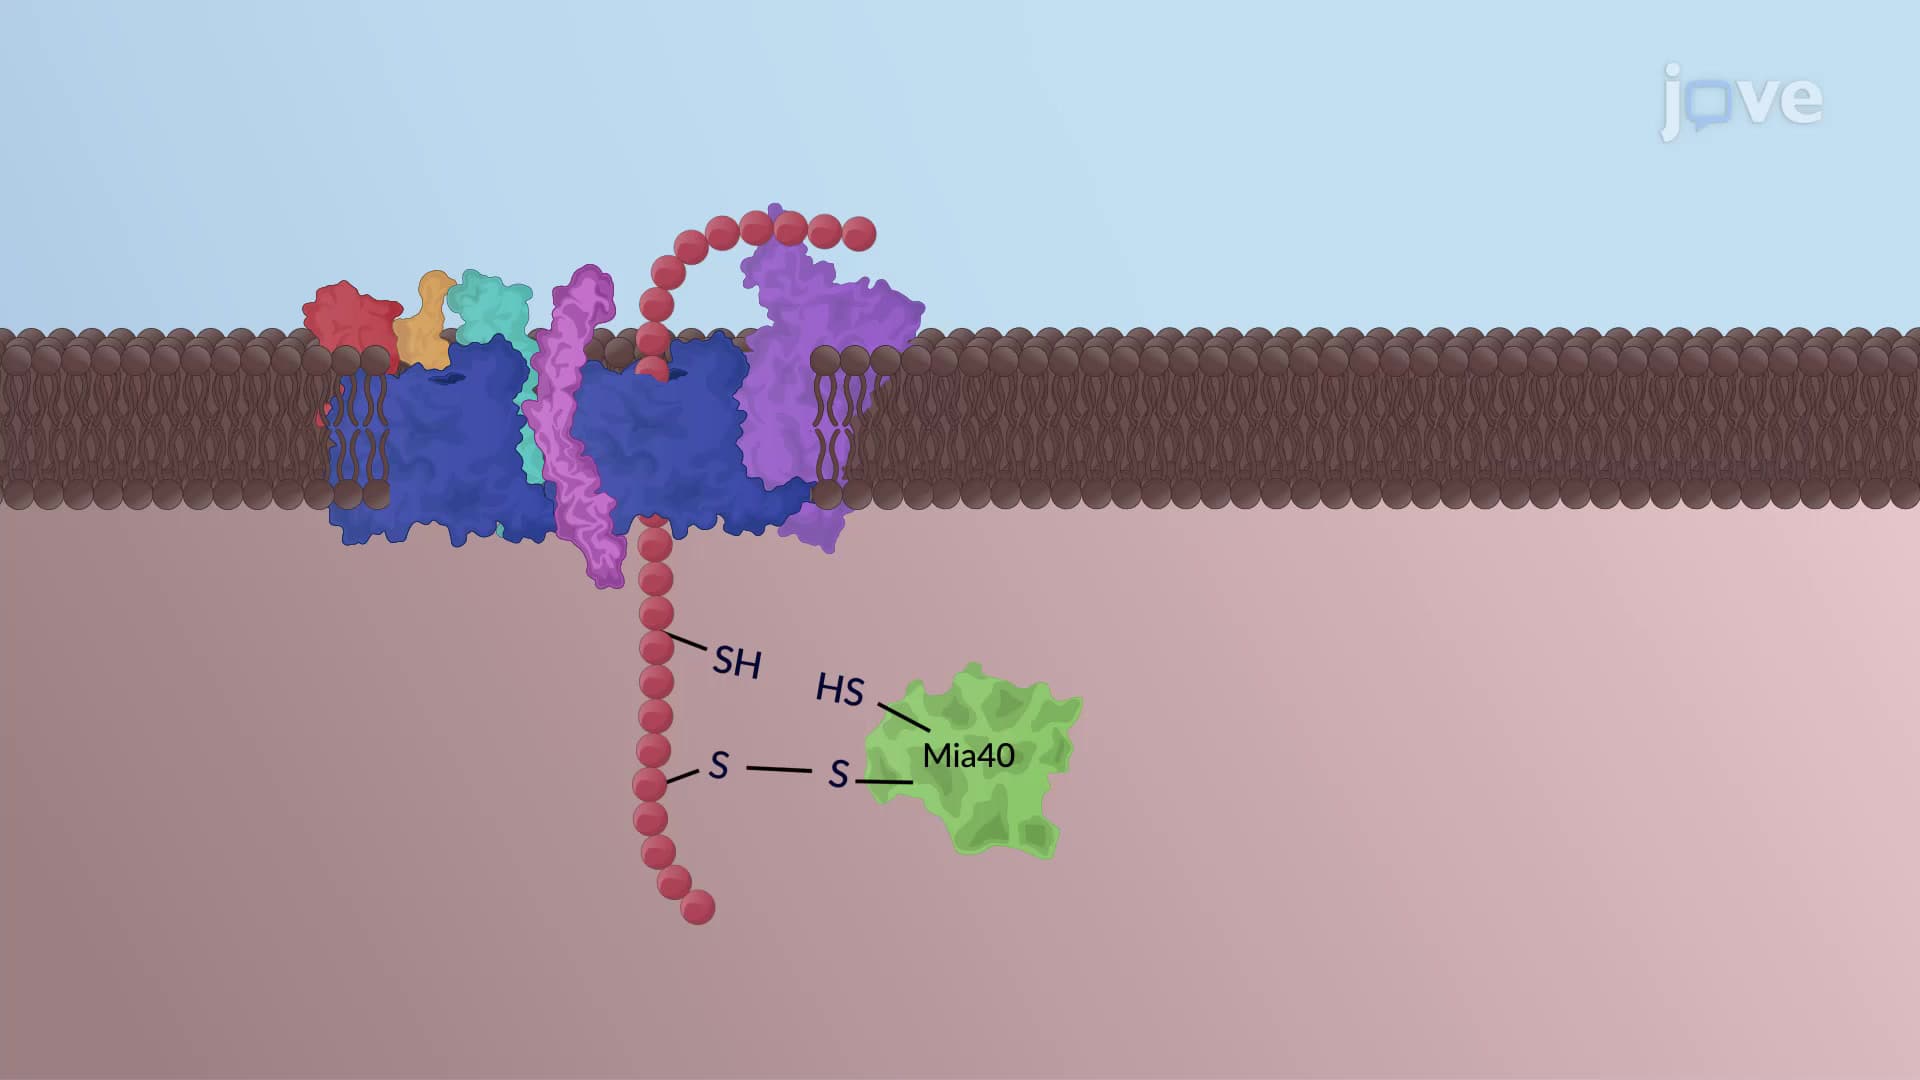
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...
 01:24
Parkinson Disease ll: Pathophysiology