Related Experiment Video
Updated: Oct 15, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
HIP: Hessian Interatomic Potentials without derivatives
Andreas Burger1,2, Luca Thiede1,2, Nikolaj Rønne3,4
1Department of Computer Science, University of Toronto, Toronto, Ontario, Canada.

Abstract:
Molecular Hessians, the second derivatives of the potential energy, are fundamental to many workflows in computational chemistry. Usually, accurate Hessians are computationally expensive to calculate and scale poorly with system size, whether computed using quantum chemistry methods or machine learning interatomic potentials. Here we introduce Hessian Interatomic Potentials (HIPs), a deep learning model that directly predicts Hessians without relying on automatic differentiation or finite differences. To do so, we construct SE(3)-equivariant, symmetric Hessians from irreducible representation (irrep) features up to degree l = 2, computed by a graph neural network. HIP Hessians are one to two orders of magnitude faster, more accurate, more memory efficient, easier to train and exhibit favorable scaling with system size. We validate our predictions across a wide range of downstream tasks, demonstrating consistently superior performance in transition state search, geometry optimization, zero-point energy corrections and vibrational analysis. We open source the HIP code and model weights.
More Related Videos

Related Concept Videos
 01:26
Thermodynamic Potentials
 03:24
Hybridization of Atomic Orbitals I
 03:35
Hybridization of Atomic Orbitals II
 01:26
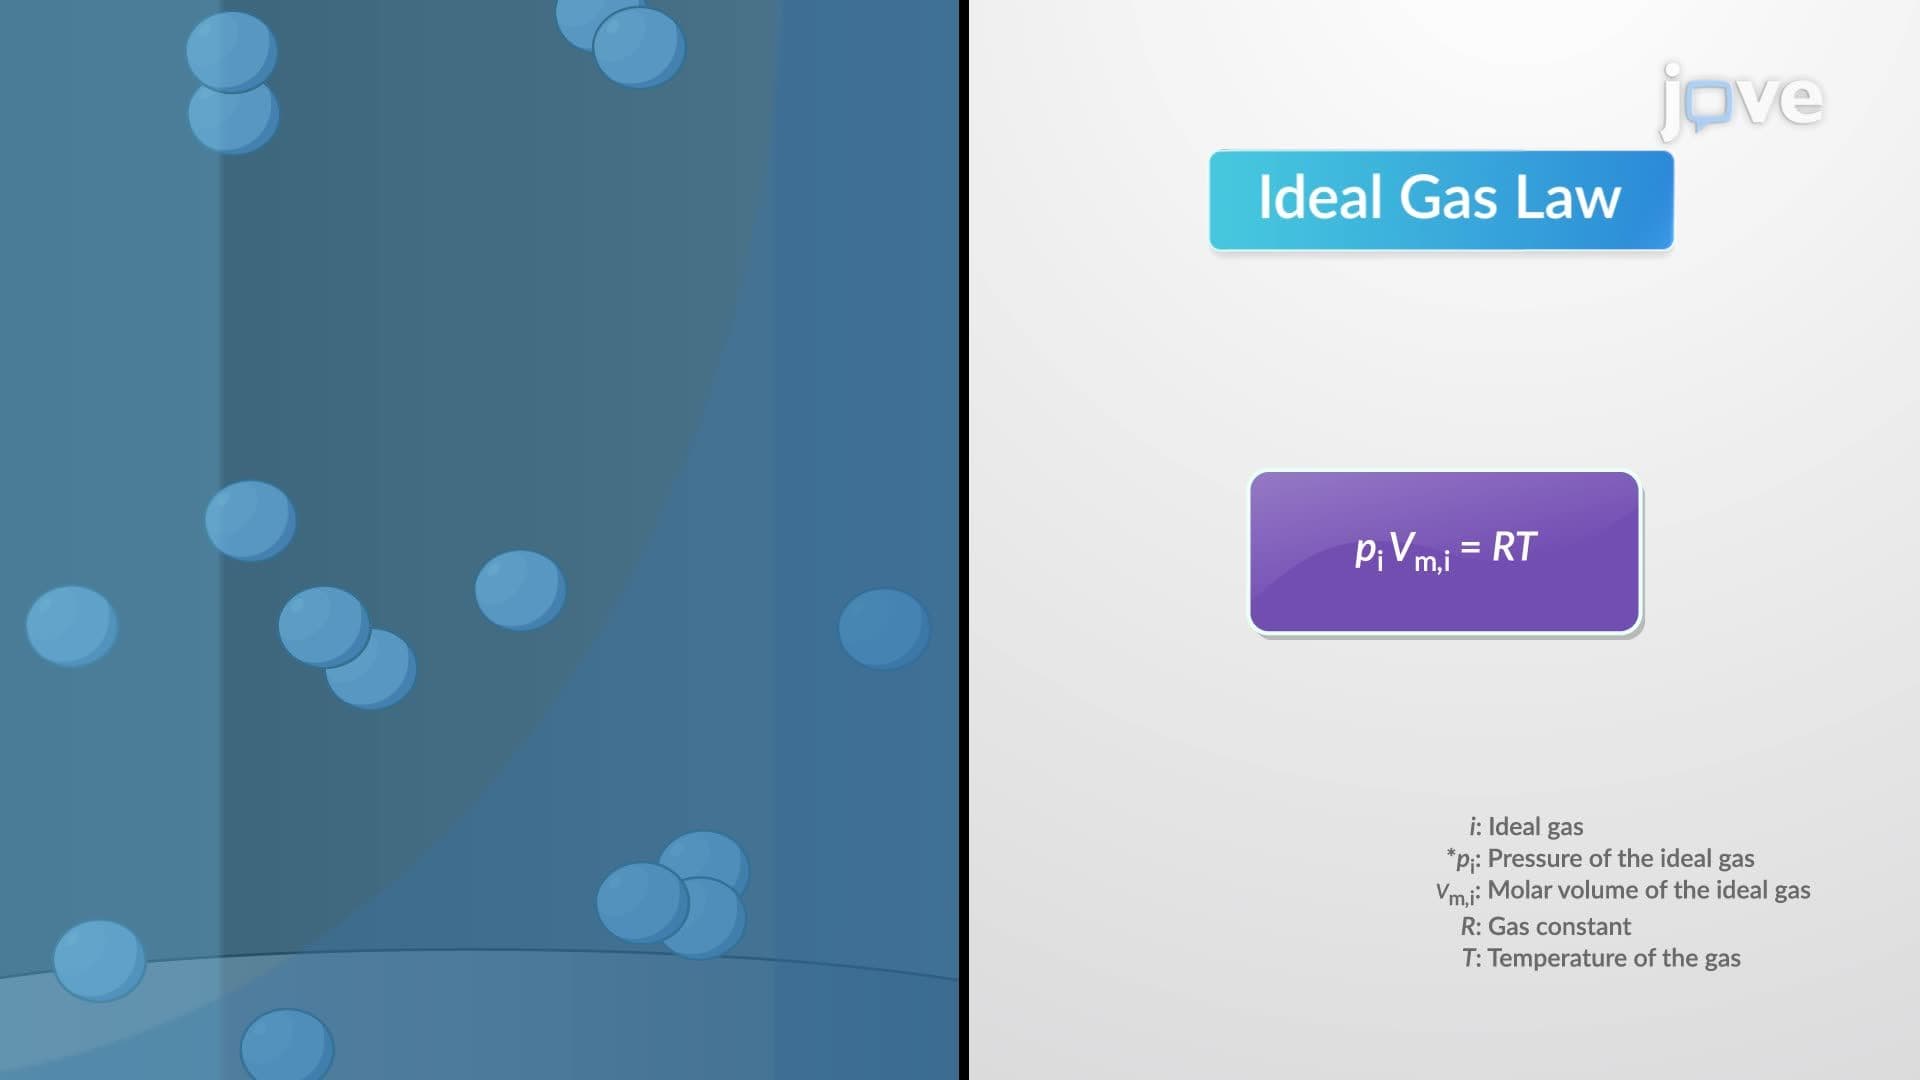
The Van der Waals Equation
 01:10
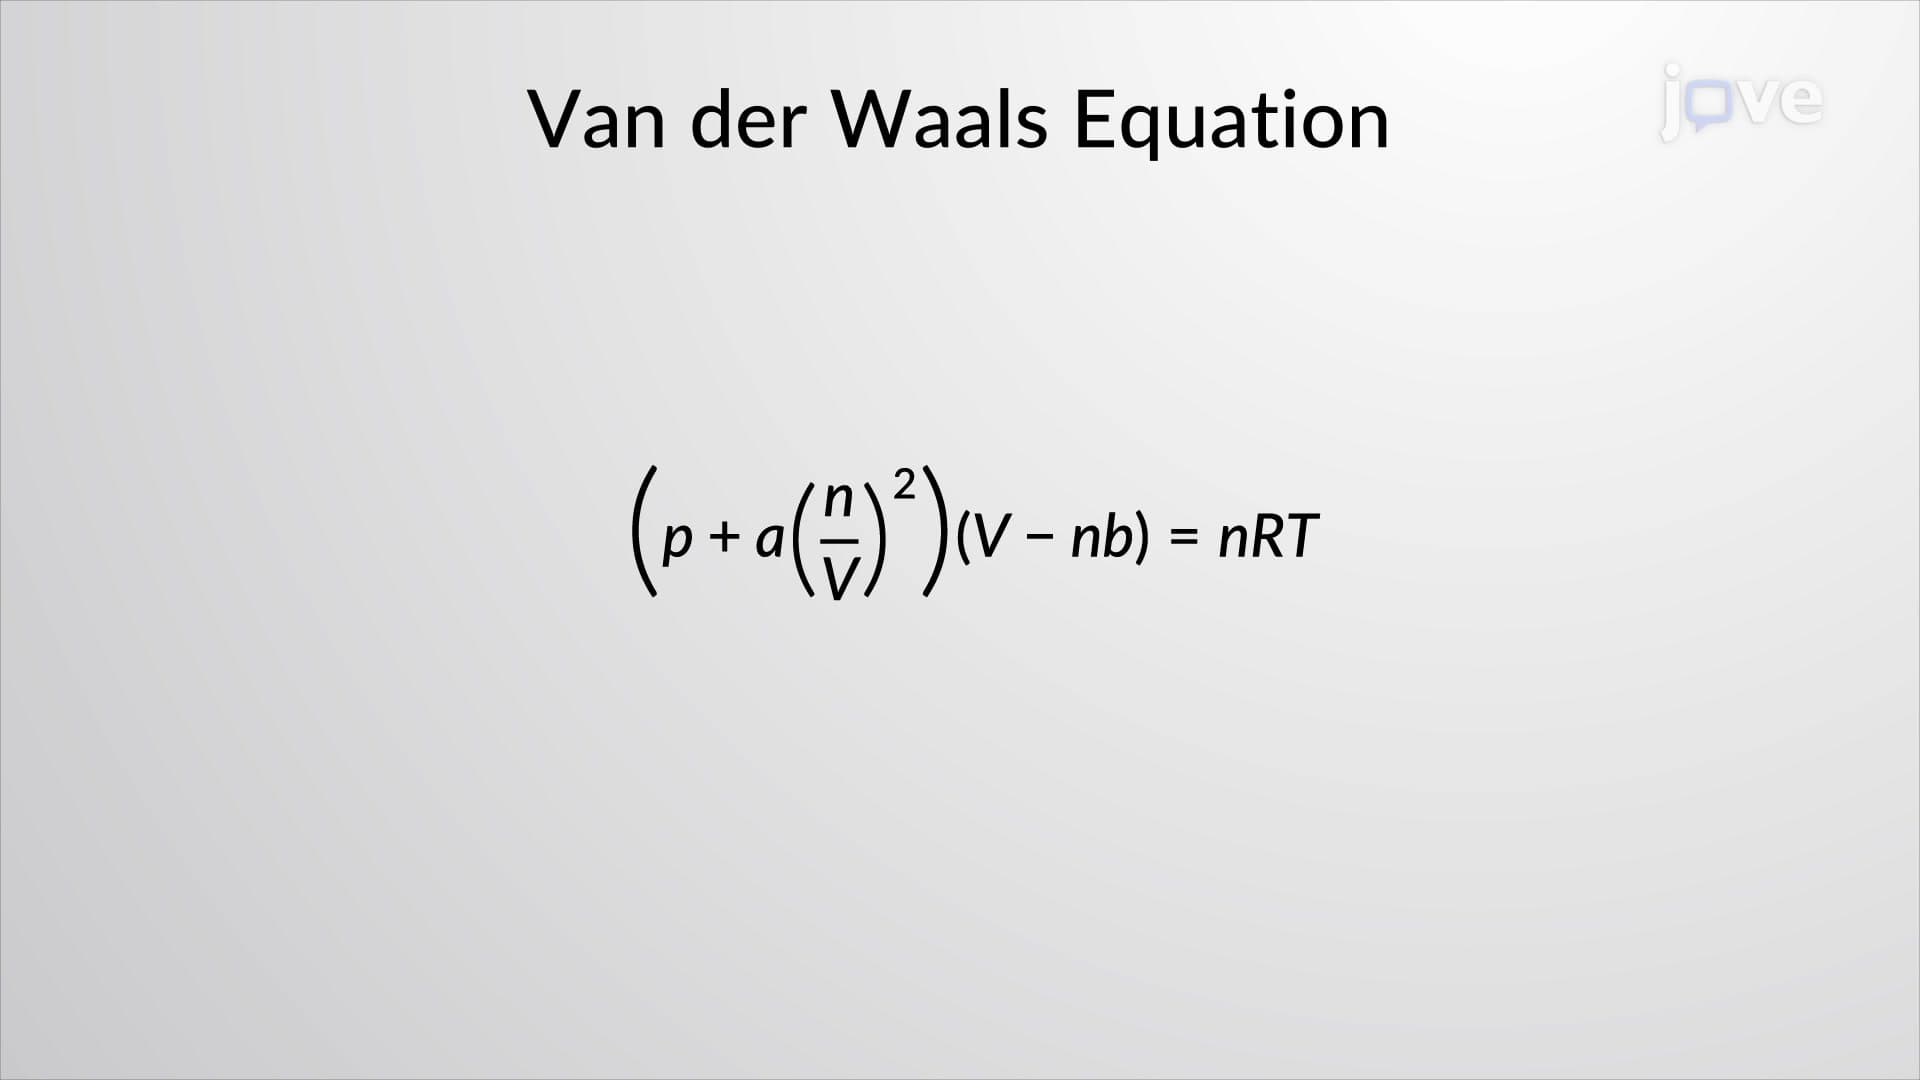
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 03:40
Hess's Law