Related Experiment Video
Updated: Oct 17, 2026
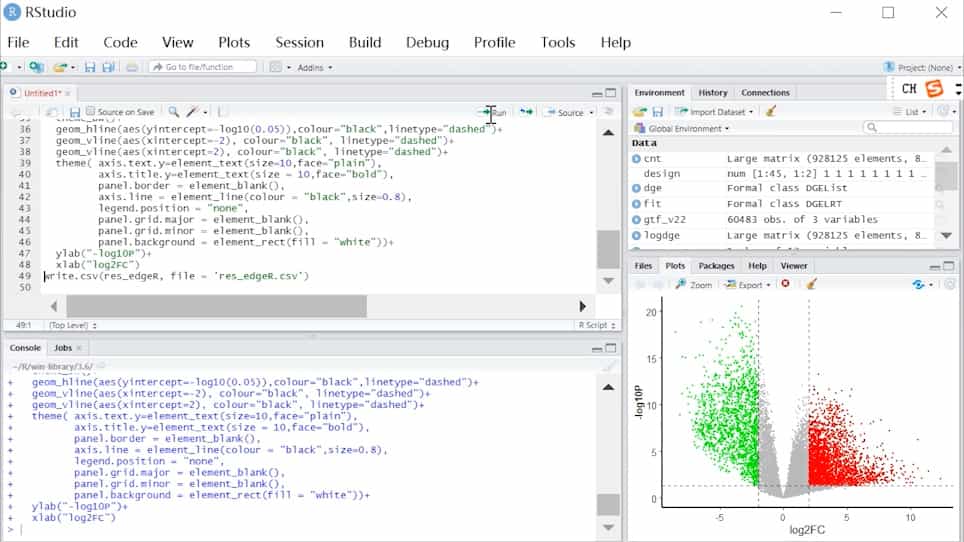
Three Differential Expression Analysis Methods for RNA Sequencing: limma, EdgeR, DESeq2
Published on: September 18, 2021
Assessing the impact of parental linear gene normalization on the performance of statistical models for circular RNA
Erda Qorri1,2, Valentin Varga1,3,4, Katalin Priskin2,5,6
1HUN-REN BRC Mutagenesis and Carcinogenesis Research Group, Institute of Genetics, HUN-REN Biological Research Centre, Szeged, Temesvári krt. 62, 6726, Hungary.

Background:
Circular RNAs (circRNAs) have emerged as promising non-invasive cancer biomarkers due to their stability, abundance in body fluids, and regulatory potential. However, circRNA differential expression analysis (DEA) remains challenging, owing to a lack of consensus on important preprocessing strategies such as filtering and normalization. While well-established bulk RNA-sequencing frameworks are commonly applied to circRNA data, newer approaches such as CIRI-DE (part of CIRI3 suite) integrate linear and circular transcript information to improve detection. Despite developments, an assessment of these integrative strategies is lacking, and the critical impact of filtering on DEA model performance has not been comprehensively evaluated.
Results:
In this study, we evaluated the impact of multiple normalization and filtering strategies on circRNA DEA using experimental datasets, including two in-house blood platelet sets and semi-parametric simulated in silico datasets. Our results emphasize the importance of selecting an appropriate filtering threshold, as overly lenient filtering can reduce model performance across datasets. We found edgeR's filterByExpr() strategy particularly effective in handling zero counts in circRNA data, while also generating the most reliable results across most datasets.Furthermore, by incorporating linear and circular information as described in CIRI-DE, most methods identified a higher number of DE circRNAs compared to circular counts alone. Notably, circRNAs identified by both CIRI-DE and the modified bulk RNA-sequencing pipelines showed substantial overlap.
Conclusion:
Our findings demonstrate that automated filtering combined with linear-aware normalization significantly enhances the sensitivity and reproducibility of circRNA DEA, providing a standardized framework for more reliable biomarker discovery in transcriptomic research.