Related Experiment Video
Updated: Oct 17, 2026
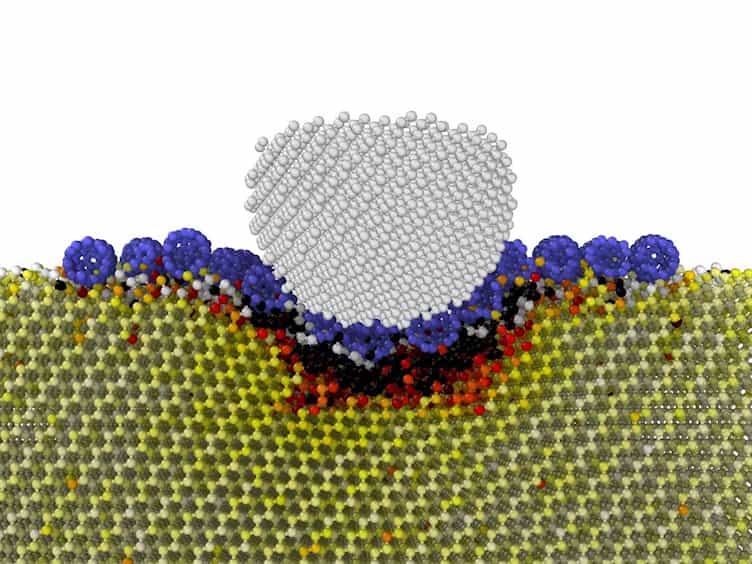
Probing C84-embedded Si Substrate Using Scanning Probe Microscopy and Molecular Dynamics
Published on: September 28, 2016
Atomistic-to-Macroscopic Mechanical Modeling of (HfNbTaZr)B2 High-Entropy Ceramics via a Developed Interatomic
Yihan Wu1, Yuan-Xin Dai2, Xingjie Chen3
1School of Mechanical Engineering and Automation, Fuzhou University, Fuzhou 350108, China.

Abstract:
Atomistic investigations of high-entropy diborides (HEBs) are currently hindered by the lack of reliable interatomic potentials. Here, we develop an analytical bond-order potential (ABOP) for the Hf/Nb/Zr/Ta/B system, parameterized via an intelligent global optimization algorithm and trained against extensive ab initio data. The developed potential accurately captures the structural, elastic, and mechanical properties of various materials, including metals, intermetallics, alloys and diborides. Additionally, the chemical short-range order (CSRO) in dynamic simulations under finite temperatures is correctly predicted. Molecular dynamics (MD) simulations reveal that HEB nanopillars exhibit brittle failure driven by shear banding, while CSRO shows limited influence on the mechanical response. Simulated nanoindentation gives modulus (435 GPa) and hardness (27.7 GPa) consistent with theoretical predictions and experiments. Furthermore, we establish a multiscale framework by extracting an atomic traction-separation law and rescaling it based on fracture energy conservation (Gc = 21.01 J/m2) for finite element cohesive zone simulations. This approach captures the brittle intergranular failure of polycrystalline HEBs, effectively bridging atomic-level mechanisms with macroscopic property predictions.

Related Concept Videos
 02:00
Molecular Models
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:26
Thermodynamic Potentials
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...