Related Experiment Video
Updated: Oct 22, 2026

Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Modeling Reactions on the Solid-Liquid Interface with Next Generation Extended Lagrangian Quantum-Based Molecular
Rae A Corrigan Grove1, Kevin G Kleiner2, Joshua Finkelstein1
1Theoretical Division, Los Alamos National Laboratory, Los Alamos, New Mexico 87545, United States.

Abstract:
We present a series of simulations of the oxygen reduction reaction (ORR) using a novel framework for atomistic simulations of surface catalysis under electrochemical bias. The framework makes use of quantum-mechanical extended Lagrangian Born-Oppenheimer molecular dynamics (XL-BOMD) simulations, which provide the speed and accuracy required for explicit atomistic treatment of both electrode and electrolyte. Simulations of solvated O2 near nitrogen-doped graphene (NG) were performed to gain insight into the ORR, and different mechanisms were observed, depending on the applied bias. Under higher bias the ORR occurred by an outer-sphere mechanism, without adsorption of O2 to NG. In this mechanism, electron transfer between the catalyst and the O2 was mediated by the solvent. Under lower bias the ORR occurred by an inner-sphere mechanism involving adsorption of O2 to NG, leading to direct electron transfer. Our extensive, all-atom quantum-mechanical molecular dynamics simulations also show clear differences between the kinetics of the ORR on this ideally polarizable electrode and commonly used kinetic theories, leading to new insights regarding mechanistic changes with varied overpotentials. Combining quantum accuracy with explicit solvation and electrostatic potential bias, XL-BOMD opens a route to predictive, atomistic insight into electrocatalytic processes, as demonstrated with the ORR.
More Related Videos
 07:31
Realistic Membrane Modeling Using Complex Lipid Mixtures in Simulation Studies
Published on: September 1, 2023
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 01:29
Liquid–Solid Solutions
 01:26
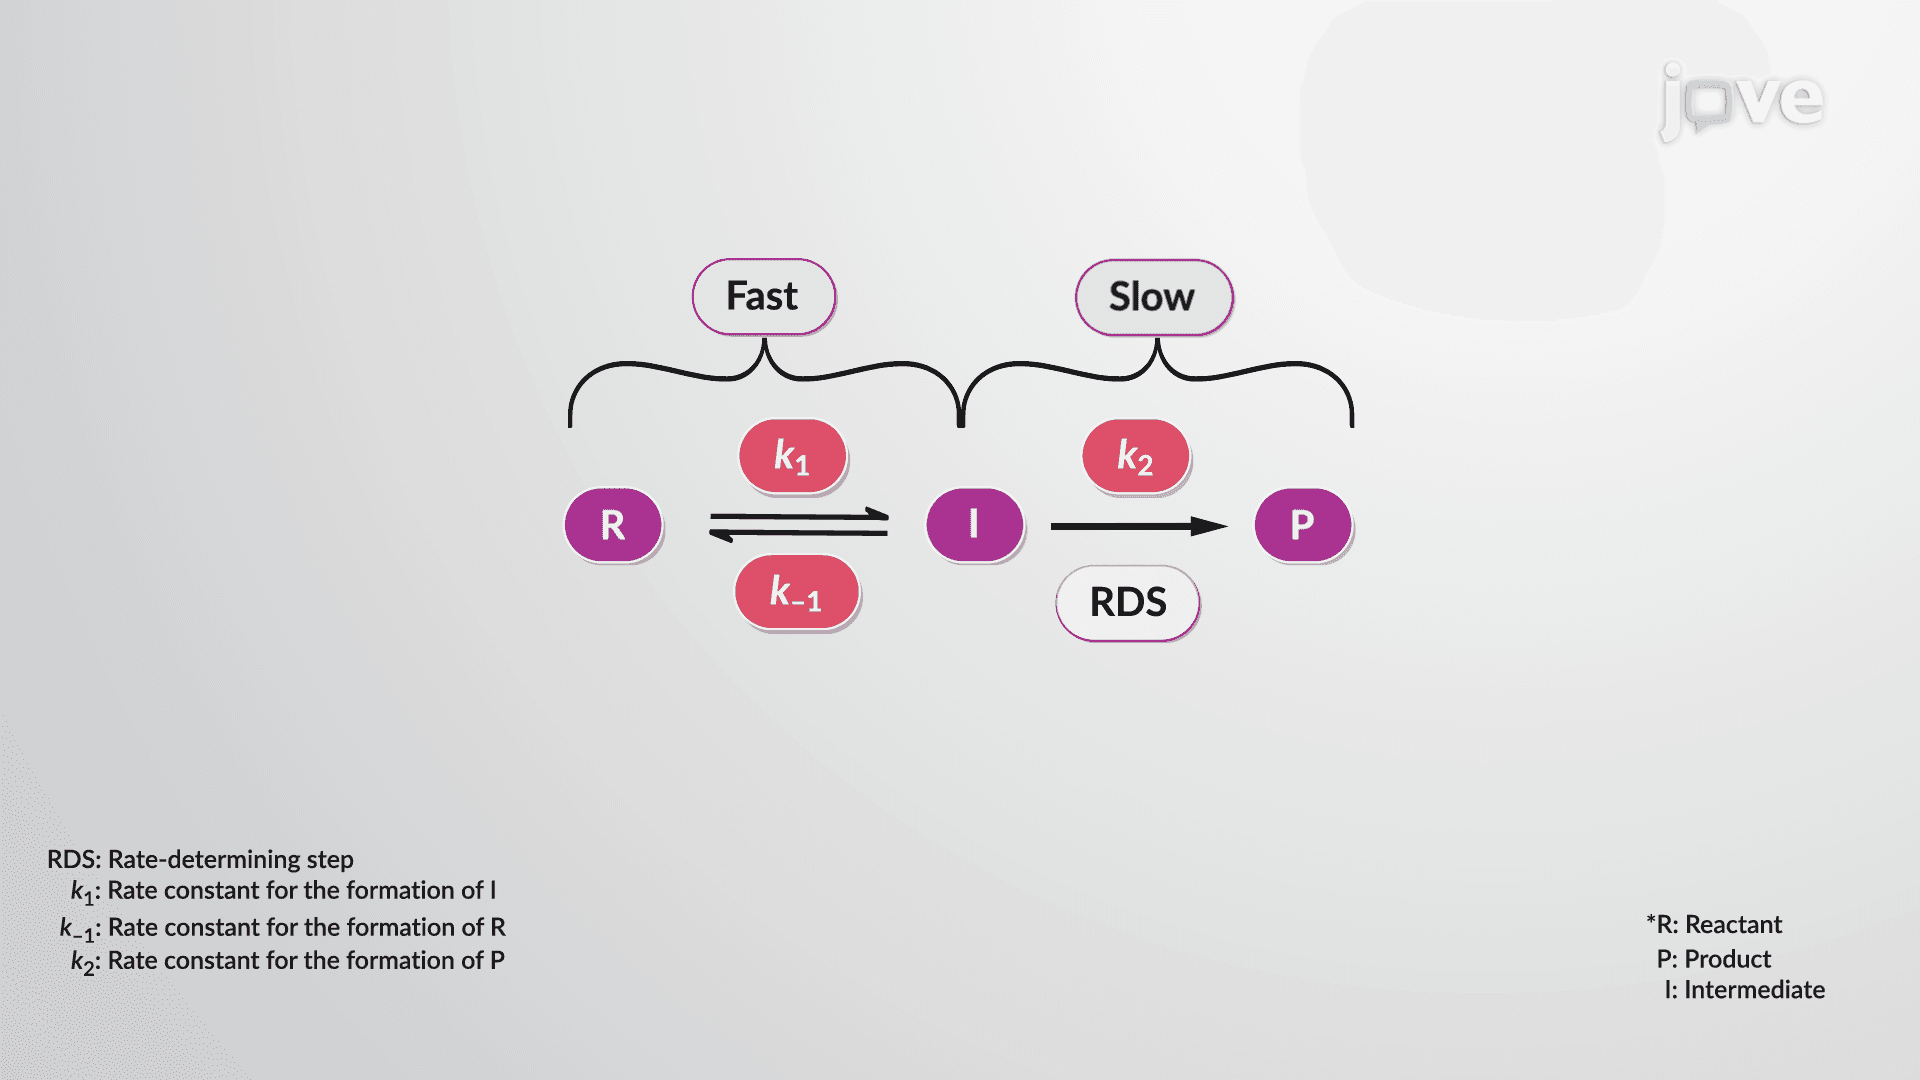
Reaction Mechanisms: The Steady-State Approximation
 01:09
Theories of Dissolution: The Danckwerts' Model and Interfacial Barrier Model
 01:27
Two Components: Liquid–Liquid Systems
 02:28
Intermolecular Forces in Solutions
When the strengths of the intermolecular forces of attraction between solute and solvent species in a solution are no different than those present in the separated components, the solution is formed with no accompanying energy change. Such a solution is called an ideal solution. A mixture of ideal gases (or gases such as helium and argon,...
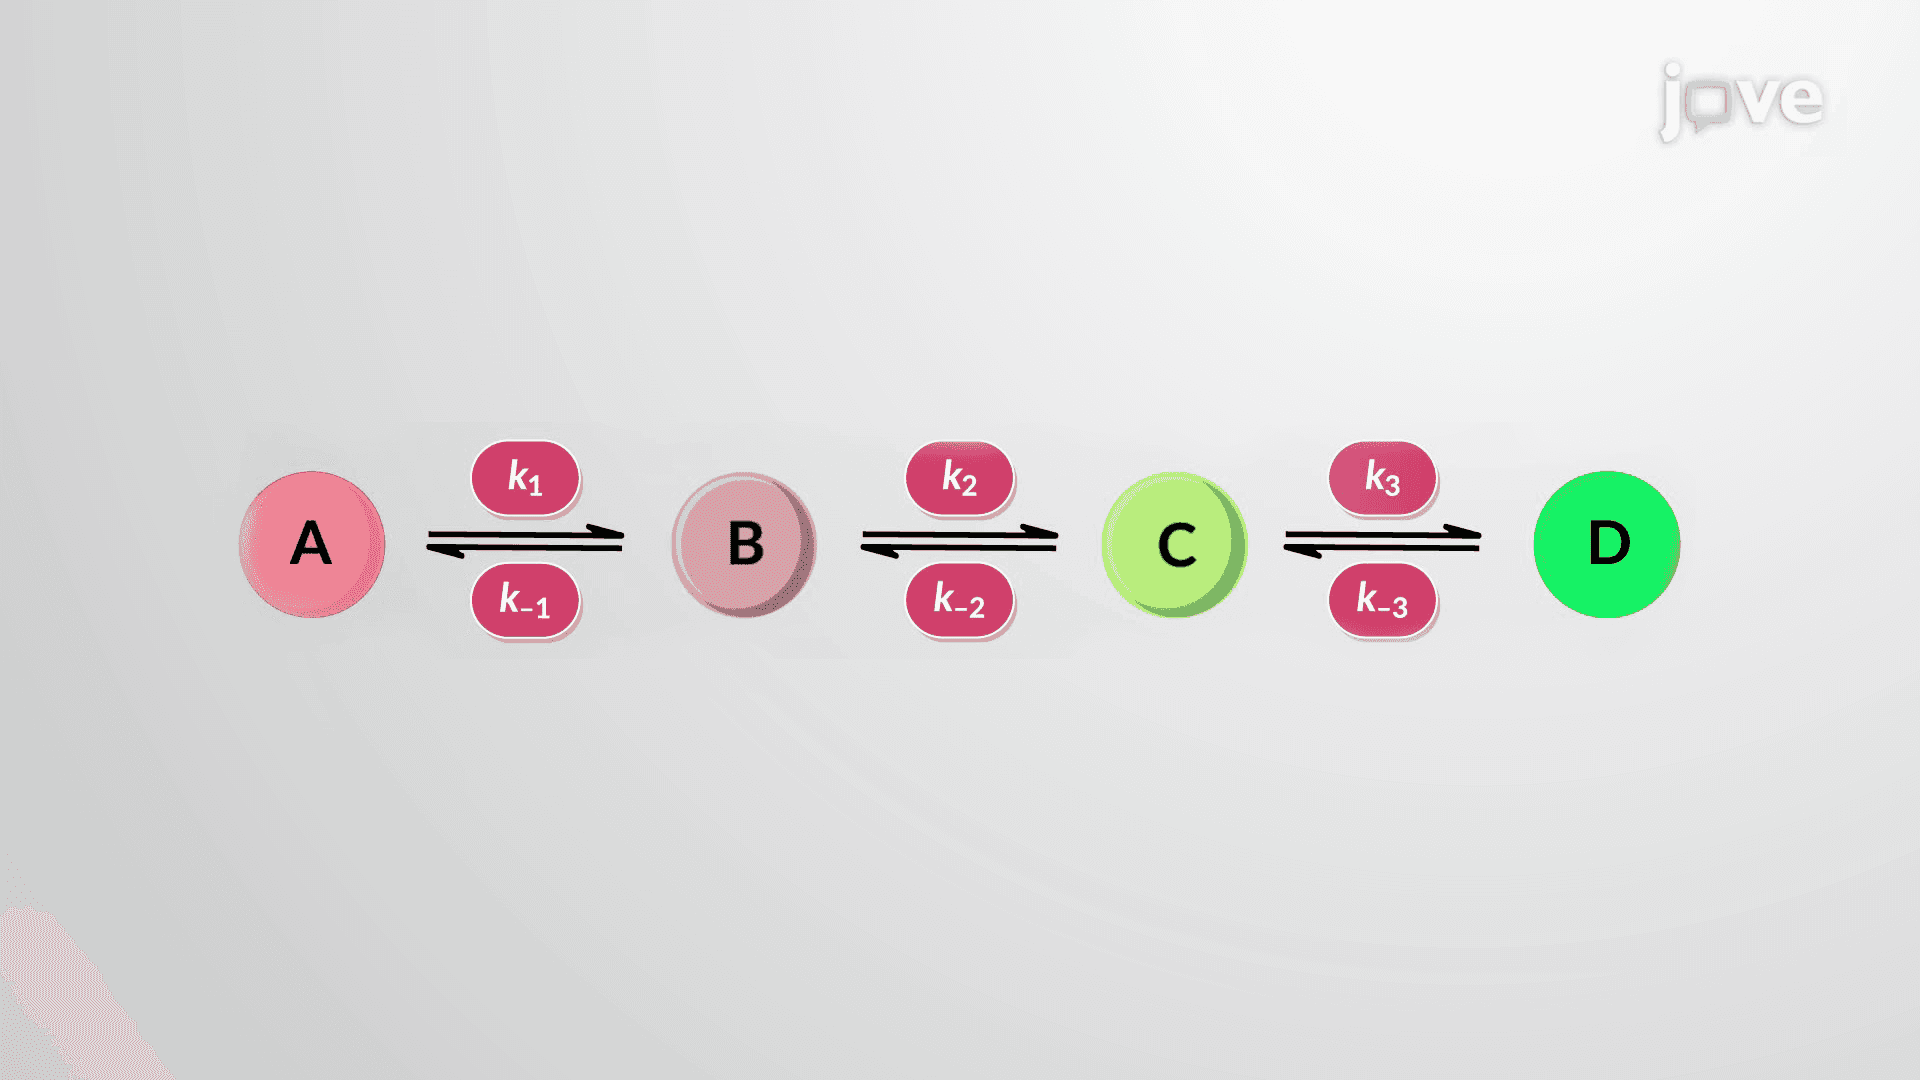 01:29
Reaction Mechanisms: Rate-limiting Step Approximation