Related Experiment Video
Updated: Oct 22, 2026
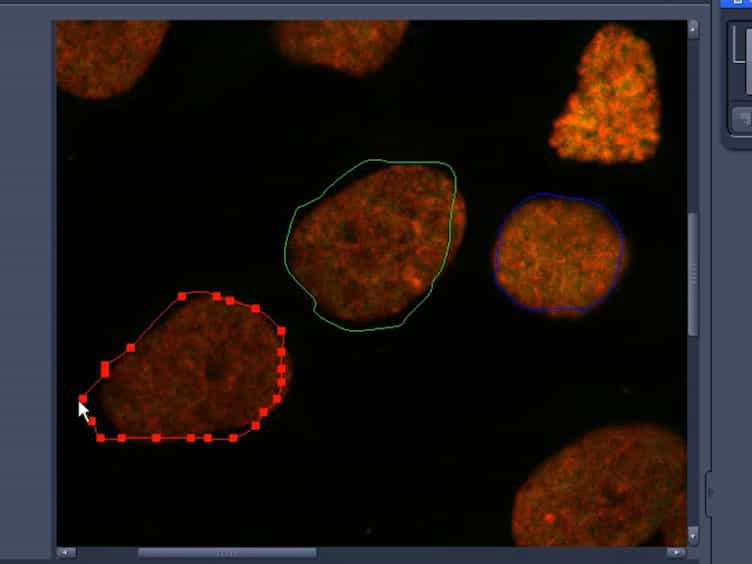
Immunostaining for DNA Modifications: Computational Analysis of Confocal Images
Published on: September 7, 2017
DNA methylation defines lineage identity yet converges among functionally related cells and is cell‑specifically
Lanlan Sun1, Dongmei Chen2, Xin Zhang3
1Key Laboratory of Biomechanics and Mechanobiology, Key Laboratory of Innovation and Transformation of Advanced Medical Devices, Ministry of Industry and Information Technology, National Medical Innovation Platform for Industry-Education Integration in Advanced Medical Devices (Interdiscipline of Medicine and Engineering), School of Engineering Medicine, Ministry of Education, Beihang University, Beijing, 100191, People's Republic of China.

Background:
DNA methylation (DNAme) is used to define cellular identity, but its ability to distinguish closely related subpopulations and its stability during in vitro culture remain to be systematically evaluated.
Results:
Lineage-scale whole-genome bisulfite sequencing (WGBS) analysis of 40 cell types across 13 organs showed that DNAme separates major lineages but converges among functionally related populations. Endothelial cells, fibroblasts, and smooth muscle cells from different organs clustered primarily by lineage rather than tissue of origin, whereas epithelial cells retained stronger organ-associated methylation signatures. Lineage-restricted analysis of endothelial cells identified 700 tissue-specific differentially methylated regions (DMRs) across seven tissues that were masked in global comparisons, indicating residual tissue-of-origin information persists within highly similar lineages. In kidney cells, in vitro passaging induced systematic methylation remodeling without erasing lineage identity. Primary and subcultured glomerular endothelial and tubular epithelial cells formed separate subclusters within their lineage groups. Genome-wide analysis identified extensive differential methylation (~ 1.88 M differentially methylated CpG sites [DMCs] in glomerular endothelial; ~2.64 M in tubular epithelial), with promoters overrepresented among hypermethylated features. Shared hypermethylated promoters were enriched for core homeostatic functions; hypomethylated promoters were enriched for differentiation and tissue-specific pathways. Motif analysis showed enrichment of bZIP and GATA factor motifs in endothelial hypomethylated regions and ETS-family motifs in epithelial cells. Cross-cell-type array analysis confirmed reproducible methylation drift across fetal skin fibroblasts, adult vascular endothelial cells, and neonatal foreskin fibroblasts, with 7,434 shared hypomethylated DMCs and 1,347 shared hypermethylated DMCs. Drift onset was cell type-dependent: adult skin fibroblasts and vascular smooth muscle cells diverged by passage 3, whereas neonatal and fetal cells remained stable through passage 10. Shared hypomethylated DMCs were enriched in partially methylated domains (PMDs) and late-replicating domains (LRDs). Cross-platform comparison detected only 44 common hypomethylated DMCs and 24 common hypermethylated DMCs between array and WGBS data, with negligible regional overlap.
Conclusions:
DNAme robustly distinguishes major lineages but converges among closely related cell types, limiting fine subpopulation resolution. Lineage-restricted analysis recovers tissue-of-origin signals otherwise obscured, whereas in vitro passaging induces cell type-dependent remodeling and reproducible methylation drift that preserves lineage identity while attenuating tissue-specific programs. No universal passage threshold ensures phenotypic stability.
Related Concept Videos
 01:21
Lineage Commitment
 03:17
Inheritance of Chromatin Structures
 00:57
Cellular Differentiation
A zygote is a...
 01:37
Epigenetic Regulation
X-chromosome...
 01:46
Epigenetic Regulation
 02:30
Genomic Imprinting and Inheritance
The expression of some genes depends on which parent passed the gene to the offspring, through a phenomenon known as...