Related Experiment Video
Updated: Oct 22, 2026
Picometer-Precision Atomic Position Tracking through Electron Microscopy
Published on: July 3, 2021
Decoding Perovskite Interfaces: Machine Learning Uncovers Geometry-Matched Dual-Site Hydrogen Bonding
Can Cheng1,2, Xinyu Gu3, Kai Sun4
1State Key Laboratory of Chemistry for NBC Hazards Protection, College of Chemistry, Fuzhou University, Fuzhou, P. R. China.

Abstract:
Molecular passivation is central to efficient and stable perovskite photovoltaics, yet interfacial molecules are still largely selected through chemical intuition and iterative experimentation. Here, an interpretable machine-learning framework is developed that extracts chemically meaningful structure-performance relationships from literature data and uses them to guide molecular screening. Using approximately 100 data points, the model quantitatively relates six molecular and material features to photovoltaic performance, while SHapley Additive exPlanations (SHAP) analysis ranks their relative contributions and identifies hydrogen-bonding capacity and adsorption strength as the two most influential factors. Guided by these data-derived relationships, nine previously unexplored molecules were screened, with 2-amino-thiazole-4-carboxamide (ATC) and 1-(2-aminoethyl)-1H-pyrrole-2,5-dione (MPD) emerging as leading candidates. Theoretical and experimental investigations show that MPD exhibits stronger Pb2 + coordination, whereas ATC forms stronger geometry-matched multisite N─H···I interactions, increasing the calculated iodide-migration barrier from 0.39 to 0.67 eV and suppressing interfacial iodide redistribution. ATC-modified cells achieve a PCE of 27.13% and a certified stabilized efficiency of 26.59%, while retaining 90% of their initial efficiency after 1460 h of unencapsulated operation. The strategy further delivers efficiencies of 24.01% and 21.14% in 20.98 cm2 rigid and 21.0 cm2 flexible modules, respectively, providing a data-to-mechanism route for rational and scalable perovskite interface engineering.
More Related Videos
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
 08:49
Multimodal Nonlinear Hyperspectral Chemical Imaging Using Line-Scanning Vibrational Sum-Frequency Generation Microscopy
Published on: December 1, 2023
Related Concept Videos
 01:04
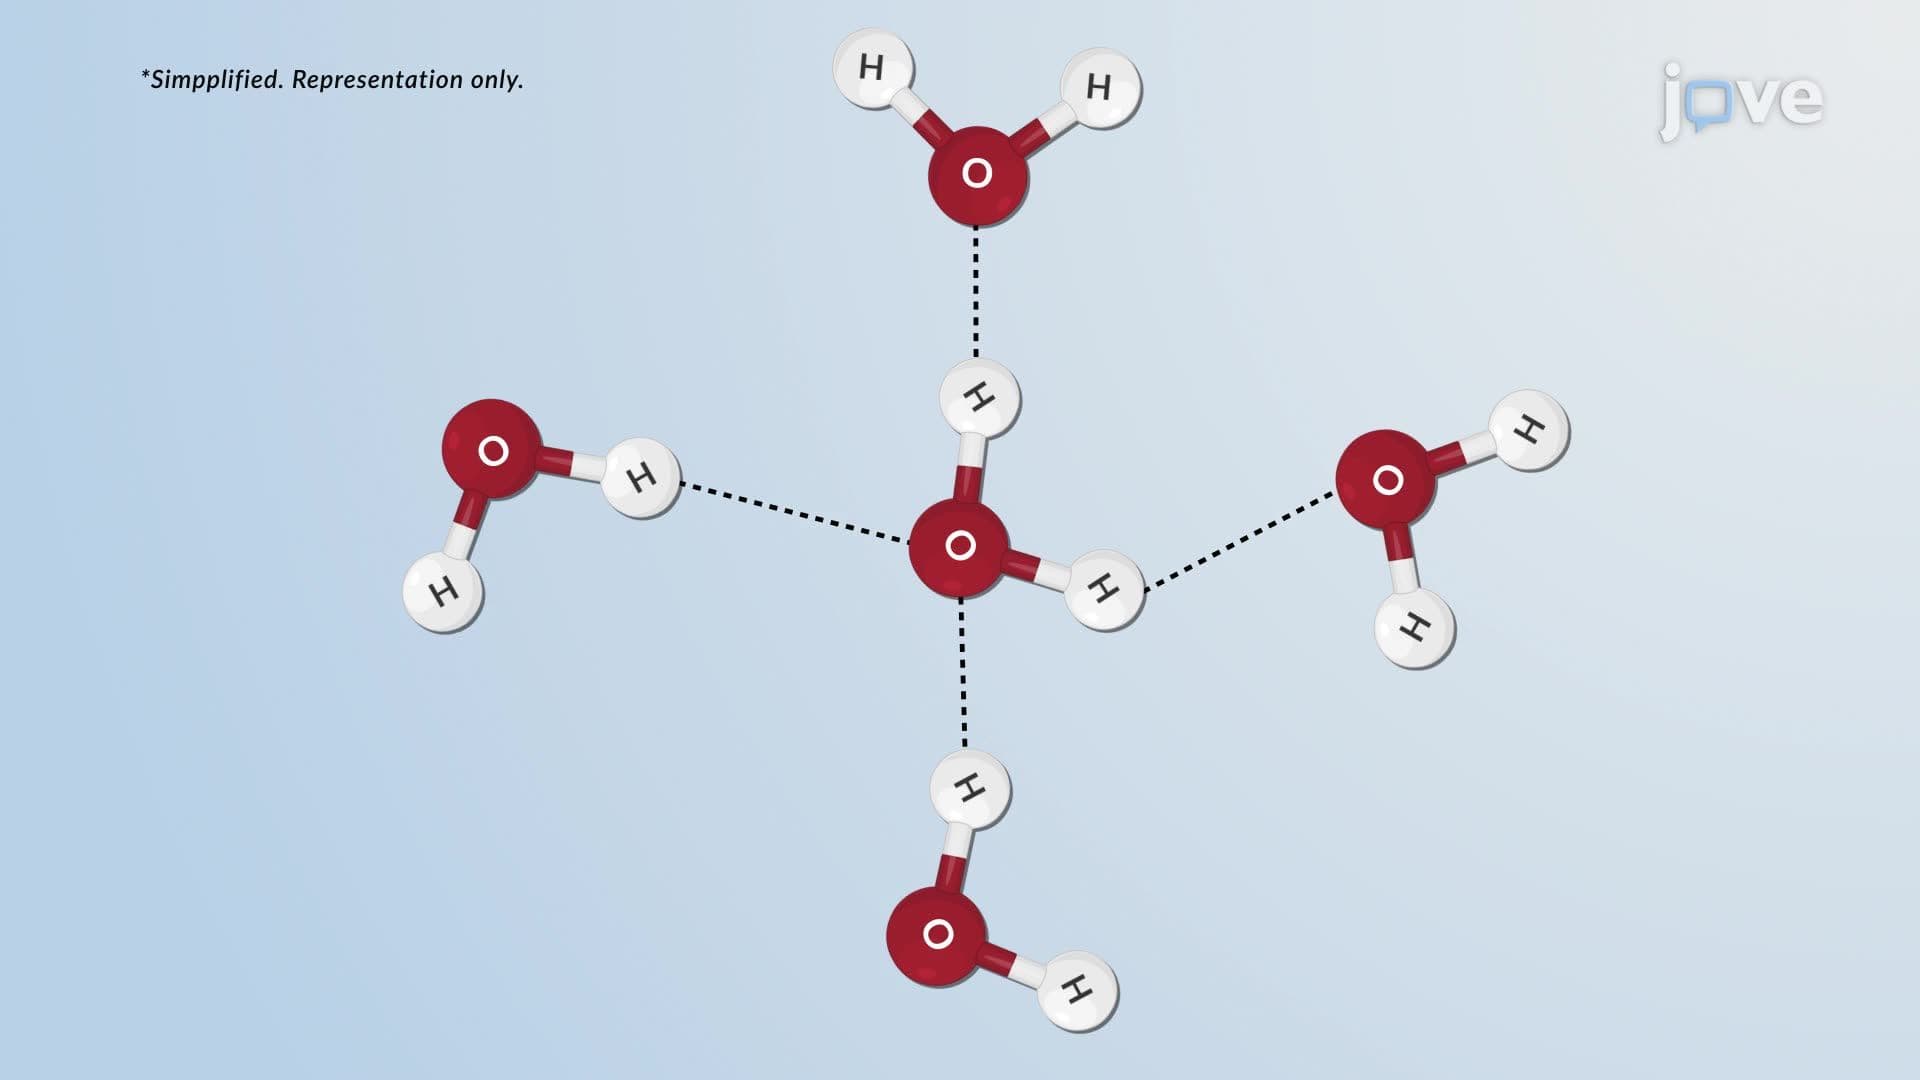
Hydrogen Bonds
 00:26
Hydrogen Bonds
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:27
Predicting Molecular Geometry
 01:29
Determination of Crystal Structures
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...