Related Experiment Video
Updated: Oct 24, 2026
High Resolution Physical Characterization of Single Metallic Nanoparticles
Published on: June 28, 2019
POMs-GibbsNet: A Deep Neural Network for the Prediction of Thermodynamic Parameters of Polyoxometalates
Jordi Buils1,2, Carles Bo1,2, David Balcells3
1Institute of Chemical Research of Catalonia (ICIQ), Tarragona, Spain.

Abstract:
Polyoxometalates (POMs) are versatile metal-oxo clusters with remarkable structural diversity and applications spanning catalysis, energy storage, and materials science. However, accurately predicting their thermodynamic properties remains computationally demanding due to the large system sizes and the cost of density functional theory (DFT) frequency calculations required to obtain Gibbs free energies. In this work, we present POMs-GibbsNet, a deep neural network designed to predict the thermal free energy correction ( ) of polyoxometalates by combining graph-based molecular representations with DFT-derived descriptors. Molecular structures were encoded using variants of the AABBA autocorrelation graph kernel, which captures atom-atom, bond-bond, and atom-bond correlations together with quantum-derived features including QTAIM descriptors. POMs-GibbsNet was trained on a dataset of 1419 POM structures spanning 19 chemical systems to obtain good results ( and RMSE = 1.9 kcal mol-1. To evaluate the performance, a benchmark against 10 classical regressors was performed. Additionally, an uncertainty study was added to the methodology. External validation using POMSimulator reaction networks comprising over 10,819 reactions shows excellent agreement between predicted and DFT reaction energies ( ), with errors typically within a few kcal mol-1. Error cancellation was also studied during this step. Notably, simplified atom-atom autocorrelation descriptors already provide robust performance while reducing model complexity. These results demonstrate that POMs-GibbsNet enables efficient estimation of thermodynamic corrections in POM systems, providing a powerful framework for accelerating the computational exploration of polyoxometalate speciation and self-assembly across large chemical spaces.
Related Concept Videos
 02:27
Predicting Molecular Geometry
 01:26
Thermodynamic Potentials
 02:24
Predicting Reaction Outcomes
 01:10
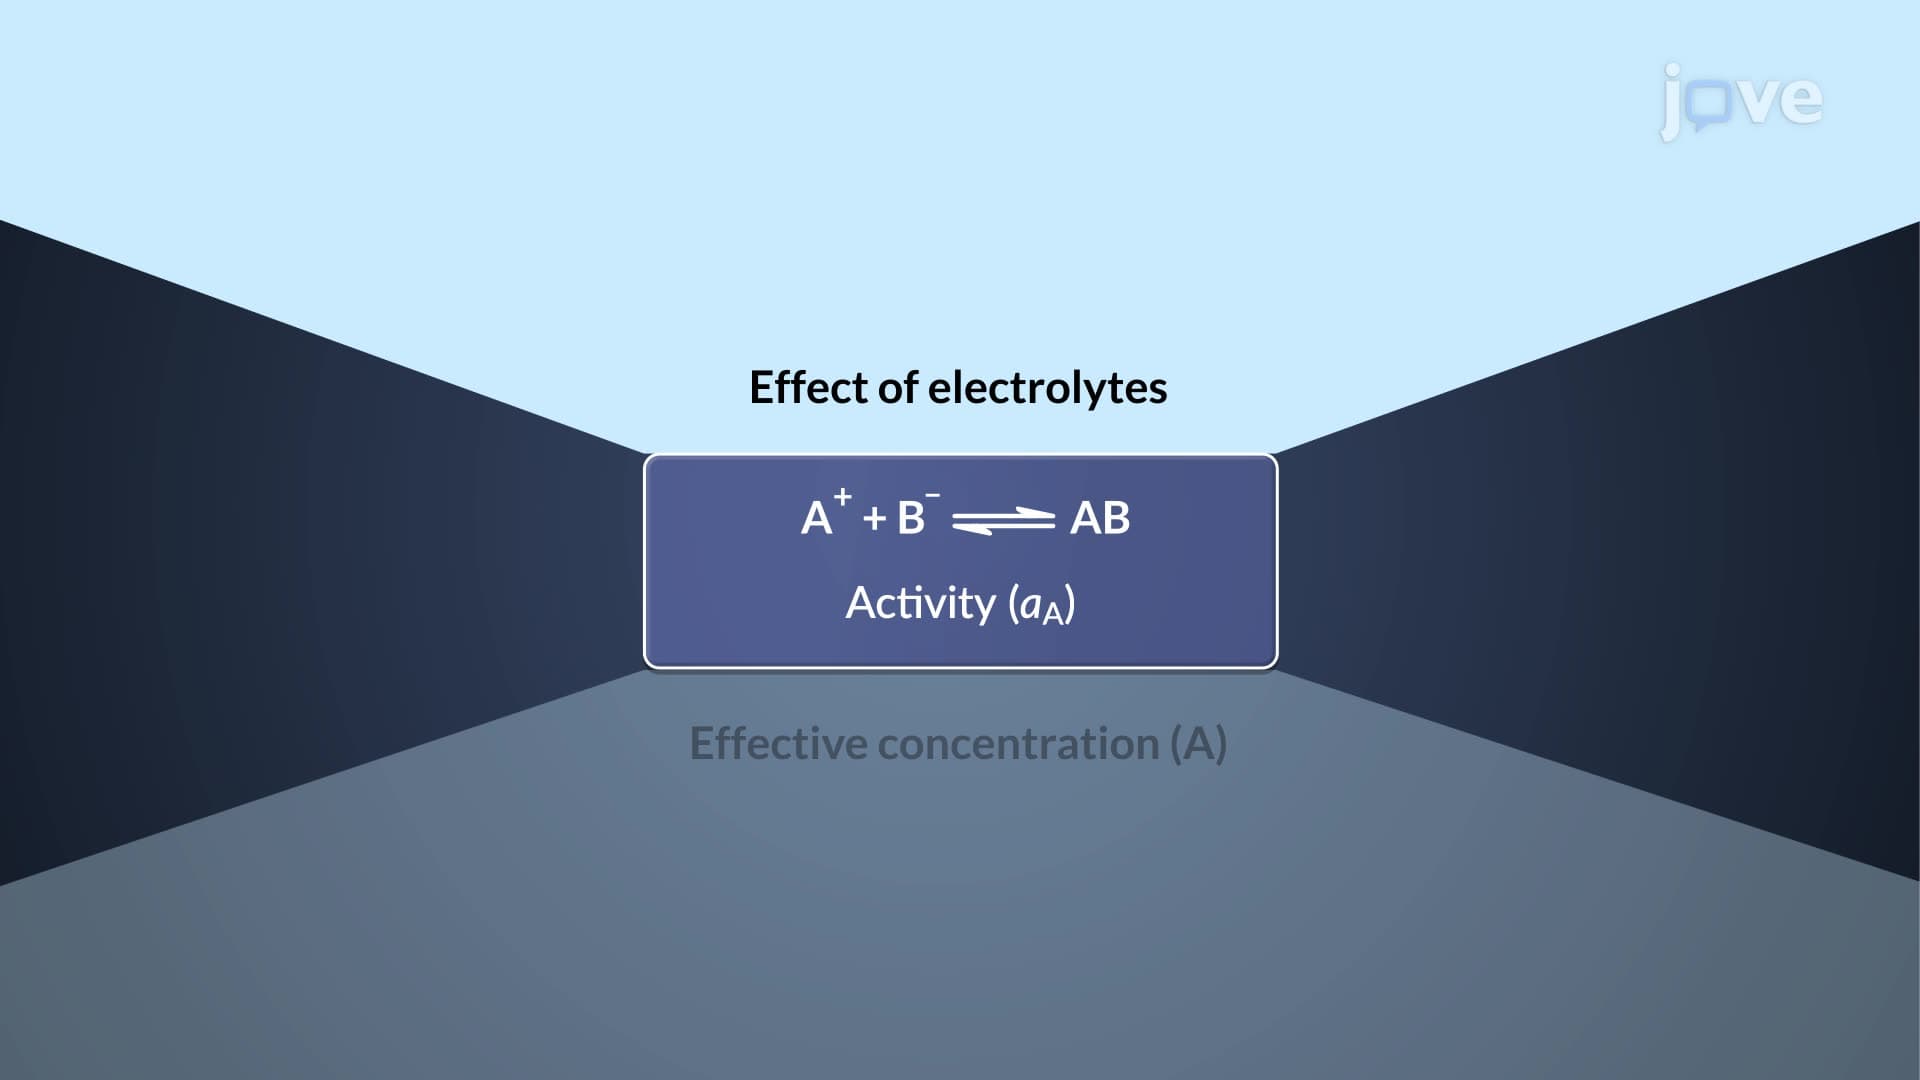
Thermodynamics: Chemical Potential and Activity
The thermodynamic equilibrium constant is more accurately defined in terms of activity rather than concentration.
 01:19
Thermodynamic Properties of Ideal Solutions