Related Experiment Video
Updated: Oct 25, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Q-Chem/CP2K software interface for projection-based, pseudoperiodic wavefunction-in-DFT embedding
Ar M Fonlon1, Elena Kolodzeiski2,3, Kay Carter-Fenk1,4,5
1Department of Chemistry and Biochemistry, The Ohio State University, Columbus, Ohio 43210, USA.

Abstract:
We report an implementation of projection-based quantum embedding that combines periodic density functional theory in the CP2K code with correlated wavefunction calculations in the Q-Chem program. Using this interface, correlated wavefunction methods can be applied to a subset of molecular orbitals, selected in an automated manner based on a user-defined list of nuclei, with an embedding potential that provides electronic coupling to the remaining orbitals that comprise the environment. Our implementation uses exact projection, without the need for any level-shift operator, and is "pseudoperiodic" in the sense that periodic boundary effects are implicit in the orbitals used in the (non-periodic) wavefunction calculation. Convergence tests demonstrate that computed properties are faithful to the corresponding periodic quantities, provided that the high-level subsystem is spatially localized and small, relative to the periodic simulation cell. Spectroscopic examples for aqueous chromophores demonstrate converged results using a very limited subset of water molecules in the simulation cell, decoupling the choice of excited-state method and basis set from the functional that is used to propagate ground-state ab initio molecular dynamics. Solvatochromic shifts for aqueous uracil are converged using equation-of-motion coupled-cluster theory, without the need for solvent molecules in the wavefunction calculation, and the embedded calculation captures solvent effects that are absent in gas-phase "microhydration" studies. Additional applications include adsorption of CO on a slab model of Si(100) and bond dissociation in a one-dimensional periodic polymer. In both of the latter cases, the system/environment partition crosses a covalent bond; these applications demonstrate that pseudoperiodic embedding can be applied to realistic systems in materials science.
More Related Videos
Related Concept Videos
 03:35
Hybridization of Atomic Orbitals II
 03:24
Hybridization of Atomic Orbitals I
 01:20
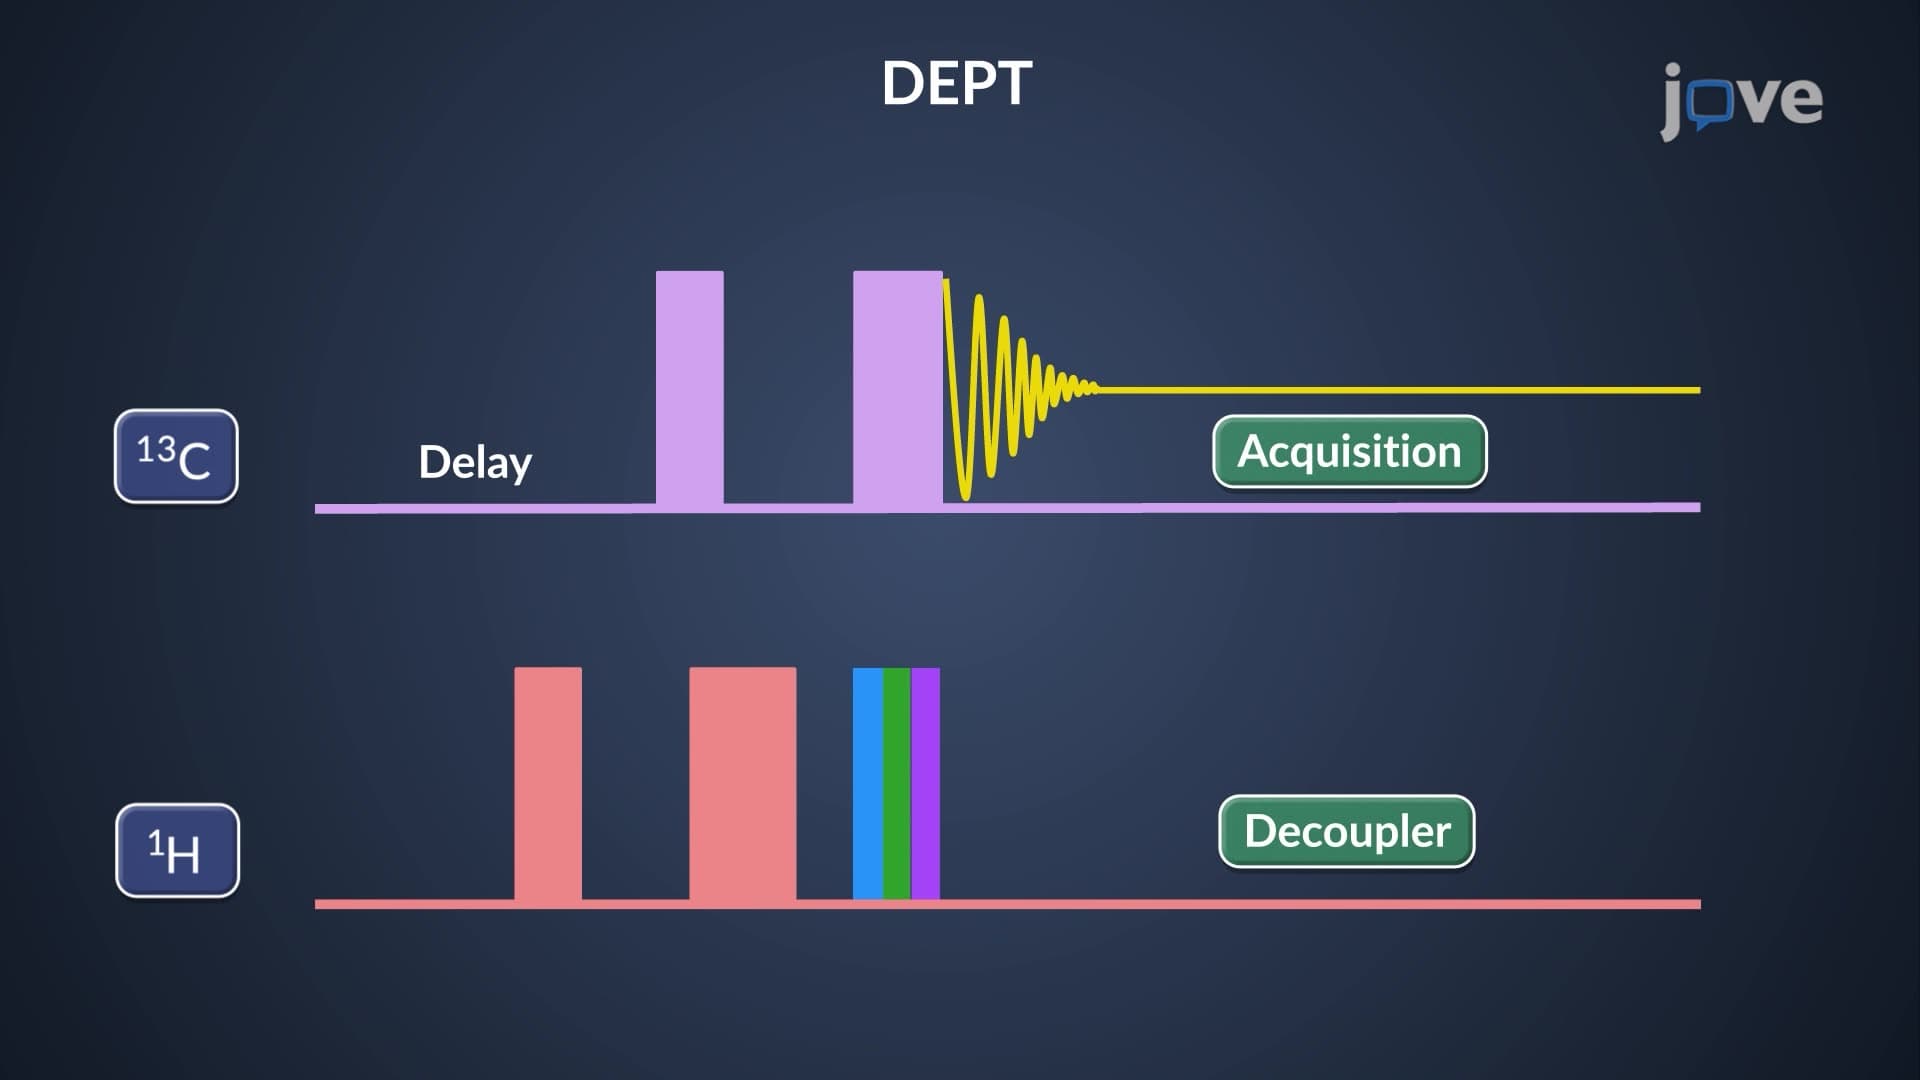
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 01:13
Graphing the Wave Function
 02:06
Newman Projections
The organic molecules rotate across the single bonds leading to numerous temporary three-dimensional structures of varying energy known as conformers.
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...