Related Experiment Video
Updated: Oct 25, 2026
Thermochemical Studies of Ni(II) and Zn(II) Ternary Complexes Using Ion Mobility-Mass Spectrometry
Published on: June 8, 2022
Zn2+ Coordination and Ligand-Exchange Dynamics in a ZnCl2-Based Deep Eutectic Electrolyte from Machine-Learning
Eudes Fileti1, Davi Fileti2, André P L de Carvalho2
1Instituto de Ciência e Tecnologia, Universidade Federal de São Paulo, São José dos Campos, São Paulo12247-014, Brazil.

Abstract:
The Zn2+ coordination shell in ZnCl2-based battery electrolytes is far more dynamic than its time-averaged structure suggests. Using a machine-learning force field (MACE) fine-tuned with density functional theory data, we simulate nanosecond-scale molecular dynamics of a ZnCl2:urea:H2O deep eutectic electrolyte across 300-400 K, a time regime inaccessible to ab initio methods and poorly captured by classical force fields. The Zn2+ first coordination shell is tetrahedral and compositionally mixed with chloride ions, urea, and water contributing comparable shares. This overall occupancy remains essentially constant with temperature, masking markedly asymmetric underlying kinetics. Residence-time analysis reveals that chloride is essentially nonexchanging on the nanosecond time scale, while water and urea exchange continuously, with residence times an order of magnitude shorter and activation energies of 12.5 and 16.2 kJ/mol, respectively. Potential of mean force and speciation analyses identify ZnCl2 as the dominant solute species and show that ligand substitution proceeds predominantly through an associative pathway. Self-diffusion coefficients and activation energies for all four constituent species further establish a mobility hierarchy consistent with this coordination picture. Together, these results provide a molecular-level rationale for the transport behavior of ZnCl2-based electrolytes and a general framework for understanding ion speciation and lability in coordinating eutectic battery electrolytes.
Related Concept Videos
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:42
Valence Bond Theory
 01:07
Ladder Diagrams: Complexation Equilibria
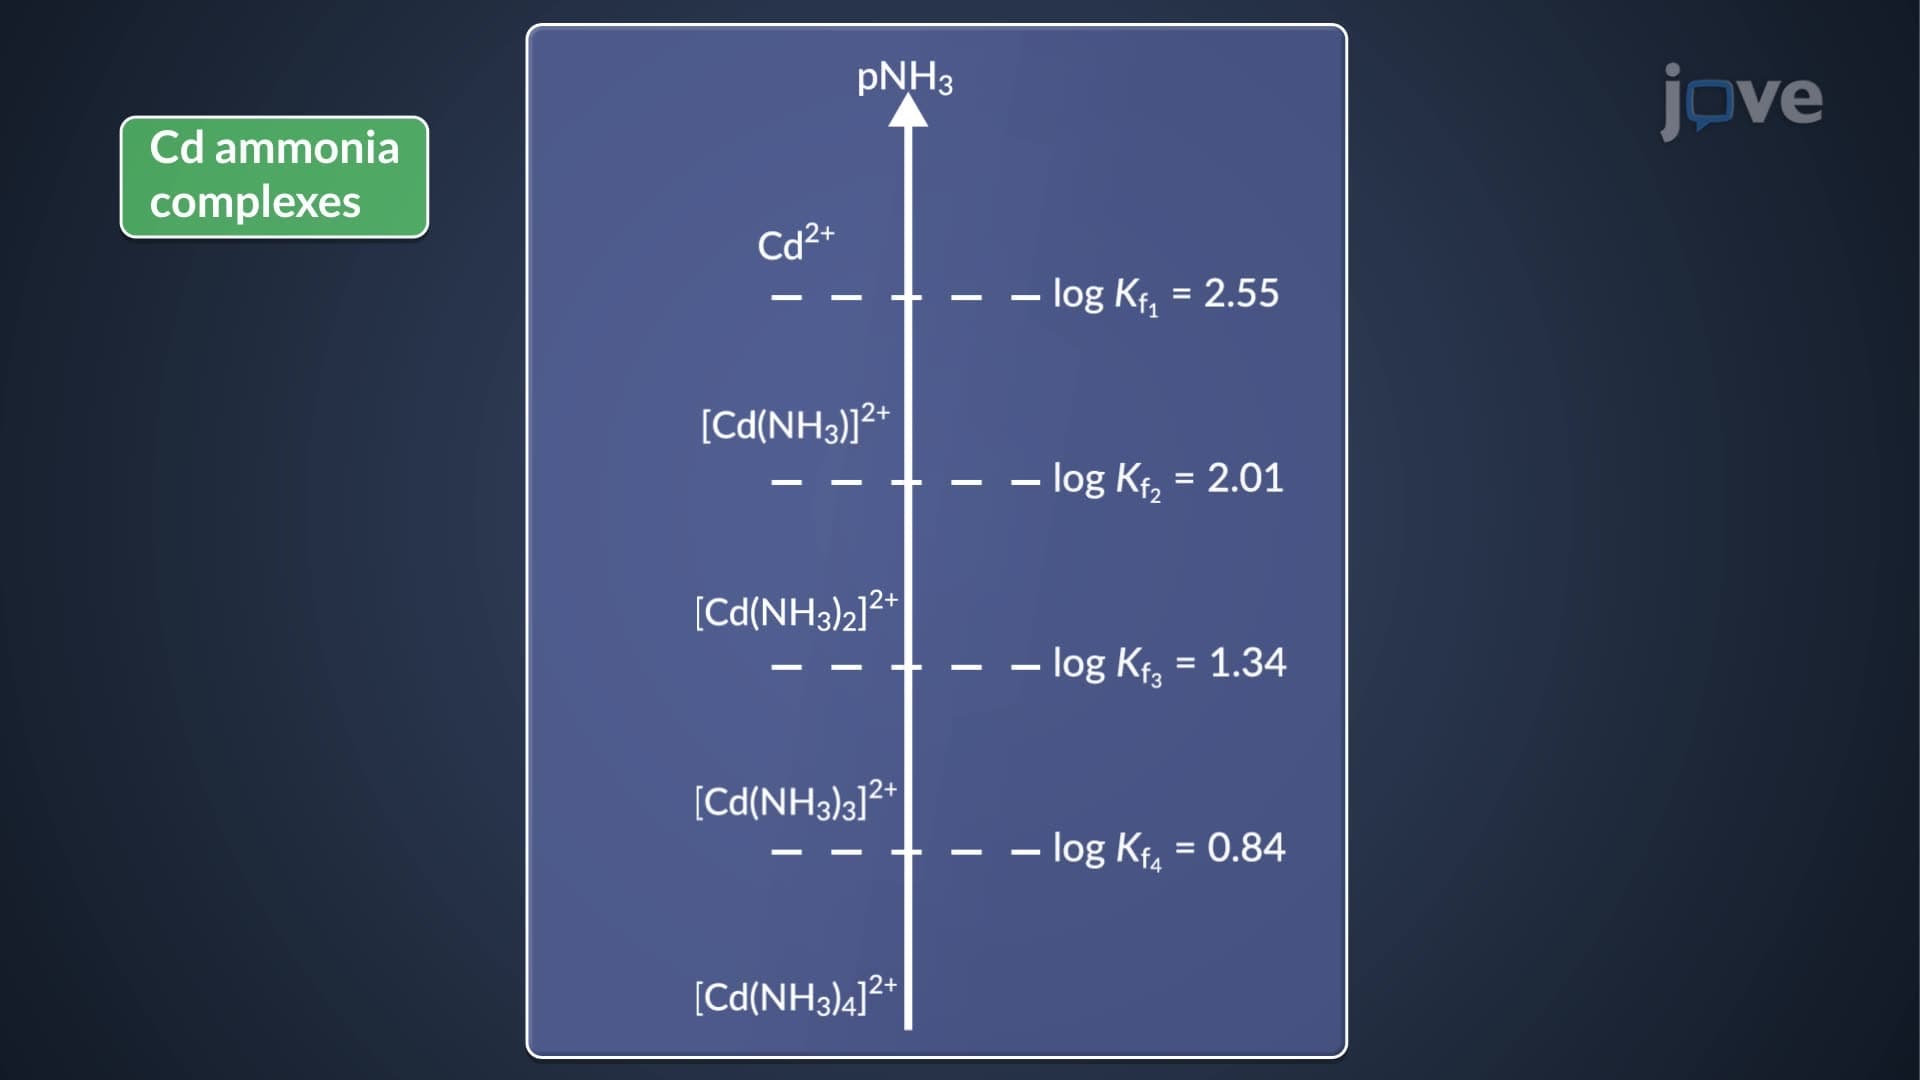
The formation constant, K1, for the formation of Cd(NH3)2+ complex from cadmium and ammonia is 3.55 × 10². Log K1 (i.e. pNH3) is 2.55, and...
 02:57
Coordination Number and Geometry
 02:51
Metal-Ligand Bonds
In these complexes, transition metals form coordinate covalent bonds, a kind of Lewis acid-base interaction in which both of the electrons in the bond are contributed by a donor (Lewis base) to an electron acceptor (Lewis acid). The Lewis acid in...