Related Experiment Video
Updated: Oct 26, 2026
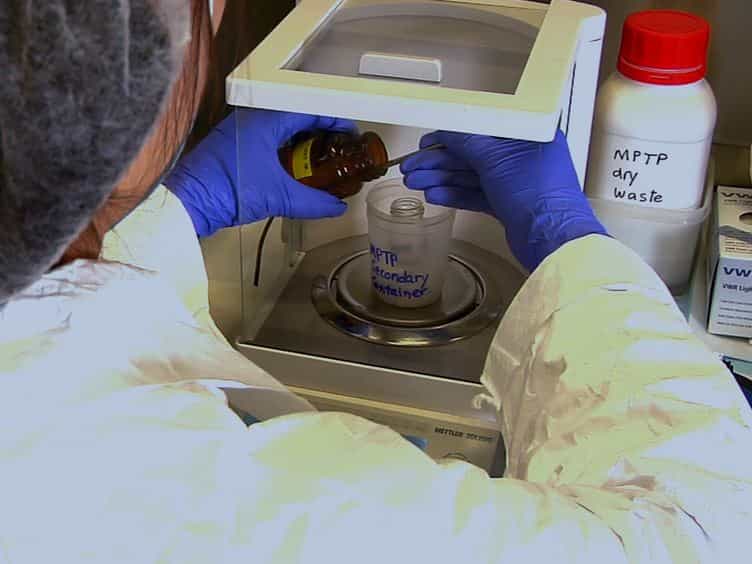
Gene-environment Interaction Models to Unmask Susceptibility Mechanisms in Parkinson's Disease
Published on: January 7, 2014
Machine Learning Approach Investigating Differentially Expressed Genes Associated With Parkinson's Disease on
Navya Ramachandran1, Bianca Maria Lewis1, Apoorva Prashanth1
1Department of Biotechnology, PES University, 100 Feet Ring Road, Banashankari 3rd Stage, Bangalore, 560085, India.

Abstract:
Parkinson's Disease (PD) is a multifactorial neurodegenerative disease, with environmental exposures such as pesticides implicated as contributing risk factors. While ensemble machine learning (ML) models have shown promise in identifying discriminating genes from transcriptomic data, their generalizability across heterogeneous toxicogenomic datasets remains poorly characterized. Here, we present a robust statistical pipeline as the primary framework for identifying PD-associated, pesticide-responsive differentially expressed genes (DEGs), developed in direct response to the limitations observed in an ensemble ML approach. The ensemble model, combining individual ML models, Random Forest, Logistic Regression and Gradient Boosting was first trained on merged GEO transcriptomic datasets following normalization and feature selection (SelectKBest, ANOVA F-test, StandardScaler). Initially, an ROC AUC (Receiver Operating Characteristic Area Under the Curve) of 0.734 with a cross-validation (CV) AUC of 0.632 was obtained with an average AUC of 0.806 +/- 0.106 across 10 different splits of train/test data points. The confusion matrix and PCA plots indicated the presence of strong batching effects driving these results. Batch correction improved CV AUC to 0.864 ± 0.142, yet cross-dataset validation collapsed to an ROC AUC of 0.523 ± 0.040, with the model consistently predicting a single class-indicating the ensemble had learned technical artefacts rather than generalizable biological signal. Given this failure of ML generalizability, an alternate, statistically grounded pipeline was developed that relied on domain-specific analysis of transcriptomics data. Differential expression was assessed using limma to determine the extent and significance of gene expression changes across pesticide-exposure datasets. Hub genes were identified through protein-protein interaction network analysis, and their associated pathways were enriched and validated against existing literature. The protein-protein interaction network was constructed and hub genes {Citation}with their respective pathways were identified. The fact that pesticides contribute to the progression of PD has been established by multiple studies. This study highlights the necessity of statistically rigorous, domain-informed approaches over black-box ML methods for identifying reliable, biologically meaningful DEGs-representing a novel methodological comparison not previously explored in pesticide-associated PD biomarker research.
Related Concept Videos
 01:24
Parkinson Disease l: Introduction
 01:24
Parkinson Disease ll: Pathophysiology
 01:47
Neural Regulation