Related Experiment Video
Updated: Oct 28, 2026
Genotyping Single Nucleotide Polymorphisms in the Mitochondrial Genome by Pyrosequencing
Published on: February 10, 2023
Structural and biochemical characterization of disease-associated mutations in the human mitochondrial processing
Danielle M Simons1, Sibei Qin2, Kim Munro3
1Department of Pharmacology & Therapeutics, McGill University, Montreal, QC, Canada; Centre de Recherche en Biologie Structurale, McGill University, Montreal, QC, Canada.

Abstract:
The mitochondrial processing peptidase (MPP) is an essential heterodimeric protease responsible for cleavage of N-terminal mitochondrial targeting sequences (N-MTSs) from most nuclear-encoded mitochondrial proteins following import into the mitochondrial matrix. Pathogenic variants in the α-subunit of MPP (PMPCA) cause a spectrum of mitochondrial disorders, including spinocerebellar ataxia type 2 (SCAR2), encephalopathy, and severe neurodevelopmental disease; however, the molecular mechanisms underlying these phenotypes remain poorly understood. Here, we combined biochemical and structural approaches to characterize five disease-associated PMPCA variants (S96L, R185W, R268W, A377T, and G515R). Enzymatic analyses revealed variant-specific defects in substrate processing, with most mutations severely impairing cleavage of the rapidly processed substrate malate dehydrogenase 2. In contrast, several variants exhibited only modest impairment in the processing of the slowly cleaved substrate PINK1. Mass photometry demonstrated reduced heterodimerization of all variants, whereas circular dichroism identified varying degrees of structural destabilization, particularly for S96L and G515R. To define the structural basis of these defects, hydrogen-deuterium exchange mass spectrometry was performed on wild type MPP and selected variants. Pathogenic mutations altered substrate-induced conformational changes within previously characterized substrate-recognition elements, including the glycine-rich loop. Furthermore, we identified a previously unrecognized region, termed the INS loop, that exhibited substrate-dependent protection uniquely in the wild type enzyme but was consistently disrupted across pathogenic variants. Together, these findings demonstrate that PMPCA mutations impair MPP function through diverse mechanisms involving protein stability, dimerization, substrate selectivity, and allosteric conformational dynamics. Our work provides a mechanistic framework for understanding genotype-phenotype relationships in PMPCA-associated disease.
Related Concept Videos
 01:48
ATP Synthase: Mechanism
 01:19
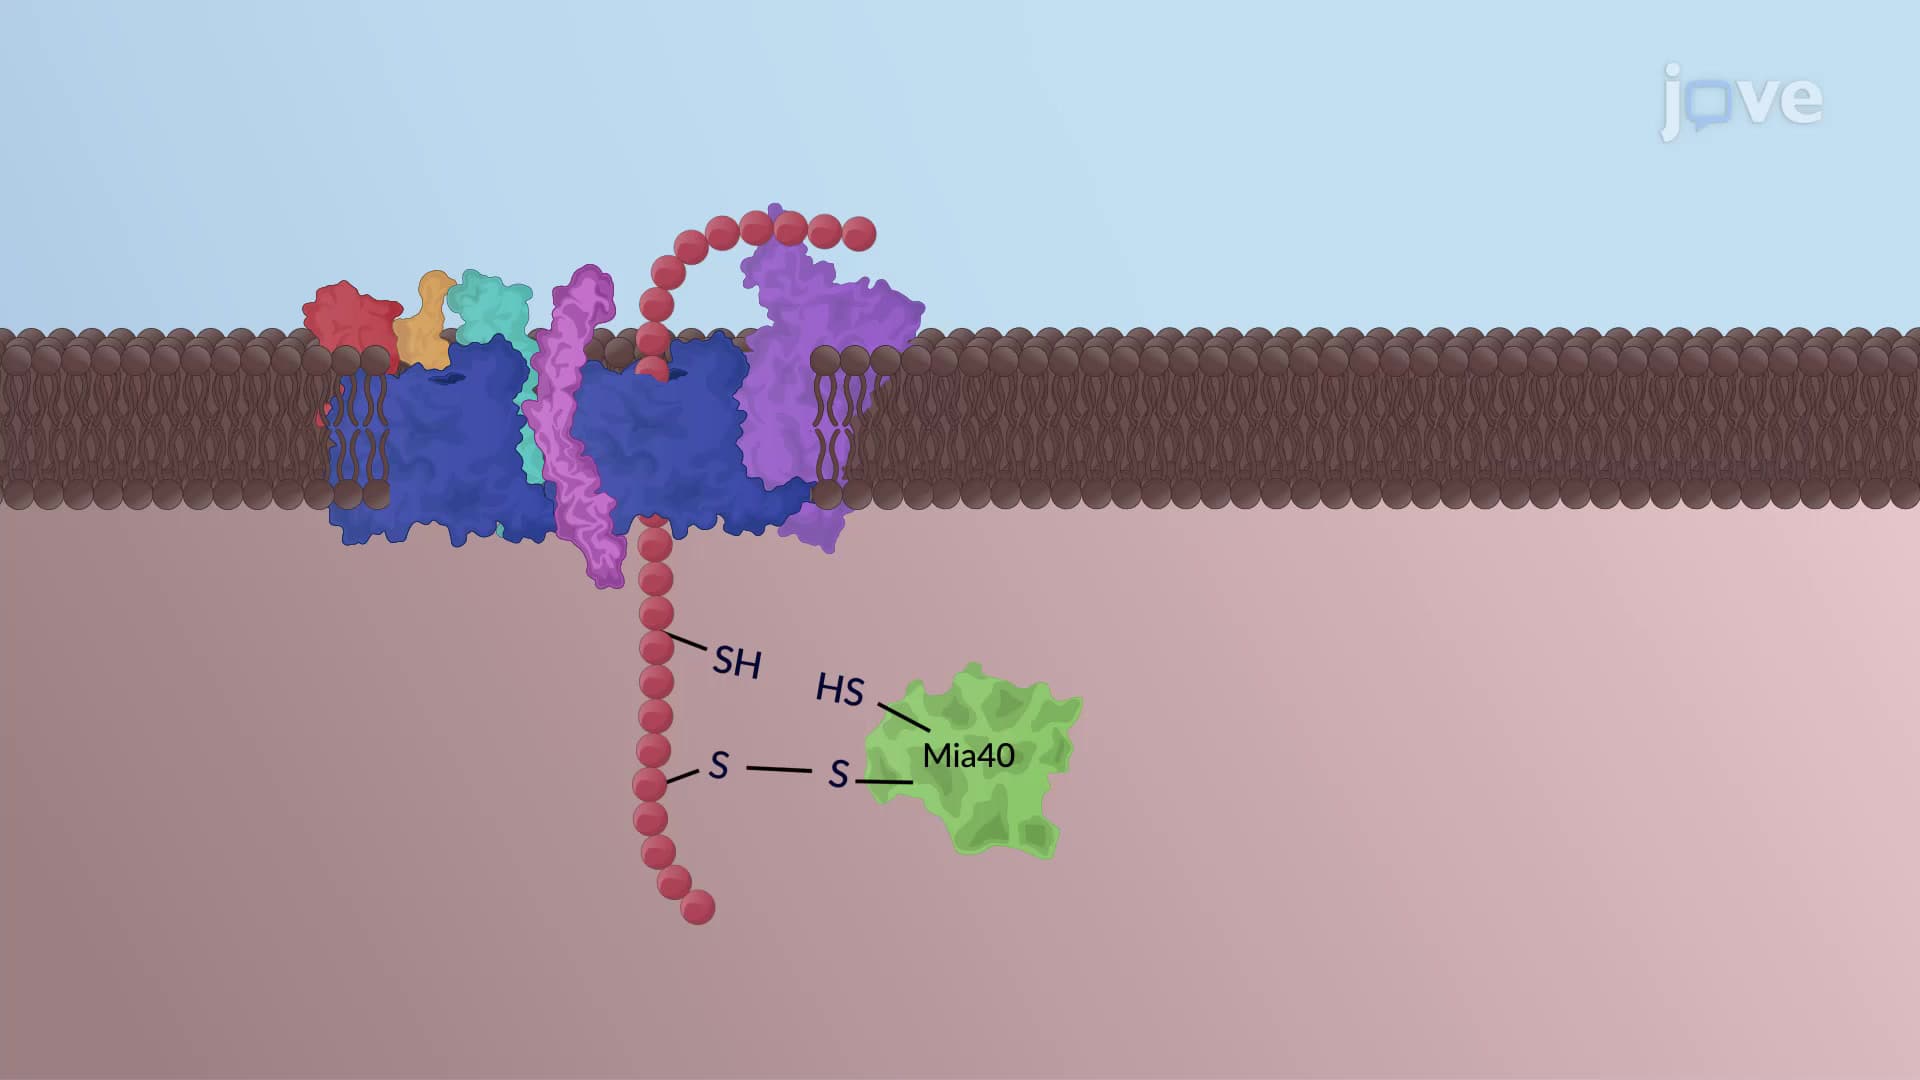
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...
 01:12
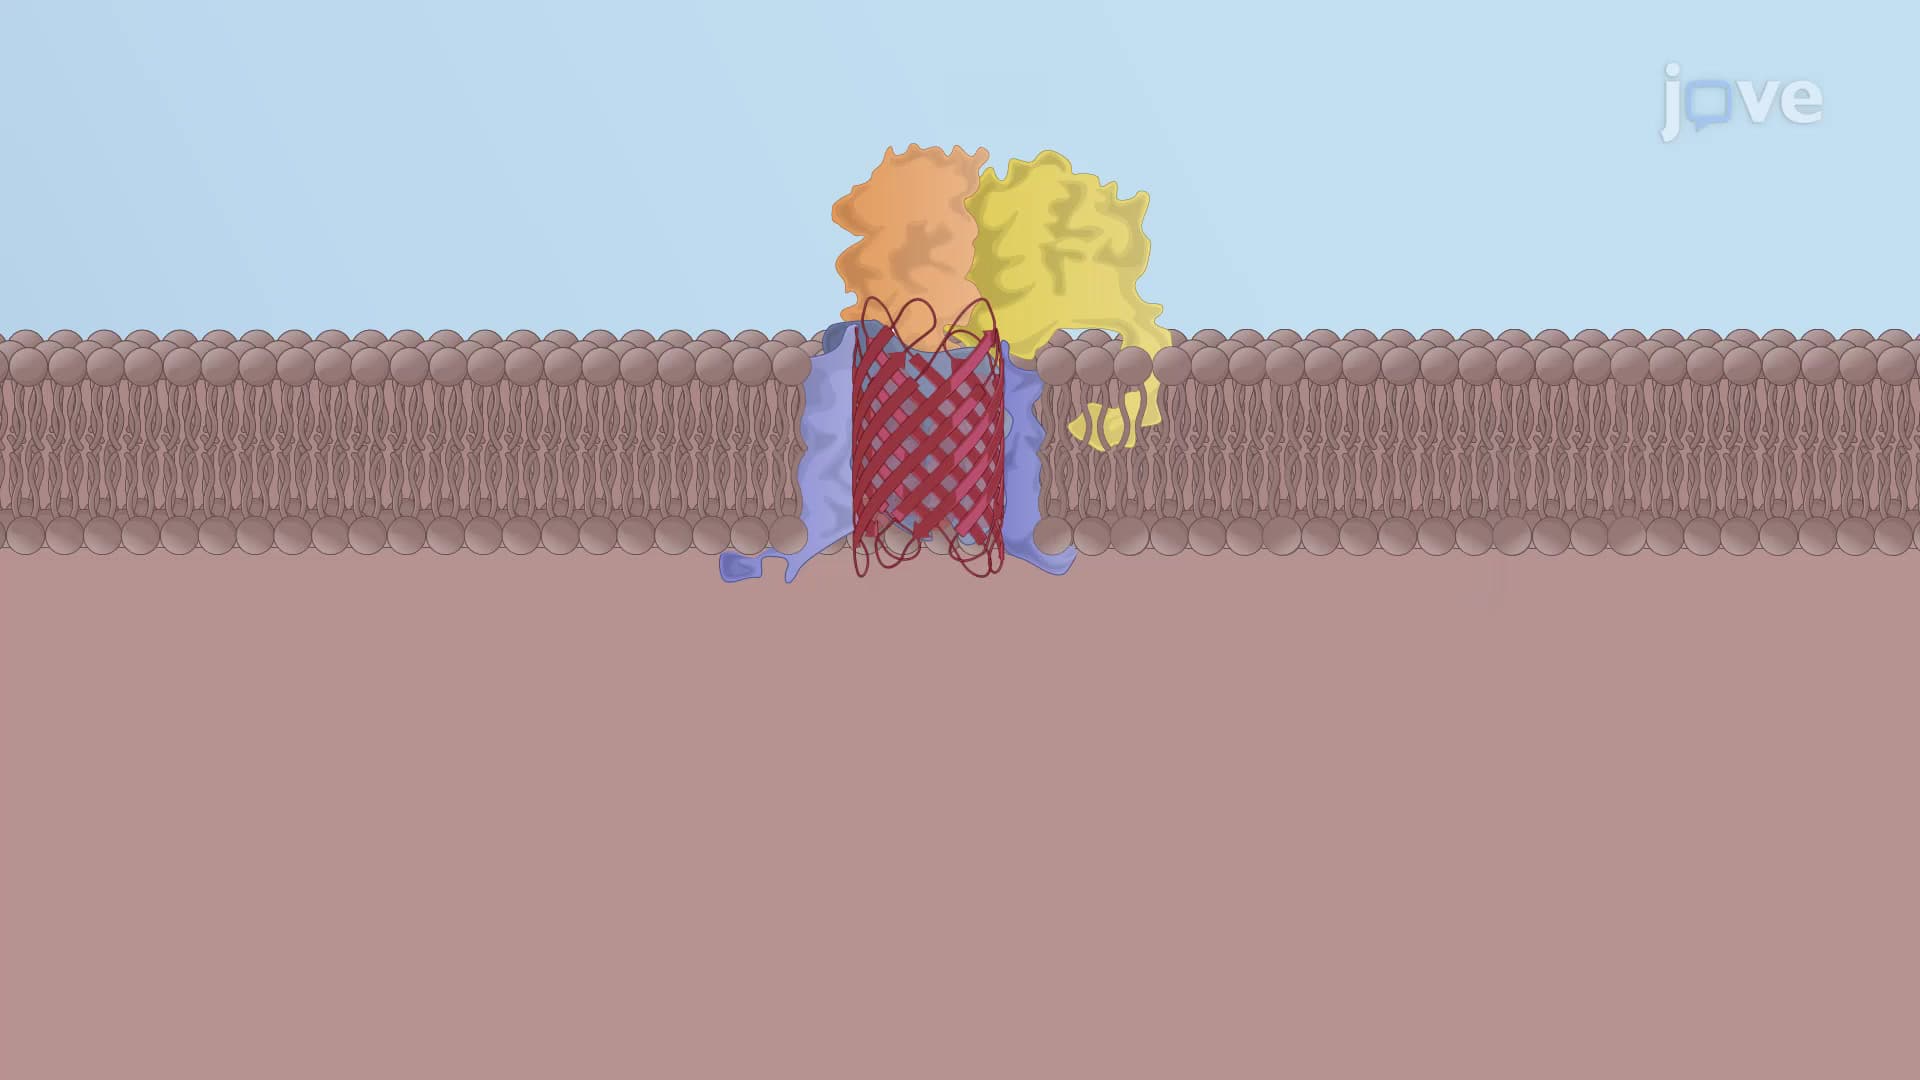
Porin Insertion in the Outer Mitochondrial Membrane
Three models describe the assembly of porins by the SAM complex and their insertion into the outer membrane. Model 1 suggests that porins are assembled outside the SAM channel as the...
 01:39
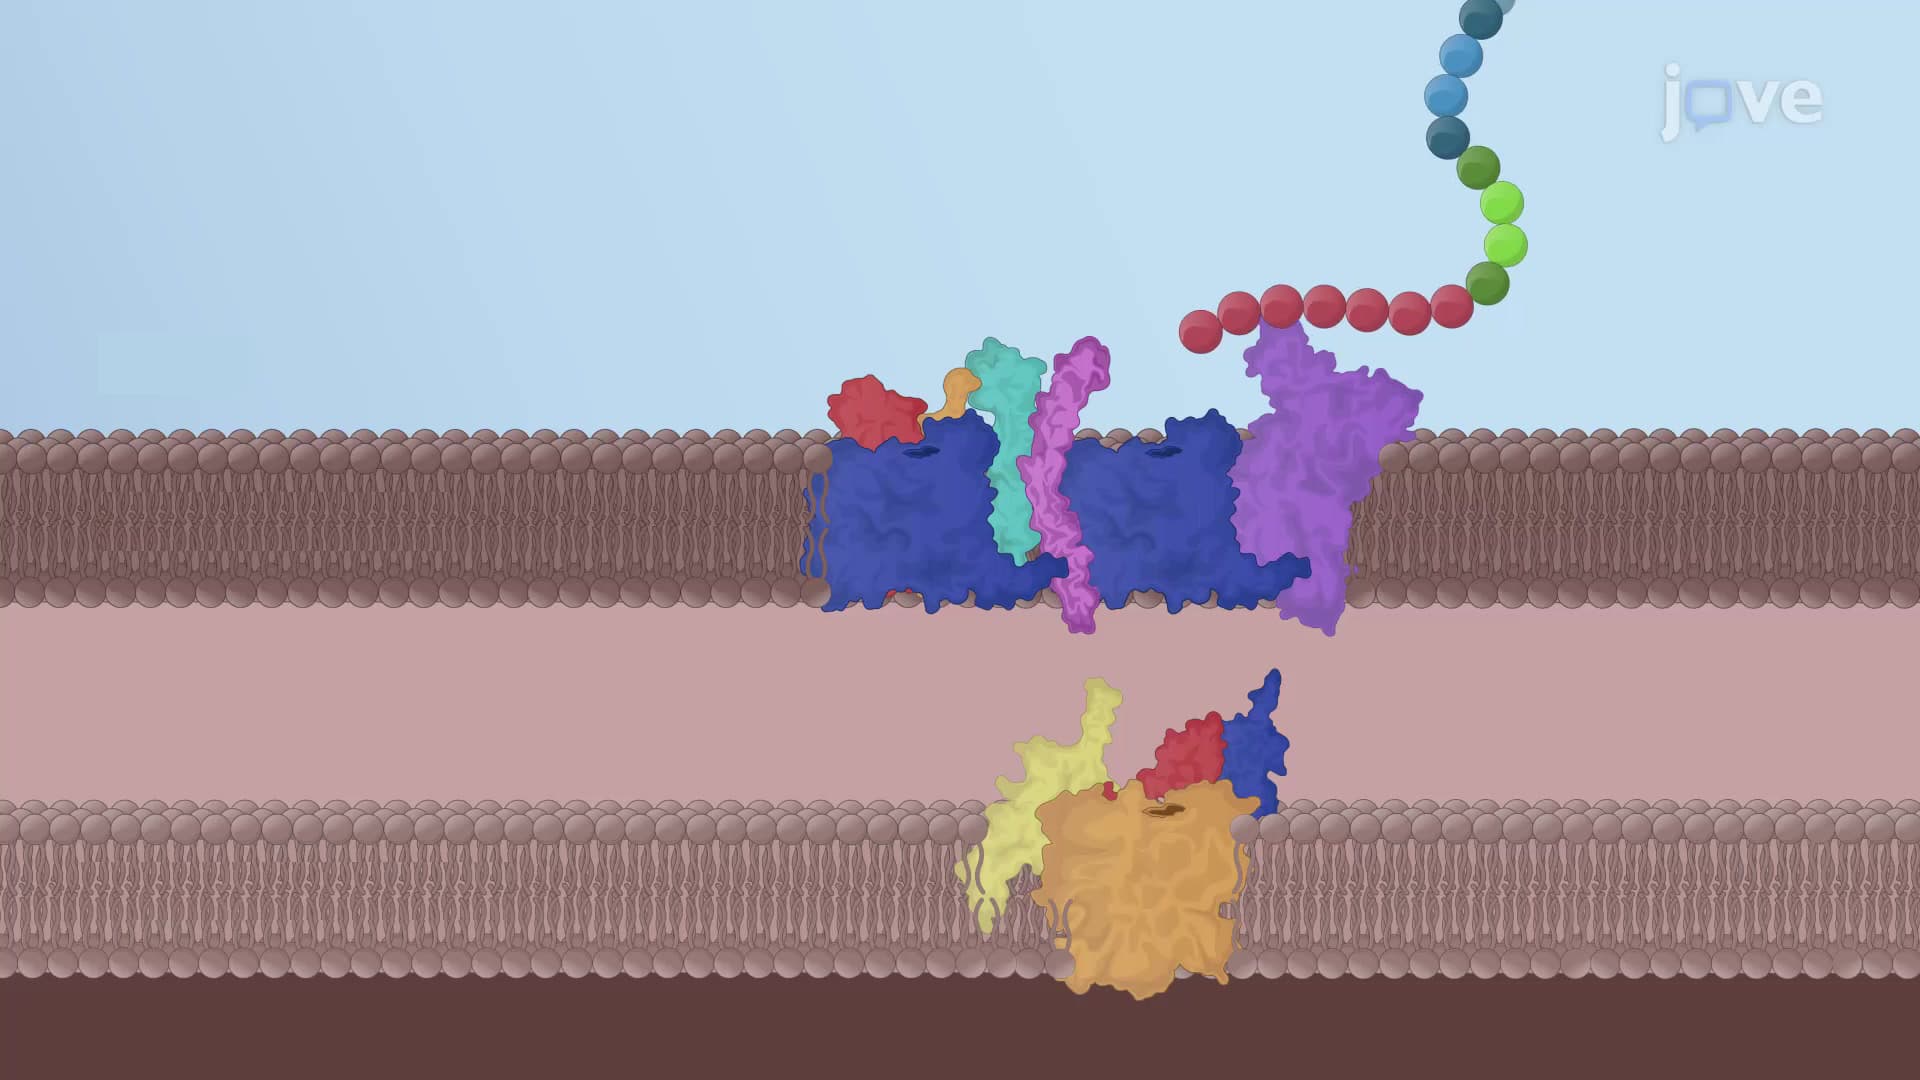
Mitochondrial Precursor Proteins
Most of the mitochondrial precursors...
 01:39
Mitochondrial Protein Sorting
Most of these mitochondrial proteins are encoded by the nucleus and imported to the mitochondria as unfolded or loosely folded precursors. Mitochondrial precursors...
 01:21
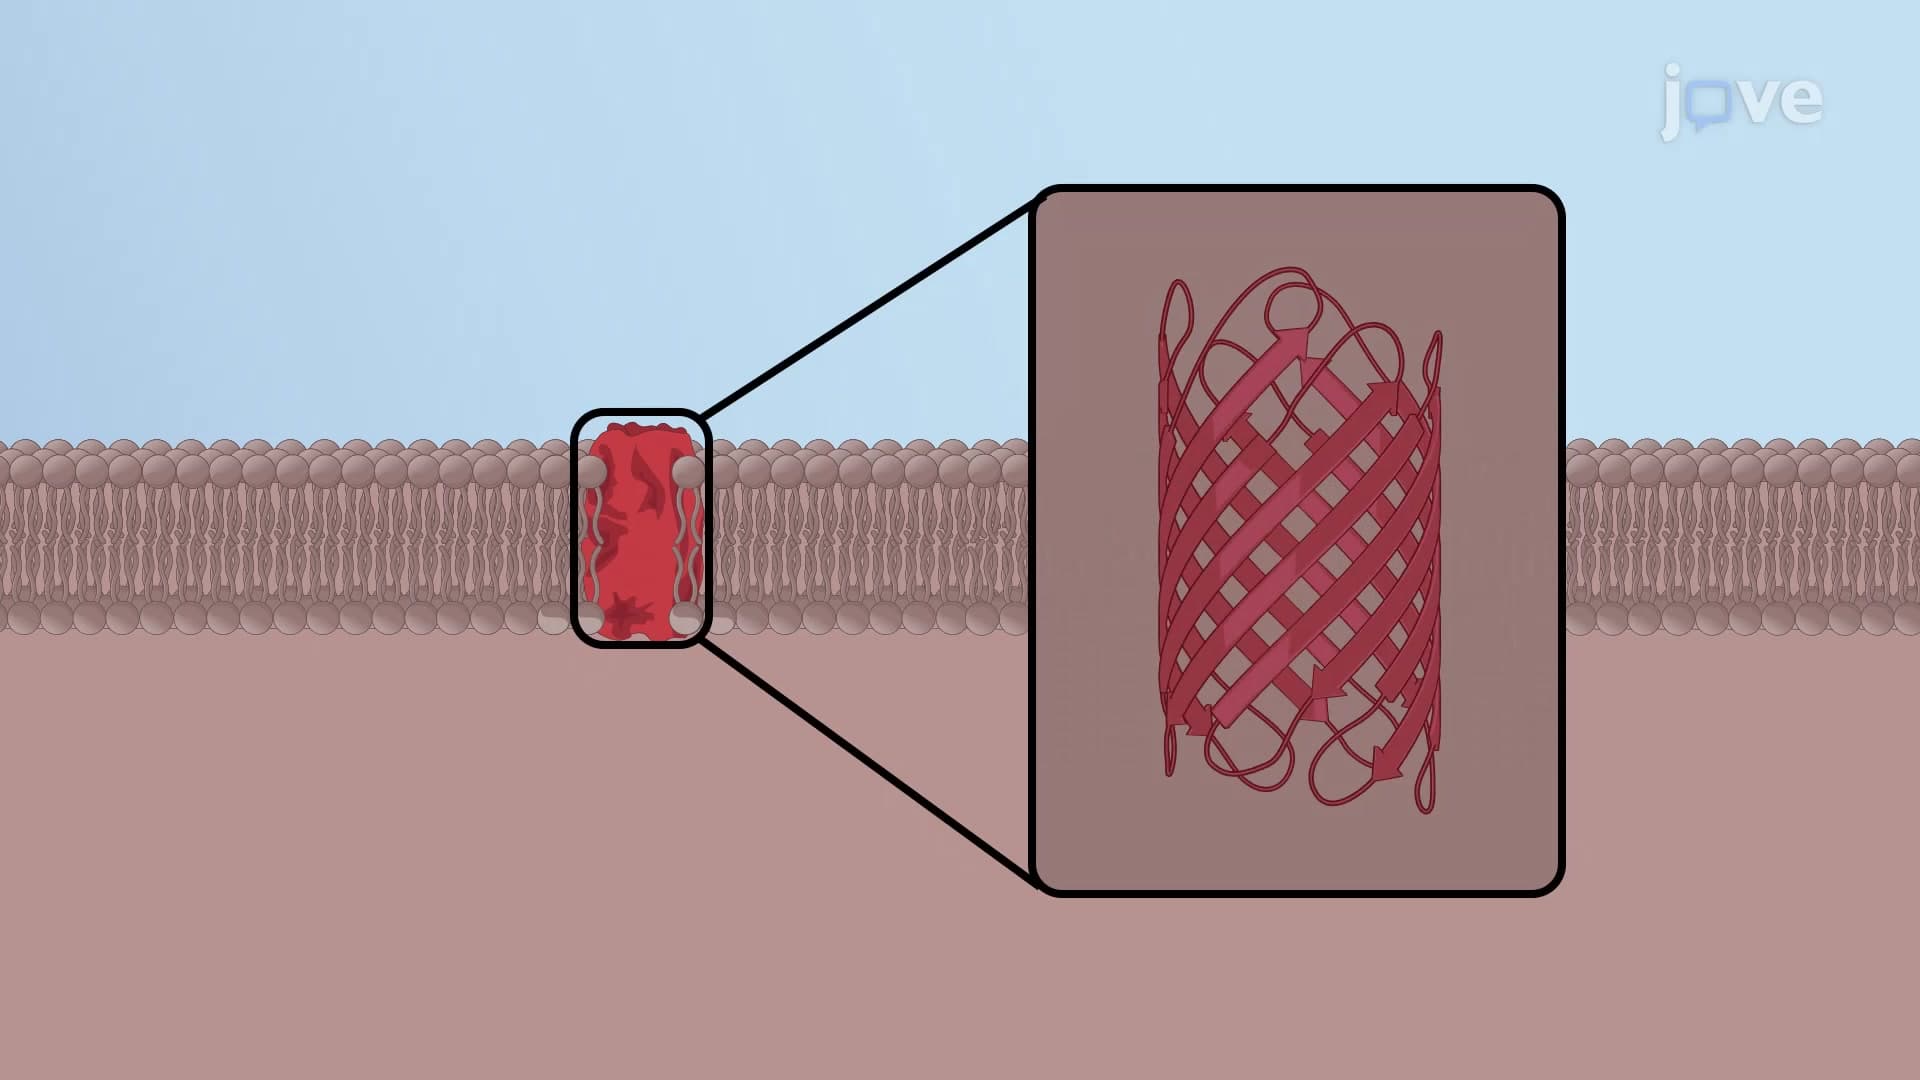
Structure of Porins