
An Improved DFTB/Force Field Model for Molecule-Rare Gas Interactions: Application to 1,4-Diazabicyclo[2.2.2]octane
Paul Guibourg1, Fernand Spiegelman1, Audrey Scognamiglio2
1Laboratoire de Chimie et Physique Quantique (LCPQ/IRSAMC), CNRS, UMR5626, Université de Toulouse, Toulouse31062, France.

Abstract:
It is often challenging to model the evolution of the properties of molecular compounds when they interact with an environment. We will present the combination of the density functional tight binding (DFTB) scheme, describing molecules, with a force field (FF) approach, describing rare gas atoms. We have extended a former model [Iftneret al.J. Chem. Phys.2014,140, 03430110.1063/1.4861431. ] by introducing an atomic-population-dependent correction to a repulsion term between the DFTB atoms and rare gas atoms. In this new model, the dispersion contribution also depends on atomic populations. This model allows, in particular, describing neutral and ionic systems and has been applied to simulate clusters of argon atoms and a 1,4-diazabicyclo[2.2.2]octan (DABCO) molecule. New reference calculations (MP2, CCSD(T)-F12) on small DABCO- Arn=1,40/+ clusters are also reported. We show that the new DFTB-FF model performances are significantly improved with respect to the former model for small aggregates, in particular with regard to cohesive energies. Finally, by coupling the new DFTB-FF model with a minima global search algorithm, namely a combination of Parallel Tempering Monte Carlo explorations and local quenches, we provide a set of stable structures for DABCO- Arn0/+ clusters in the range n = 1-50 atoms pointing out the size evolution of geometric features and energetic properties. Ionization potentials are shown to decrease with size, consistent with new experimental measurements. This decrease is shown to undergo a transition after the completion of the first solvation shell.
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 11:38
In situ FTIR Spectroscopy as a Tool for Investigation of Gas/Solid Interaction: Water-Enhanced CO2 Adsorption in UiO-66 Metal-Organic Framework
Published on: February 1, 2020
Related Concept Videos
 04:01
Real Gases: Effects of Intermolecular Forces and Molecular Volume Deriving Van der Waals Equation
 02:36
Molecular Geometry and Dipole Moments
 01:10
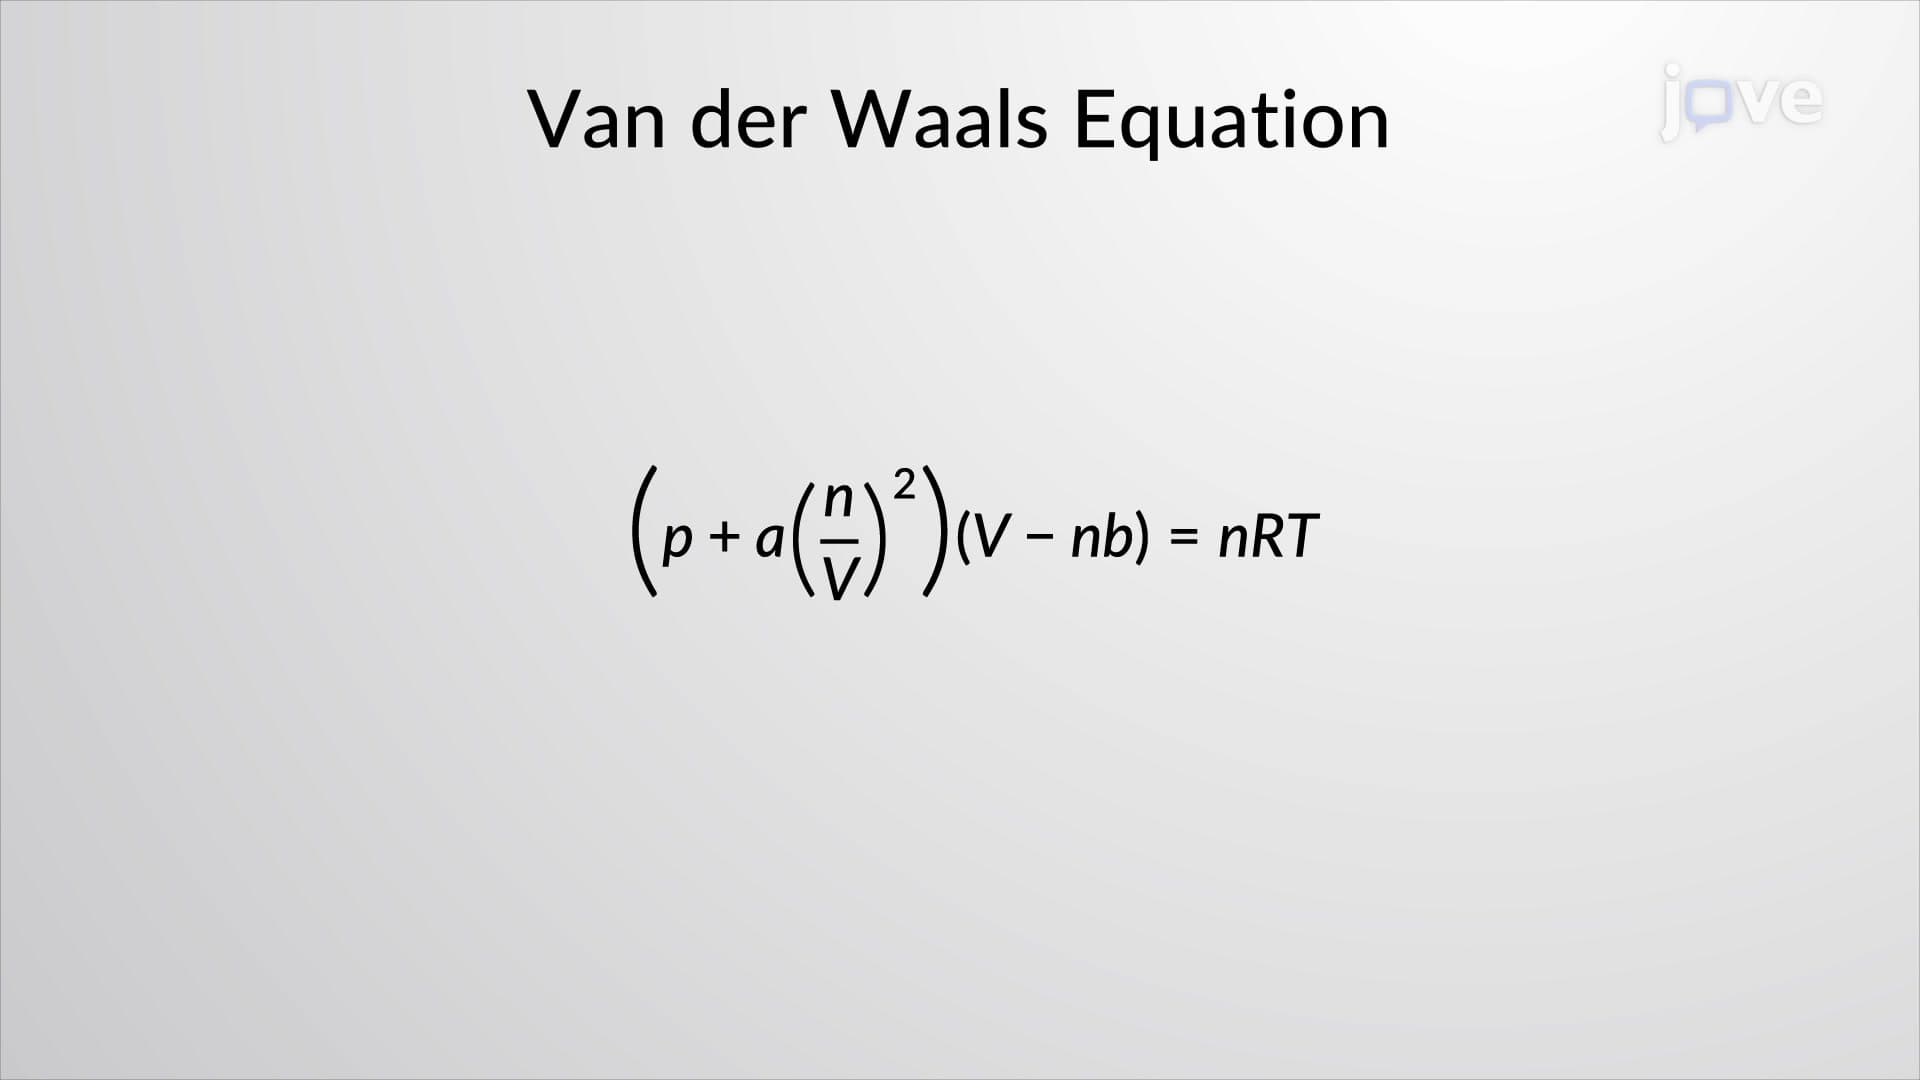
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 03:24
Hybridization of Atomic Orbitals I
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 01:18
Structure of Benzene: Molecular Orbital Model