Related Experiment Video
Updated: Oct 28, 2026
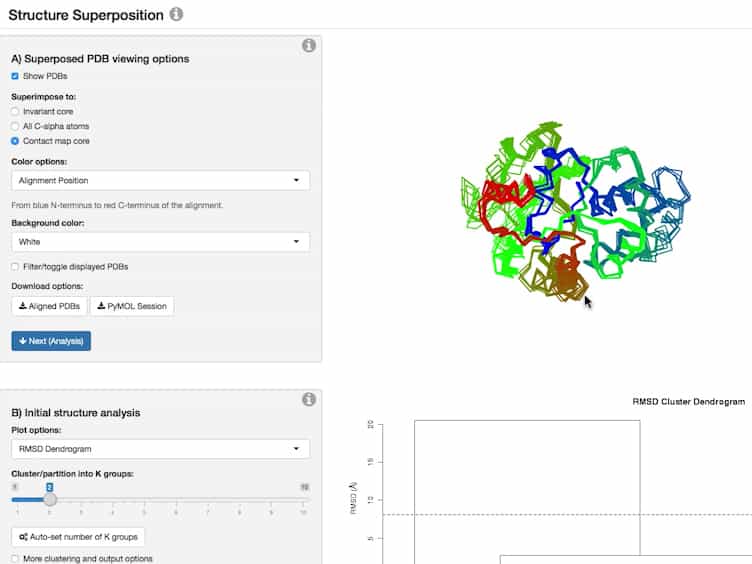
Investigating Protein Sequence-structure-dynamics Relationships with Bio3D-web
Published on: July 16, 2017
Comparative Conformational Sampling of Apo Adenylate Kinase by Enhanced Molecular Dynamics and a Sequence-Based
Qingling Zhao1,2, Ting Cui1,2, Ying Li1,2
1Library of Beijing Union University, Beijing 100101, China.

Abstract:
Conformational sampling remains a major challenge in molecular dynamics (MD) simulations of proteins with large-scale functional motions. Here, we use apo Escherichia coli adenylate kinase (AdK) as a benchmark for the open-closed transition and compare nine sampling protocols under a uniform explicit-solvent budget of 300 ns per method. The methods include unbiased MD, metadynamics, targeted MD (TMD), accelerated MD, simulated tempering, interpolation-based MD, and BioEmu. Closed-state sampling was evaluated relative to a reference-closed basin defined from ligand-stripped PDB 1AKE. TMD was the only method that both reached and maintained the closed reference, whereas unbiased MD did not enter this basin within 300 ns. Metadynamics increased sampling along the domain-distance coordinate but also promoted excessive LID opening, while simulated tempering showed a similar tendency. BioEmu generated closed-like conformations, but these states were not maintained during subsequent short MD simulations. Overall, the results show that collective-variable coverage does not necessarily correspond to stable target state occupancy and that different sampling methods provide different types of conformational information. These findings highlight the importance of selecting sampling strategies according to the specific scientific objective.